
A generic force field for simulating native protein structures using dissipative particle dynamics
Rakesh Vaiwala1, K Ganapathy Ayappa1,2
1Department of Chemical Engineering, Indian Institute of Science, Bangalore 560012, India. ayappa@iisc.ac.in.

Abstract:
A coarse-grained force field for molecular dynamics simulations of the native structures of proteins in a dissipative particle dynamics (DPD) framework is developed. The parameters for bonded interactions are derived by mapping the bonds and angles for 20 amino acids onto target distributions obtained from fully atomistic simulations in explicit solvent. A dual-basin potential is introduced for stabilizing backbone angles, to cover a wide spectrum of protein secondary structures. The backbone dihedral potential enables folding of the protein from an unfolded initial state to the folded native structure. The proposed force field is validated by evaluating the structural properties of several model peptides and proteins including the SARS-CoV-2 fusion peptide, consisting of α-helices, β-sheets, loops and turns. Detailed comparisons with fully atomistic simulations are carried out to assess the ability of the proposed force field to stabilize the different secondary structures present in proteins. The compact conformations of the native states were evident from the radius of gyration and the high intensity peaks of the root mean square deviation histograms, which were found to be within 0.4 nm. The Ramachandran-like energy landscape on the phase space of backbone angles (θ) and dihedrals (ϕ) effectively captured the conformational phase space of α-helices at ∼(ϕ = 50°,θ = 90°) and β-strands at ∼(ϕ = ±180°,θ = 90-120°). Furthermore, the residue-residue native contacts were also well reproduced by the proposed DPD model. The applicability of the model to multidomain complexes was assessed using lysozyme and a large α-helical bacterial pore-forming toxin, cytolysin A. Our study illustrates that the proposed force field is generic, and can potentially be extended for efficient in silico investigations of membrane bound polypeptides and proteins using DPD simulations.
More Related Videos
 09:17
Structure-Based Simulation and Sampling of Transcription Factor Protein Movements along DNA from Atomic-Scale Stepping to Coarse-Grained Diffusion
Published on: March 1, 2022
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Related Concept Videos
 01:13
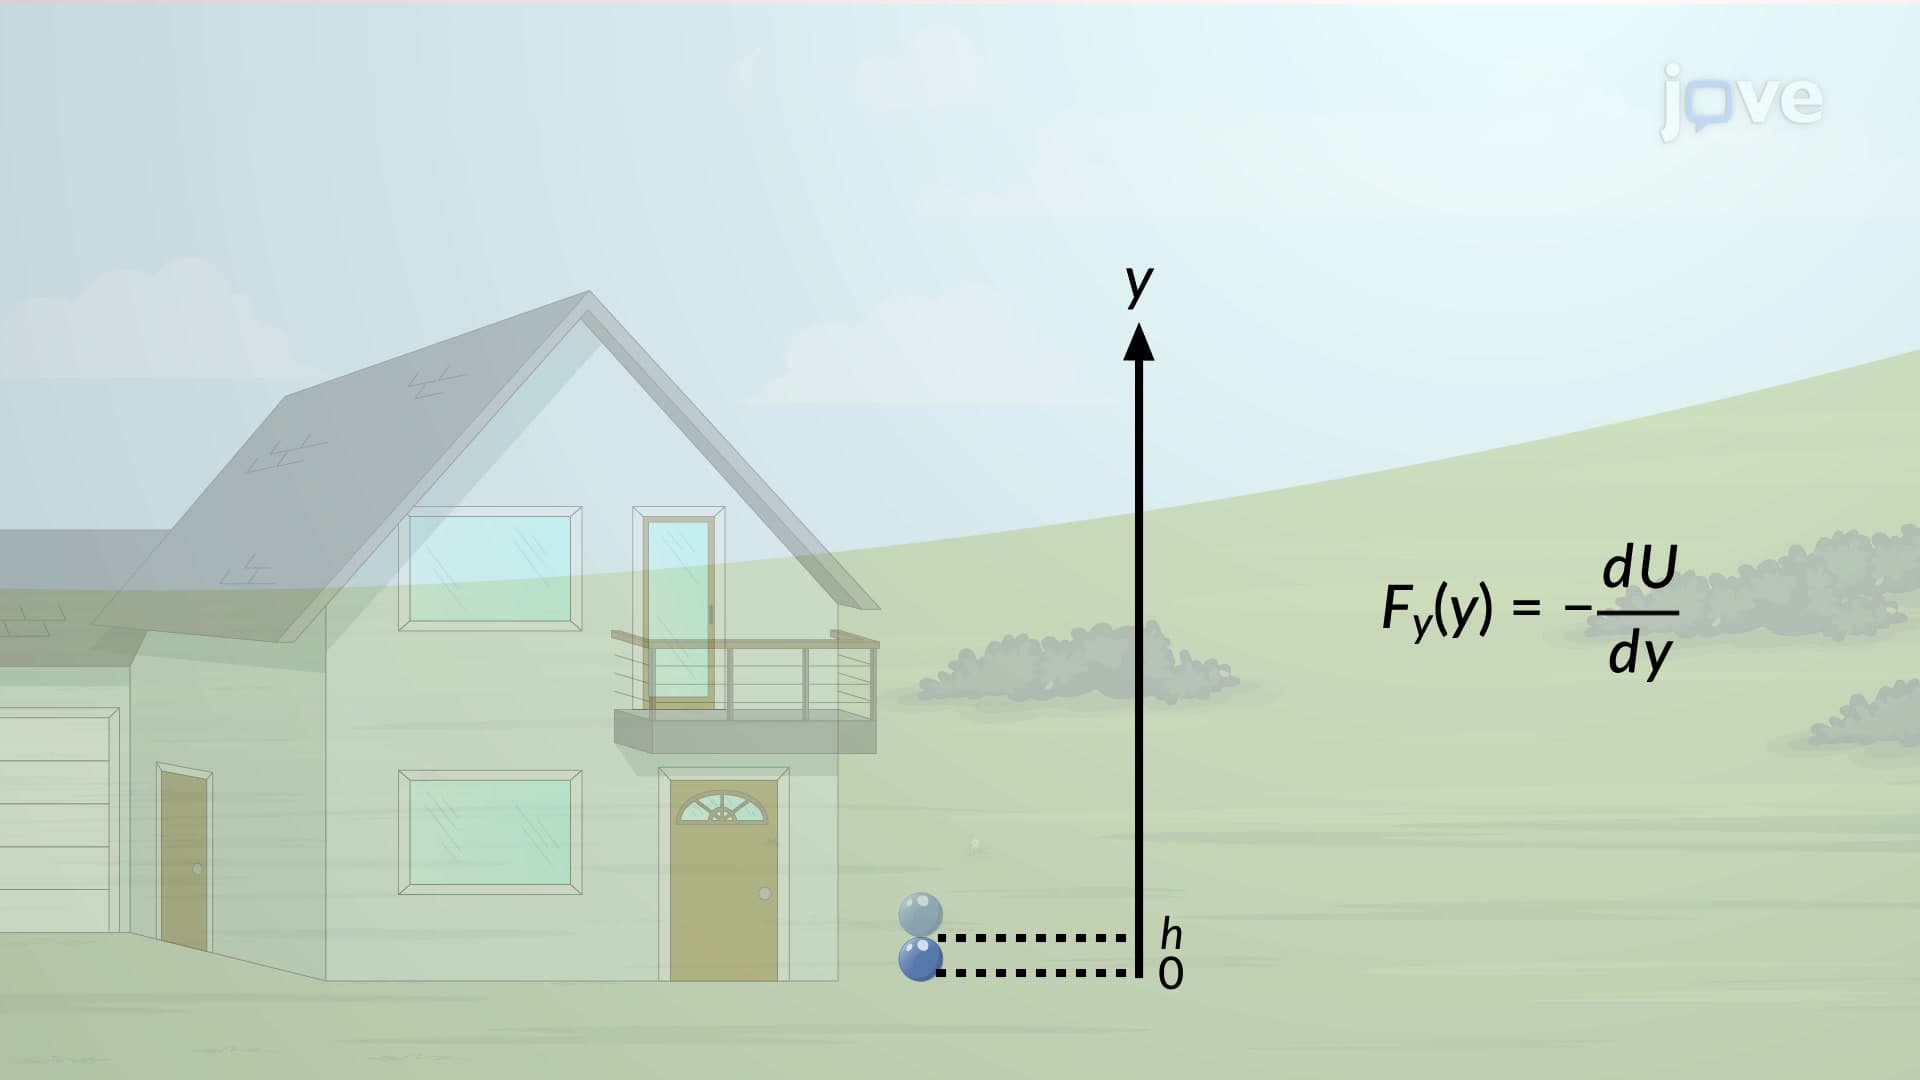
Force and Potential Energy in One Dimension
 01:19
Protein Dynamics in Living Cells
Fluorescent recovery after photobleaching (FRAP) is a fluorescent-protein-based detection technique used to quantify protein movement rates within the cell. This method exposes a small portion of the cell to an intense laser beam. The laser beam causes permanent photobleaching of the fluorophore-tagged proteins in the exposed region. As the bleached...
 01:30
Three-Dimensional Force System