An accurate single descriptor for ion-π interactions
Zhangyun Liu1, Zheng Chen1, Jinyang Xi1
1Collaborative Innovation Center of Chemistry for Energy Materials, Key Laboratory of Molecular Catalysis and Innovative Materials, MOE Key Laboratory of Computational Physical Sciences, Department of Chemistry, Fudan University, Shanghai 200433, China.

Abstract:
Non-covalent interactions between ions and π systems play an important role in molecular recognition, catalysis and biology. To guide the screen and design for artificial hosts, catalysts and drug delivery, understanding the physical nature of ion-π complexes via descriptors is indispensable. However, even with multiple descriptors that contain the leading term of electrostatic and polarized interactions, the quantitative description for the binding energies (BEs) of ion-π complexes is still lacking because of the intrinsic shortcomings of the commonly used descriptors. Here, we have shown that the impartment of orbital details into the electrostatic energy (coined as OEE) makes an excellent single descriptor for BEs of not only spherical, but also multiply-shaped, ion-π systems, highlighting the importance of an accurate description of the electrostatic interactions. Our results have further demonstrated that OEEs from a low-level method could be calibrated to BEs from a high-level method, offering a powerful practical strategy for an accurate prediction of a set of ion-π interactions.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 06:53
Photoelectron Imaging of Anions Illustrated by 310 Nm Detachment of F−
Published on: July 27, 2018
Related Concept Videos
 01:27
π Electron Effects on Chemical Shift: Overview
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:18
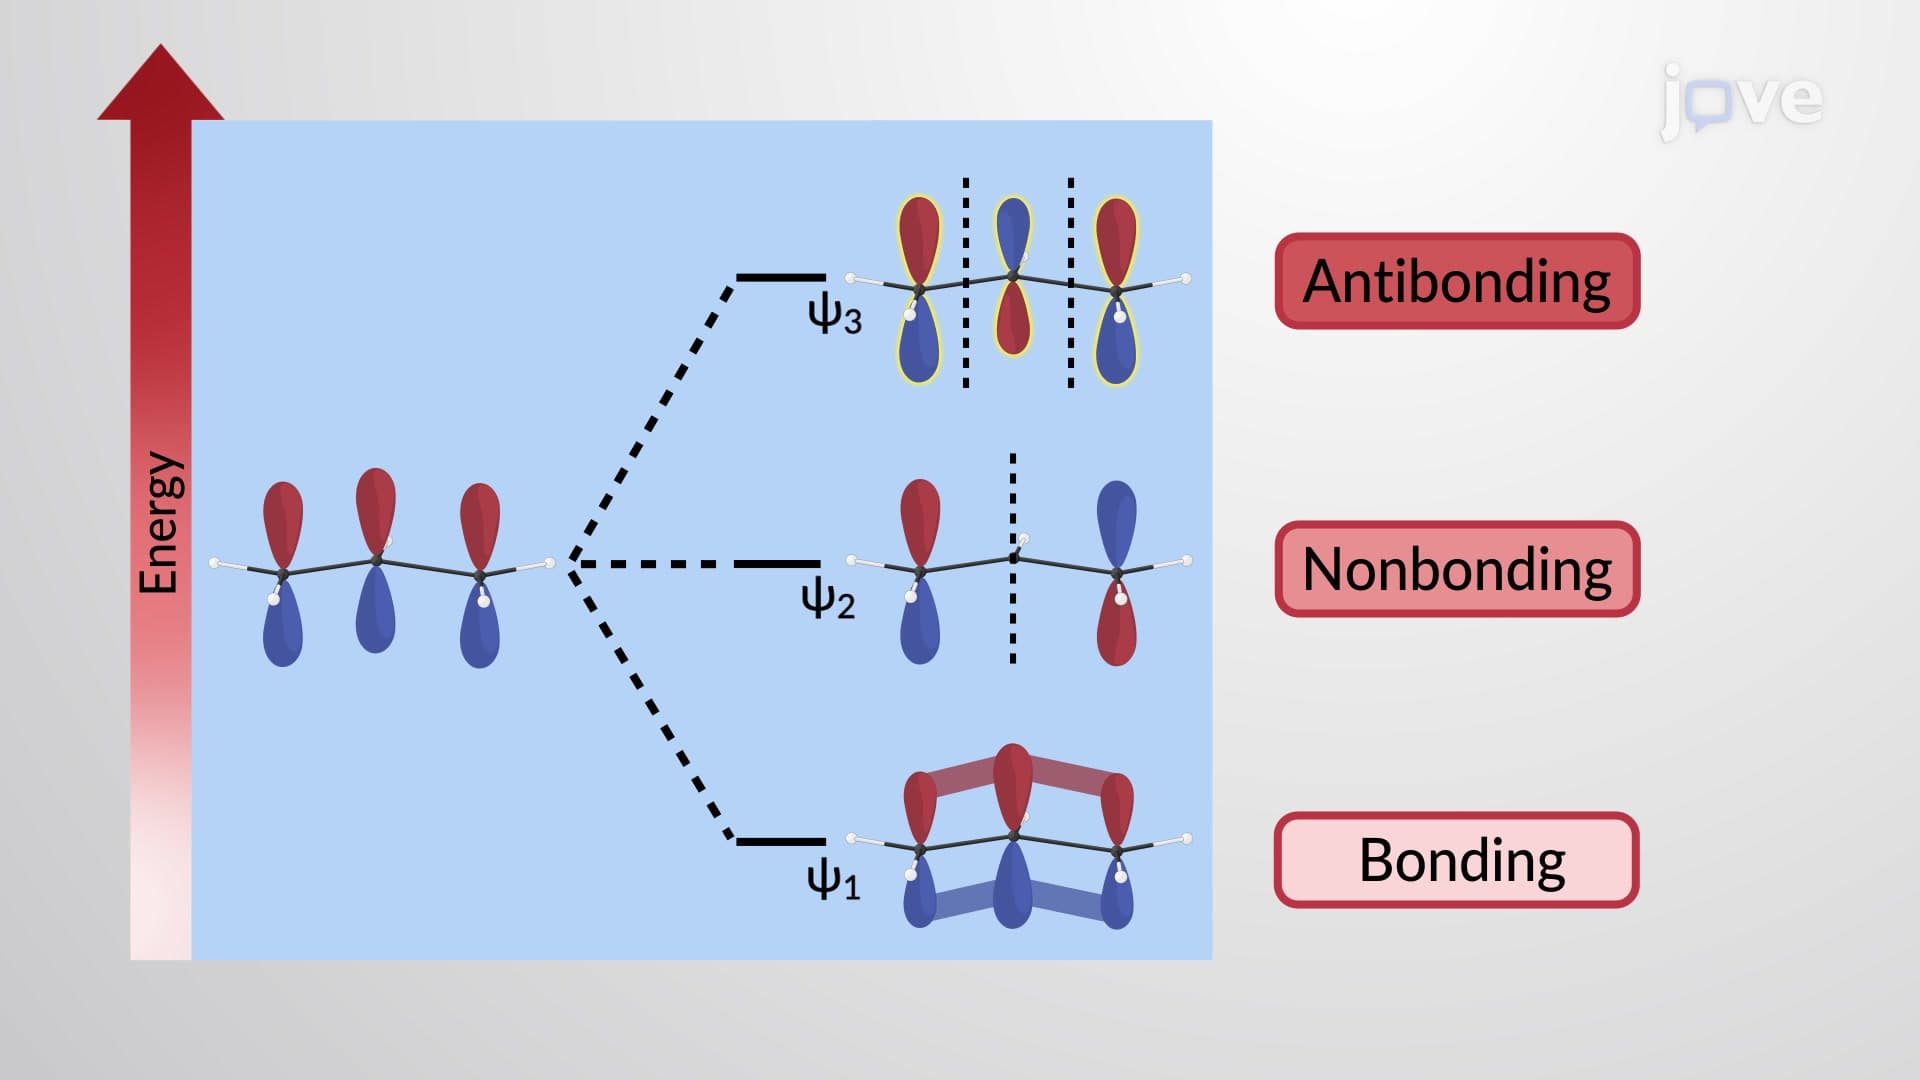
π Molecular Orbitals of the Allyl Cation and Anion
 03:10
Ionic Radii
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:35
Molecular Orbital Theory I