Related Experiment Video
Updated: Oct 13, 2025


Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Improved electronic structure prediction of chalcopyrite semiconductors from a semilocal density functional based on
Arghya Ghosh1, Subrata Jana2,3, Manish K Niranjan1
1Department of Physics, Indian Institute of Technology, Hyderabad, India.

Abstract:
The correct treatment ofdelectrons is of prime importance in order to predict the electronic properties of the prototype chalcopyrite semiconductors. The effect ofdstates is linked with the anion displacement parameteru, which in turn influences the bandgap of these systems. Semilocal exchange-correlation functionals which yield good structural properties of semiconductors and insulators often fail to predict reasonableubecause of the underestimation of the bandgaps arising from the strong interplay betweendelectrons. In the present study, we show that the meta-generalized gradient approximation (meta-GGA) obtained from the cuspless hydrogen density (MGGAC) (2019Phys. Rev.B 100 155140) performs in an improved manner in apprehending the key features of the electronic properties of chalcopyrites, and its bandgaps are comparative to that obtained using state-of-art hybrid methods. Moreover, the present assessment also shows the importance of the Pauli kinetic energy enhancement factor,α= (τ-τ)/τunifin describing thedelectrons in chalcopyrites. The present study strongly suggests that the MGGAC functional within semilocal approximations can be a better and preferred choice to study the chalcopyrites and other solid-state systems due to its superior performance and significantly low computational cost.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 07:24
Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:12
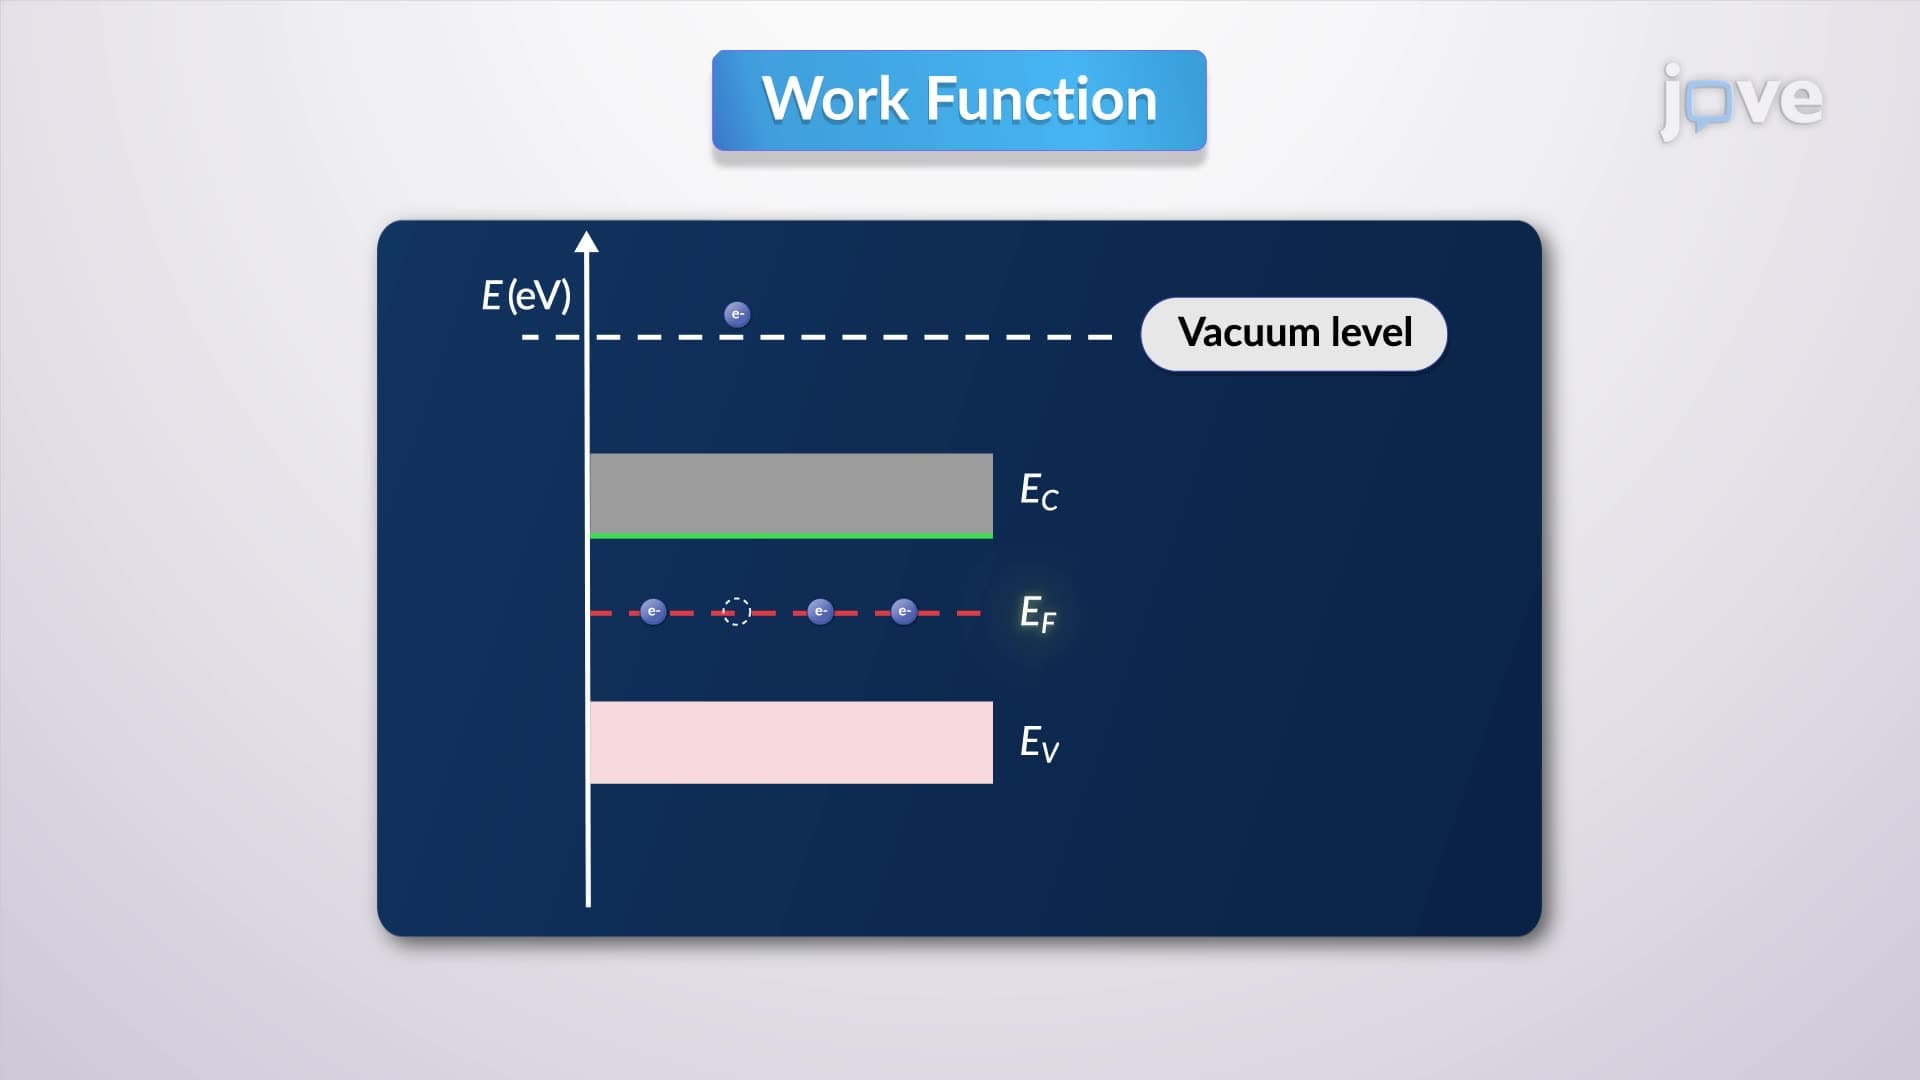
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 01:27
π Electron Effects on Chemical Shift: Overview
 02:27
Predicting Molecular Geometry
 03:06
The Pauli Exclusion Principle