Related Experiment Video
Updated: Oct 12, 2025
Functional Reconstitution and Channel Activity Measurements of Purified Wildtype and Mutant CFTR Protein
Published on: March 9, 2015
CFTR Protein: Not Just a Chloride Channel?
Laurence S Hanssens1, Jean Duchateau2, Georges J Casimir1,2
1Department of Pediatric Pulmonology and Cystic Fibrosis Clinic, Hôpital Universitaire des Enfants Reine Fabiola, Université Libre de Bruxelles (ULB), Avenue J.J. Crocq 15, 1020 Brussels, Belgium.

Abstract:
Cystic fibrosis (CF) is a recessive genetic disease caused by mutations in a gene encoding a protein called Cystic Fibrosis Transmembrane Conductance Regulator (CFTR). The CFTR protein is known to acts as a chloride (Cl-) channel expressed in the exocrine glands of several body systems where it also regulates other ion channels, including the epithelial sodium (Na+) channel (ENaC) that plays a key role in salt absorption. This function is crucial to the osmotic balance of the mucus and its viscosity. However, the pathophysiology of CF is more challenging than a mere dysregulation of epithelial ion transport, mainly resulting in impaired mucociliary clearance (MCC) with consecutive bronchiectasis and in exocrine pancreatic insufficiency. This review shows that the CFTR protein is not just a chloride channel. For a long time, research in CF has focused on abnormal Cl- and Na+ transport. Yet, the CFTR protein also regulates numerous other pathways, such as the transport of HCO3-, glutathione and thiocyanate, immune cells, and the metabolism of lipids. It influences the pH homeostasis of airway surface liquid and thus the MCC as well as innate immunity leading to chronic infection and inflammation, all of which are considered as key pathophysiological characteristics of CF.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:20
Cotranslational Protein Translocation
Sec61 channel partners for cotranslational translocation
During cotranslational translocation, the Sec61 channel partners with the signal recognition particle (SRP), the signal recognition particle receptor (SR), and the ribosomes to transport the nascent polypeptide chain...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:29
Insertion of Multi-pass Transmembrane Proteins in the RER
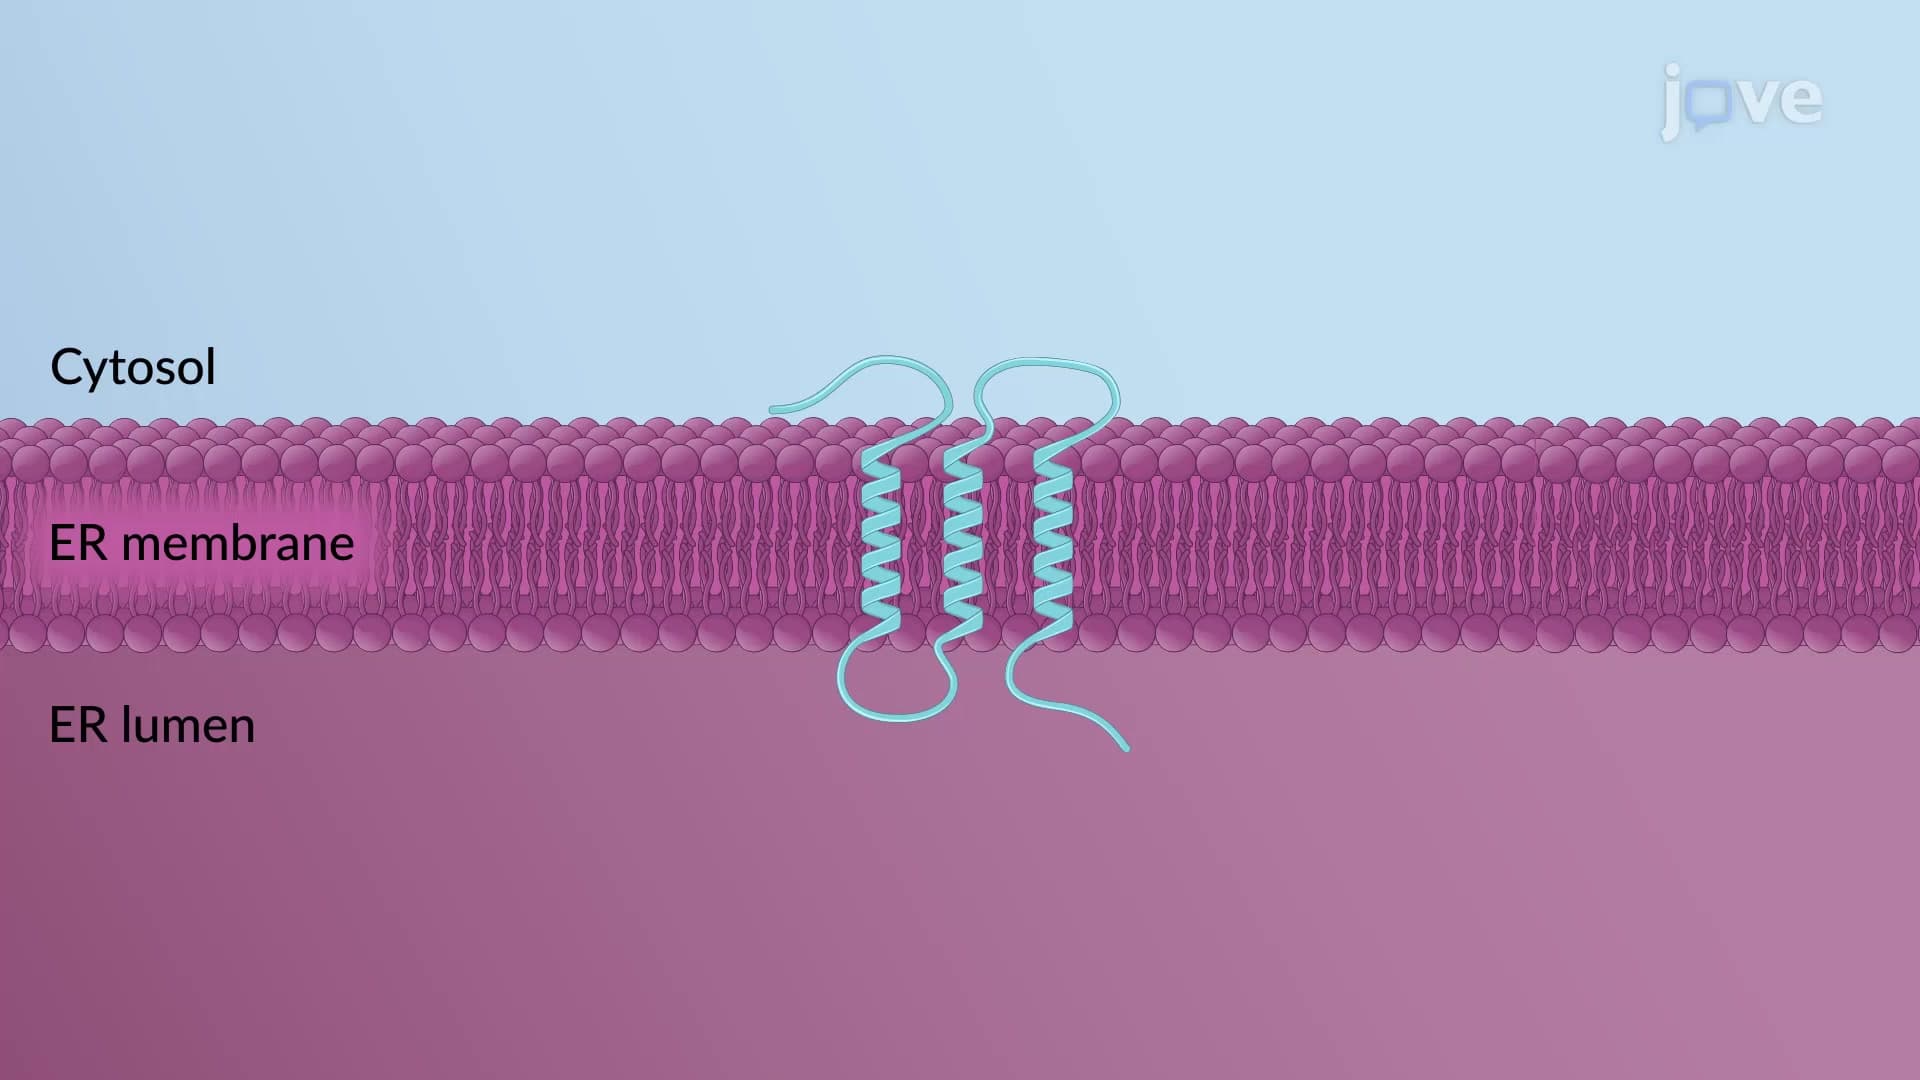
The multipass transmembrane proteins are the type IV integral membrane proteins with multiple topogenic sequences determining their spatial arrangement in the ER membrane. Nearly all multipass proteins lack a cleavable signal sequence and use...
 01:28
Protein Translocation Machinery on the ER Membrane
Sec61 protein conducting channel
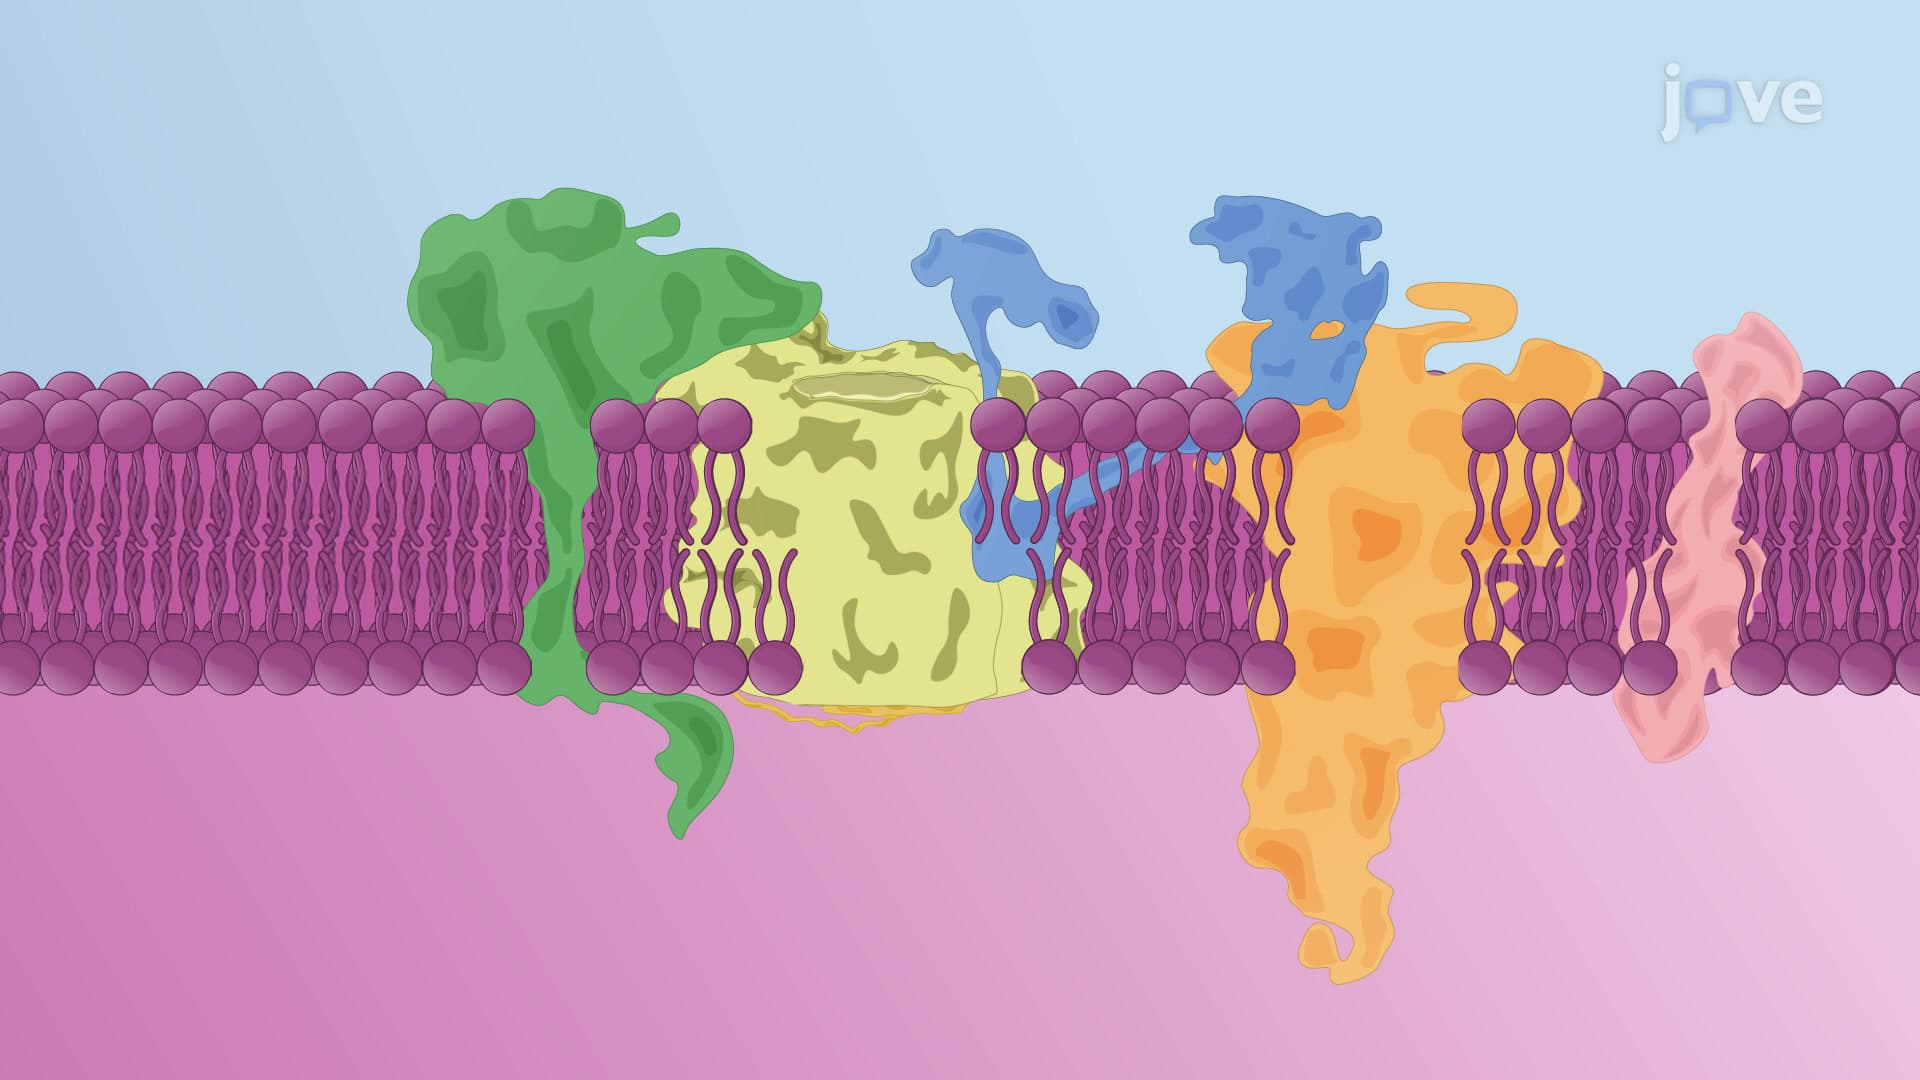
In eukaryotes, the translocon complex comprises a core heterotrimeric translocator channel called the Sec61 complex. This channel includes three transmembrane proteins, Sec61α, Sec61β, and Sec61γ, and is the largest subunit of the...
 01:27
Post-translational Translocation of Proteins to the RER
Targeting proteins to the ER
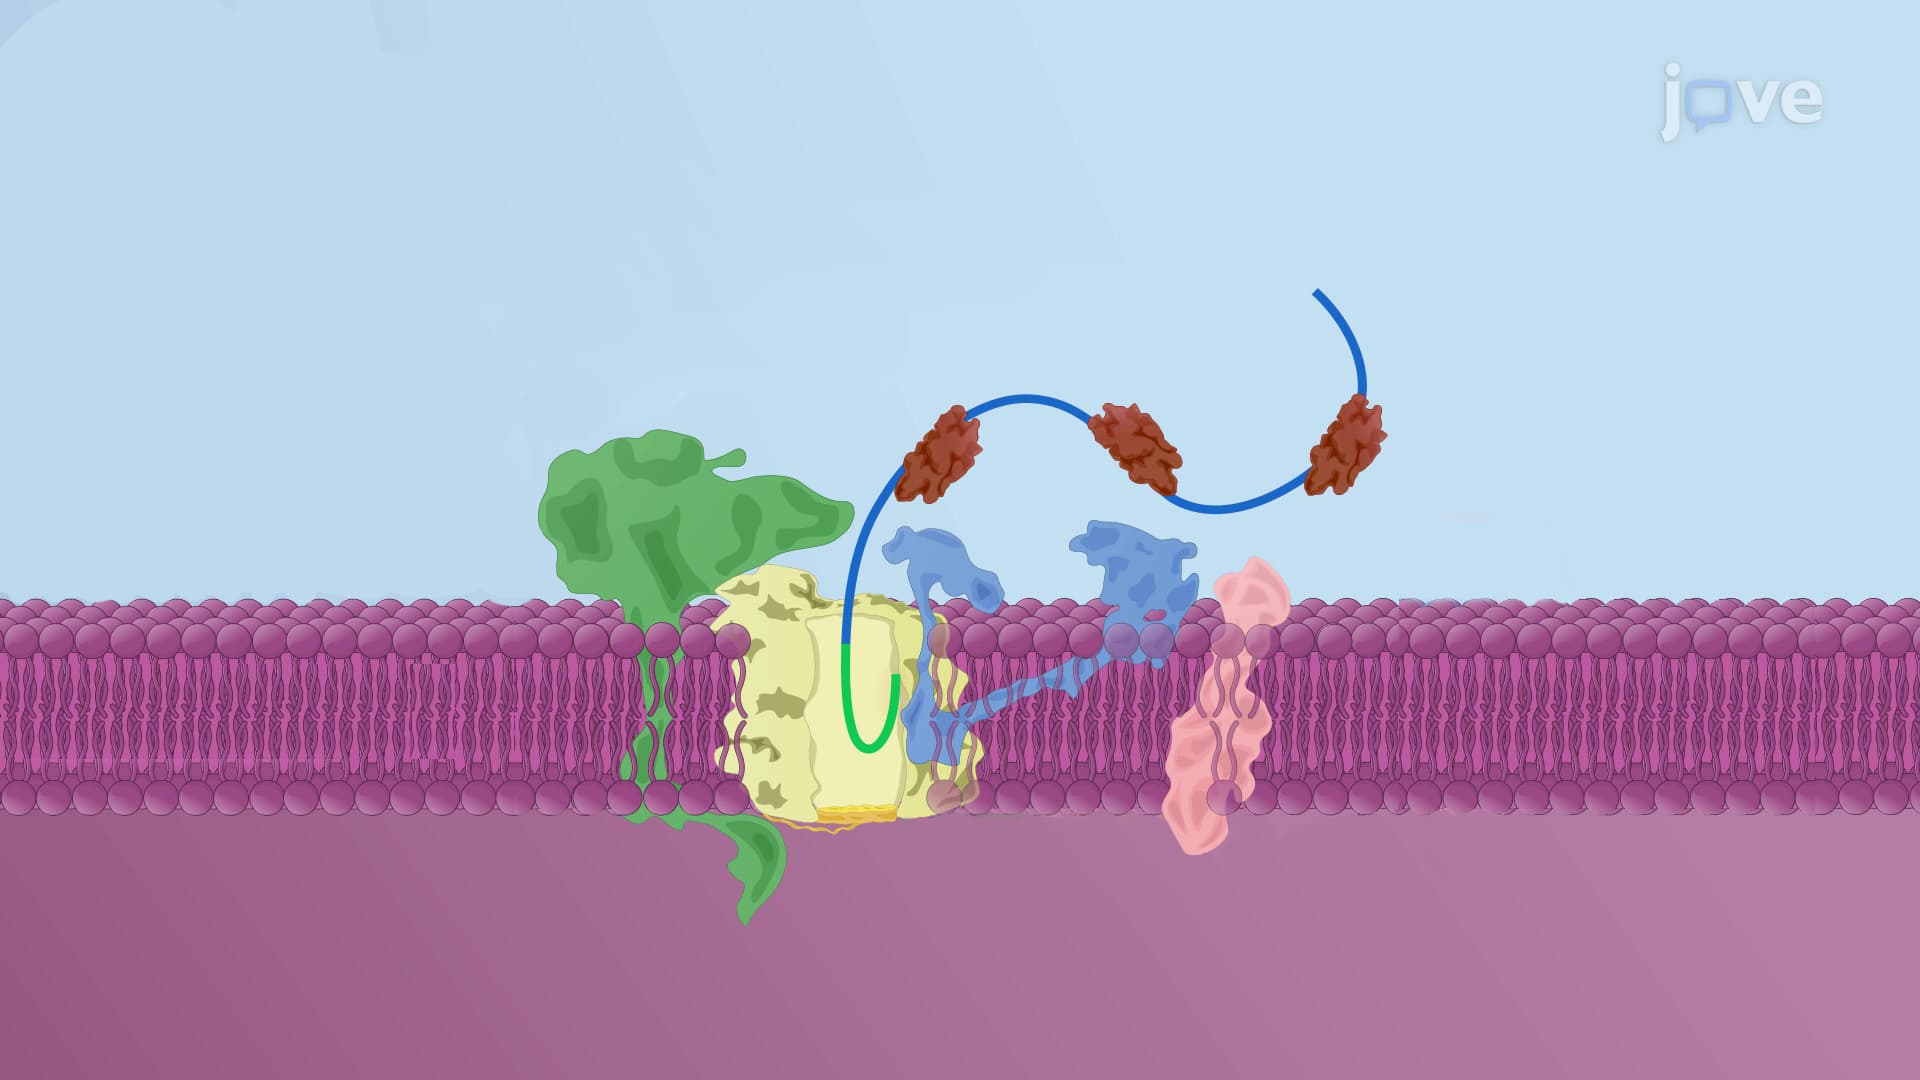
Hsp40 and Hsp70 chaperone molecules bind the translated proteins in the cytosol to prevent their folding. The chaperone binding helps to keep the signal...