Related Experiment Video
Updated: Oct 8, 2025
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
Generating novel molecule for target protein (SARS-CoV-2) using drug-target interaction based on graph neural network
Amit Ranjan1, Shivansh Shukla1, Deepanjan Datta1
1Department of Computer Science and Engineering, Indian Institute of Technology Patna, Patna, 801103 India.

Abstract:
The transmittable spread of viral coronavirus (SARS-CoV-2) has resulted in a significant rise in global mortality. Due to lack of effective treatment, our aim is to generate a highly potent active molecule that can bind with the protein structure of SARS-CoV-2. Different machine learning and deep learning approaches have been proposed for molecule generation; however, most of these approaches represent the drug molecule and protein structure in 1D sequence, ignoring the fact that molecules are by nature in 3D structure, and because of this many critical properties are lost. In this work, a framework is proposed that takes account of both tertiary and sequential representations of molecules and proteins using Gated Graph Neural Network (GGNN), Knowledge graph, and Early Fusion approach. The generated molecules from GGNN are screened using Knowledge Graph to reduce the search space by discarding the non-binding molecules before being fed into the Early Fusion model. Further, the binding affinity score of the generated molecule is predicted using the early fusion approach. Experimental result shows that our framework generates valid and unique molecules with high accuracy while preserving the chemical properties. The use of a knowledge graph claims that the entire generated dataset of molecules was reduced by roughly 96% while retaining more than 85% of good binding desirable molecules and the rejection of more than 99% of fruitless molecules. Additionally, the framework was tested with two of the SARS-CoV-2 viral proteins: RNA-dependent-RNA polymerase (RdRp) and 3C-like protease (3CLpro).
More Related Videos
 08:31
Biosensor-based High Throughput Biopanning and Bioinformatics Analysis Strategy for the Global Validation of Drug-protein Interactions
Published on: December 1, 2020
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
 01:26
Drug Discovery: Overview
 02:04
Protein-protein Interfaces
 01:26
Targets for Drug Action: Overview
Receptors are either membrane-spanning or intracellular proteins, which upon binding a ligand, get activated and transmit the signal downstream to elicit a response. Drugs bind receptors, either mimicking the action of endogenous ligands or blocking the receptor activity to bring about a modified response. Nearly 35% of approved drugs target the G...
 01:15
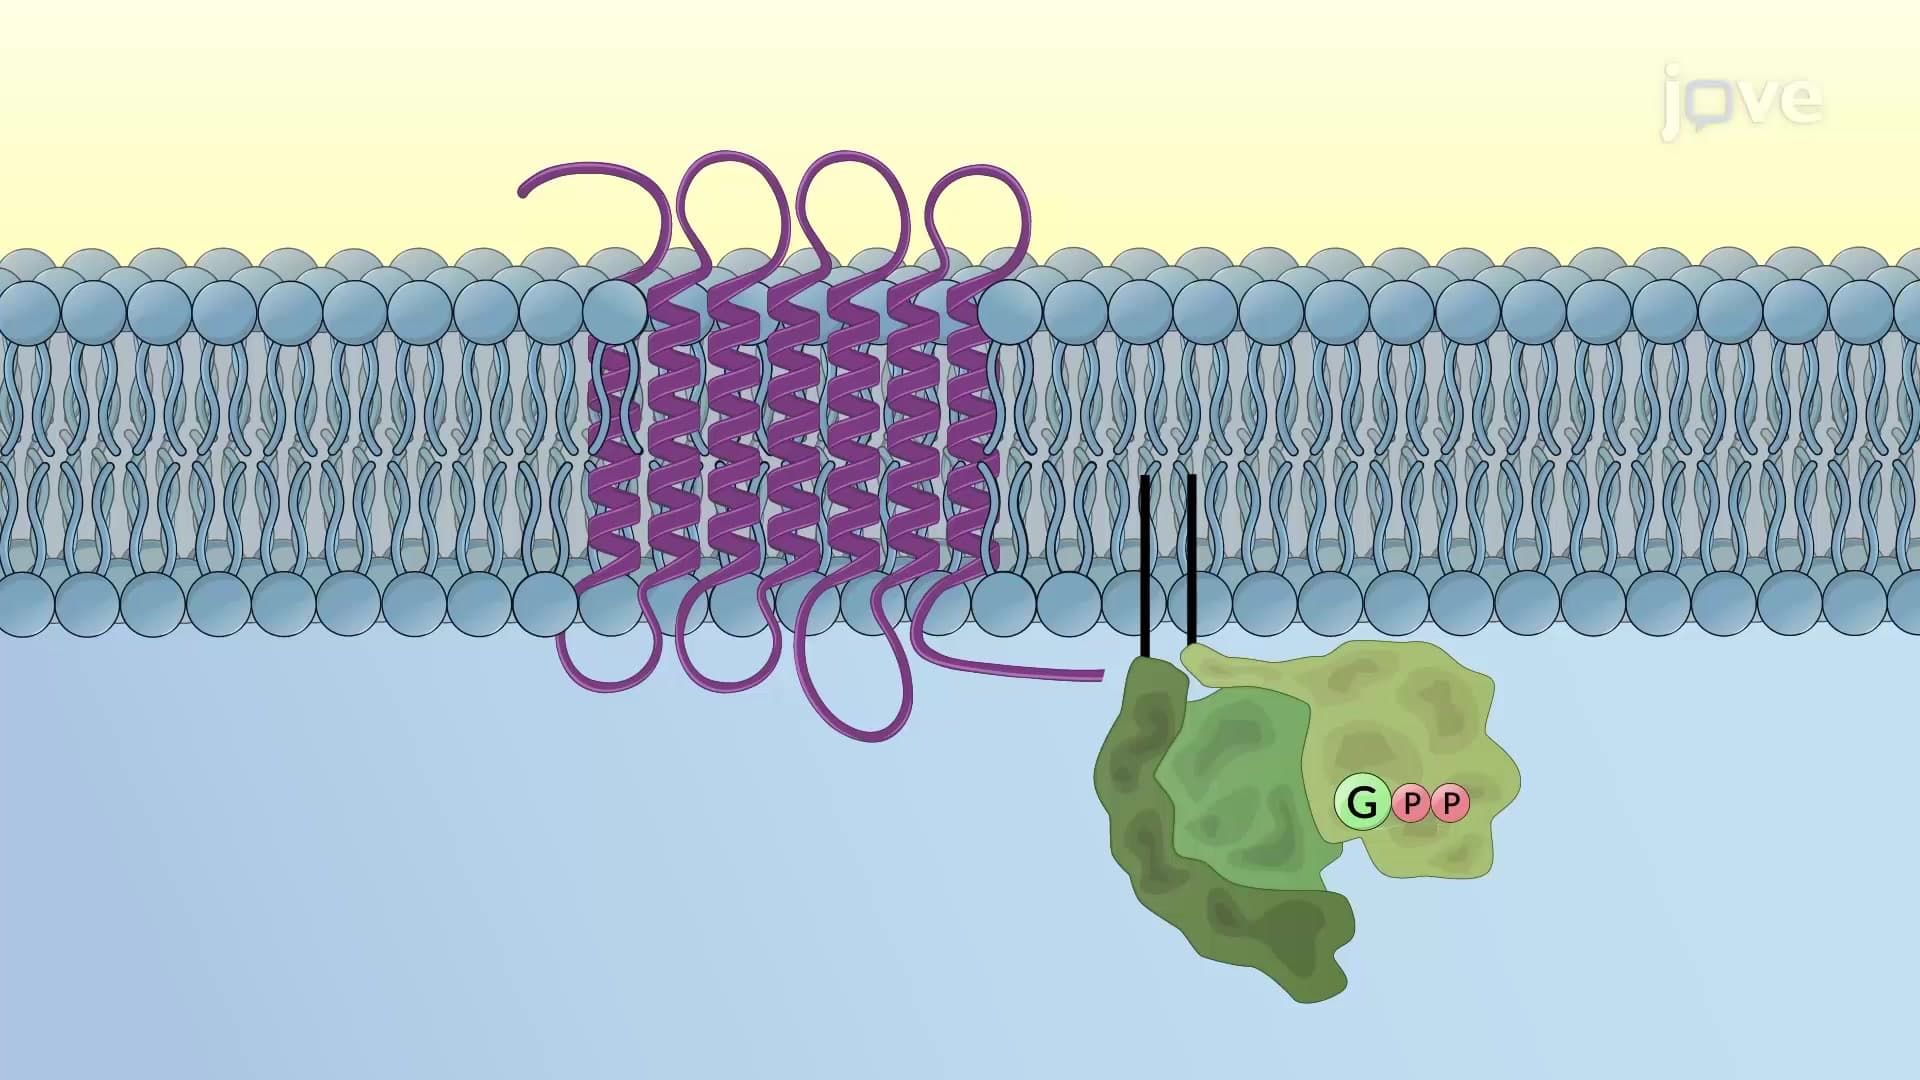
G Protein-coupled Receptors
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...