Related Experiment Video
Updated: Oct 8, 2025


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Accurate X-ray Absorption Spectra near L- and M-Edges from Relativistic Four-Component Damped Response Time-Dependent
Lukas Konecny1, Jan Vicha2, Stanislav Komorovsky3
1Hylleraas Centre for Quantum Molecular Sciences, Department of Chemistry, University of Tromsø - The Arctic University of Norway, 9037 Tromsø, Norway.

Abstract:
The simulation of X-ray absorption spectra requires both scalar and spin-orbit (SO) relativistic effects to be taken into account, particularly near L- and M-edges where the SO splitting of core p and d orbitals dominates. Four-component Dirac-Coulomb Hamiltonian-based linear damped response time-dependent density functional theory (4c-DR-TDDFT) calculates spectra directly for a selected frequency region while including the relativistic effects variationally, making the method well suited for X-ray applications. In this work, we show that accurate X-ray absorption spectra near L2,3- and M4,5-edges of closed-shell transition metal and actinide compounds with different central atoms, ligands, and oxidation states can be obtained by means of 4c-DR-TDDFT. While the main absorption lines do not change noticeably with the basis set and geometry, the exchange-correlation functional has a strong influence with hybrid functionals performing the best. The energy shift compared to the experiment is shown to depend linearly on the amount of Hartee-Fock exchange with the optimal value being 60% for spectral regions above 1000 eV, providing relative errors below 0.2% and 2% for edge energies and SO splittings, respectively. Finally, the methodology calibrated in this work is used to reproduce the experimental L2,3-edge X-ray absorption spectra of [RuCl2(DMSO)2(Im)2] and [WCl4(PMePh2)2], and resolve the broad bands into separated lines, allowing an interpretation based on ligand field theory and double point groups. These results support 4c-DR-TDDFT as a reliable method for calculating and analyzing X-ray absorption spectra of chemically interesting systems, advance the accuracy of state-of-the art relativistic DFT approaches, and provide a reference for benchmarking more approximate techniques.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:12
Author Spotlight: Exploring Light-Driven Chemical Reactions and Energy-Harnessing Devices in Photochemical Research
Published on: February 16, 2024
Related Concept Videos
 01:14
Molecular Spectroscopy: Absorption and Emission
 01:23
Atomic Spectroscopy: Absorption, Emission, and Fluorescence
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 01:13
Atomic Absorption Spectroscopy: Radiation and Light Sources
Two common narrow-range 'line' sources used in AAS are hollow-cathode lamps (HCLs) and...
 01:30
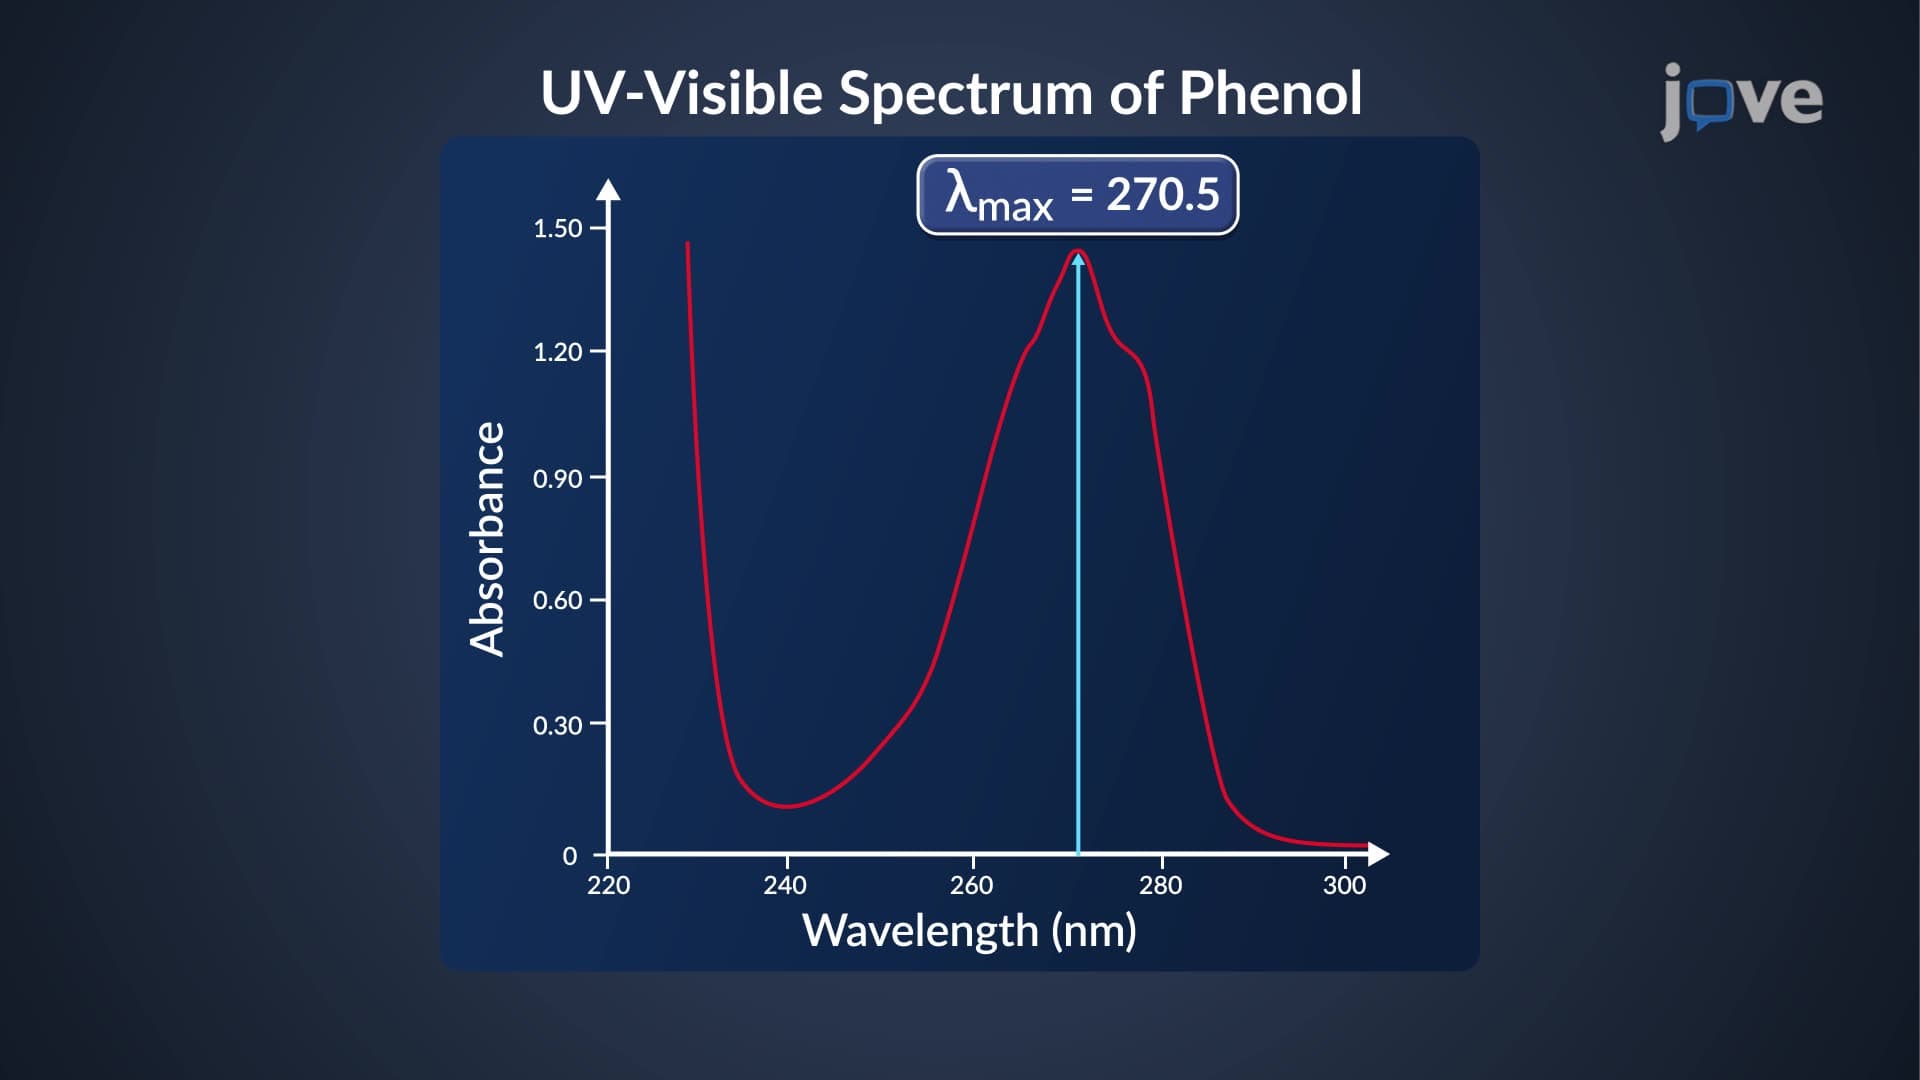
UV–Vis Spectrum
The UV–Vis spectrum of a molecule is the plot of its absorbance versus wavelength. The plot is drawn by taking molar...