Related Experiment Video
Updated: Jun 25, 2026


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
A Comparison between Enrichment Optimization Algorithm (EOA)-Based and Docking-Based Virtual Screening
Jacob Spiegel1, Hanoch Senderowitz1
1Department of Chemistry, Bar-Ilan University, Ramat-Gan 5290002, Israel.

Abstract:
Virtual screening (VS) is a well-established method in the initial stages of many drug and material design projects. VS is typically performed using structure-based approaches such as molecular docking, or various ligand-based approaches. Most docking tools were designed to be as global as possible, and consequently only require knowledge on the 3D structure of the biotarget. In contrast, many ligand-based approaches (e.g., 3D-QSAR and pharmacophore) require prior development of project-specific predictive models. Depending on the type of model (e.g., classification or regression), predictive ability is typically evaluated using metrics of performance on either the training set (e.g.,QCV2) or the test set (e.g., specificity, selectivity or QF1/F2/F32). However, none of these metrics were developed with VS in mind, and consequently, their ability to reliably assess the performances of a model in the context of VS is at best limited. With this in mind we have recently reported the development of the enrichment optimization algorithm (EOA). EOA derives QSAR models in the form of multiple linear regression (MLR) equations for VS by optimizing an enrichment-based metric in the space of the descriptors. Here we present an improved version of the algorithm which better handles active compounds and which also takes into account information on inactive (either known inactive or decoy) compounds. We compared the improved EOA in small-scale VS experiments with three common docking tools, namely, Glide-SP, GOLD and AutoDock Vina, employing five molecular targets (acetylcholinesterase, human immunodeficiency virus type 1 protease, MAP kinase p38 alpha, urokinase-type plasminogen activator, and trypsin I). We found that EOA consistently outperformed all docking tools in terms of the area under the ROC curve (AUC) and EF1% metrics that measured the overall and initial success of the VS process, respectively. This was the case when the docking metrics were calculated based on a consensus approach and when they were calculated based on two different sets of single crystal structures. Finally, we propose that EOA could be combined with molecular docking to derive target-specific scoring functions.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 14:34
A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:26
Drug Discovery: Overview
 01:16
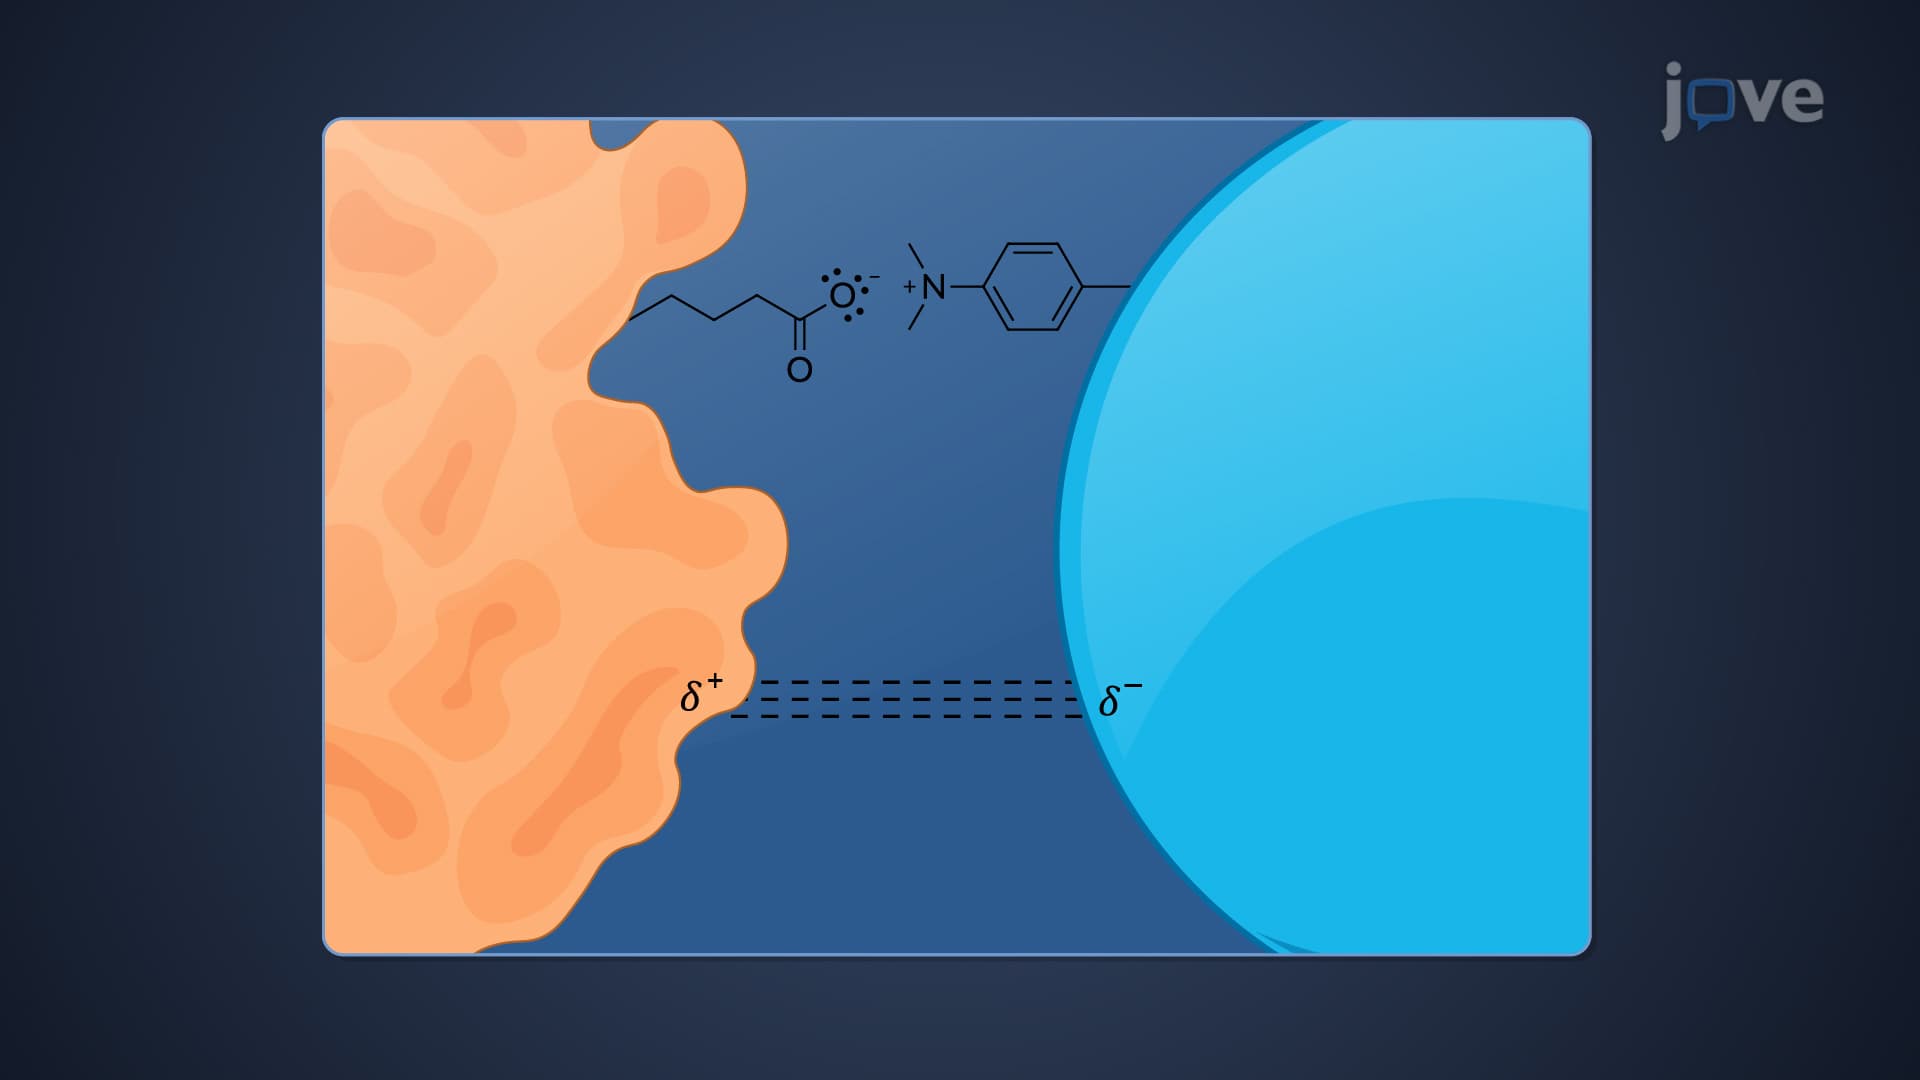
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:22
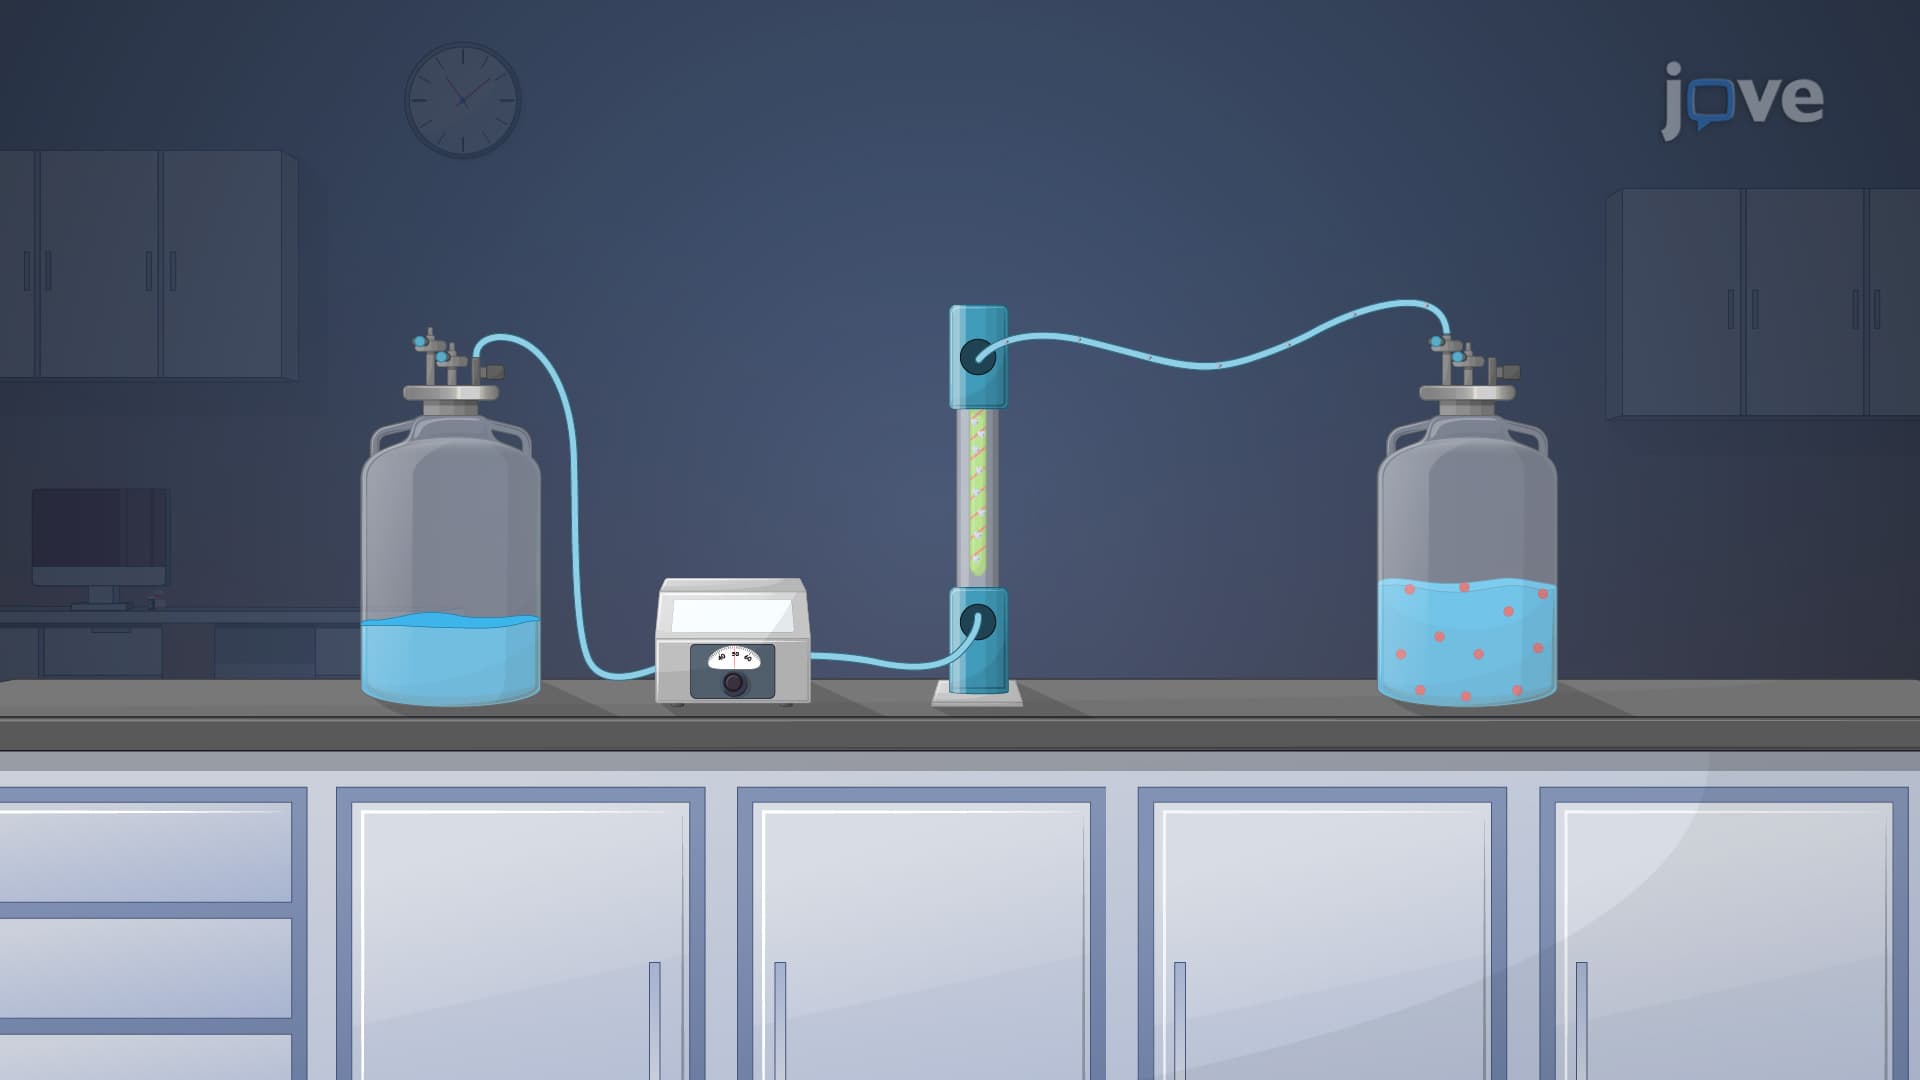
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...