
Clinical Outcome and Underlying Genetic Cause of Functional Terminal Complement Pathway Deficiencies in a Multicenter
Annalie Shears1, Cathal Steele2, Jamie Craig3
1NIHR Paediatric Academic Clinical Fellow, University of Manchester, Manchester, UK.

Background:
Terminal complement pathway deficiencies often present with severe and recurrent infections. There is a lack of good-quality data on these rare conditions. This study investigated the clinical outcome and genetic variation in a large UK multi-center cohort with primary and secondary terminal complement deficiencies.
Methods:
Clinicians from seven UK centers provided anonymised demographic, clinical, and laboratory data on patients with terminal complement deficiencies, which were collated and analysed.
Results:
Forty patients, median age 19 (range 3-62) years, were identified with terminal complement deficiencies. Ten (62%) of 16 patients with low serum C5 concentrations had underlying pathogenic CFH or CFI gene variants. Two-thirds were from consanguineous Asian families, and 80% had an affected family member. The median age of the first infection was 9 years. Forty-three percent suffered meningococcal serotype B and 43% serotype Y infections. Nine (22%) were treated in intensive care for meningococcal septicaemia. Two patients had died, one from intercurrent COVID-19. Twenty-one (52%) were asymptomatic and diagnosed based on family history. All but one patient had received booster meningococcal vaccines and 70% were taking prophylactic antibiotics.
Discussion:
The genetic etiology and clinical course of patients with primary and secondary terminal complement deficiency are variable. Patients with low antigenic C5 concentrations require genetic testing, as the low level may reflect consumption secondary to regulatory defects in the pathway. Screening of siblings is important. Only half of the patients develop septicaemia, but all should have a clear management plan.
More Related Videos
 08:22
A Novel Strategy Combining Array-CGH, Whole-exome Sequencing and In Utero Electroporation in Rodents to Identify Causative Genes for Brain Malformations
Published on: December 1, 2017
 05:53
Candidate Gene Testing in Clinical Cohort Studies with Multiplexed Genotyping and Mass Spectrometry
Published on: June 21, 2018
Related Concept Videos
 01:27
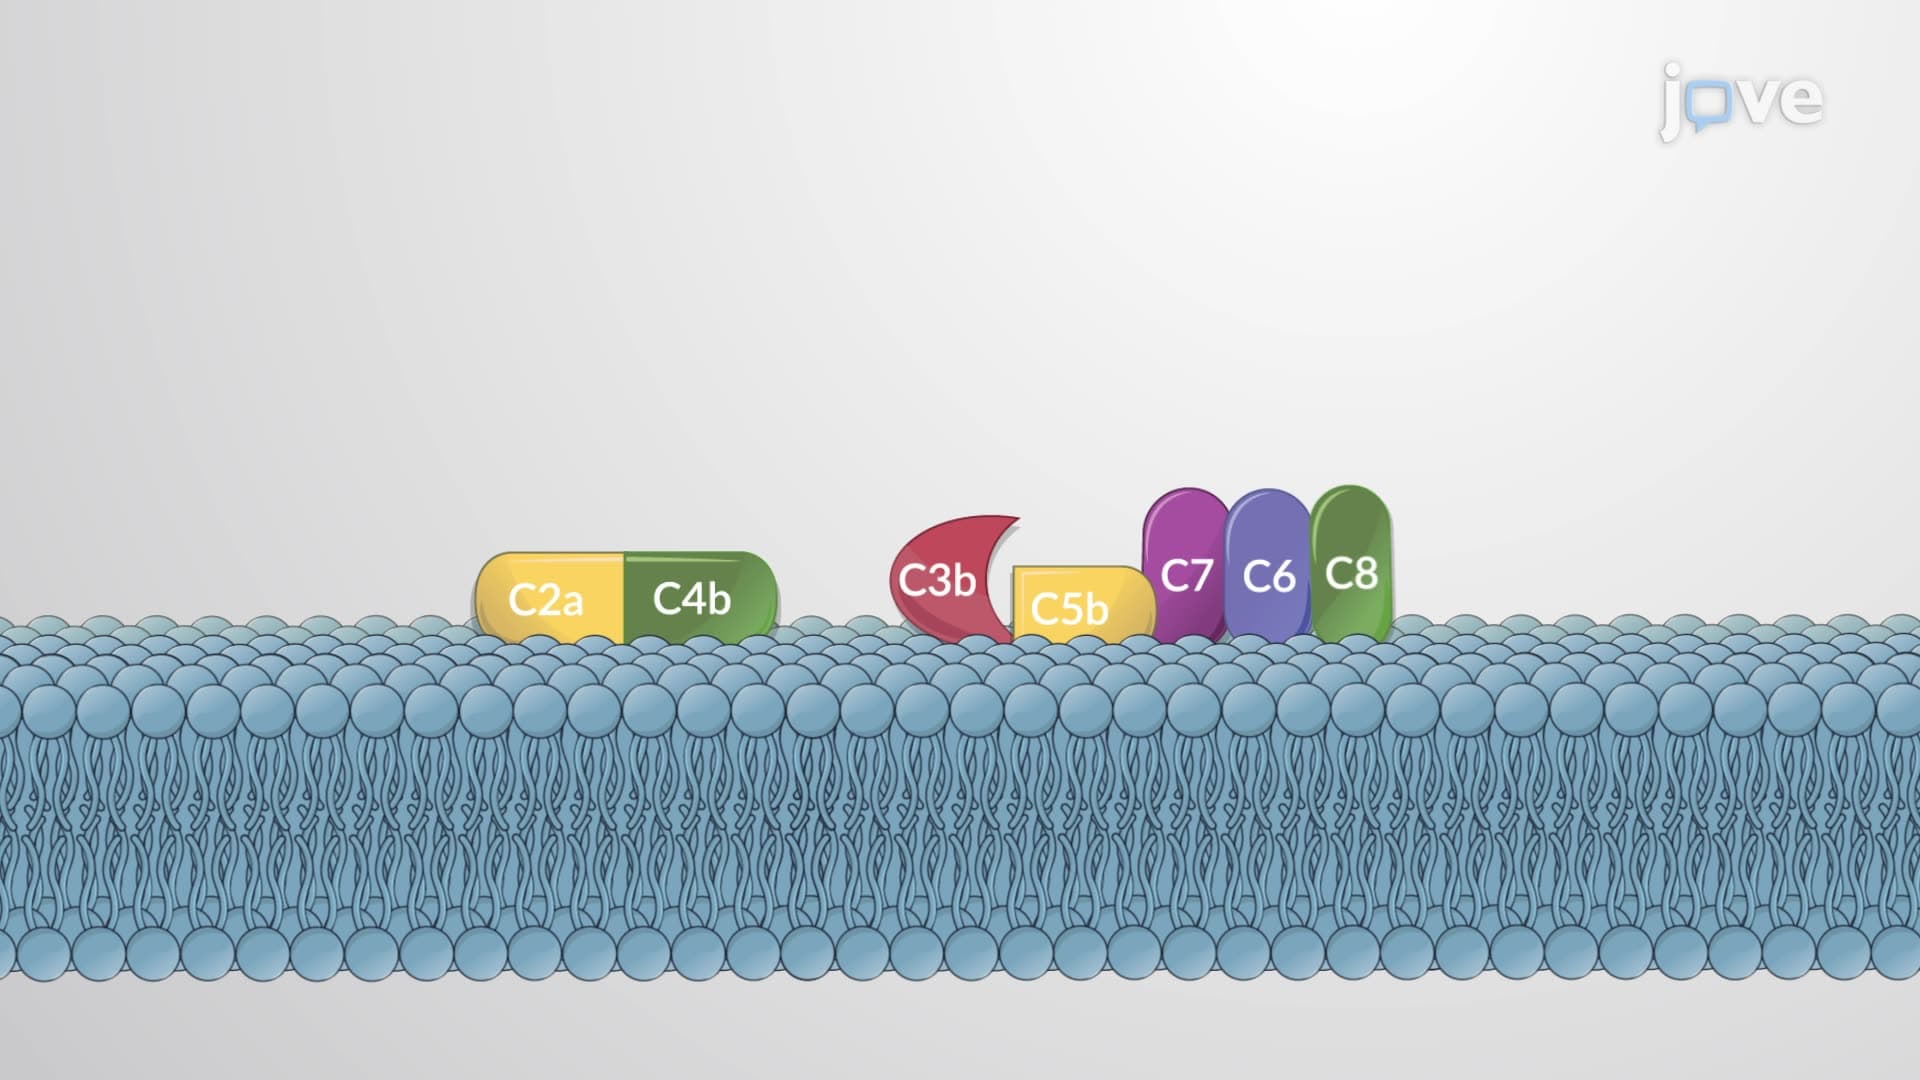
Complement System
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 00:49
Complementation Tests
Organisms heterozygous for different mutations are crossed pairwise in all combinations. If present on different genes, the mutations can complement each other by providing the missing...