Related Experiment Video
Updated: Oct 4, 2025


In Vivo Functional Study of Disease-associated Rare Human Variants Using Drosophila
Published on: August 20, 2019
De Novo ATP1A1 Variants in an Early-Onset Complex Neurodevelopmental Syndrome
Maike F Dohrn1, Adriana P Rebelo1, Siddharth Srivastava1
1From the Dr. John T. Macdonald Foundation (M.F.D., A.P.R., S.Z.), Department of Human Genetics and John P. Hussman Institute for Human Genomics, University of Miami, Miller School of Medicine, FL; Department of Neurology (M.F.D.), Medical Faculty, RWTH Aachen University Hospital, Aachen, Germany; Department of Neurology (S.S., S.M.), Boston Children's Hospital, Harvard Medical School, MA; Department of Translational Medicine (G.C., N.B.-P.), Federico II University; Telethon Institute of Genetics and Medicine (G.C., N.B.-P.), Pozzuoli, Naples, Italy; Department of Pediatrics and Rare Disorders (R.S.), Wroclaw Medical University, Poland; Illumina Inc (A.M., R.T.), San Diego, CA; Division of Pediatric Genetics (D.B.), Department of Genetics, Medical College of Wisconsin, Milwaukee; Department of Clinical Genetics (I.L.), Erasmus MC, University Medical Center Rotterdam; Department of Neurology (R.F.N.), Eramus MC, Medical Center Rotterdam; and Amphia Hospital (C.A.-T.), Breda, the Netherlands.

Abstract:
ATP1A1 encodes the α1 subunit of the sodium-potassium ATPase, an electrogenic cation pump highly expressed in the nervous system. Pathogenic variants in other subunits of the same ATPase, encoded by ATP1A2 or ATP1A3, are associated with syndromes such as hemiplegic migraine, dystonia, or cerebellar ataxia. Worldwide, only 16 families have been reported carrying pathogenic ATP1A1 variants to date. Associated phenotypes are axonal neuropathies, spastic paraplegia, and hypomagnesemia with seizures and intellectual disability. By whole exome or genome sequencing, we identified 5 heterozygous ATP1A1 variants, c.674A>G;p.Gln225Arg, c.1003G>T;p.Gly335Cys, c.1526G>A;p.Gly509Asp, c.2152G>A;p.Gly718Ser, and c.2768T>A;p.Phe923Tyr, in 5 unrelated children with intellectual disability, spasticity, and peripheral, motor predominant neuropathy. Additional features were sensory loss, sleep disturbances, and seizures. All variants occurred de novo and are absent from control populations (MAF GnomAD = 0). Affecting conserved amino acid residues and constrained regions, all variants have high pathogenicity in silico prediction scores. In HEK cells transfected with ouabain-insensitive ATP1A1 constructs, cell viability was significantly decreased in mutants after 72h treatment with the ATPase inhibitor ouabain, demonstrating loss of ATPase function. Replicating the haploinsufficiency mechanism of disease with a gene-specific assay provides pathogenicity information and increases certainty in variant interpretation. This study further expands the genotype-phenotype spectrum of ATP1A1.
More Related Videos
 08:22
A Novel Strategy Combining Array-CGH, Whole-exome Sequencing and In Utero Electroporation in Rodents to Identify Causative Genes for Brain Malformations
Published on: December 1, 2017
 08:04
Identification and Classification of Position-specific GABAA Receptor Subunit Missense Variants for Their Role In Hippocampal Pyramidal Neurons
Published on: June 6, 2025
Related Concept Videos
 01:48
ATP Synthase: Mechanism
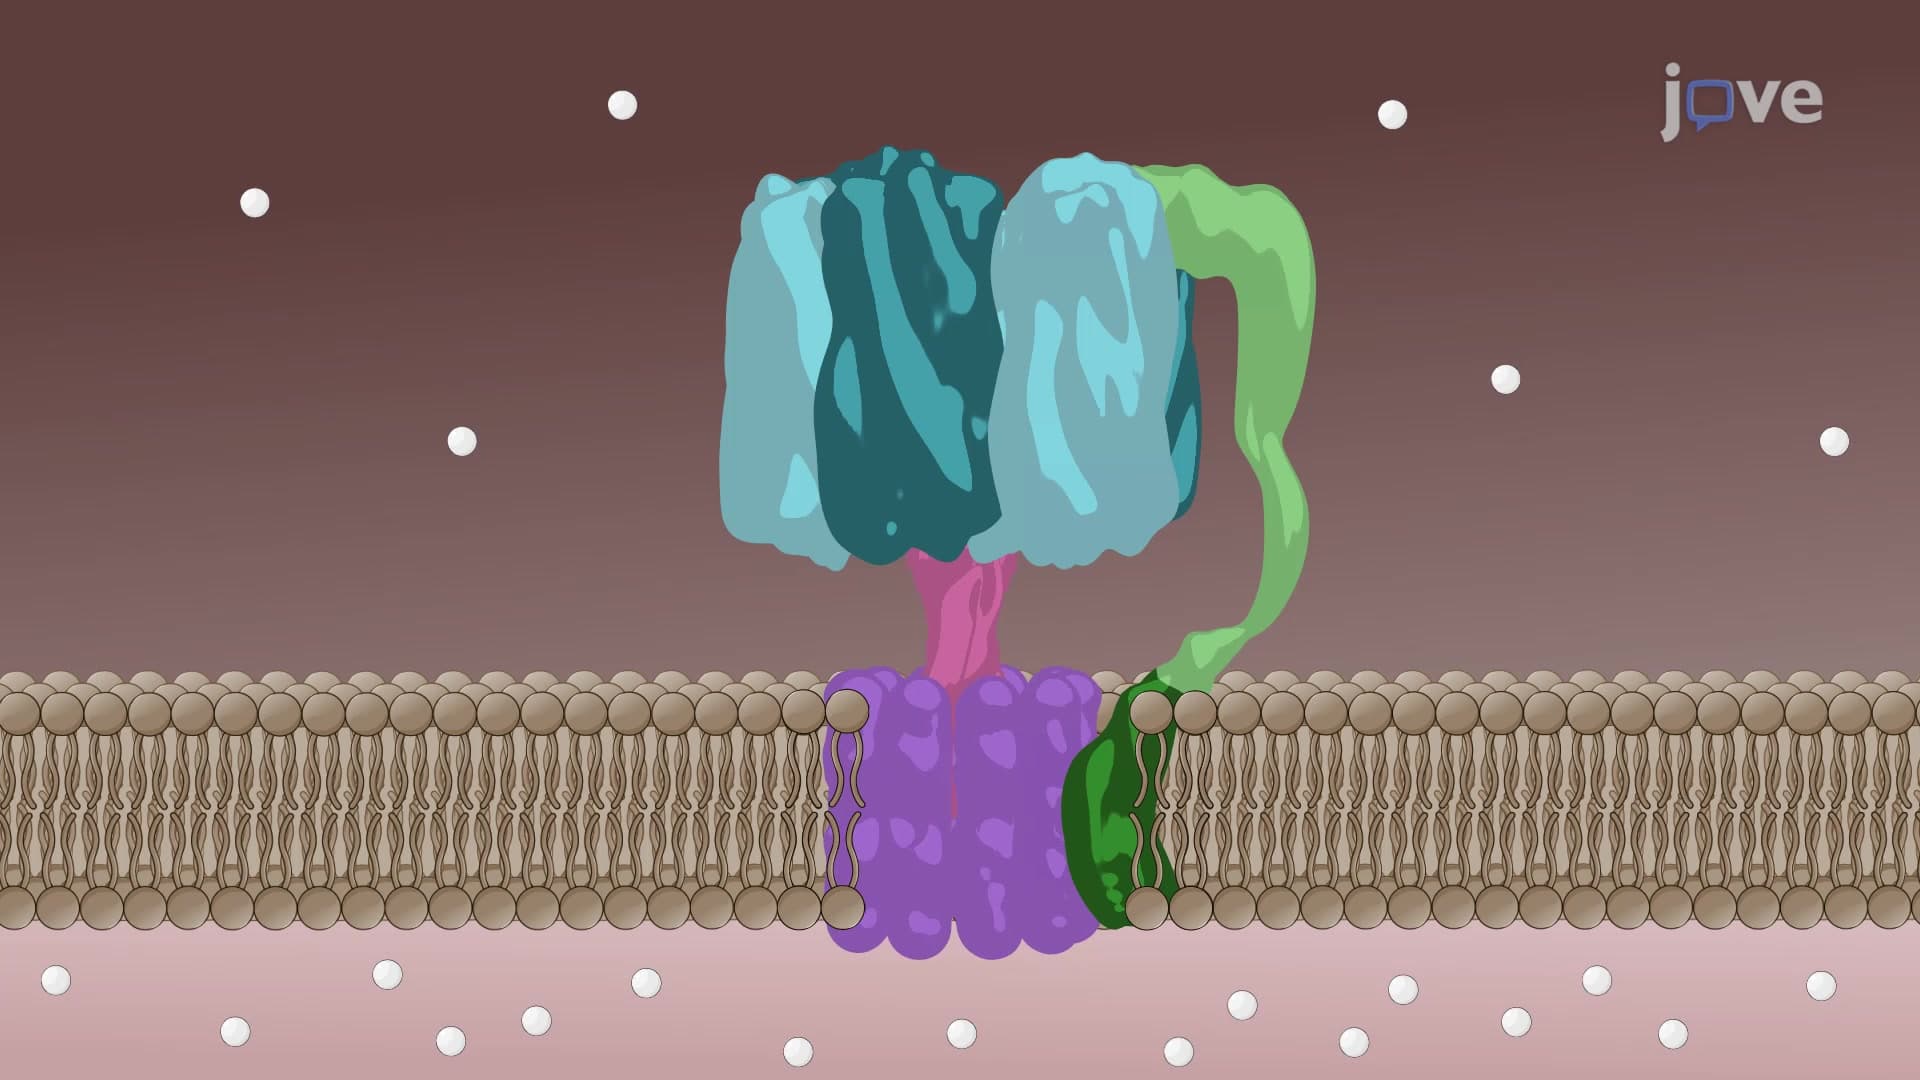 01:18
ATP Synthase: Structure
 01:43
Sex-linked Disorders
 01:19
Autism Spectrum Disorder
These core symptoms manifest differently among individuals, ranging from mild to severe. The disorder's complexity extends beyond its clinical presentation, encompassing a diverse range of biological, cognitive, and sociocultural influences.
 01:20
Antiepileptic Drugs: Modulators of Neurotransmitter Release Mediated by SV2A Protein
SV2A is a transmembrane glycoprotein located predominantly in the brain, modulating the release of neurotransmitters for neuronal communication. Both levetiracetam and brivaracetam exhibit a high affinity for...
 02:59
Animal Mitochondrial Genetics