Related Experiment Video
Updated: Oct 2, 2025


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Combined Free-Energy Calculation and Machine Learning Methods for Understanding Ligand Unbinding Kinetics
Magd Badaoui1,2, Pedro J Buigues2, Dénes Berta2
1Department of Chemistry, King's College London, London SE1 1DB, United Kingdom.

Abstract:
The determination of drug residence times, which define the time an inhibitor is in complex with its target, is a fundamental part of the drug discovery process. Synthesis and experimental measurements of kinetic rate constants are, however, expensive and time consuming. In this work, we aimed to obtain drug residence times computationally. Furthermore, we propose a novel algorithm to identify molecular design objectives based on ligand unbinding kinetics. We designed an enhanced sampling technique to accurately predict the free-energy profiles of the ligand unbinding process, focusing on the free-energy barrier for unbinding. Our method first identifies unbinding paths determining a corresponding set of internal coordinates (ICs) that form contacts between the protein and the ligand; it then iteratively updates these interactions during a series of biased molecular dynamics (MD) simulations to reveal the ICs that are important for the whole of the unbinding process. Subsequently, we performed finite-temperature string simulations to obtain the free-energy barrier for unbinding using the set of ICs as a complex reaction coordinate. Importantly, we also aimed to enable the further design of drugs focusing on improved residence times. To this end, we developed a supervised machine learning (ML) approach with inputs from unbiased "downhill" trajectories initiated near the transition state (TS) ensemble of the string unbinding path. We demonstrate that our ML method can identify key ligand-protein interactions driving the system through the TS. Some of the most important drugs for cancer treatment are kinase inhibitors. One of these kinase targets is cyclin-dependent kinase 2 (CDK2), an appealing target for anticancer drug development. Here, we tested our method using two different CDK2 inhibitors for the potential further development of these compounds. We compared the free-energy barriers obtained from our calculations with those observed in available experimental data. We highlighted important interactions at the distal ends of the ligands that can be targeted for improved residence times. Our method provides a new tool to determine unbinding rates and to identify key structural features of the inhibitors that can be used as starting points for novel design strategies in drug discovery.
More Related Videos
Related Concept Videos
 02:18
The Equilibrium Binding Constant and Binding Strength
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:22
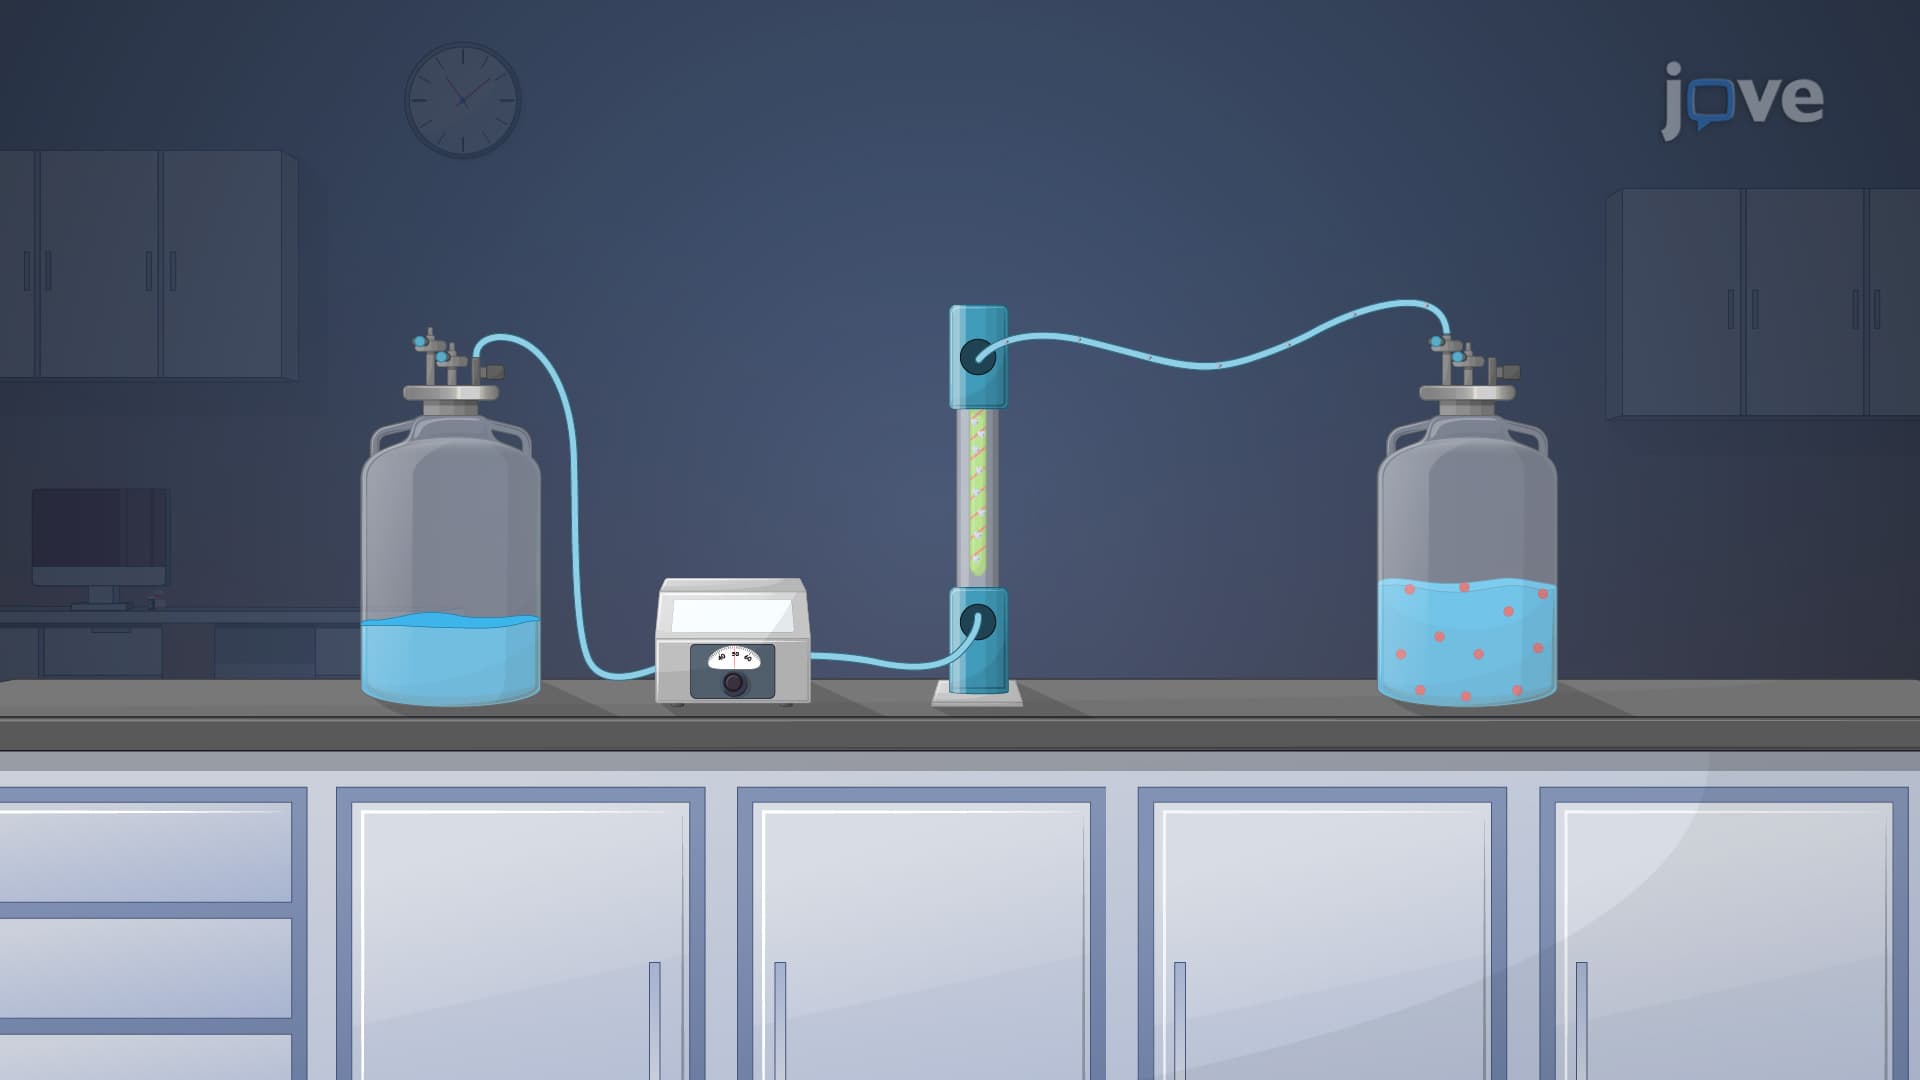
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 00:49
Ligand Binding and Linkage
 01:16
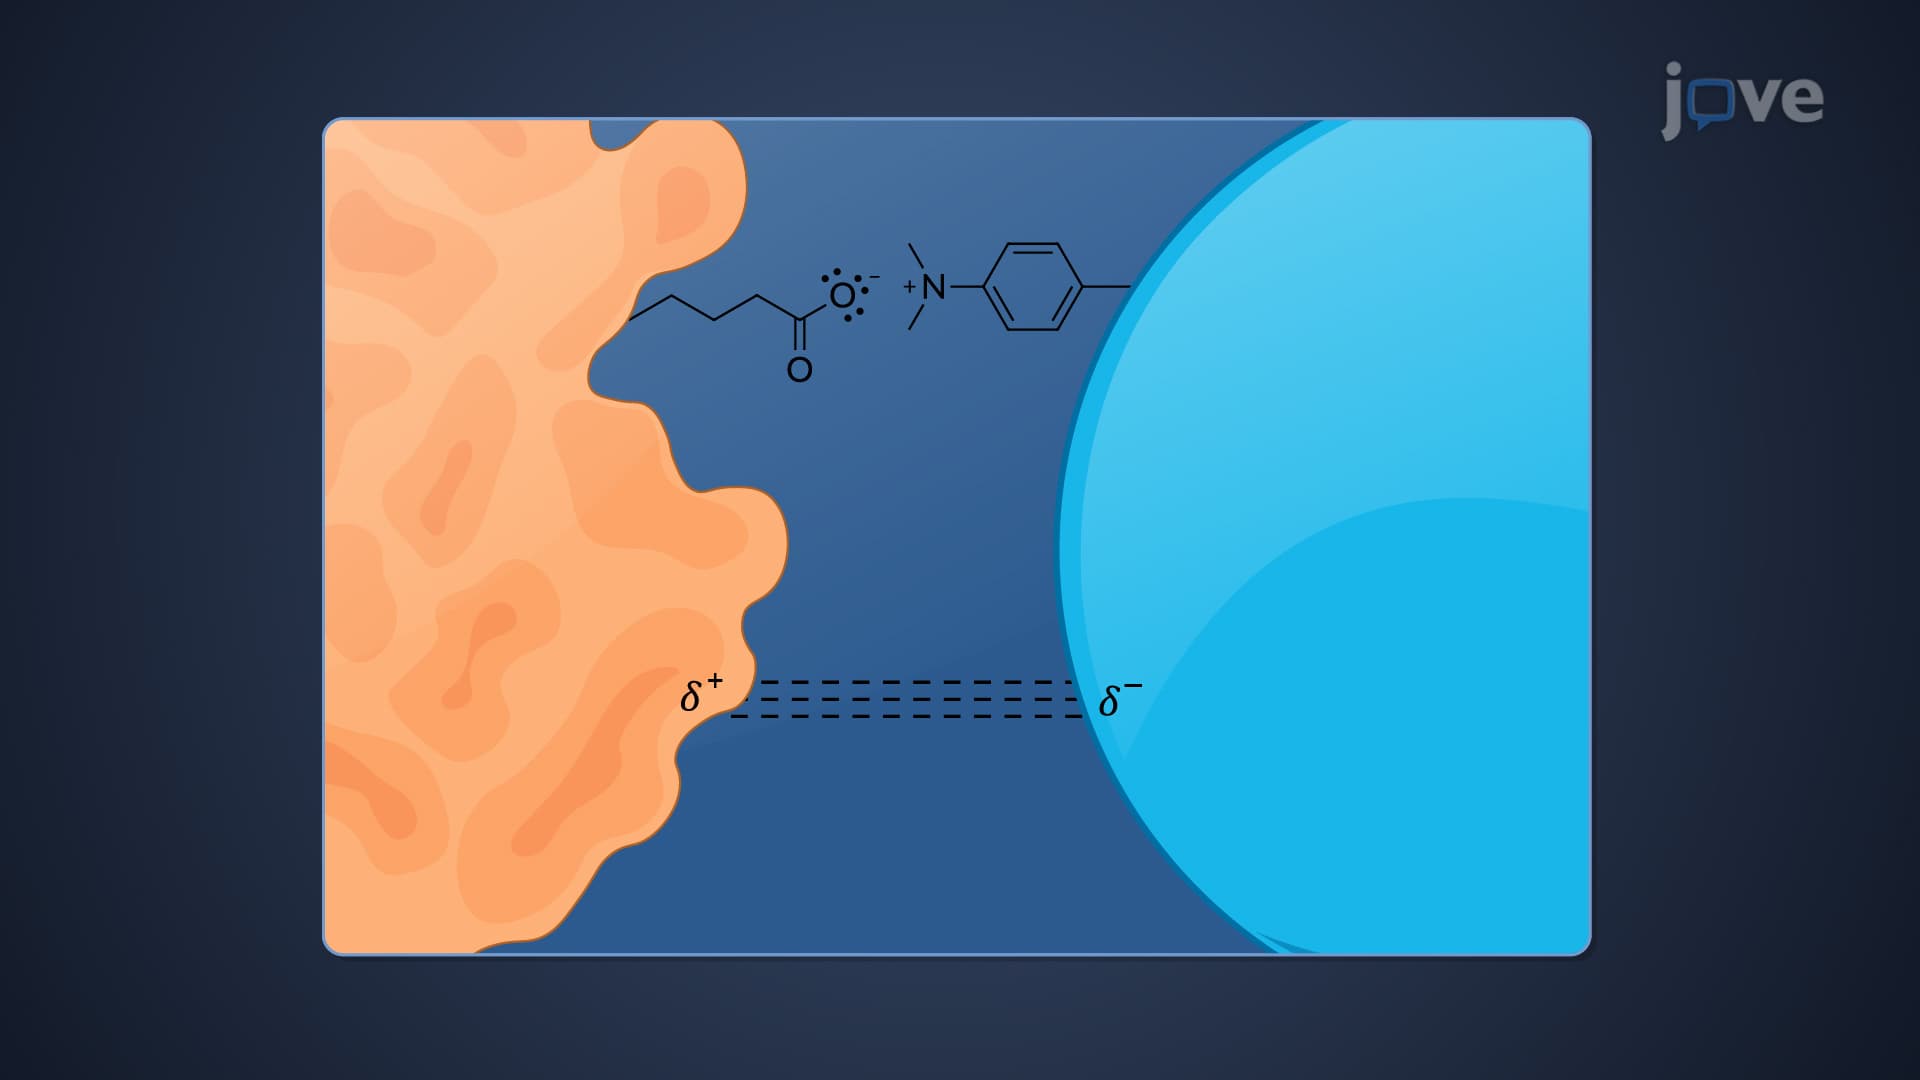
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...