Related Experiment Video
Updated: Oct 2, 2025


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Developing end-point methods for absolute binding free energy calculation using the Boltzmann-quasiharmonic model
Lauren Wickstrom1, Emilio Gallicchio2,3,4, Lieyang Chen3,4,5
1Borough of Manhattan Community College, The City University of New York, Department of Science, New York, New York, USA.

Abstract:
Understanding the physical forces underlying receptor-ligand binding requires robust methods for analyzing the binding thermodynamics. In end-point binding free energy methods the binding free energy is naturally decomposable into physically intuitive contributions such as the solvation free energy and configurational entropy that can provide insights. Here we present a new end-point method called EE-BQH (Effective Energy-Boltzmann-Quasiharmonic) which combines the Boltzmann-Quasiharmonic model for configurational entropy with different solvation free energy methods, such as the continuum solvent PBSA model and the integral equation-based 3D-RISM, to estimate the absolute binding free energy. We compare EE-BQH with other treatments of configurational entropy such as Quasiharmonic models in internal coordinates (QHIC) and in Cartesian coordinates (QHCC), and Normal Mode analysis (NMA), by testing them on the octa acids host-guest complexes from the SAMPL8 blind challenge. The accuracies in the calculated absolute binding free energies strongly depend on the configurational entropy and solvation free energy methods used. QHIC and BQH yield the best agreements with the established potential of mean force (PMF) estimates, with R2 of ∼0.7 and mean unsigned error of ∼1.7 kcal mol-1. These results from the end-point calculations are also in similar agreement with experiments. While 3D-RISM in combination with QHIC or BQH lead to reasonable correlations with the PMF results and experiments, the calculated absolute binding free energies are underestimated by ∼5 kcal mol-1. While the binding is accompanied by a significant reduction in the ligand translational/rotational entropy, the change in the torsional entropy in these host-guest systems is slightly positive. Compared with BQH, QHIC underestimates the reduction of configurational entropy because of the non-Gaussian probability distributions in the ligand rotation and a small number of torsions. The study highlights the crucial role of configurational entropy in determining binding and demonstrates the potential of using the new end-point method to provide insights in more complex protein-ligand systems.
More Related Videos

Related Concept Videos
 02:18
The Equilibrium Binding Constant and Binding Strength
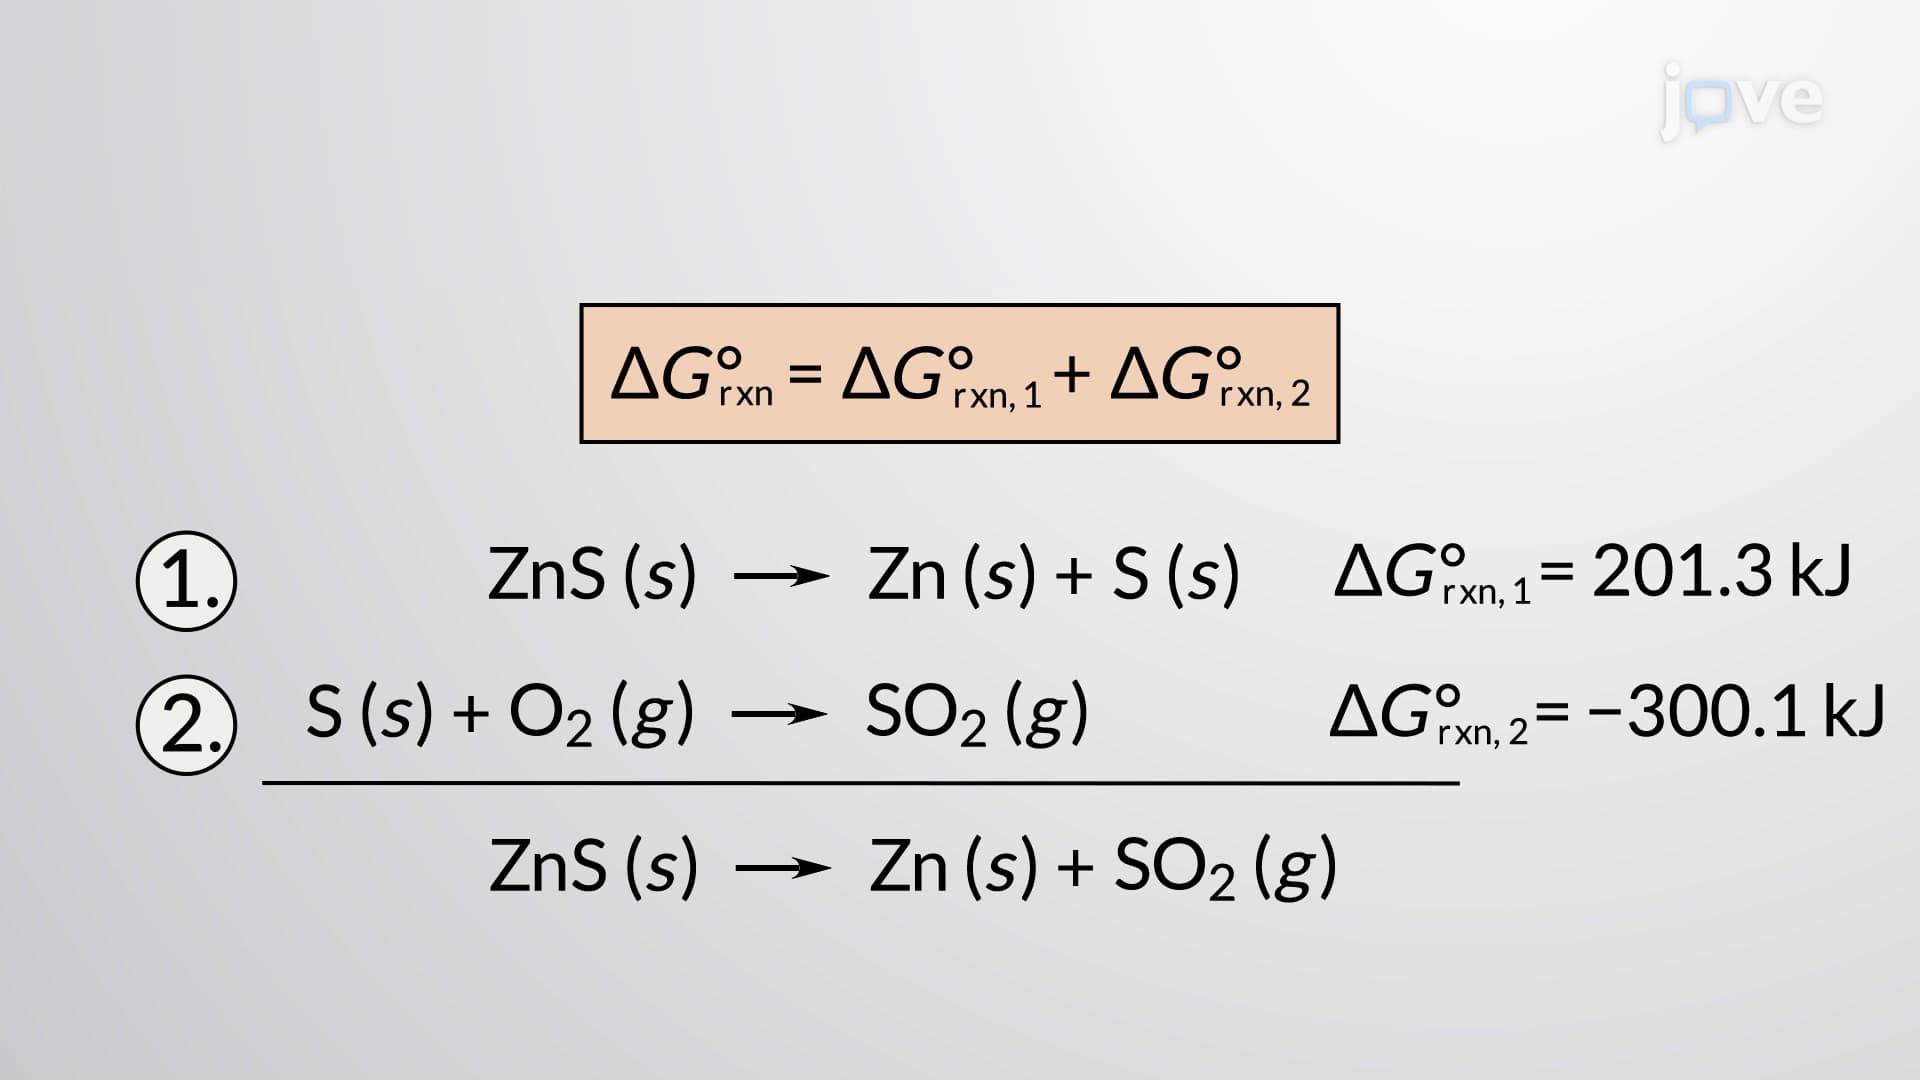 02:49
Calculating Standard Free Energy Changes
 01:26
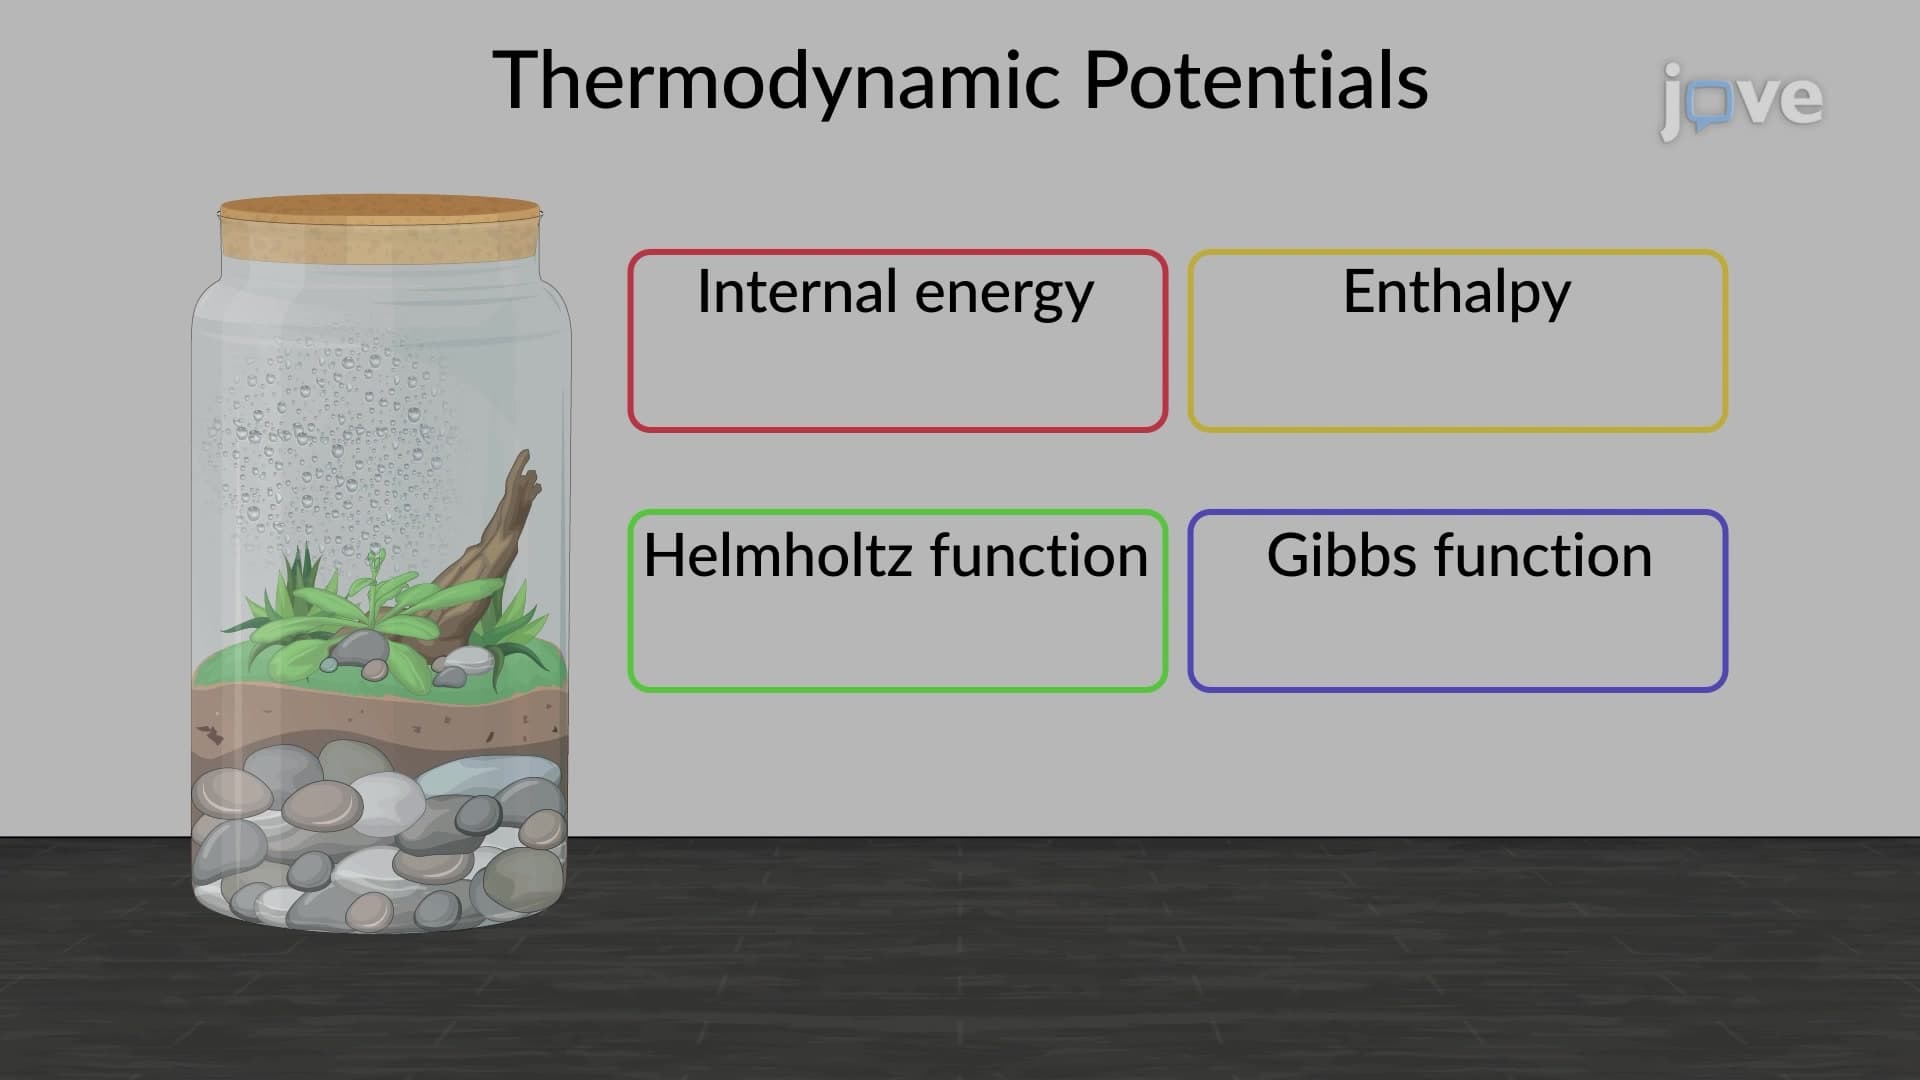
Thermodynamic Potentials
 02:44
The Born-Haber Cycle
 02:13
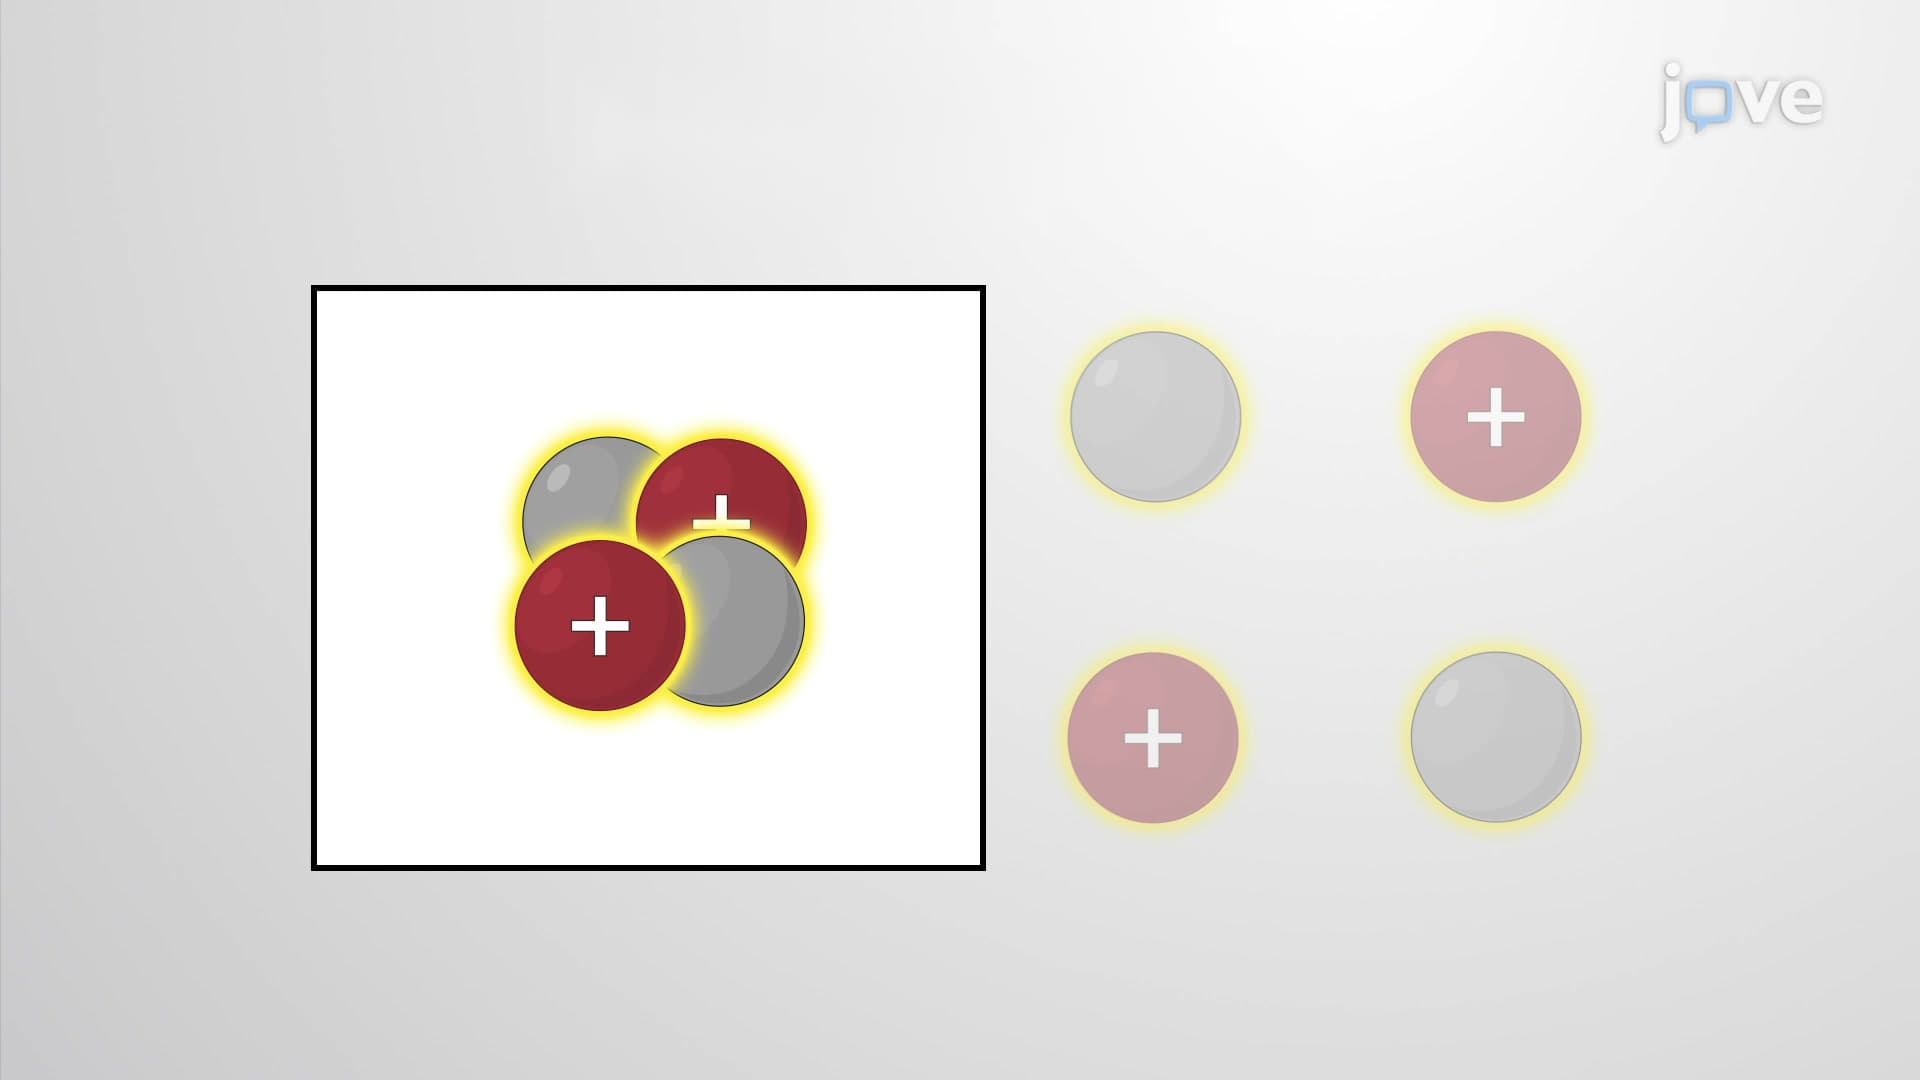
Nuclear Binding Energy
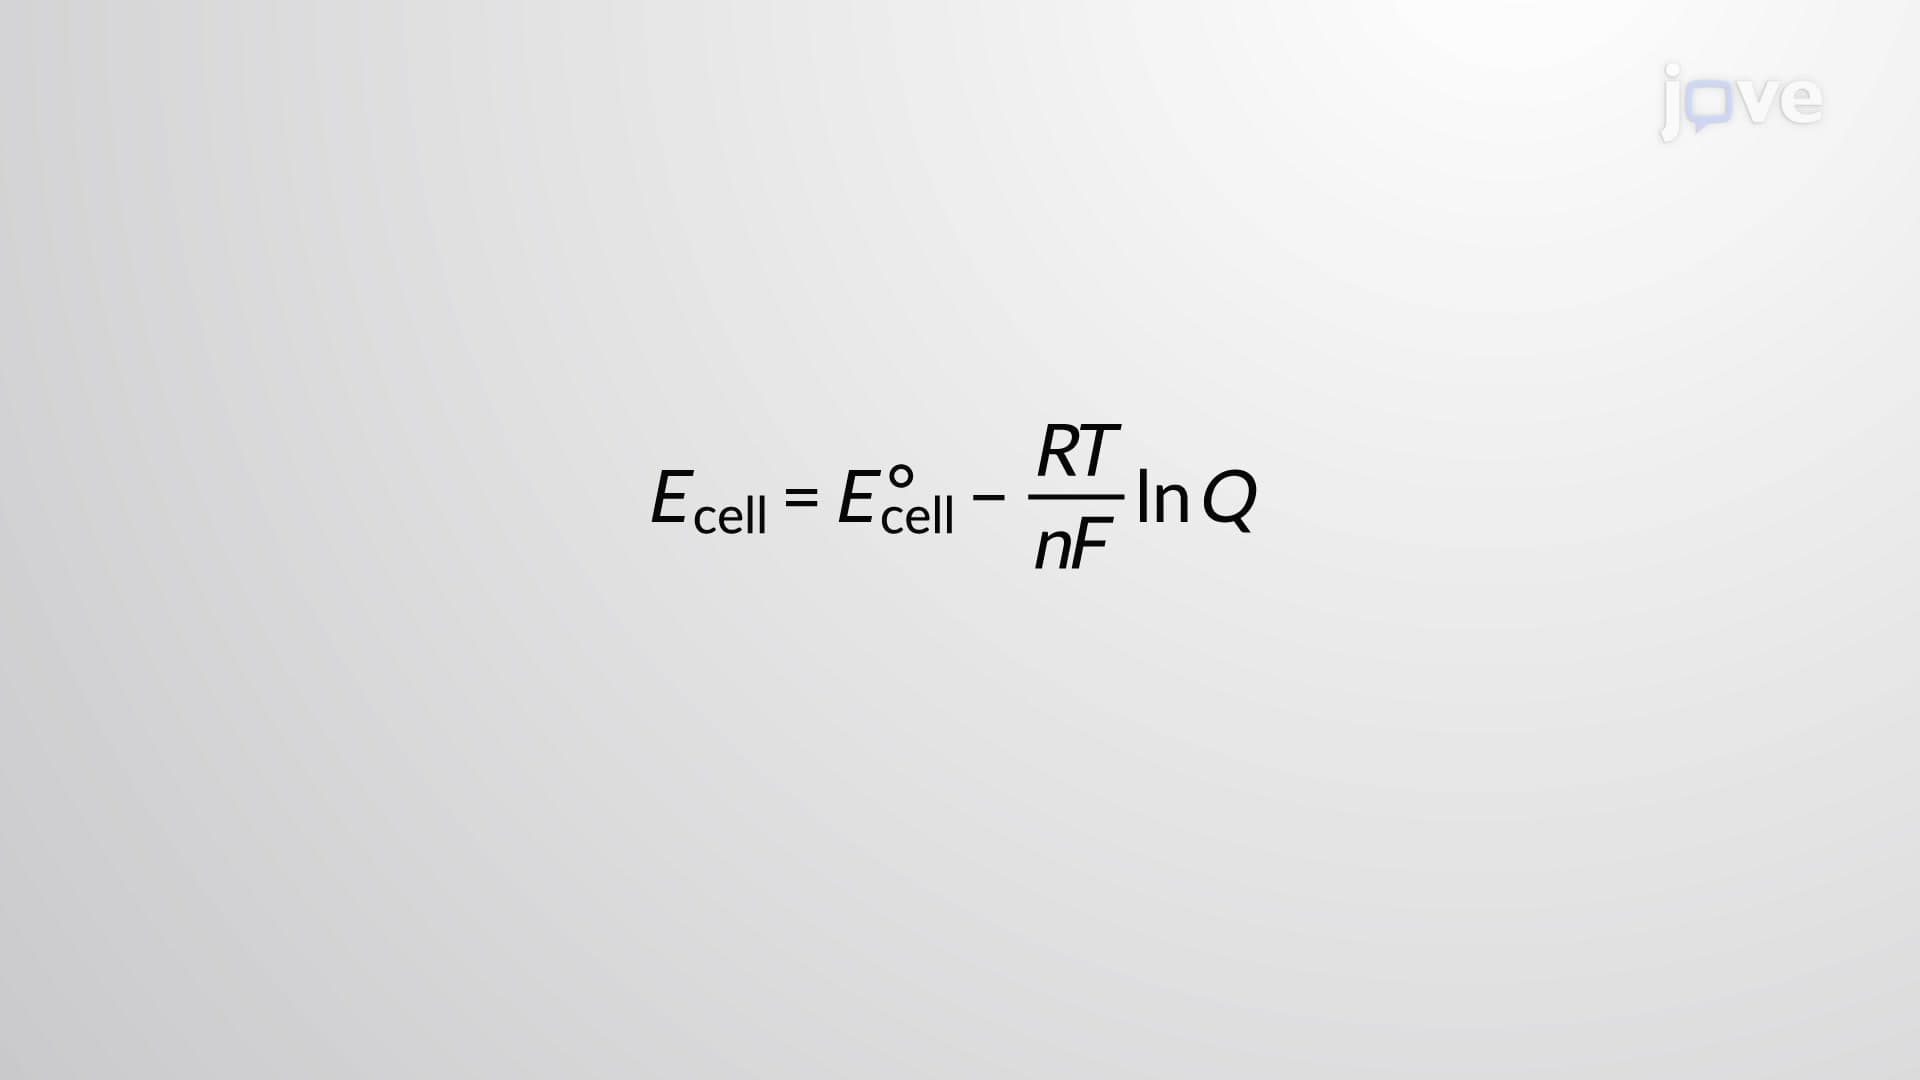 02:59
The Nernst Equation
The interconnection between standard cell potentials and various thermodynamic parameters such as the standard free energy change ΔG° and equilibrium constant K has been previously explored. For example, a redox reaction involving zinc(II) and tin(II) ions at 1 M concentration with Eºcell = +0.291 V and ΔG° = −56.2 kJ is spontaneous.