Related Experiment Video
Updated: Sep 30, 2025

Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Single Excitation Energies Obtained from the Ensemble "HOMO-LUMO Gap": Exact Results and Approximations
Tim Gould1, Zahed Hashimi1, Leeor Kronik2
1Queensland Micro- and Nanotechnology Centre, Griffith University, Nathan, QLD 4111, Australia.

Abstract:
In calculations based on density functional theory, the "HOMO-LUMO gap" (difference between the highest occupied and lowest unoccupied molecular orbital energies) is often used as a low-cost, ad hoc approximation for the lowest excitation energy. Here we show that a simple correction based on rigorous ensemble density functional theory makes the HOMO-LUMO gap exact in principle and significantly more accurate in practice. The introduced perturbative ensemble density functional theory approach predicts different and useful values for singlet-singlet and singlet-triplet excitations, using semilocal and hybrid approximations. Excitation energies are similar in quality to time-dependent density functional theory, especially at high fractions of exact exchange. The approach therefore offers an easy-to-implement and low-cost route to robust prediction of molecular excitation energies.
Related Concept Videos
 02:13
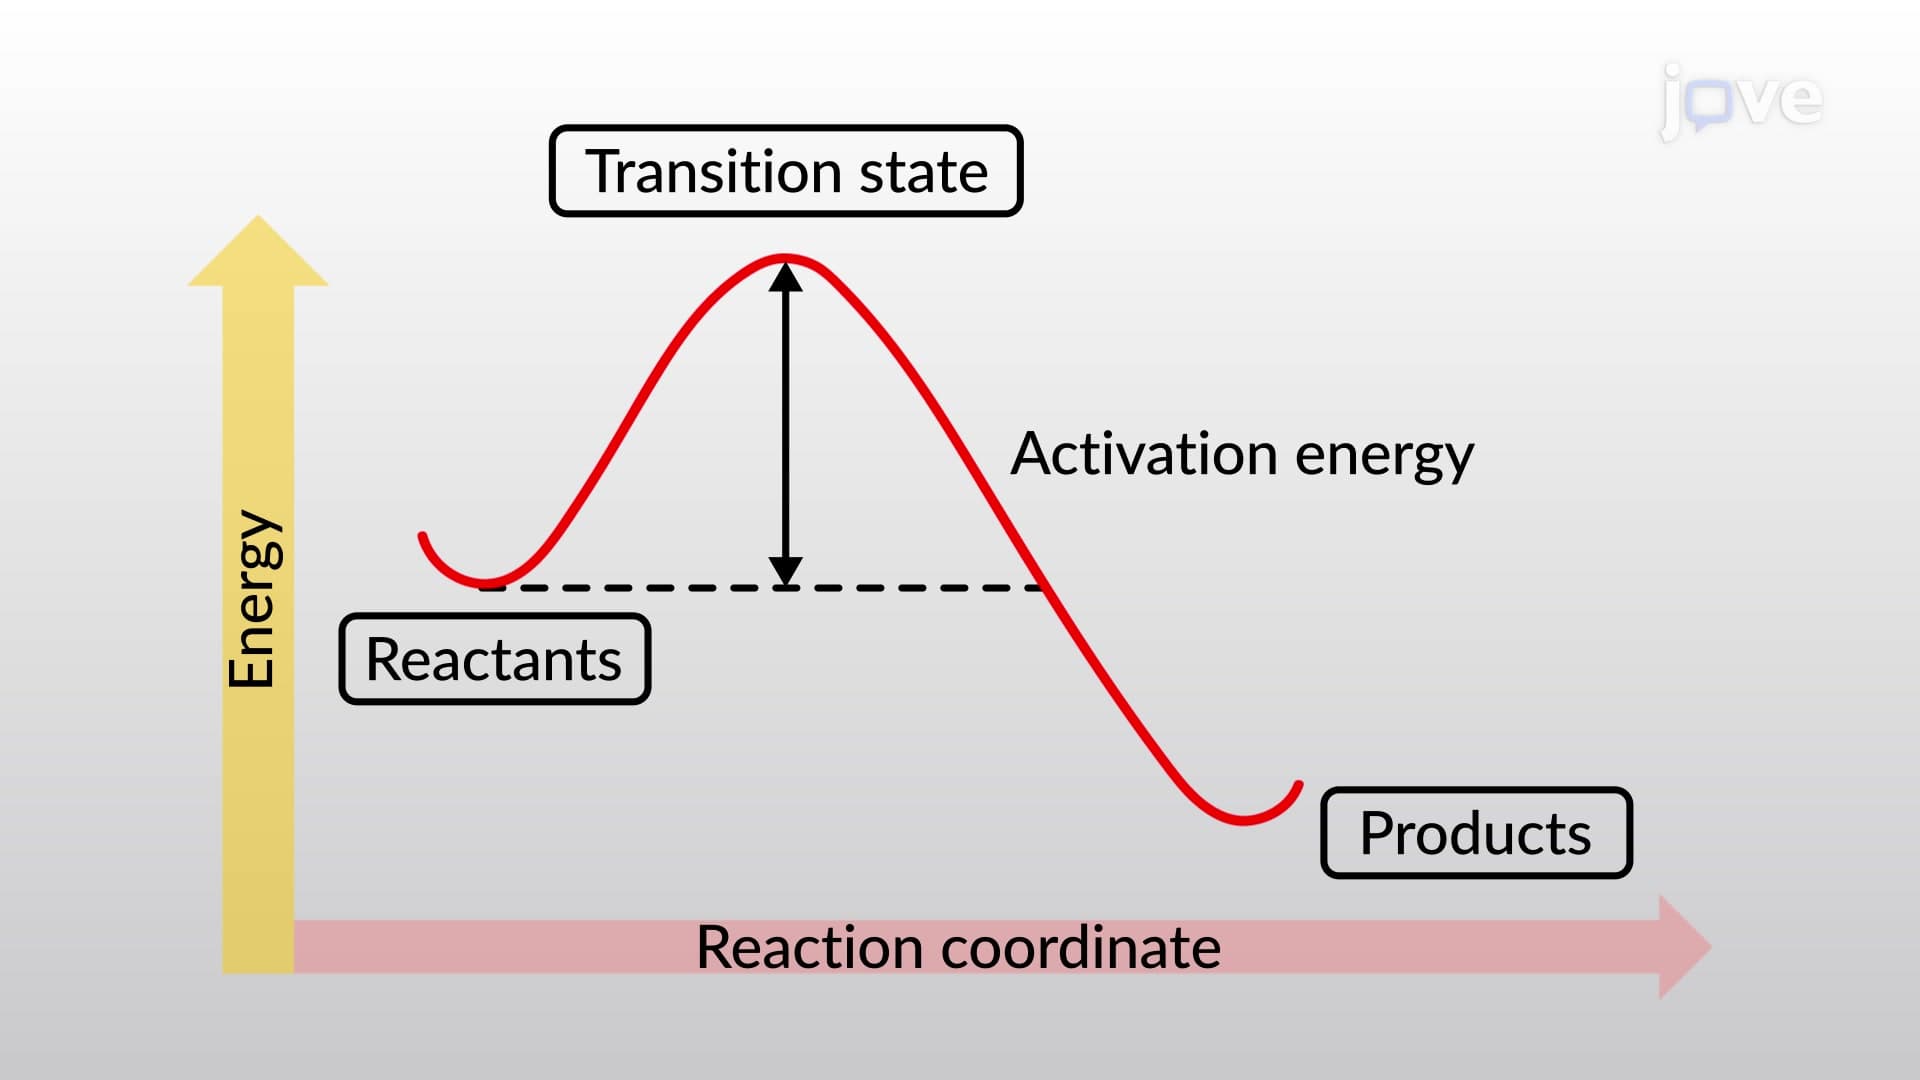
Bond Dissociation Energy and Activation Energy
 01:13
¹H NMR Signal Multiplicity: Splitting Patterns
 01:12
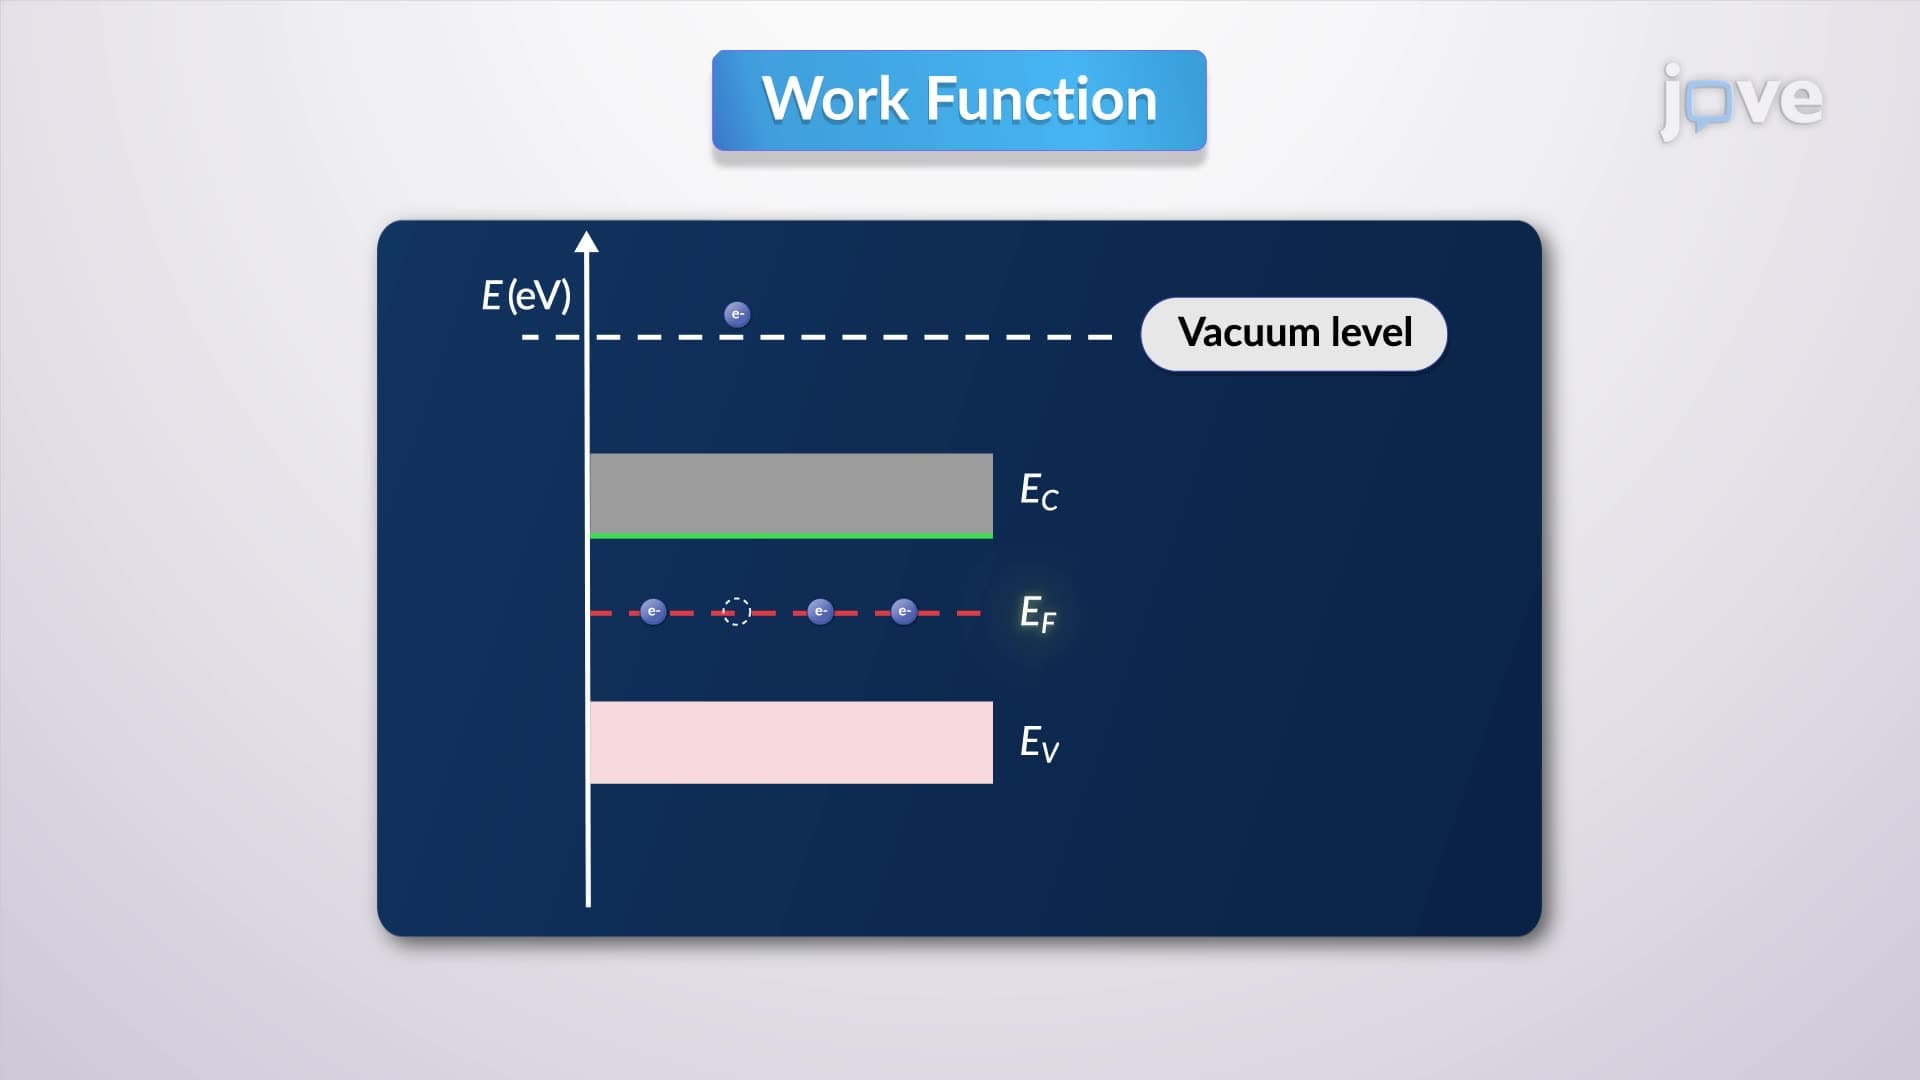
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 01:17
Electron Paramagnetic Resonance (EPR) Spectroscopy: Organic Radicals
 02:13
Energy Diagrams, Transition States, and Intermediates
 01:21
Energy Associated With a Charge Distribution