Related Experiment Video
Updated: Sep 27, 2025
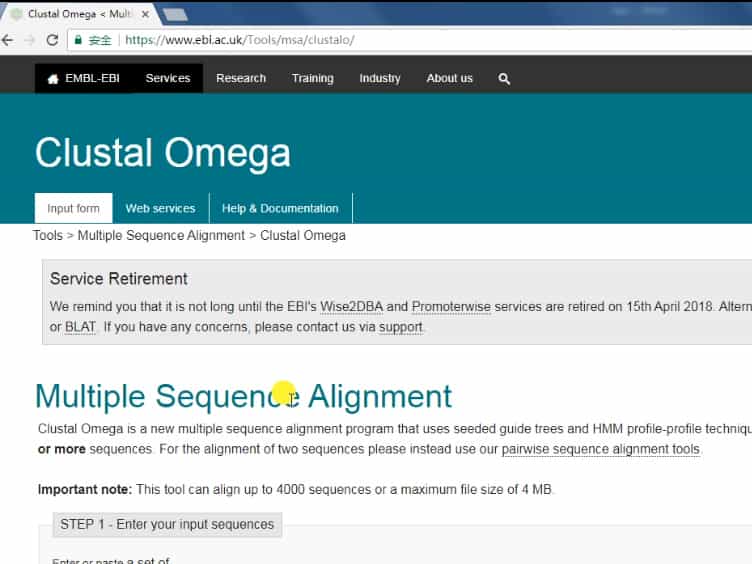
Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
SaAlign: Multiple DNA/RNA sequence alignment and phylogenetic tree construction tool for ultra-large datasets and
Ziyuan Wang1, Junjie Tan2, Yanling Long3
1Key Laboratory of Bio-Resource and Eco-Environment of Ministry of Education, College of Life Sciences, Sichuan University, Chengdu 610064, Sichuan, PR China.

Abstract:
Multiple DNA/RNA sequence alignment is an important fundamental tool in bioinformatics, especially for phylogenetic tree construction. With DNA-sequencing improvements, the amount of bioinformatics data is constantly increasing, and various tools need to be iterated constantly. Mitochondrial genome analyses of multiple individuals and species require bioinformatics software; therefore, their performances need to be optimized. To improve the alignment of ultra-large datasets and ultra-long sequences, we optimized a dynamic programming algorithm using longest common substring methods. Ultra-large test DNA datasets, containing sequences of different lengths, some over 300 kb (kilobase), revealed that the Multiple DNA/RNA Sequence Alignment Tool Based on Suffix Tree (SaAlign) saved time and computational space. It outperformed the existing technical tools, including MAFFT and HAlign-II. For mitochondrial genome datasets having limited numbers of sequences, MAFFT performed the required tasks, but it could not handle ultra-large mitochondrial genome datasets for core dump error. We implement a multiple DNA/RNA sequence alignment tool based on Center Star strategy and use suffix array algorithm to optimize the spatial and time efficiency. Nowadays, whole-genome research and NGS technology are becoming more popular, and it is necessary to save computational resources for laboratories. That software is of great significance in these aspects, especially in the study of the whole-mitochondrial genome of plants.
Related Concept Videos
 02:54
Evolutionary Relationships through Genome Comparisons
 01:29
Modern Molecular Taxonomy
 03:21
Phylogenetic Trees
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
 01:57
Sanger Sequencing
 02:51
Multi-species Conserved Sequences
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...