
Electronic, optical and thermoelectric properties of Fe2ZrP compound determined via first-principles calculations
Esmaeil Pakizeh1, Jaafar Jalilian2, Mahnaz Mohammadi3
1Faculty of Petroleum and Gas, Yasouj University Gachsaran 75813-56001 Iran e.pakizeh@yu.ac.ir esmaeil_pakizeh@yahoo.com.

Abstract:
In this study, based on the density functional theory and semi-classical Boltzmann transport theory, we investigated the structural, thermoelectric, optical and phononic properties of the Fe2ZrP compound. The results of the electronic band structure analysis indicate that Fe2ZrP is an indirect band gap semiconductor in the spin-down state with the band gap of 0.48 eV. Thermoelectric properties in the temperature range of 300-800 K were calculated. Fe2ZrP exhibits the high Seebeck coefficient of 512 μV K-1 at room temperature along with the huge power factor of 19.21 × 10¹¹ W m-1 K-2 s-1 at 800 K, suggesting Fe2ZrP as a potential thermoelectric material. The Seebeck coefficient decreased with an increase in temperature, and the highest value was obtained for p-type doped Fe2ZrP when the optimum carrier concentration was 0.22 × 1023 cm-3; the n-type doped Fe2ZrP had high electrical conductivity than the p-type doped Fe2ZrP. Thermal conductivity increased with an increase in chemical potential. Optical calculations illustrated that there was a threshold in the imaginary dielectric function for the spin-down channel. Spin-dependent optical calculations showed that the intraband contributions affected only the spin-up optical spectra due to the free-electron effects. Generally, the results confirmed that the intraband contribution had the main role in the optical spectra in the low energy infra-red and visible ranges of light. We also presented the phononic properties and found that these materials were dynamically stable.
Related Concept Videos
 01:31
Ferromagnetism
 01:18
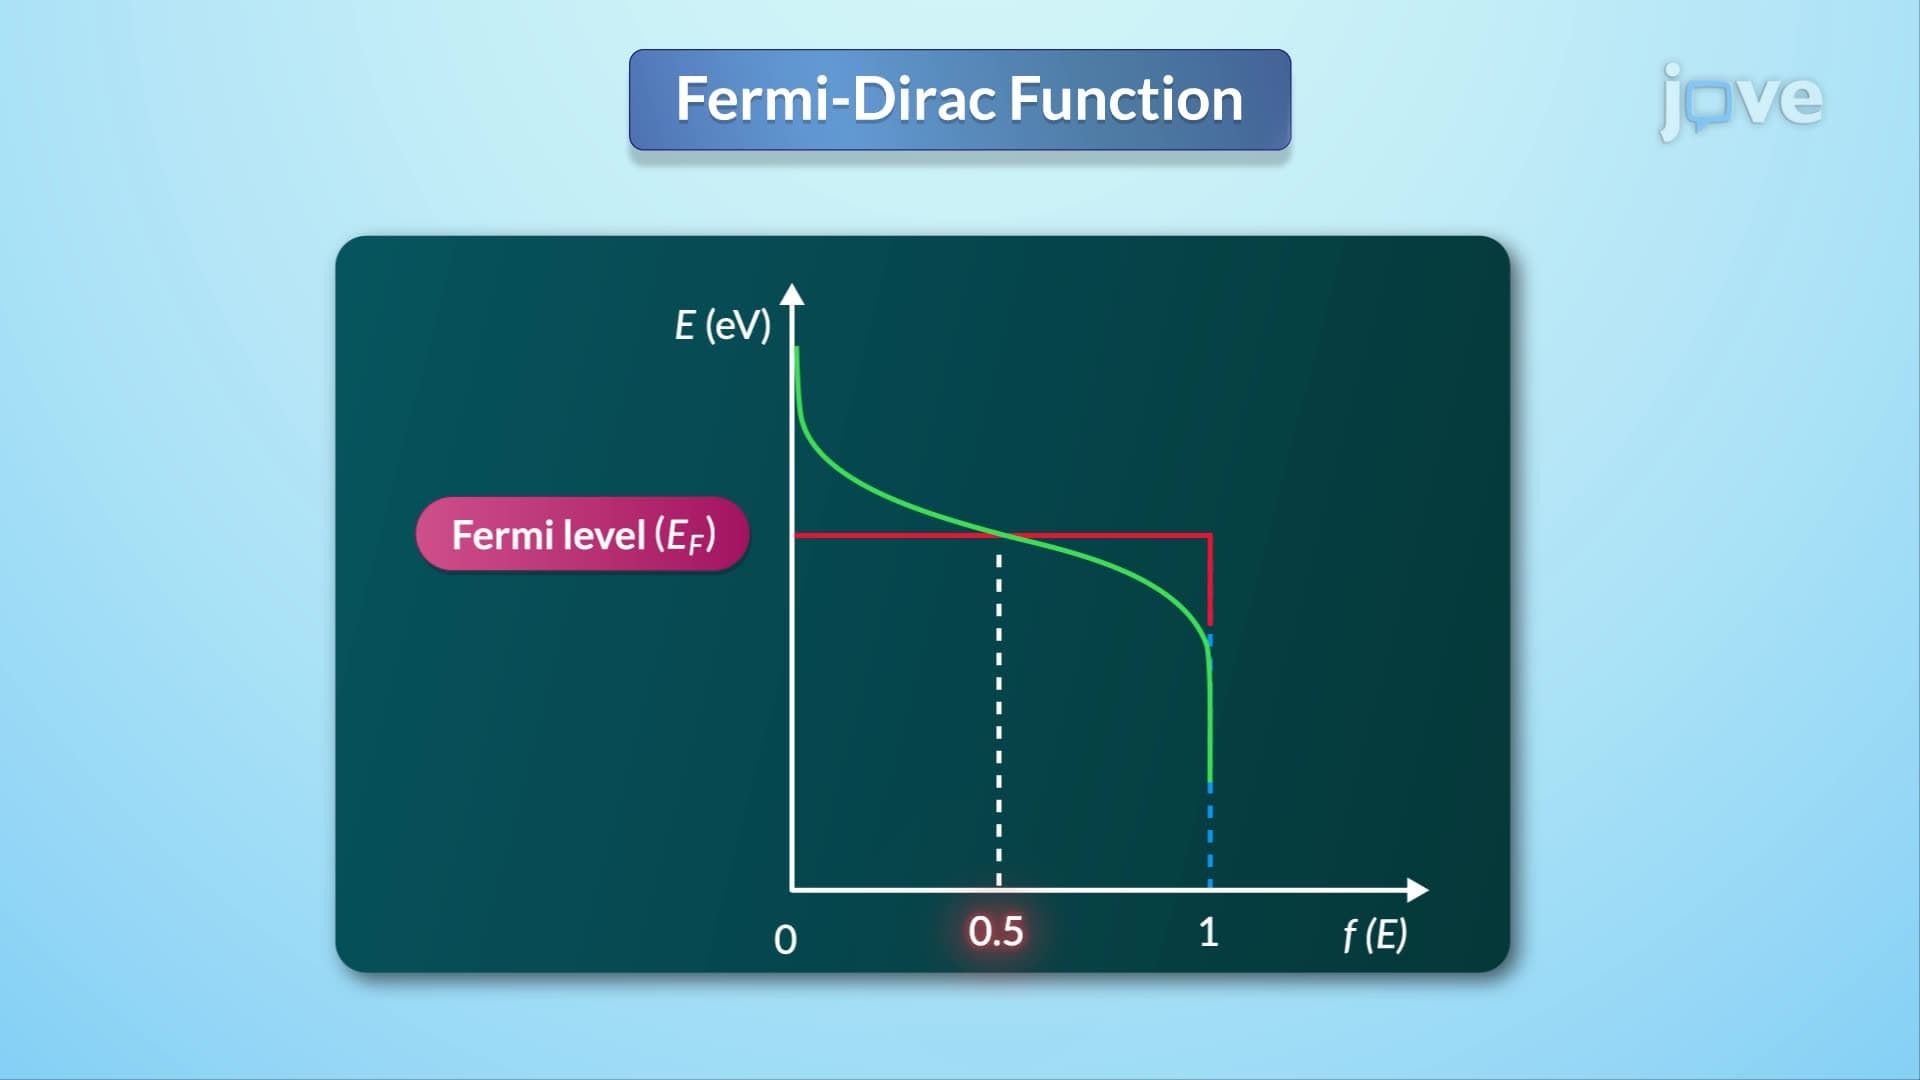
Fermi Level
At absolute zero temperature, electrons fill all energy states up to the Fermi level, leaving upper states empty. As the temperature rises,...
 01:26
Thermodynamic Potentials
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Properties of Transition Metals
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...