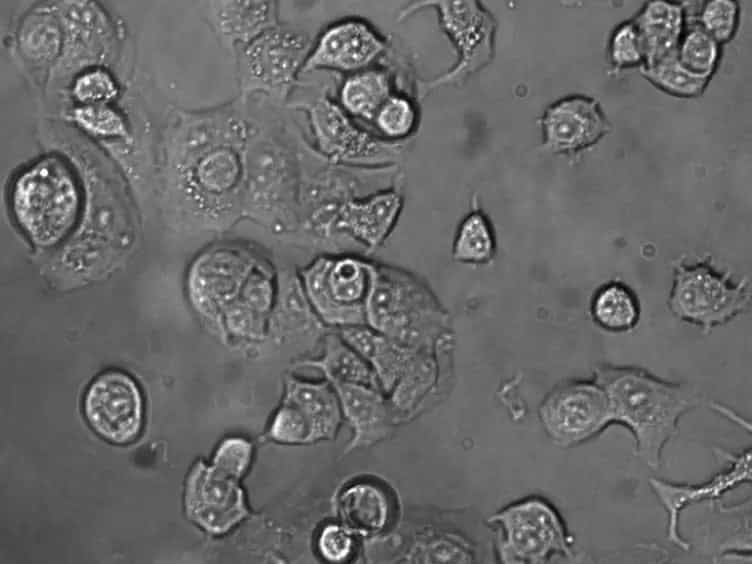

Inhibitory role of TRIP-Br1 oncoprotein in anticancer drug-mediated programmed cell death via mitophagy activation
Samil Jung1, Davaajargal Myagmarjav1, Taeyeon Jo1
1Division of Biological Sciences, Sookmyung Women's University, Seoul, 14310, South Korea.

Abstract:
Chemotherapy has been widely used as a clinical treatment for cancer over the years. However, its effectiveness is limited because of resistance of cancer cells to programmed cell death (PCD) after treatment with anticancer drugs. To elucidate the resistance mechanism, we initially focused on cancer cell-specific mitophagy, an autophagic degradation of damaged mitochondria. This is because mitophagy has been reported to provide cancer cells with high resistance to anticancer drugs. Our data showed that TRIP-Br1 oncoprotein level was greatly increased in the mitochondria of breast cancer cells after treatment with various anticancer drugs including staurosporine (STS), the main focus of this study. STS treatment increased cellular ROS generation in cancer cells, which triggered mitochondrial translocation of TRIP-Br1 from the cytosol via dephosphorylation of TRIP-Br1 by protein phosphatase 2A (PP2A). Up-regulated mitochondrial TRIP-Br1 suppressed cellular ROS levels. In addition, TRIP-Br1 rapidly removed STS-mediated damaged mitochondria by activating mitophagy. It eventually suppressed STS-mediated PCD via degradation of VDACI, TOMM20, and TIMM23 mitochondrial membrane proteins. TRIP-Br1 enhanced mitophagy by increasing expression levels of two crucial lysosomal proteases, cathepsins B and D. In conclusion, TRIP-Br1 can suppress the sensitivity of breast cancer cells to anticancer drugs by activating autophagy/mitophagy, eventually promoting cancer cell survival.
Insights
The oncoprotein TRIP-Br1 enhances breast cancer cell survival by activating mitophagy, a process that removes damaged mitochondria and suppresses programmed cell death (PCD) in response to chemotherapy.
Area of Science:
- Cell Biology
- Molecular Oncology
- Cancer Research
Background:
- Chemotherapy is a cornerstone of cancer treatment, but its efficacy is often compromised by cancer cell resistance.
- Cancer cells resist chemotherapy partly by evading programmed cell death (PCD).
- Mitophagy, the selective degradation of damaged mitochondria, is implicated in conferring drug resistance to cancer cells.
Purpose of the Study:
- To investigate the role of the oncoprotein TRIP-Br1 in chemoresistance.
- To elucidate the mechanism by which cancer cells resist chemotherapy-induced PCD.
- To determine the involvement of mitophagy in TRIP-Br1-mediated chemoresistance.
Main Methods:
- Utilized breast cancer cell models treated with staurosporine (STS) to induce cell death.
- Assessed TRIP-Br1 expression and localization in mitochondria following STS treatment.
- Investigated the impact of TRIP-Br1 on reactive oxygen species (ROS) levels, mitophagy, and mitochondrial protein degradation.
- Measured the expression of lysosomal proteases cathepsins B and D.
Main Results:
- TRIP-Br1 levels increased in mitochondria of breast cancer cells post-chemotherapy.
- STS treatment induced ROS, triggering TRIP-Br1 translocation to mitochondria via dephosphorylation by protein phosphatase 2A (PP2A).
- Mitochondrial TRIP-Br1 suppressed ROS, enhanced mitophagy, and degraded key mitochondrial proteins (VDAC1, TOMM20, TIMM23), inhibiting STS-induced PCD.
- TRIP-Br1 upregulated cathepsins B and D, crucial for lysosomal function in mitophagy.
Conclusions:
- TRIP-Br1 promotes breast cancer cell survival by activating mitophagy and suppressing chemotherapy-induced PCD.
- TRIP-Br1 functions as a key mediator of chemoresistance through mitochondrial quality control.
- Targeting TRIP-Br1 or mitophagy pathways may represent a therapeutic strategy to overcome chemoresistance in breast cancer.
Related Concept Videos
 01:31
The Intrinsic Apoptotic Pathway
 01:17
The Extrinsic Apoptotic Pathway
 02:57
Targeted Cancer Therapies
There are several types of targeted therapies against...
 02:23
Abnormal Proliferation
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 02:34
Inhibition of Cdk Activity