Related Experiment Video
Updated: Sep 2, 2025
Enhancing Tumor Content through Tumor Macrodissection
Published on: February 12, 2022
Hemophagocytic lymphohistiocytosis as an onset of diffuse large B-cell lymphoma: A case report
Yueqing Cao1,2,3, Lang Zou1,2,3, Hao Zhou1,2,3
1Department of Hematology, Xiangya Hospital, Central South University, Changsha, Hunan 410008, P.R. China.

A 53-year-old male presented with a 1-month history of hyperpyrexia. The clinical manifestations revealed hemophagocytic lymphohistiocytosis (HLH). Although a lymph node biopsy could not be obtained, a bone marrow biopsy revealed the activated B-cell subtype of diffuse large B-cell lymphoma (DLBCL). After being treated with HLH-1994 (dexamethasone and etoposide), a rituximab-containing chemotherapy and target agents involving bortezomib, the patient achieved remission. To understand the molecular profile of patient, next-generation sequencing and MYD88 L265P mutation examinations were performed, and the patient was determined to be positive for the MYD88 L265P mutation. Reports of DLBCL with plasmacytic differentiation and a MYD88 innate immune signal transduction adaptor L265P mutation concurrent with HLH are rare. Early recognition, precise diagnosis and timely therapy are pivotal in improving patient prognosis. Furthermore, molecular profiling enables researchers to develop potential therapies aimed at the activated NF-κB and endoplasmic reticulum stress signaling pathways. The present study highlights this pathogenesis and provides suggestions for further individualized therapeutics.
A 53-year-old male presented with a 1-month history of hyperpyrexia. The clinical manifestations revealed hemophagocytic lymphohistiocytosis (HLH). Although a lymph node biopsy could not be obtained, a bone marrow biopsy revealed the activated B-cell subtype of diffuse large B-cell lymphoma (DLBCL). After being treated with HLH-1994 (dexamethasone and etoposide), a rituximab-containing chemotherapy and target agents involving bortezomib, the patient achieved remission. To understand the molecular profile of patient, next-generation sequencing and MYD88 L265P mutation examinations were performed, and the patient was determined to be positive for the MYD88 L265P mutation. Reports of DLBCL with plasmacytic differentiation and a MYD88 innate immune signal transduction adaptor L265P mutation concurrent with HLH are rare. Early recognition, precise diagnosis and timely therapy are pivotal in improving patient prognosis. Furthermore, molecular profiling enables researchers to develop potential therapies aimed at the activated NF-κB and endoplasmic reticulum stress signaling pathways. The present study highlights this pathogenesis and provides suggestions for further individualized therapeutics.
Related Concept Videos
 01:16
Primary Lymphoid Organs
The red bone marrow is a soft, spongy tissue nestled in the interior of long bones such as the humerus and femur. It is the site...
 01:15
Secondary Lymphoid Organs
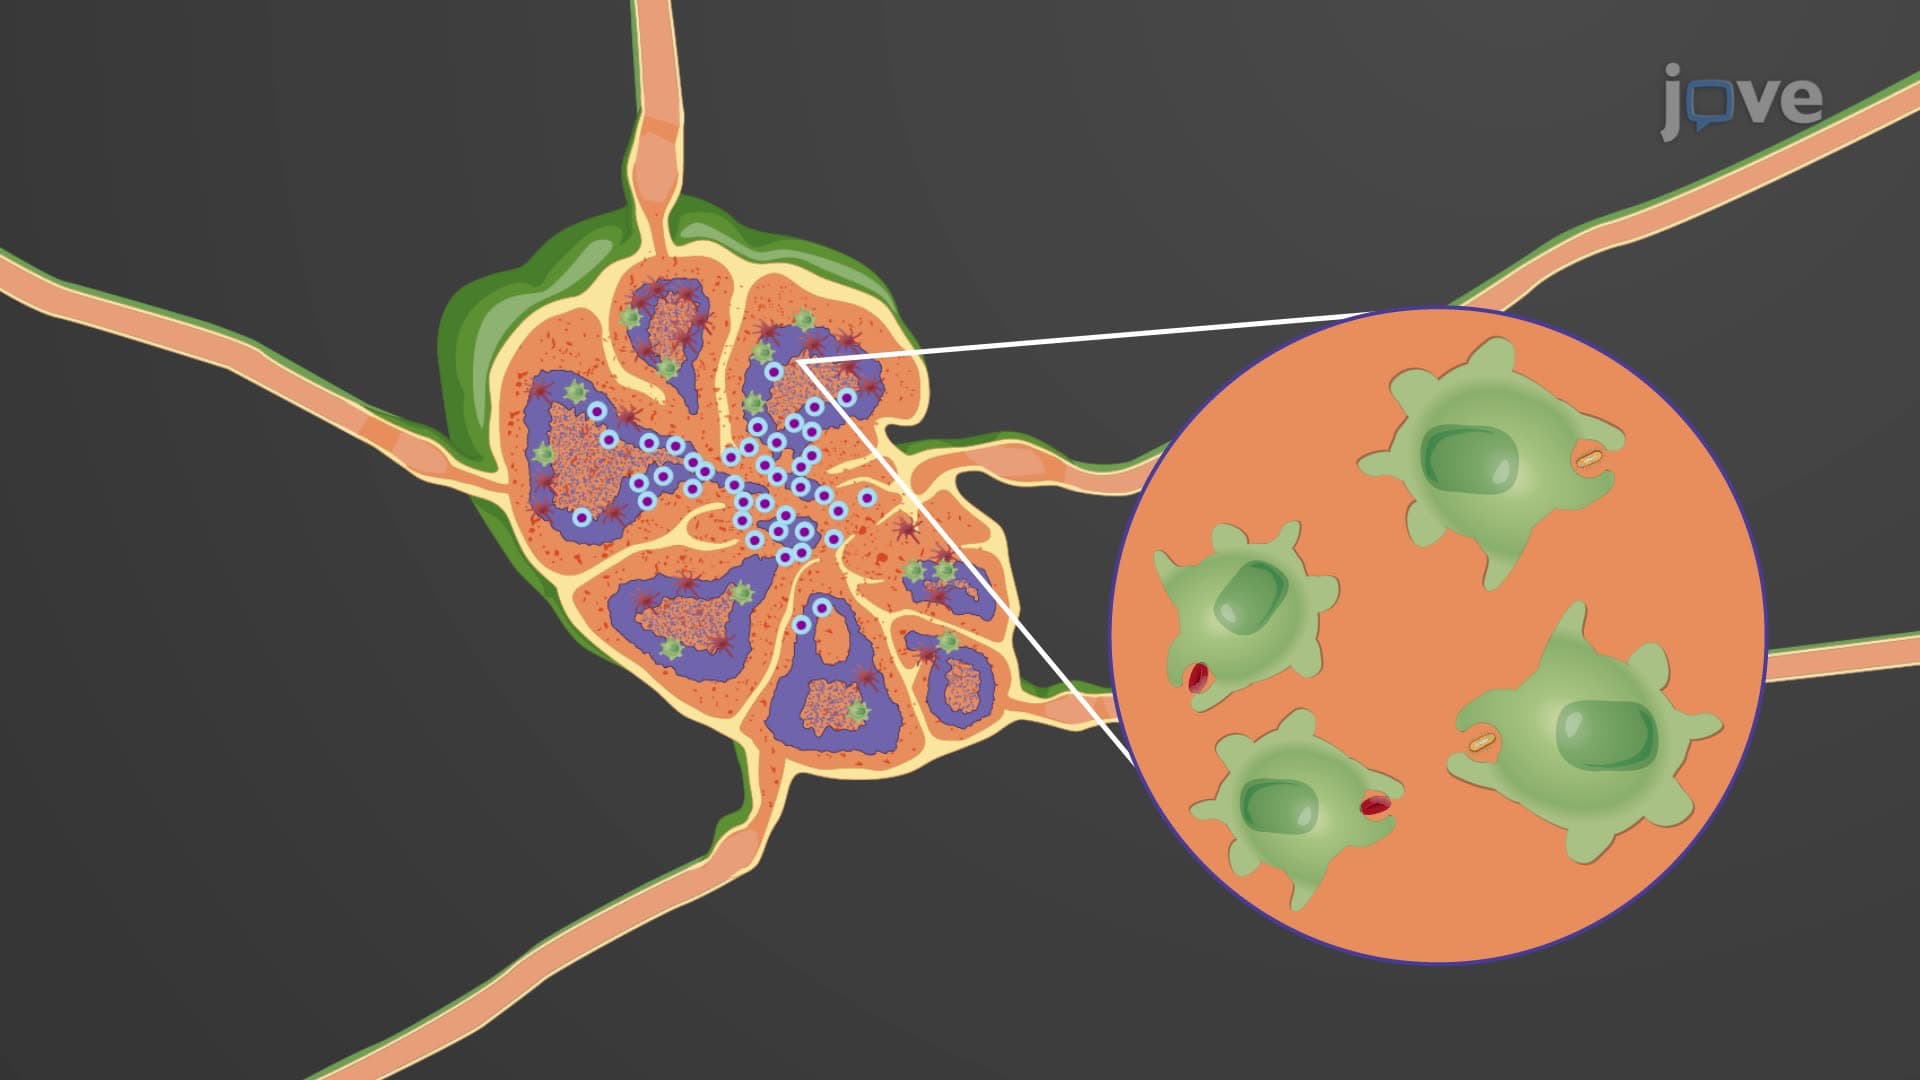
The spleen is a vital organ in the lymphatic system, nestled in the upper left side of the abdomen. It is composed of two primary regions: the red pulp and the white pulp, each having distinct functions. The red pulp performs a significant role in blood filtration. It efficiently purges the blood of old or damaged red blood cells and...