Related Experiment Video
Updated: Aug 29, 2025
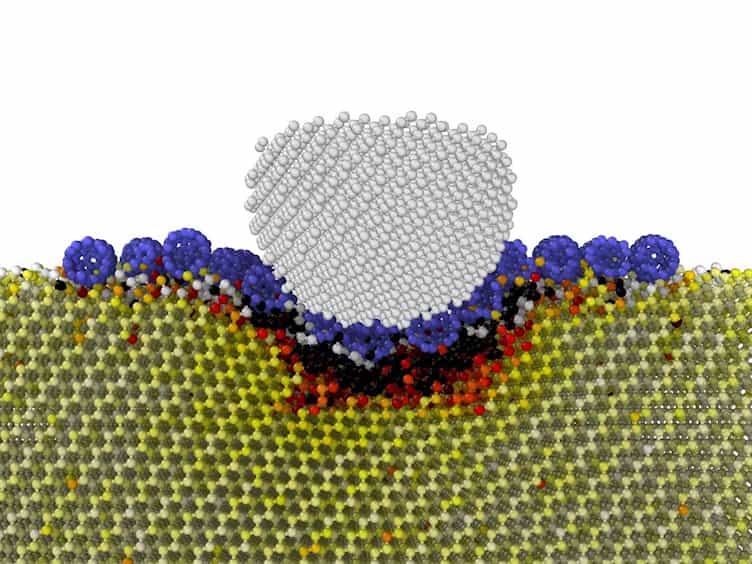

Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Structures and localized vibrational states of defects in graphite by tight-binding calculations
Rong Cheng1,2, Wen-Cai Lu1,2, K M Ho2
1College of Physics, Qingdao University, Qingdao, Shandong 266071, China.

Abstract:
The structural and vibrational properties of pristine graphite and point defects in graphite are studied by tight-binding (TB) calculations using a three-center TB potential model. We showed that the three-center TB potential without "ad hoc" van der Waals interaction corrections can accurately describe the inter-layer distance of graphite and the lowest-energy structures and stabilities of typical point defects in graphite. The results from our TB calculations are in good agreement with those from density-functional theory calculations with van der Waals interaction corrections. We also investigated the vibrational properties to gain better understanding on the localization of vibrational states induced by the point defects. Our calculation results show that although localized or quasi-localized vibrational modes can be found in all defected graphite, the localization induced by Frenkel pair, dual-vacancy, and dual-interstitial defects is much stronger. Atomic displacements associated with the localized vibrational modes induced by these three point defects are also analyzed.
More Related Videos
 11:42
Fabrication of Gate-tunable Graphene Devices for Scanning Tunneling Microscopy Studies with Coulomb Impurities
Published on: July 24, 2015
 06:57
Theoretical Calculation and Experimental Verification for Dislocation Reduction in Germanium Epitaxial Layers with Semicylindrical Voids on Silicon
Published on: July 17, 2020
Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:18
Network Covalent Solids
To break or to melt a covalent network solid, covalent bonds must be broken. Because covalent bonds are relatively strong, covalent network solids are typically...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:22
Structures of Solids
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...