Related Experiment Video
Updated: Aug 25, 2025
Measurement of the Pressure-volume Curve in Mouse Lungs
Published on: January 27, 2015
Variability of clinically measured lung clearance index in children with cystic fibrosis
Bettina S Frauchiger1, Kathryn A Ramsey1, Jakob Usemann2,3
1Department of Pediatrics, Division of Pediatric Respiratory Medicine, Inselspital, Bern University Hospital, University of Bern, Bern, Switzerland.

Rationale:
The lung clearance index (LCI) is increasingly being used in the clinical surveillance of patients with cystic fibrosis (CF). However, there are limited data on long-term variability and physiologically relevant changes in LCI during routine clinical surveillance.
Objectives:
To evaluate the long-term variability of LCI and propose a threshold for a physiologically relevant change.
Methods:
In children aged 4-18 years with CF, LCI was measured every 3 months as part of routine clinical surveillance during 2011-2020 in two centers. The variability of LCI during periods of clinical stability was assessed using mixed-effects models and was used to identify thresholds for physiologically relevant changes.
Results:
Repeated LCI measurements of acceptable quality (N = 858) were available in 100 patients with CF; for 74 patients, 399 visits at clinical stability were available. The variability of repeated LCI measurements over time expressed as the coefficient of variation (CV%) was 7.4%. The upper limit of normal (ULN) for relative changes in LCI between visits was 19%.
Conclusion:
We report the variability of LCI in children and adolescents with CF during routine clinical surveillance. According to our data, a change in LCI beyond 19% may be considered physiologically relevant. These findings will help guide clinical decisions according to LCI changes.
Related Concept Videos
 00:56
One-Compartment Open Model for IV Bolus Administration: Estimation of Clearance
In the one-compartment open model for intravenous (IV) bolus administration, clearance is estimated by dividing the elimination rate by the plasma drug concentration. This equation leverages the elimination rate constant and the apparent...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:24
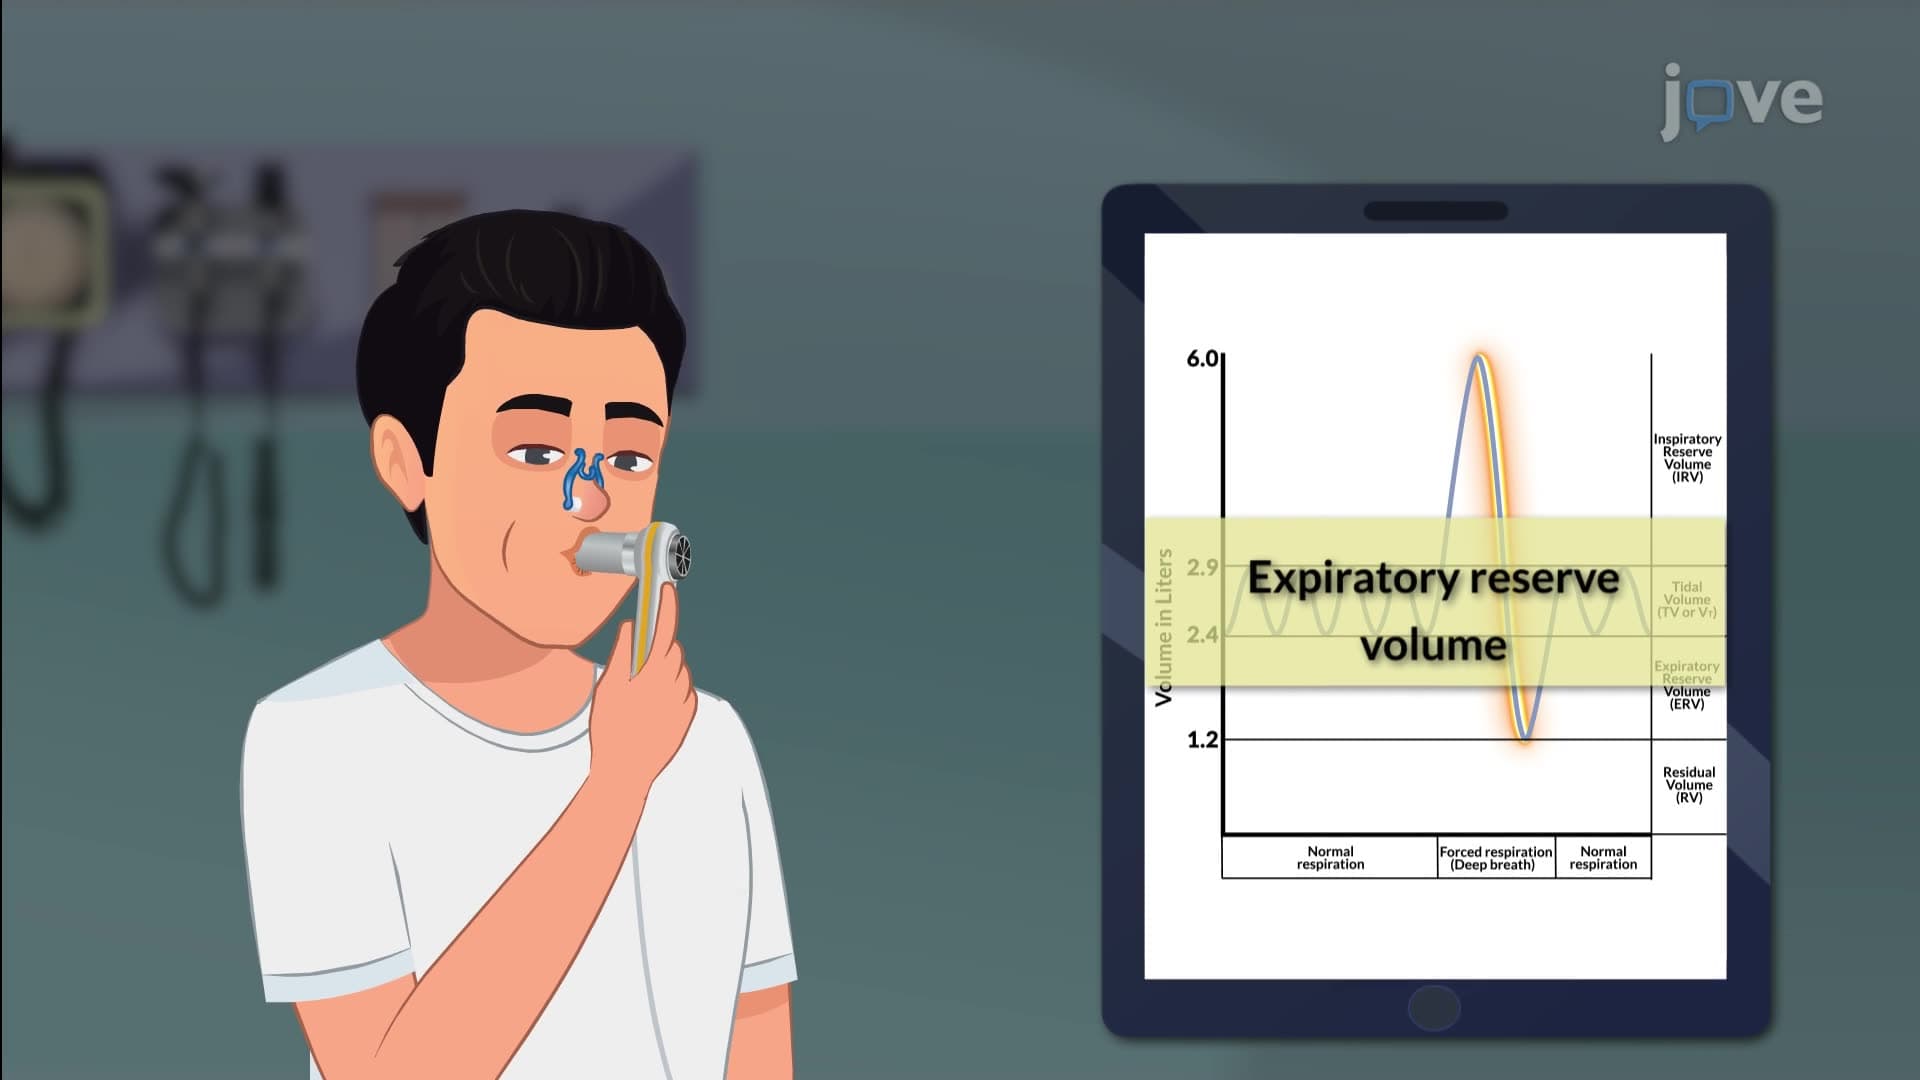
Respiratory Capacities
One key metric is the Inspiratory Capacity (IC), which represents the maximum amount of air that can be inhaled with full effort. IC is calculated by summing the tidal volume and inspiratory reserve volume, typically ranging from 2.4 to 3.6 liters.
The Functional Residual Capacity (FRC) represents the air in the...
 01:15
Respiratory Volumes
Tidal Volume (TV) Tidal volume (TV) is the air inhaled or exhaled in a...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
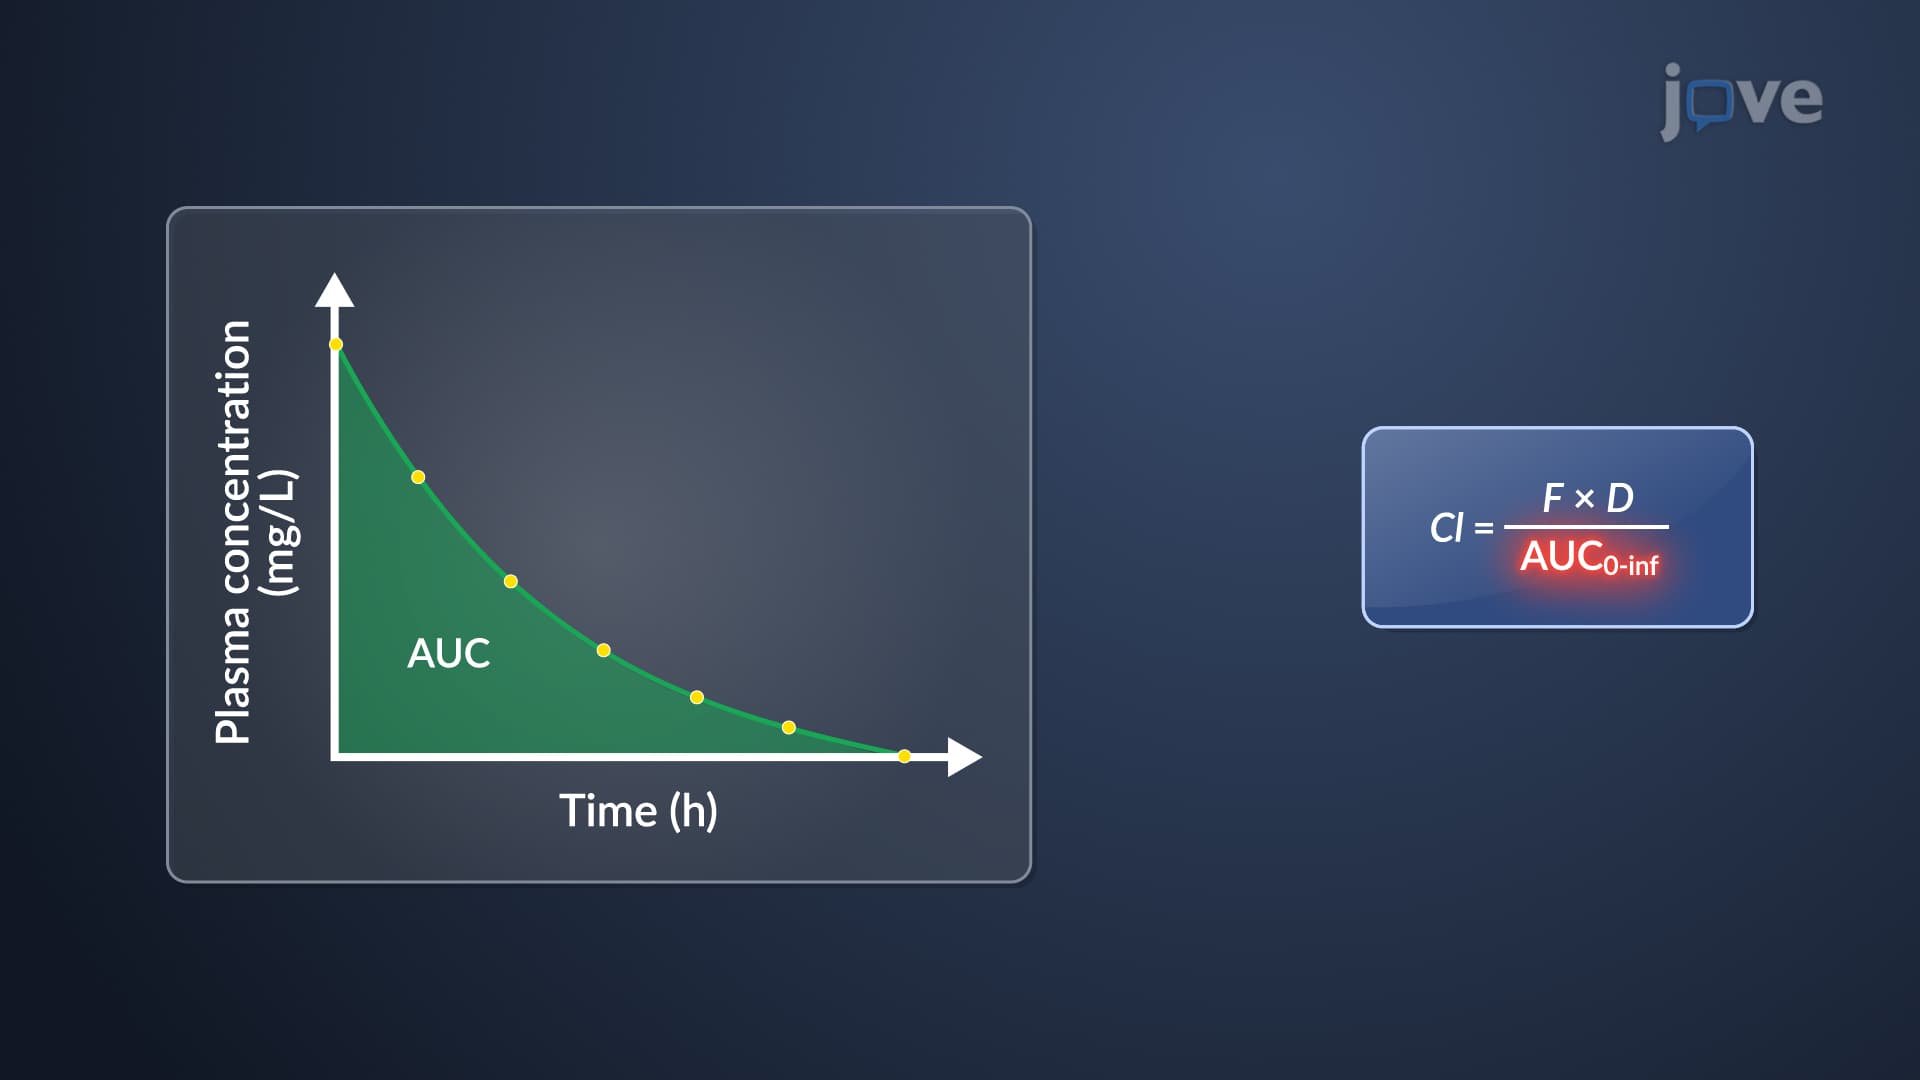 01:17
Clearance Models: Noncompartmental Models
The noncompartmental approach capitalizes on extensive sampling data, correlating the volume of distribution to systemic exposure and the administered dosage. This method enables...