Related Experiment Video
Updated: Aug 24, 2025
Photoelectron Imaging of Anions Illustrated by 310 Nm Detachment of F−
Published on: July 27, 2018
Valence molecular orbitals and cationic structures of 2-fluoropyridine by high-resolution ion spectroscopy and
1Forensic Chemistry Division, National Forensic Service, Wonju 26460, Republic of Korea.

Abstract:
The alteration of the valence molecular orbitals' ordering of halopyridine molecules, by the introduction of a halogen atom(s) as substituent on the pyridine ring, has spurred an extensive interest for their investigation. Herein, the effect of a fluorine substituent on the two outermost orbitals of pyridine was elucidated by investigating the photoionization dynamics of 2-fluoropyridine (2-FP), considering that the geometrical changes with respect to the neutral geometry induced by adiabatic ionic transition affect the vacuum ultraviolet mass-analyzed threshold ionization (VUV-MATI) spectrum. The adiabatic ionization energy associated with the 0-0 band on the measured high-resolution VUV-MATI spectrum was determined to be 9.6702 ± 0.0004 eV (77 995 ± 3 cm-1), which differs considerably from the 9.401 eV by two-color ionization spectroscopy. Franck-Condon simulation of the MATI spectrum corresponded quantitatively with the experimental results. Interestingly, among the forbidden transitions under CS symmetry, an out-of-plane ring-bending mode resulting from the warped cationic structure of 2-FP with C1 symmetry was discovered. Rigorously, among the unassigned peaks, the first prominent peak at 78 532 cm-1 should rather be assigned as the origin of the excited electronic state (D1) of the 2-FP cation, in accordance with time-dependent density functional theory calculations. Natural bond orbital analysis led to the conclusion that such observations could be induced by electron removal from the highest occupied molecular orbital (HOMO) consisting of the π orbital of the pyridine ring and lone-pair orbital of the fluorine atom or from the HOMO-1 of the molecular non-bonding orbitals, to generate the two proximate electronic states of the cation.
More Related Videos
 14:11
Synthesis of pH Dependent Pyrazole, Imidazole, and Isoindolone Dipyrrinone Fluorophores using a Claisen-Schmidt Condensation Approach
Published on: June 10, 2021
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:42
Valence Bond Theory
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 02:27
Predicting Molecular Geometry
 01:18
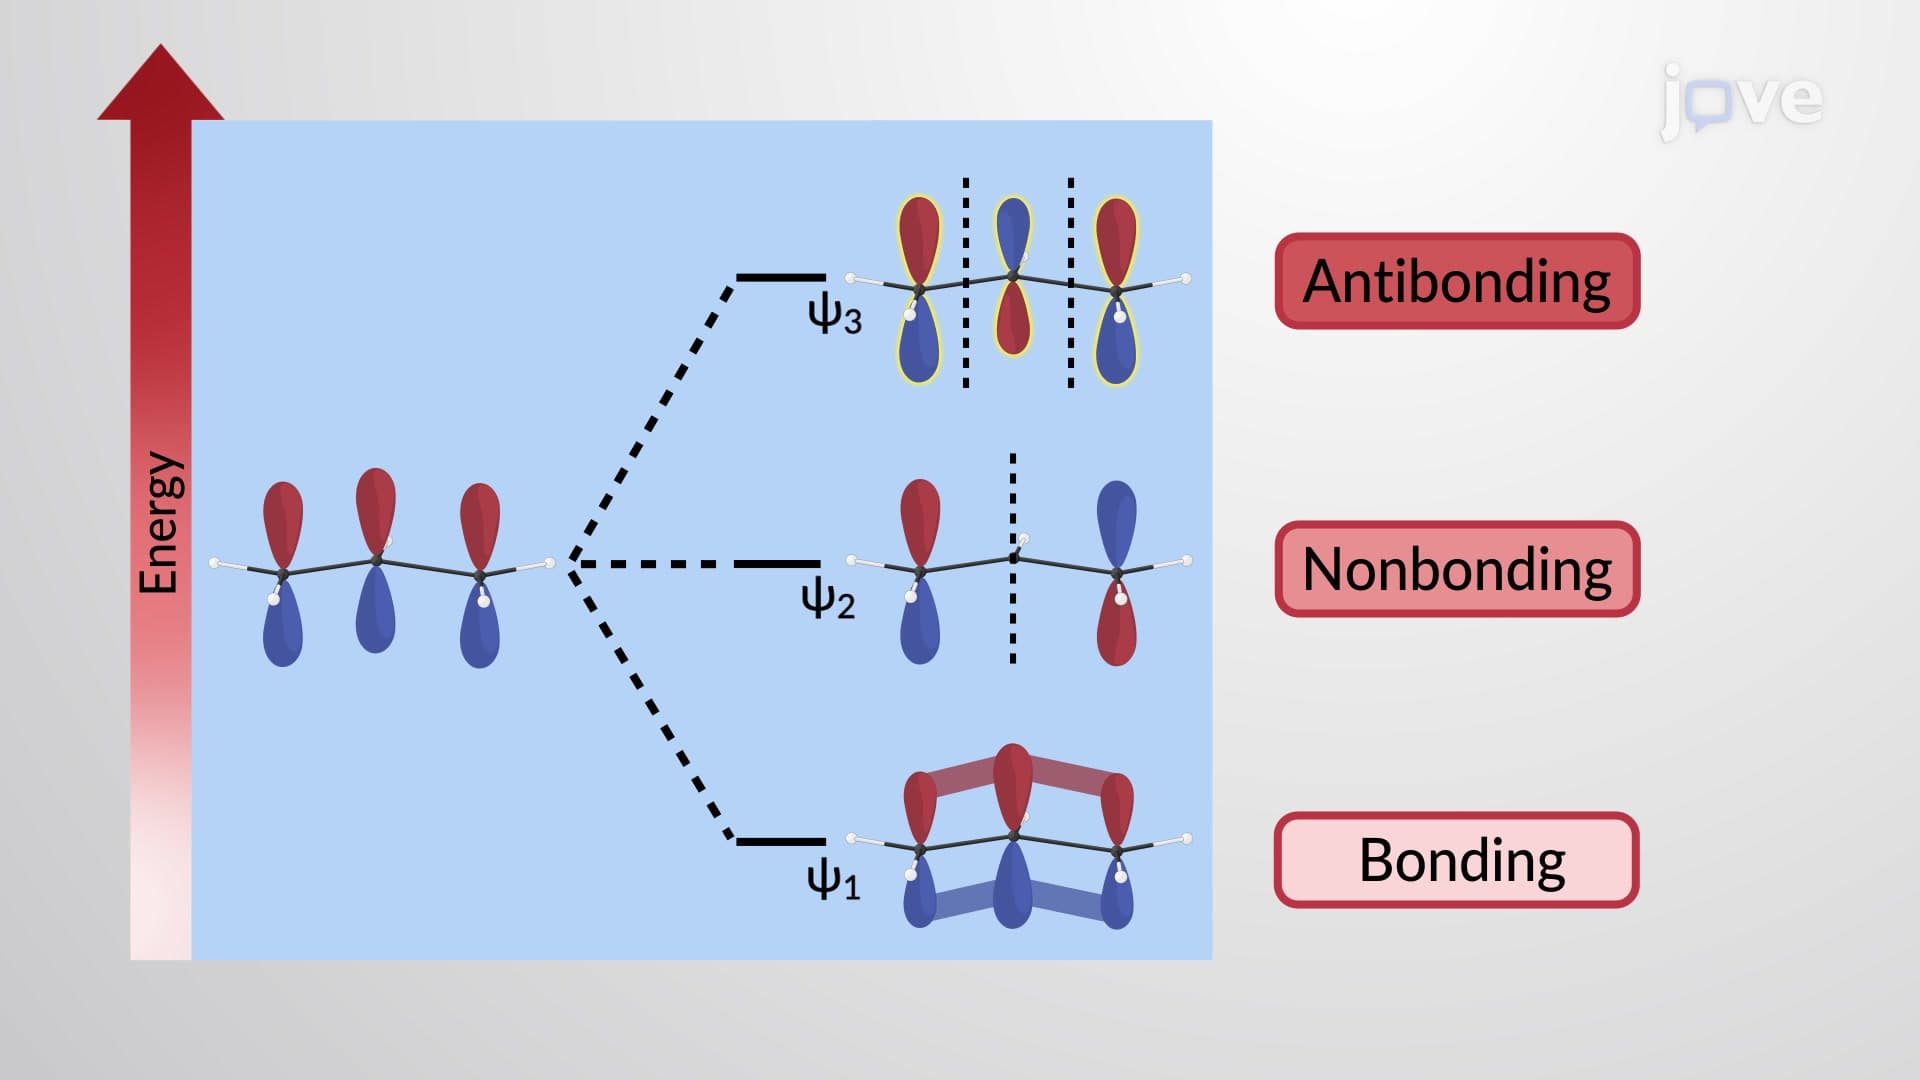
π Molecular Orbitals of the Allyl Cation and Anion
 03:35
Hybridization of Atomic Orbitals II