Related Experiment Video
Updated: Aug 14, 2025
Engineering Antiviral Agents via Surface Plasmon Resonance
Published on: June 14, 2022
Insight into free energy and dynamic cross-correlations of residue for binding affinity of antibody and receptor
Wei Lim Chong1,2,3, Patchareenart Saparpakorn1,2, Chak Sangma1,2
1Department of Chemistry, Faculty of Science, Kasetsart University, Bangkok, 10900, Thailand.

Abstract:
SARS-CoV-2 virus continues to evolve and mutate causing most of the mutated variants resist to many of the therapeutic monoclonal antibodies (mAbs). Despite several mAbs retained neutralizing capability for Omicron BA.1 and BA.2, reduction in neutralization potency was reported. Hence, effort of searching for mAb that is broader in neutralization breadth without losing the neutralizing ability is continued. MW06 was reported with capability in neutralizing most of the variants of concern (VOC) and it binds to the conserved region (left flank) near epitope mAb sotrovimab (S309). In this study, binding affinity of mAb MW06 and its cocktail formulation with MW05 for receptor binding domain (RBD) SARS-CoV-2 virus was investigated under molecular dynamics simulations (MDs). Binding free energies computed by Molecular Mechanics Generalised Born Surface Area (MM-GBSA) algorithm predicted the binding affinity of MW06 for RBD BA.1 (-53 kcal/mol) as strong as RBD wildtype (-58 kcal/mol) while deterioration was observed for RBD BA.2 (-43 kcal/mol). Alike S309 and MW06, simulated cocktail mAb (MW05 and MW06)-RBD interactions suggested the neutralizing capability of the cocktail formulation for RBD BA.1 and BA.2 reduced. Meanwhile, residue pairs that favour the communication between the mAb and RBD have been identified by decomposing the free energy per pairwise residue basis. Apart from understanding the effects of mutation occurred in the RBD region on human angiotensin-converting enzyme 2 (hACE2) binding, impact of heavily mutated RBD on mAb-RBD interactions was investigated in this study as well. In addition to energetic profile obtained from MDs, plotting the dynamics cross-correlation map of the mAb-RBD complex under elastic network model (ENM) was aimed to understand the cross-correlations between residue fluctuations. It allows simple and rapid analysis on the motions or dynamics of the protein residues of mAbs and RBD in complex. Protein residues having correlated motions are normally part of the structural domains of the protein and their respective motions and protein function are related. Motion of mutated RBD residues and mAb residues was less correlated while their respective interactions energy computed to be higher. The combined techniques of MDs and ENM offered simplicity in understanding dynamics and energy contribution that explain binding affinity of mAb-RBD complexes.
Related Concept Videos
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:16
Protein-Drug Binding: Mechanism and Kinetics
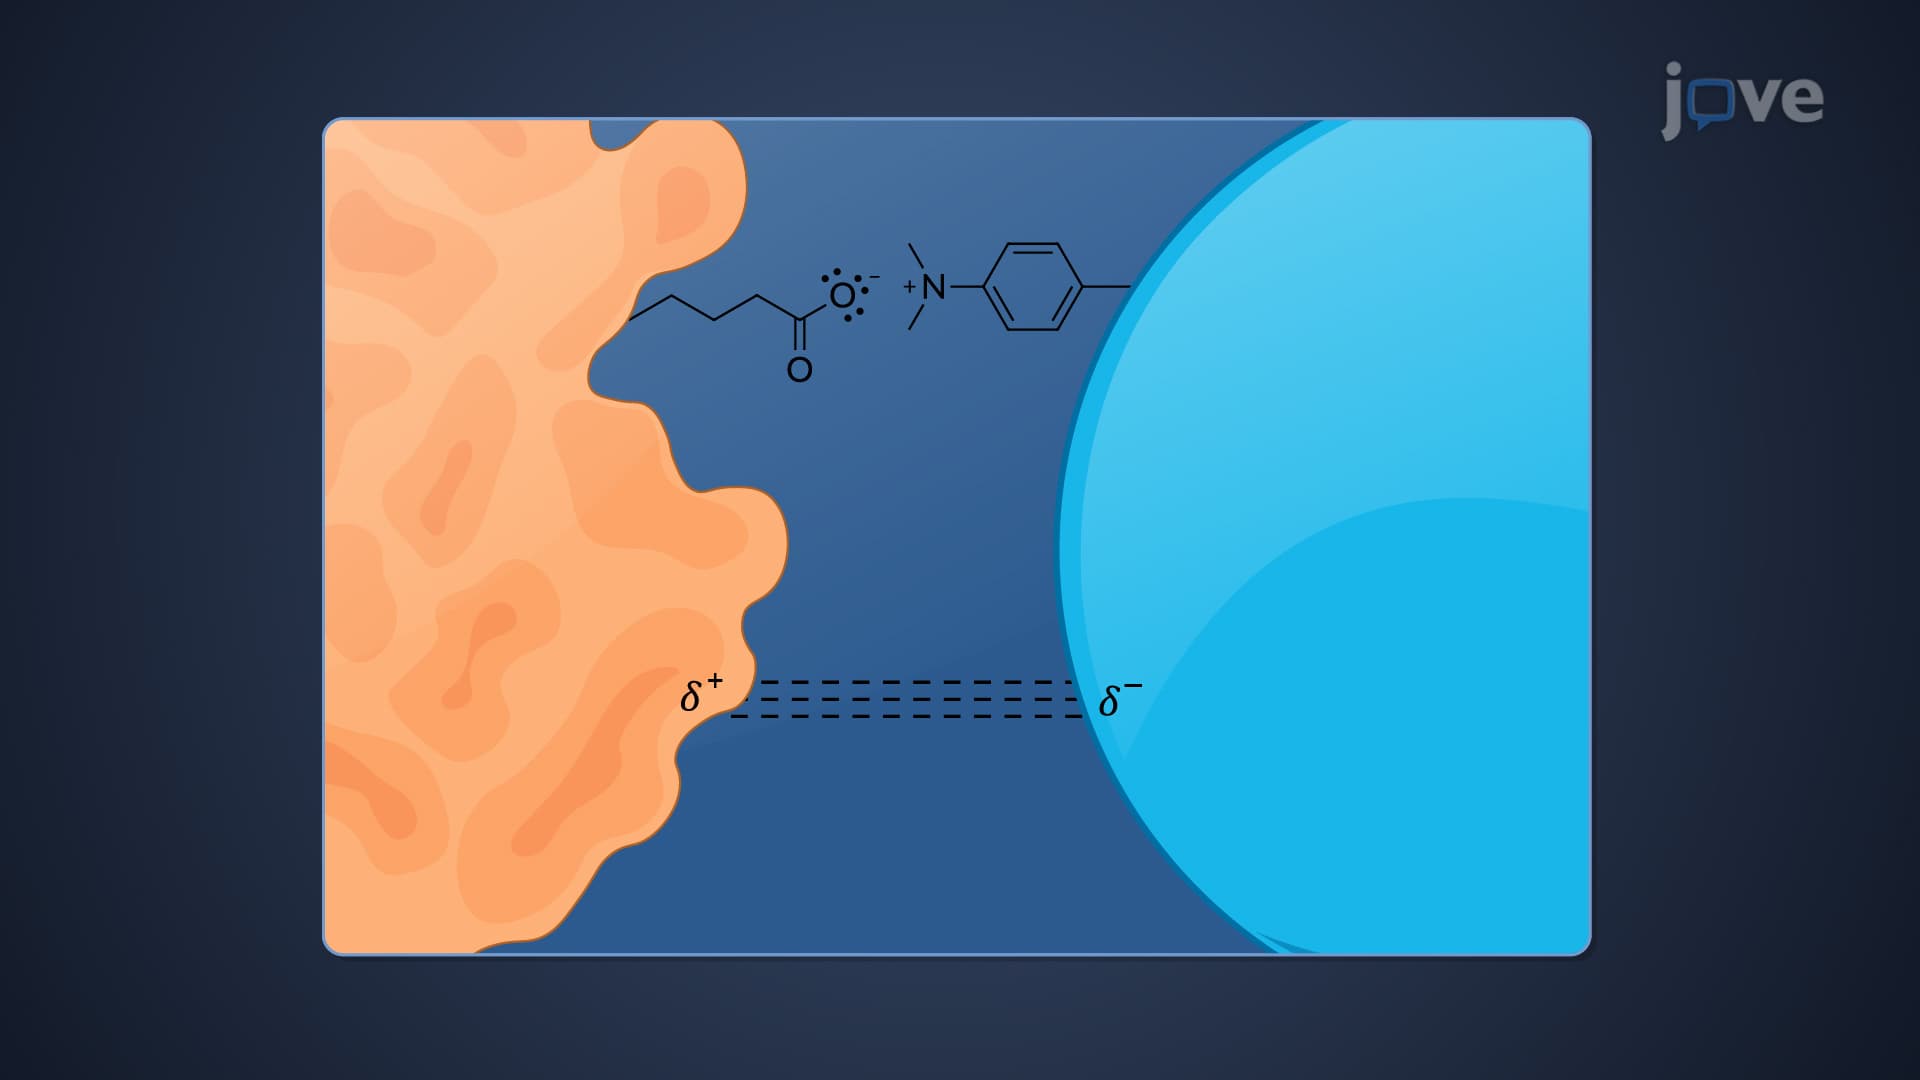
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...