Related Experiment Video
Updated: Aug 12, 2025

Nuclear Magnetic Resonance Spectroscopy for the Identification of Multiple Phosphorylations of Intrinsically Disordered Proteins
Published on: December 27, 2016
Improving the accuracy of 31P NMR chemical shift calculations by use of scaling methods
William H Hersh1,2, Tsz-Yeung Chan1,2
1Department of Chemistry and Biochemistry, Queens College, Queens, NY 11367-1597, USA.

Abstract:
Calculation of 31P NMR chemical shifts for a series of tri- and tetracoordinate phosphorus compounds using several basis sets and density functional theory (DFT) functionals gave a modest fit to experimental chemical shifts, but an excellent linear fit when plotted against the experimental values. The resultant scaling methods were then applied to a variety of "large" compounds previously selected by Latypov et al. and a set of stereoisomeric and unusual compounds selected here. No one method was best for all structural types. For compounds that contain P-P bonds and P-C multiple bonds, the Latypov et al. method using the PBE0 functional was best (mean absolute deviation/root mean square deviation (MAD/RMSD) = 6.9/8.5 ppm and 6.6/8.2 ppm, respectively), but for the full set of compounds gave higher deviations (MAD/RMSD = 8.2/12.3 ppm), and failed by over 60 ppm for a three-membered phosphorus heterocycle. Use of the M06-2X functional for both the structural optimization and NMR chemical shift calculation was best overall for the compounds without P-C multiple bonds (MAD/RMSD = 5.4/7.1 ppm), but failed by 30-49 ppm for compounds having any P-C multiple-bond character. Failures of these magnitudes have not been reported previously for these widely used functionals. These failures were then used to screen a variety of recommended functionals, leading to better overall methods for calculation of these chemical shifts: optimization with the M06-2X functional and NMR calculation with the PBE0 or ωB97x-D functionals gave values for MAD/RMSD = 6.9/8.5 ppm and 6.8/9.1 ppm, respectively, over an experimental chemical shift range of -181 to 356 ppm. Due to the unexplained failures observed, we recommend use of more than one method when looking at novel structures.
More Related Videos
 10:52
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
 09:32
A New Straightforward Method for Lipophilicity logP Measurement using 19F NMR Spectroscopy
Published on: January 30, 2019
Related Concept Videos
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 01:15
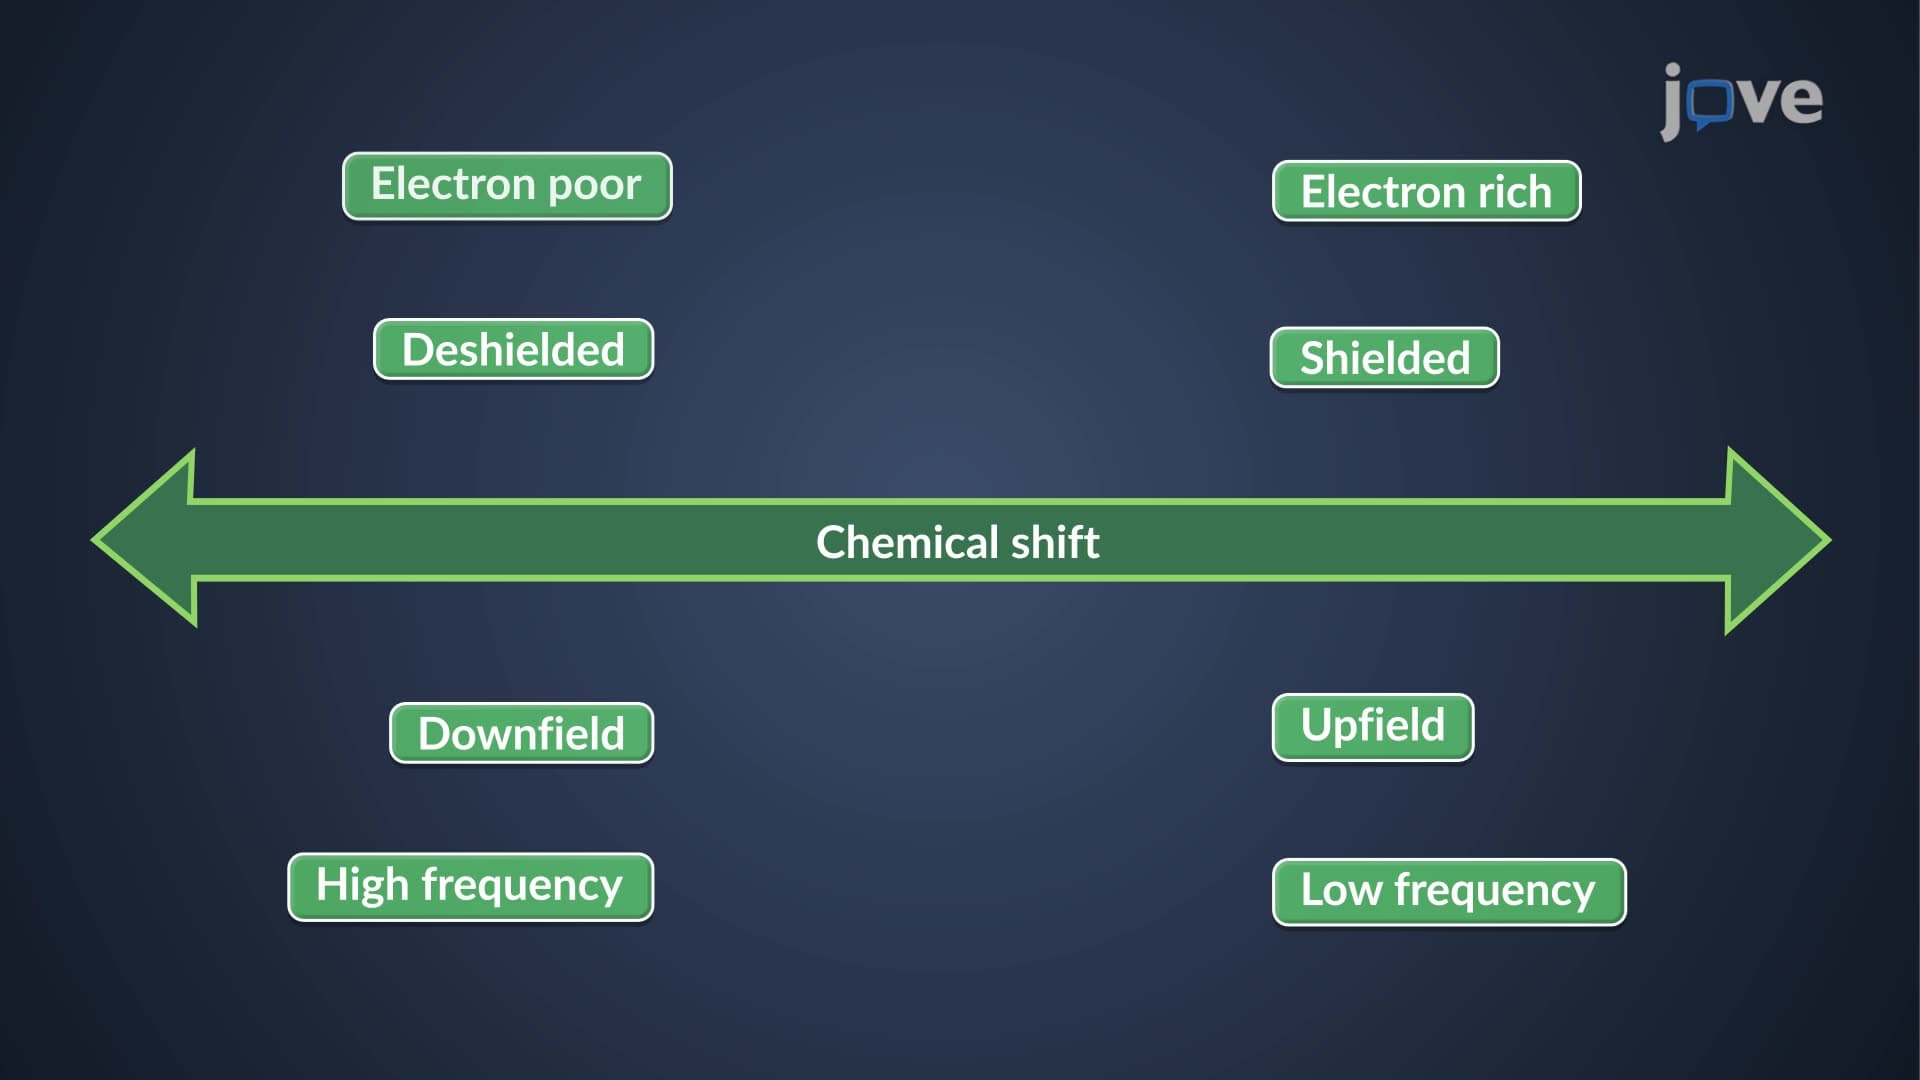
NMR Spectroscopy: Chemical Shift Overview
For instance, the proton...
 01:16
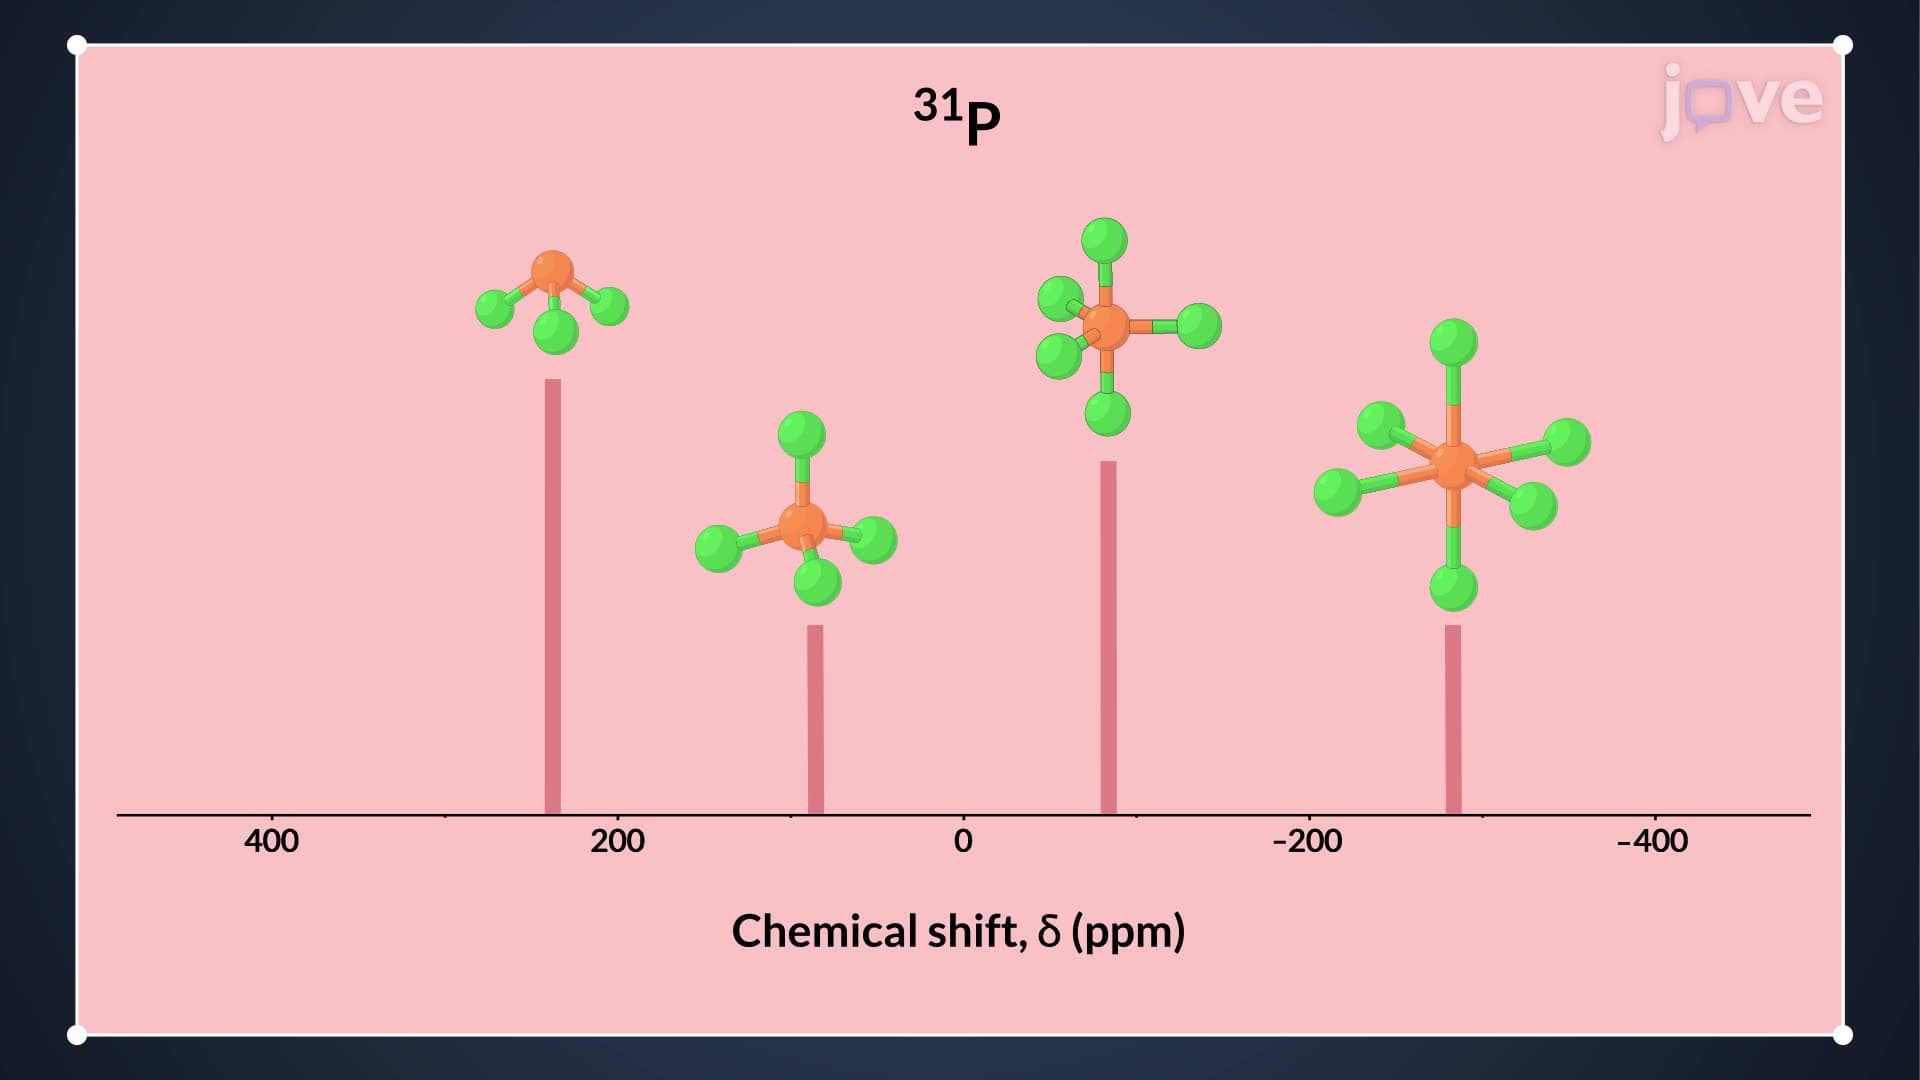
Other Nuclides: 31P, 19F, 15N NMR
While fluorine-19 and phosphorous-31 have high natural abundances (100%) and positive gyromagnetic ratios, nitrogen-15 has a low natural abundance and a negative gyromagnetic ratio. However, nitrogen-15 is still preferred over nitrogen-14 (which has a...
 01:14
NMR Spectrometers: Resolution and Error Correction
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 00:58
¹H NMR Chemical Shift Equivalence: Enantiotopic and Diastereotopic Protons
In chiral compounds such as 2-butanol, replacing the methylene hydrogens at C3 produces a pair of...