Related Experiment Video
Updated: Aug 10, 2025

A Virtual Machine Platform for Non-Computer Professionals for Using Deep Learning to Classify Biological Sequences of Metagenomic Data
Published on: September 25, 2021
Characterization of Klebsiella pneumoniae bacteriophages, KP1 and KP12, with deep learning-based structure prediction
Youngju Kim1,2, Sang-Mok Lee3, Linh Khanh Nong3
1Optipharm Inc., Cheongju-si, Chungcheongbuk-do, Republic of Korea.

Abstract:
Concerns over Klebsiella pneumoniae resistance to the last-line antibiotic treatment have prompted a reconsideration of bacteriophage therapy in public health. Biotechnological application of phages and their gene products as an alternative to antibiotics necessitates the understanding of their genomic context. This study sequenced, annotated, characterized, and compared two Klebsiella phages, KP1 and KP12. Physiological validations identified KP1 and KP12 as members of Myoviridae family. Both phages showed that their activities were stable in a wide range of pH and temperature. They exhibit a host specificity toward K. pneumoniae with a broad intraspecies host range. General features of genome size, coding density, percentage GC content, and phylogenetic analyses revealed that these bacteriophages are distantly related. Phage lytic proteins (endolysin, anti-/holin, spanin) identified by the local alignment against different databases, were subjected to further bioinformatic analyses including three-dimensional (3D) structure prediction by AlphaFold. AlphaFold models of phage lysis proteins were consistent with the published X-ray crystal structures, suggesting the presence of T4-like and P1/P2-like bacteriophage lysis proteins in KP1 and KP12, respectively. By providing the primary sequence information, this study contributes novel bacteriophages for research and development pipelines of phage therapy that ultimately, cater to the unmet clinical and industrial needs against K. pneumoniae pathogens.
Insights
Novel bacteriophages KP1 and KP12 targeting Klebsiella pneumoniae were characterized. Their genomic context and lytic proteins were analyzed, offering new tools for phage therapy development against antibiotic-resistant bacteria.
Area of Science:
- Microbiology and Virology
- Biotechnology and Bioinformatics
Background:
- Rising antibiotic resistance in Klebsiella pneumoniae necessitates alternative treatments.
- Bacteriophage therapy is a promising alternative, requiring detailed phage genomic understanding.
Purpose of the Study:
- To sequence, annotate, characterize, and compare two Klebsiella phages, KP1 and KP12.
- To analyze the genomic context and lytic proteins of these novel phages for therapeutic applications.
Main Methods:
- Genomic sequencing and annotation of KP1 and KP12.
- Physiological stability and host specificity assays.
- Bioinformatic analysis of lytic proteins, including 3D structure prediction using AlphaFold.
Main Results:
- KP1 and KP12 are Myoviridae family members, stable across a wide pH/temperature range.
- Both phages exhibit K. pneumoniae specificity with broad intraspecies host range.
- Phylogenetic analysis shows distant relation; AlphaFold models confirm T4-like and P1/P2-like lysis proteins.
Conclusions:
- KP1 and KP12 represent novel bacteriophages with potential for phage therapy development.
- This study provides essential genomic data for K. pneumoniae phage therapy research and development.
Related Concept Videos
 01:26
DNA Bacteriophages
 01:30
Lytic Cycle of Bacteriophages
 00:43
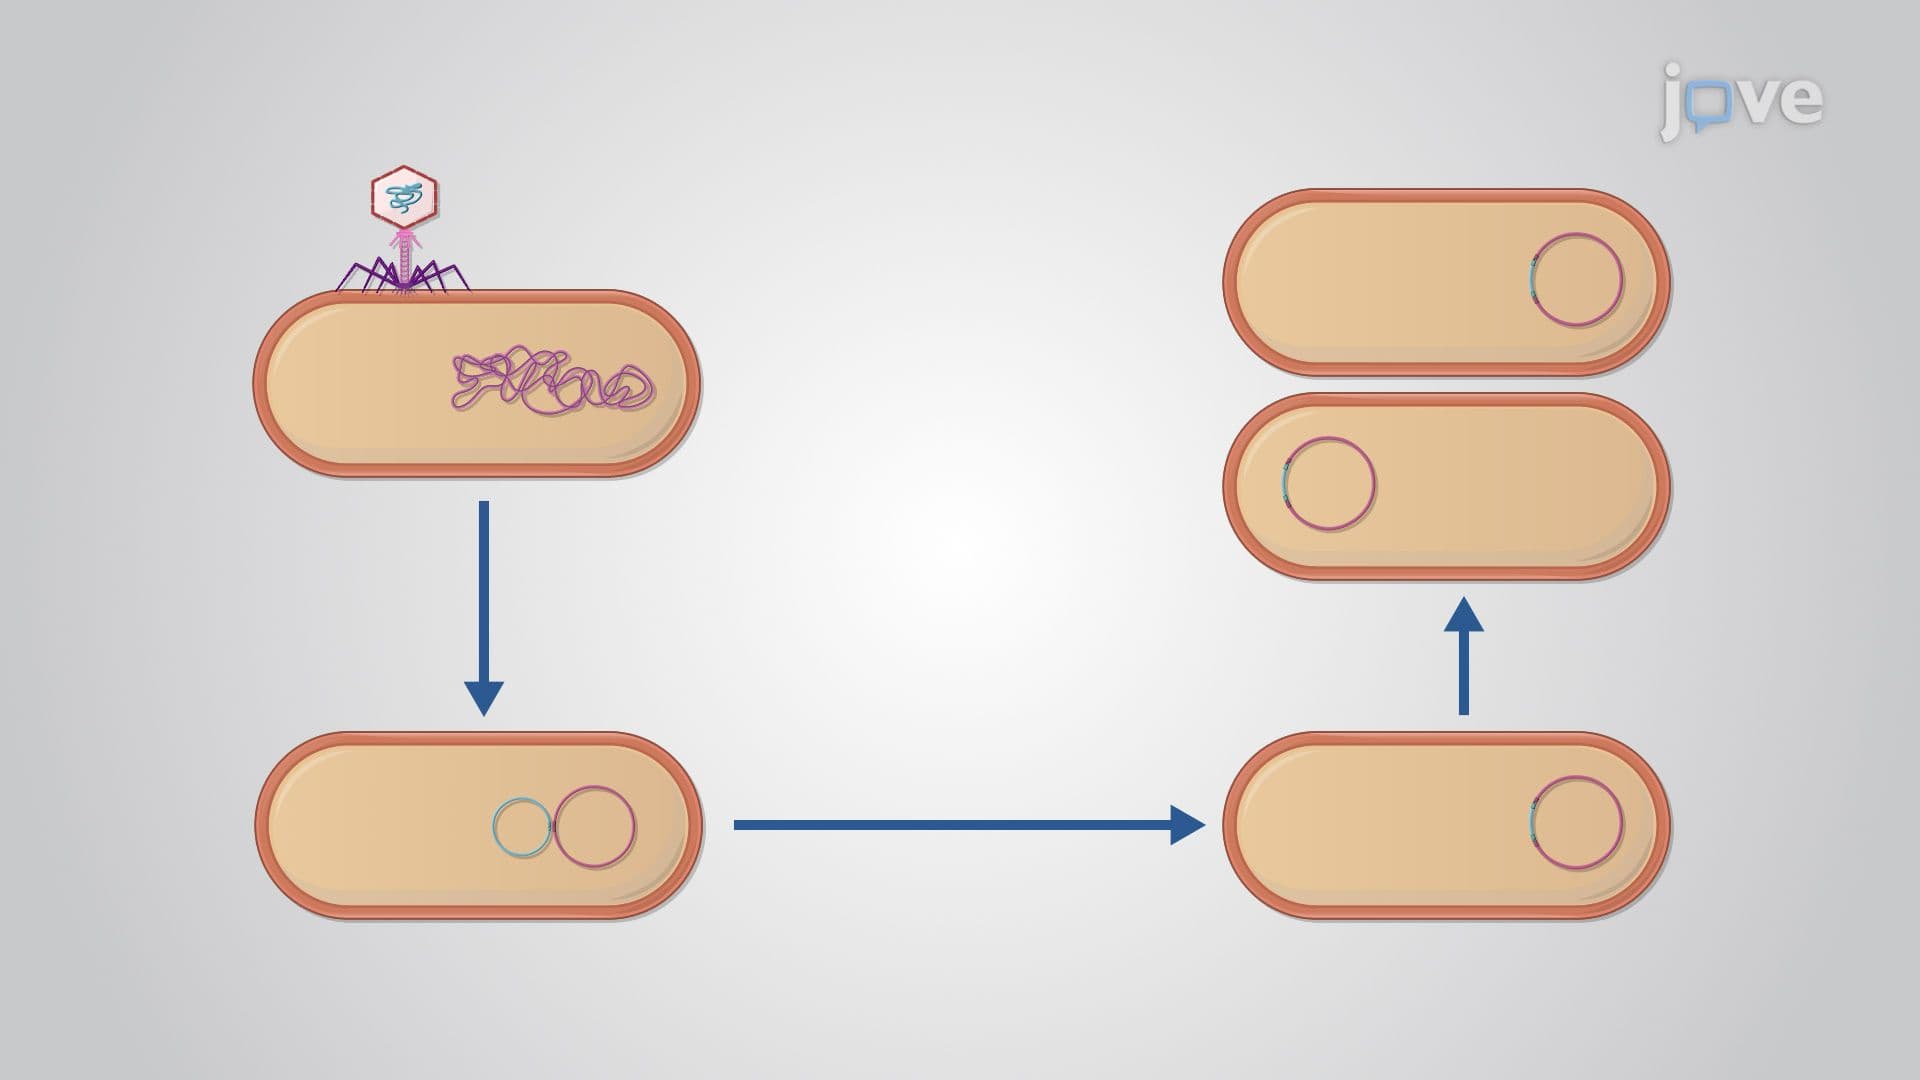
Lysogenic Cycle of Bacteriophages
 00:56
Viral Structure
 01:26
Size and Structure of Viral Genomes
 01:16
Viral Replication: Lysogenic Cycle