Related Experiment Video
Updated: Aug 9, 2025

Using Cyclic Voltammetry, UV-Vis-NIR, and EPR Spectroelectrochemistry to Analyze Organic Compounds
Published on: October 18, 2018
Evaluation of tight-binding DFT performance for the description of organic photochromes properties
Corentin Poidevin1, Gwenhaël Duplaix-Rata1, Karine Costuas1
1Univ Rennes, CNRS, ISCR (Institut des Sciences Chimiques de Rennes) - UMR 6226, F-35000 Rennes, France.

Abstract:
Photochromic molecules are widely studied and developed for their many potential applications. To optimize the required properties through theoretical models, a considerable chemical space is to be explored, and their environment in devices is to be accounted for.. To this end, cheap and reliable computational methods can be powerful tools to steer synthetic developments. As ab initio methods remain costly for extensive studies (in terms of the size of the system and/or number of molecules), semiempirical methods such as density functional tight-binding (TB) could offer a good compromise between accuracy computational cost. However, these approaches necessitate benchmarking on the families of compounds of interest. Thus, the aim of the present study is to evaluate the accuracy of several key features calculated with TB methods (DFTB2, DFTB3, GFN2-xTB, and LC-DFTB2) for three sets of photochromic organic molecules: azobenzene (AZO), norbornadiene/quadricyclane (NBD/QC), and dithienylethene (DTE) derivatives. The features considered here are the optimized geometries, the difference in energy between the two isomers (ΔE), and of the energies of the first relevant excited states. All the TB results are compared to those obtained with DFT methods and state-of-the-art electronic structure calculation methods: DLPNO-CCSD(T) for ground states and DLPNO-STEOM-CCSD for excited states. Our results show that, overall, DFTB3 is the TB method leading to the best results for the geometries and the ΔE values and can be used alone for these purposes for NBD/QC and DTE derivatives. Single point calculations at the r2SCAN-3c level using TB geometries allow circumventing the deficiencies of the TB methods in the AZO series. For electronic transition calculations, the range separated LC-DFTB2 method is the most accurate TB method tested for AZO and NBD/QC derivatives, in close agreement with the reference.
More Related Videos
 06:08
Time-resolved Photophysical Characterization of Triplet-harvesting Organic Compounds at an Oxygen-free Environment Using an iCCD Camera
Published on: December 27, 2018
 08:51
Scale-up Chemical Synthesis of Thermally-activated Delayed Fluorescence Emitters Based on the Dibenzothiophene-S,S-Dioxide Core
Published on: October 24, 2017
Related Concept Videos
 01:29
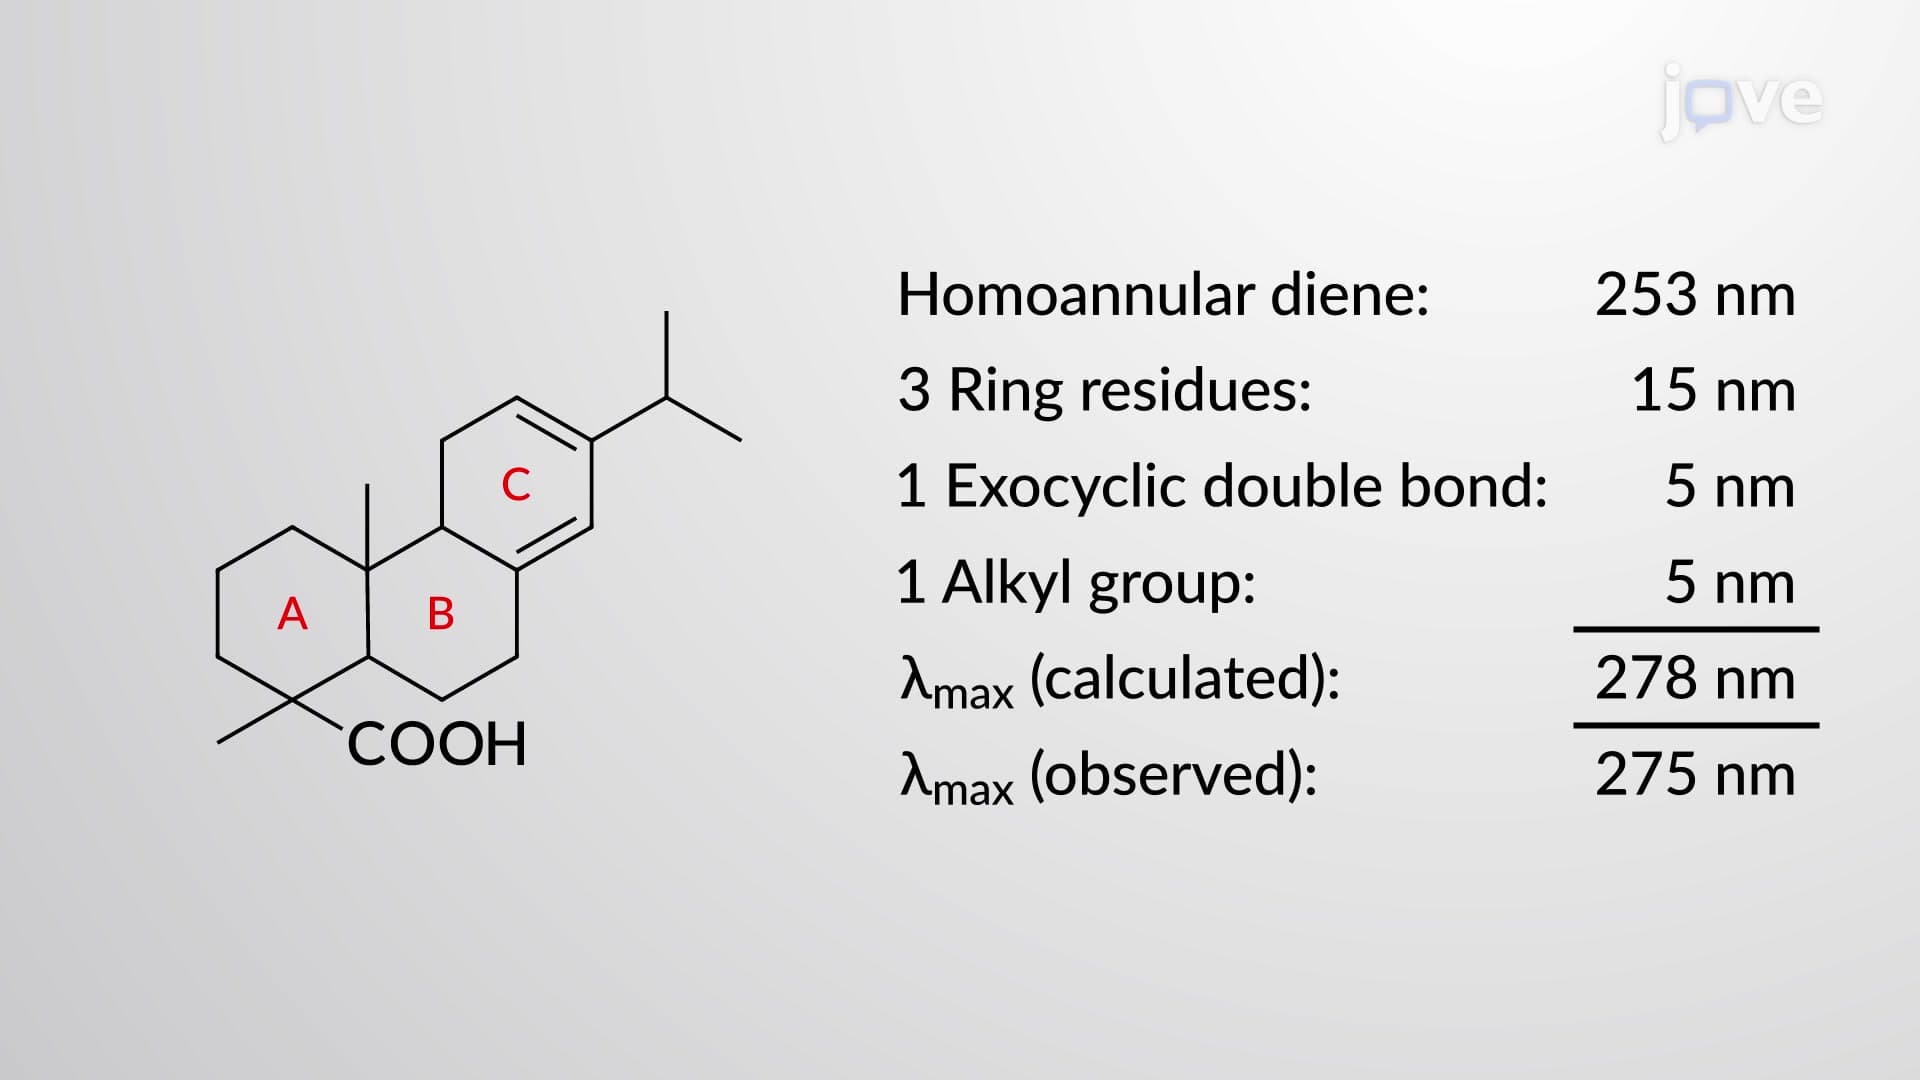
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is the extent...
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 02:13
Energy Diagrams, Transition States, and Intermediates
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds