Structure-based drug design with geometric deep learning
Related Experiment Video
Updated: Aug 8, 2025

Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
Clemens Isert1, Kenneth Atz1, Gisbert Schneider2
1ETH Zurich, Department of Chemistry and Applied Biosciences, Vladimir-Prelog-Weg 4, Zurich, 8093, Switzerland.

Abstract:
Structure-based drug design uses three-dimensional geometric information of macromolecules, such as proteins or nucleic acids, to identify suitable ligands. Geometric deep learning, an emerging concept of neural-network-based machine learning, has been applied to macromolecular structures. This review provides an overview of the recent applications of geometric deep learning in bioorganic and medicinal chemistry, highlighting its potential for structure-based drug discovery and design. Emphasis is placed on molecular property prediction, ligand binding site and pose prediction, and structure-based de novo molecular design. The current challenges and opportunities are highlighted, and a forecast of the future of geometric deep learning for drug discovery is presented.

Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:26
Drug Discovery: Overview
 02:27
Predicting Molecular Geometry
 02:40
Ligand Binding Sites
 01:15
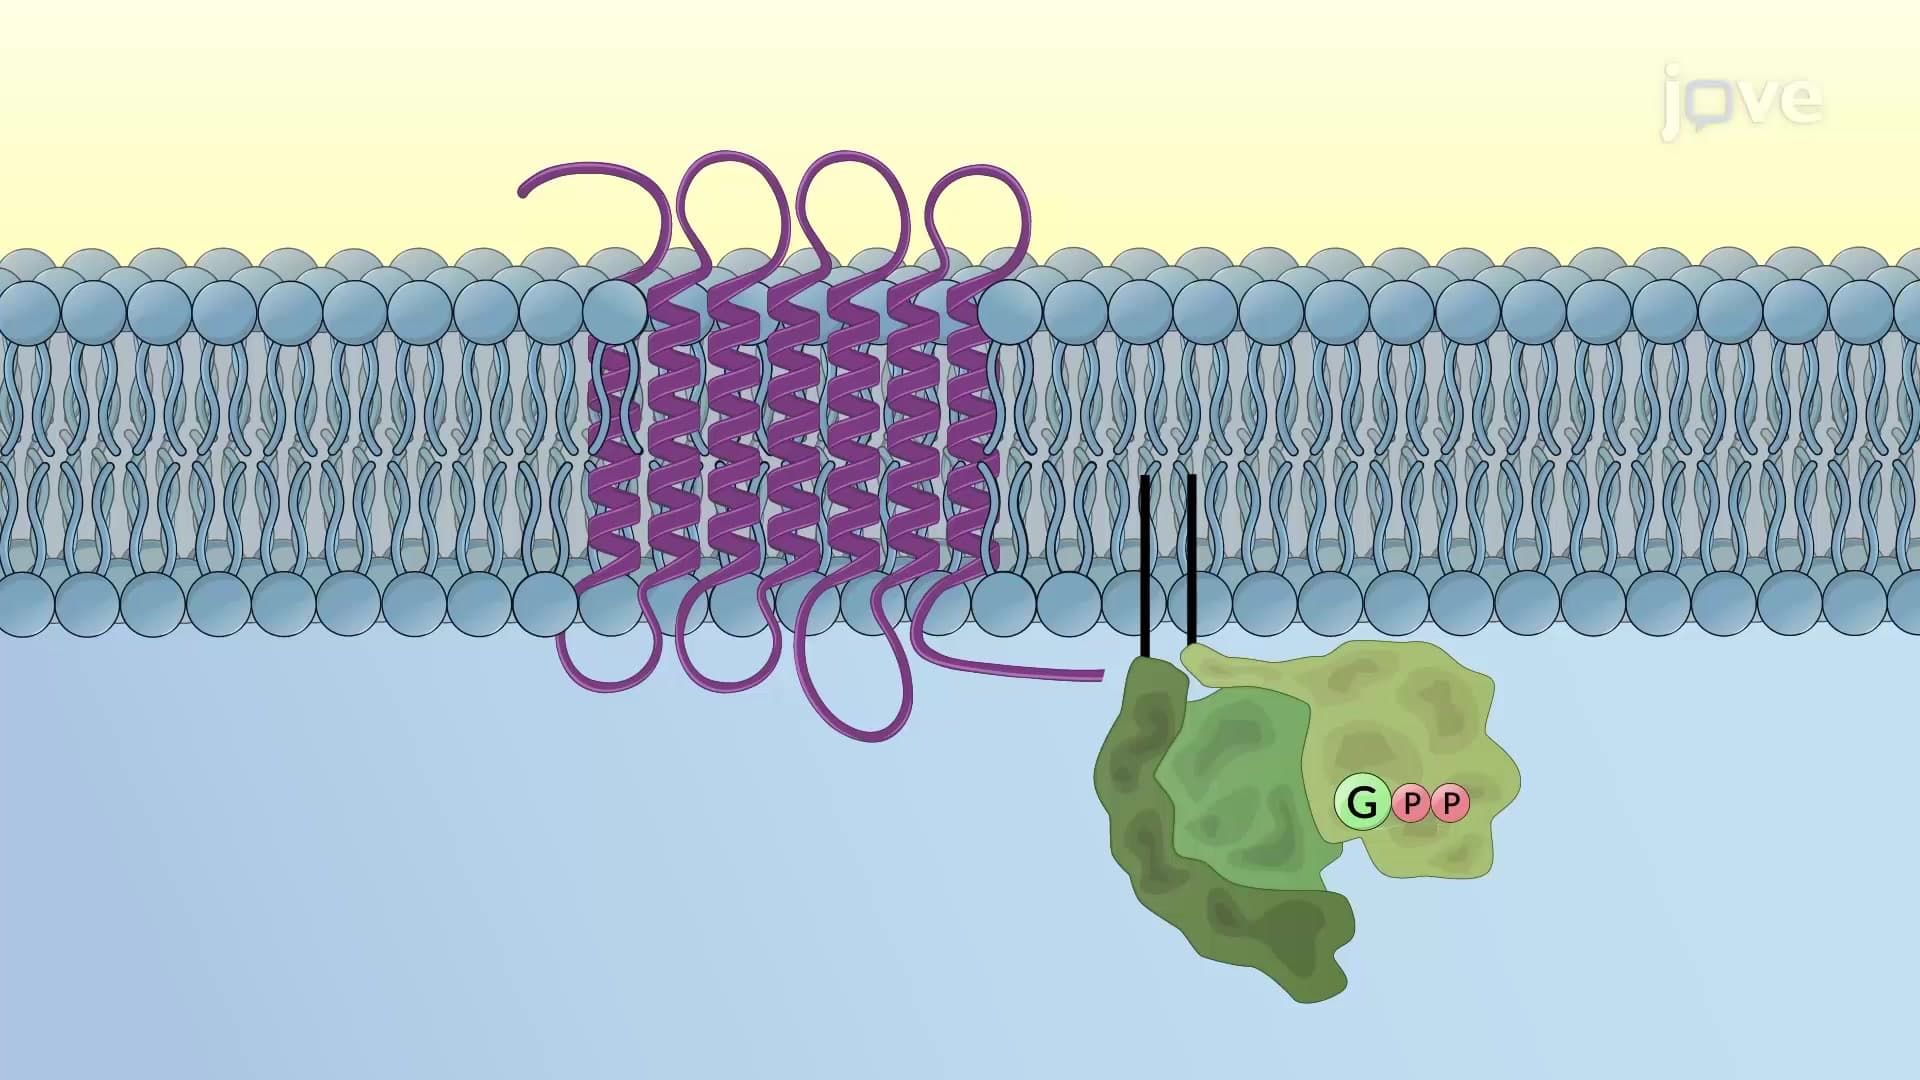
G Protein-coupled Receptors
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...