

Comparing Ab Initio Molecular Dynamics and a Semiclassical Grand Canonical Scheme for the Electric Double Layer of
Jun Huang1,2, Yufan Zhang2, Mengru Li1
1Institute of Theoretical Chemistry, Ulm University, 89081 Ulm, Germany.

Abstract:
The theoretical modeling of metal/water interfaces centers on an appropriate configuration of the electric double layer (EDL) under grand canonical conditions. In principle, ab initio molecular dynamics (AIMD) simulations would be the appropriate choice for treating the competing water-water and water-metal interactions and explicitly considering the atomic and electronic degrees of freedom. However, this approach only allows simulations of relatively small canonical ensembles over a limited period (shorter than 100 ps). On the other hand, computationally efficient semiclassical approaches can treat the EDL model based on a grand canonical scheme by averaging the microscopic details. Thus, an improved description of the EDL can be obtained by combining AIMD simulations and semiclassical methods based on a grand canonical scheme. By taking the Pt(111)/water interface as an example, we compare these approaches in terms of the electric field, water configuration, and double-layer capacitance. Furthermore, we discuss how the combined merits of the approaches can contribute to advances in EDL theory.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 01:27
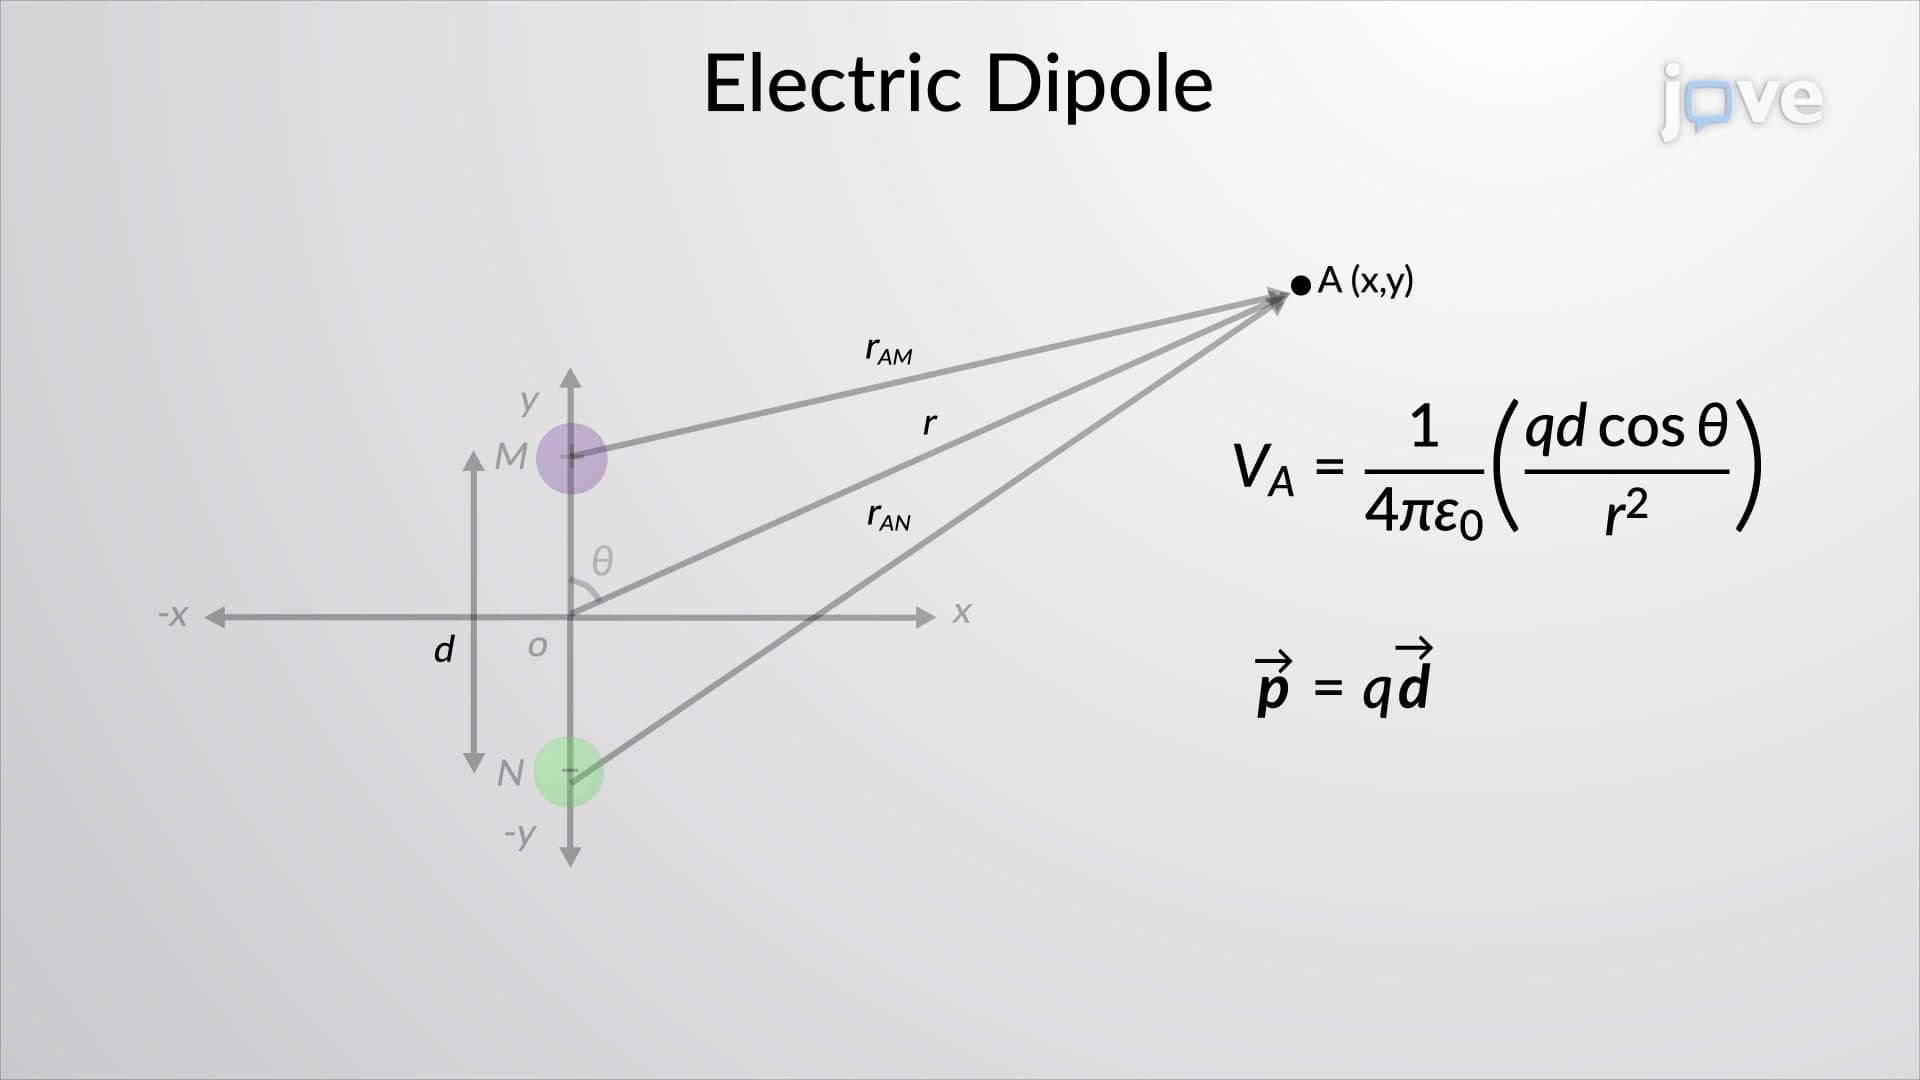
Calculations of Electric Potential II
Consider a...
 01:27
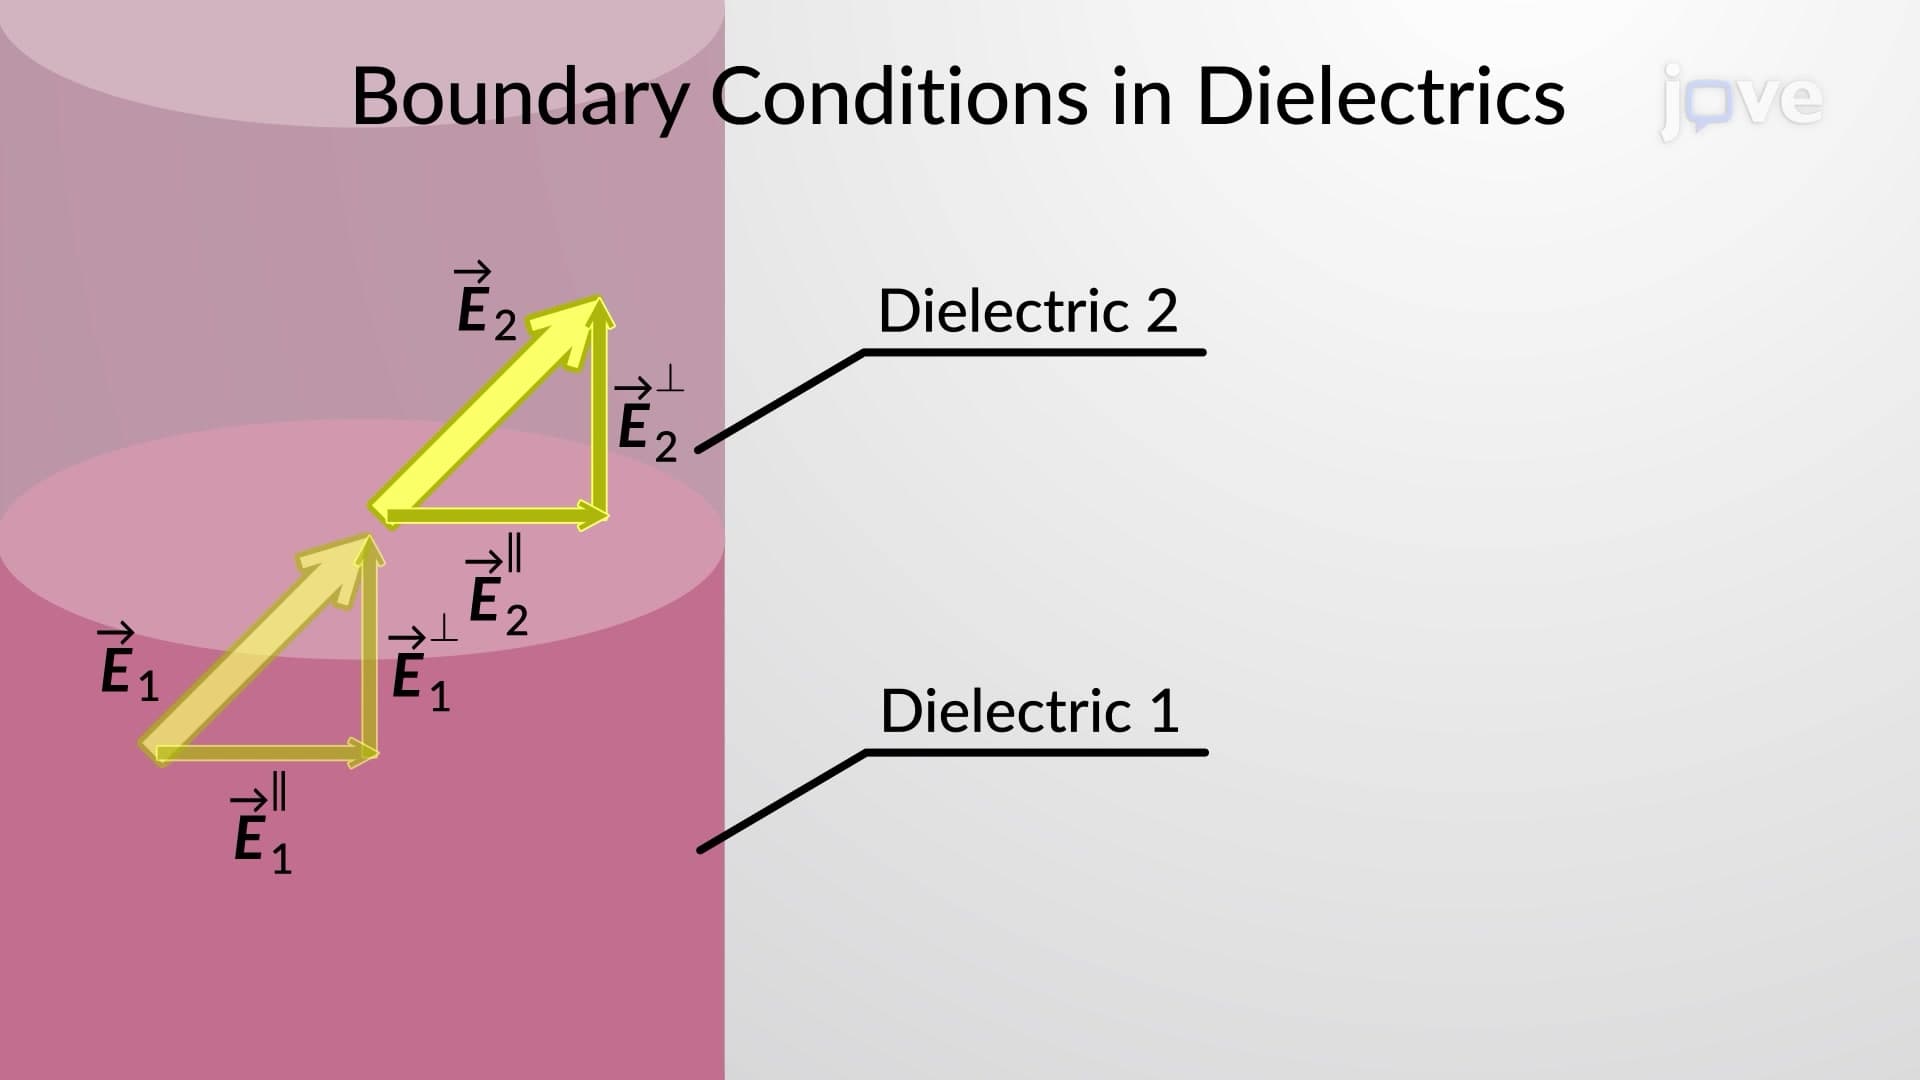
Electrostatic Boundary Conditions in Dielectrics
Consider a case where both the mediums across a boundary are two different dielectric materials. Recall that the electric field and electric displacement are proportional and related through the material's...
 01:10
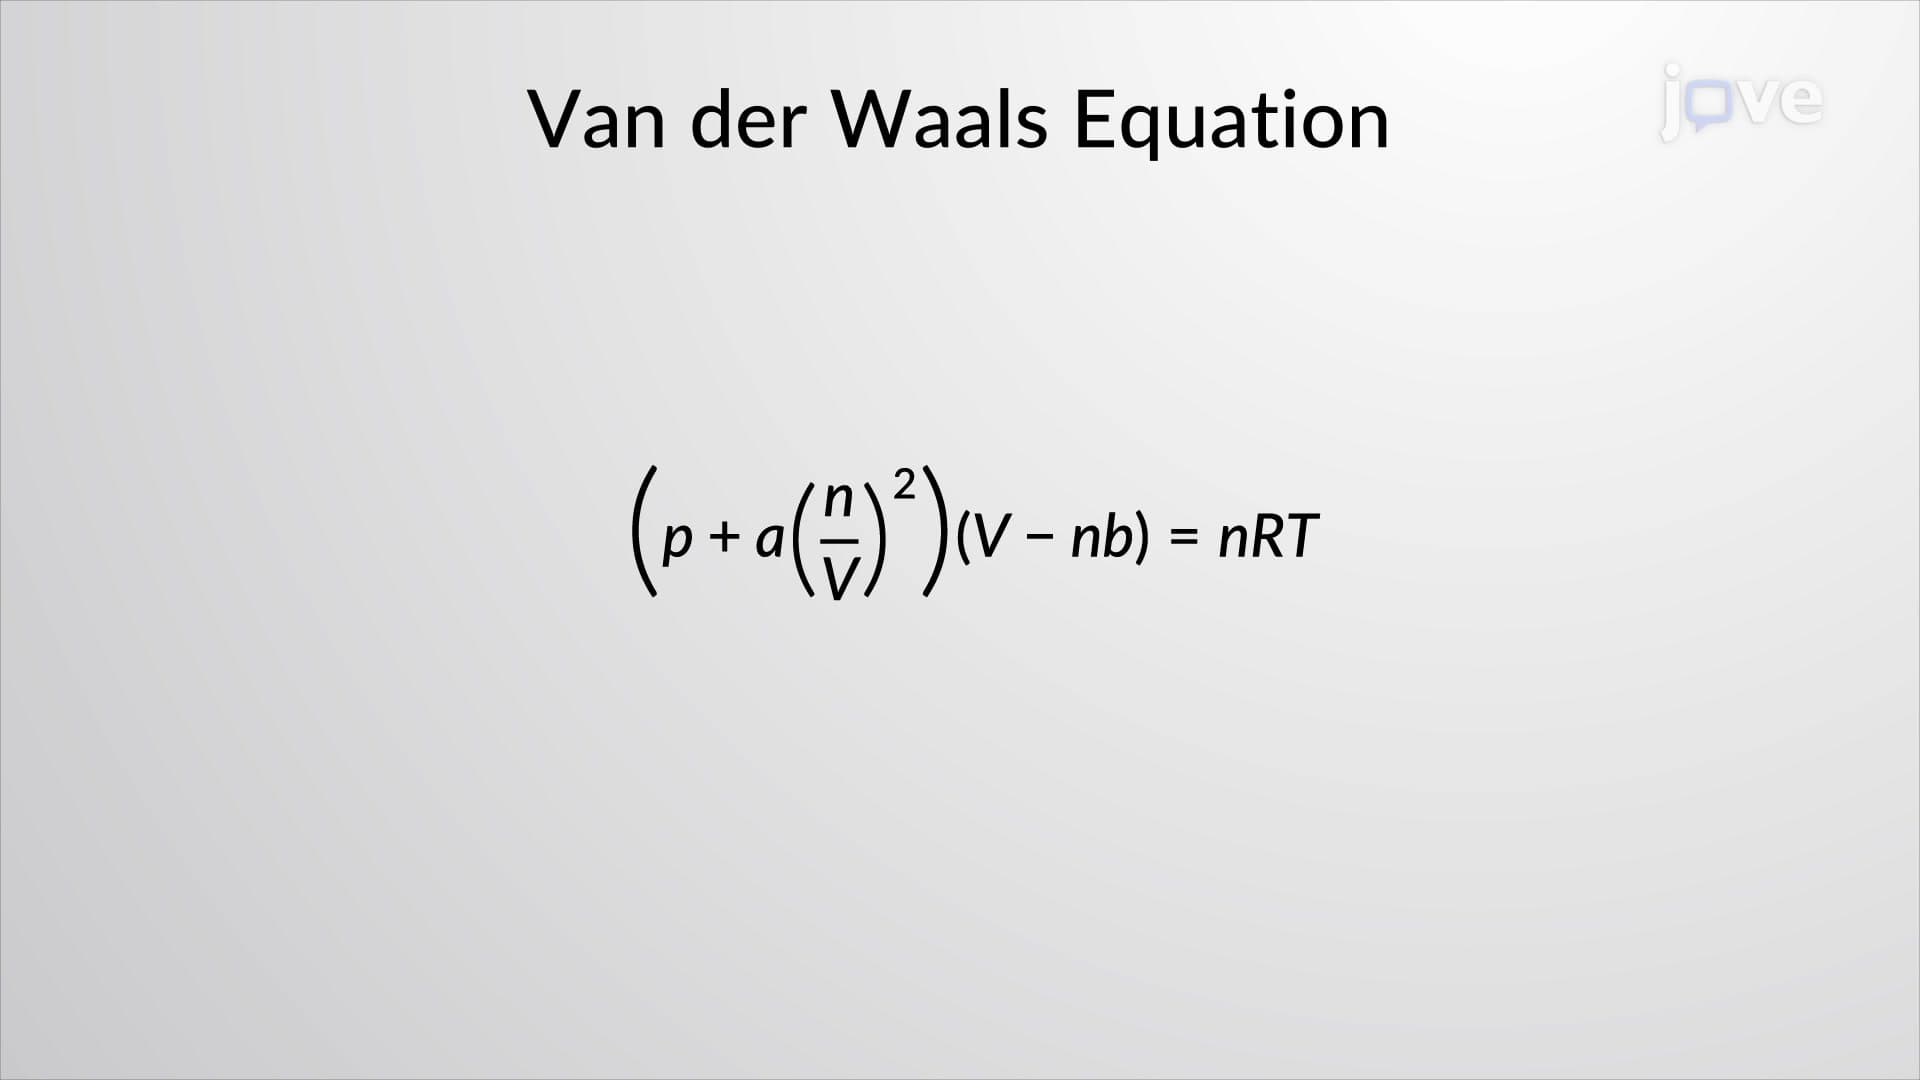
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:06
Interfacial Electrochemical Methods: Overview
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding