Related Experiment Video
Updated: Aug 8, 2025
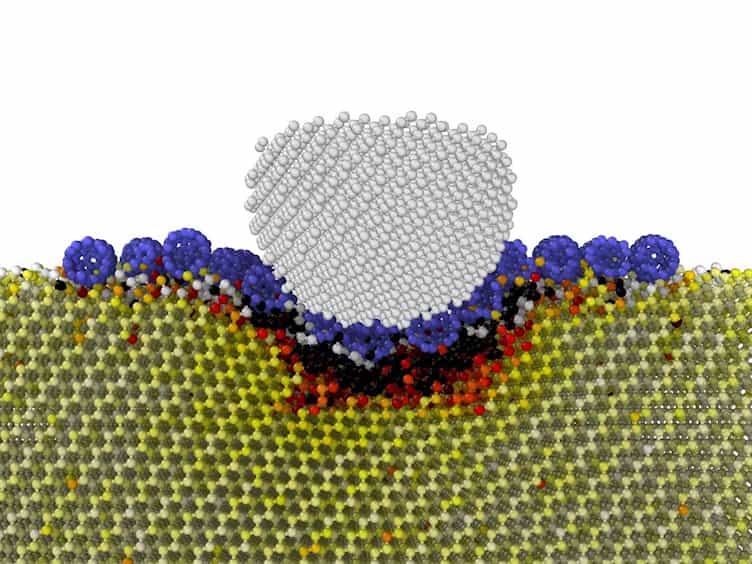

Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
A highly accurate full-dimensional ab initio potential surface for the rearrangement of methylhydroxycarbene
Heng-Ding Wang1, Yan-Lin Fu1, Bina Fu1
1State Key Laboratory of Molecular Reaction Dynamics and Center for Theoretical and Computational Chemistry, Dalian Institute of Chemical Physics, Chinese Academy of Sciences, Dalian 116023, China. zhangdh@dicp.ac.cn.

Abstract:
We report here a full-dimensional machine learning global potential surface (PES) for the rearrangement of methylhydroxycarbene (H3C-C-OH, 1t). The PES is trained with the fundamental invariant neural network (FI-NN) method on 91 564 ab initio energies calculated at the UCCSD(T)-F12a/cc-pVTZ level of theory, covering three possible product channels. FI-NN PES has the correct symmetry properties with respect to permutation of four identical hydrogen atoms and is suitable for dynamics studies of the 1t rearrangement. The averaged root mean square error (RMSE) is 11.4 meV. Six important reaction pathways, as well as the energies and vibrational frequencies at the stationary geometries on these pathways are accurately preproduced by our FI-NN PES. To demonstrate the capacity of the PES, we calculated the rate coefficient of hydrogen migration in -CH3 (path A) and hydrogen migration of -OH (path B) with instanton theory on this PES. Our calculations predicted the half-life of 1t to be 95 min, which is excellent in agreement with experimental observations.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:07
Radicals: Electronic Structure and Geometry
Accordingly, the structure of a trivalent radical lies between the geometries of carbocations and carbanions. An sp2-hybridized carbocation is trigonal planar, while an sp3-hybridized carbanion is trigonal pyramidal. Here, the difference in geometry is...
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 01:21
[3,3] Sigmatropic Rearrangement of 1,5-Dienes: Cope Rearrangement
 02:10
Carbocations
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...