Related Experiment Video
Updated: Aug 4, 2025
Engineering Antiviral Agents via Surface Plasmon Resonance
Published on: June 14, 2022
Identifying promising druggable binding sites and their flexibility to target the receptor-binding domain of
M Ghoula1, S Naceri1, S Sitruk1
1Université Paris Cité, CNRS, INSERM, Unité de Biologie Fonctionnelle et Adaptative, F-75013 Paris, France.

Abstract:
The spike protein of severe acute respiratory syndrome coronavirus 2 (SARS-CoV-2) is crucial for viral infection. The interaction of its receptor-binding domain (RBD) with the human angiotensin-converting enzyme 2 (ACE2) protein is required for the virus to enter the host cell. We identified RBD binding sites to block its function with inhibitors by combining the protein structural flexibility with machine learning analysis. Molecular dynamics simulations were performed on unbound or ACE2-bound RBD conformations. Pockets estimation, tracking and druggability prediction were performed on a large sample of simulated RBD conformations. Recurrent druggable binding sites and their key residues were identified by clustering pockets based on their residue similarity. This protocol successfully identified three druggable sites and their key residues, aiming to target with inhibitors for preventing ACE2 interaction. One site features key residues for direct ACE2 interaction, highlighted using energetic computations, but can be affected by several mutations of the variants of concern. Two highly druggable sites, located between the spike protein monomers interface are promising. One weakly impacted by only one Omicron mutation, could contribute to stabilizing the spike protein in its closed state. The other, currently not affected by mutations, could avoid the activation of the spike protein trimer.
More Related Videos
 08:31
Biosensor-based High Throughput Biopanning and Bioinformatics Analysis Strategy for the Global Validation of Drug-protein Interactions
Published on: December 1, 2020
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 02:04
Protein-protein Interfaces
 01:15
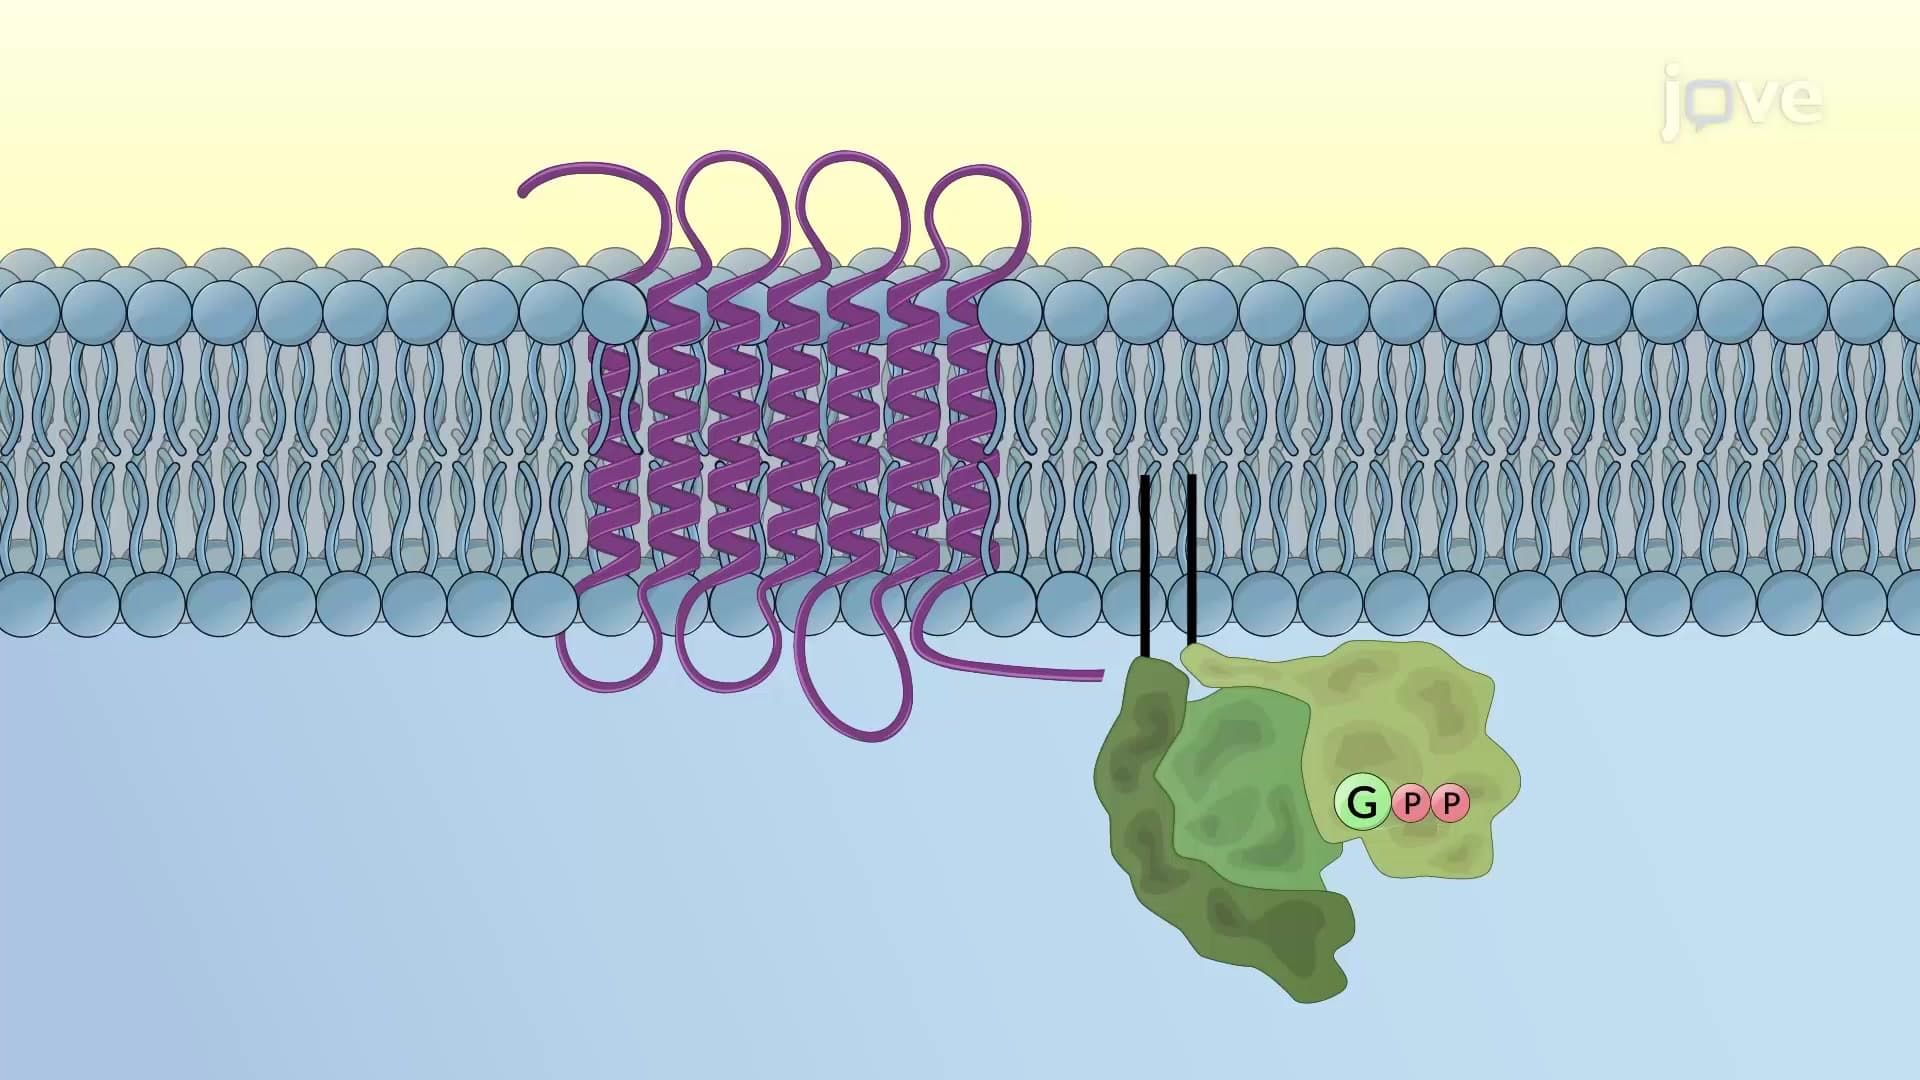
G Protein-coupled Receptors
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 00:49
Ligand Binding and Linkage