

Revisited relativistic Dirac-Hartree-Fock X-ray scattering factors. II. Chemically relevant cations and selected
Shiroye Olukayode1, Charlotte Froese Fischer2, Anatoliy Volkov1
1Department of Chemistry and Computational Science Program, Middle Tennessee State University, Murfreesboro, TN 37132, USA.

Abstract:
The previously described approach for determination of the relativistic atomic X-ray scattering factors (XRSFs) at the Dirac-Hartree-Fock level [Olukayode et al. (2023). Acta Cryst. A79, 59-79] has been used to evaluate the XRSFs for a total of 318 species including all chemically relevant cations [Greenwood & Earnshaw (1997). Chemistry of the Elements], six monovalent anions (O-, F-, Cl-, Br-, I-, At-), the ns1np3 excited (valence) states of carbon and silicon, and several exotic cations (Db5+, Sg6+, Bh7+, Hs8+ and Cn2+) for which the chemical compounds have been recently identified, thus significantly extending the coverage relative to all the earlier studies. Unlike the data currently recommended by the International Union of Crystallography (IUCr) [Maslen et al. (2006). International Tables for Crystallography, Vol. C, Section 6.1.1, pp. 554-589], which originate from different levels of theory including the non-relativistic Hartree-Fock and correlated methods, as well as the relativistic Dirac-Slater calculations, the re-determined XRSFs come from a uniform treatment of all species within the same relativistic B-spline Dirac-Hartree-Fock approach [Zatsarinny & Froese Fischer (2016). Comput. Phys. Comm. 202, 287-303] that includes the Breit interaction correction and the Fermi nuclear charge density model. While it was not possible to compare the quality of the generated wavefunctions with that from the previous studies due to a lack (to the best of our knowledge) of such data in the literature, a careful comparison of the total electronic energies and the estimated atomic ionization energies with experimental and theoretical values from other studies instils confidence in the quality of the calculations. A combination of the B-spline approach and a fine radial grid allowed for a precise determination of the XRSFs for each species in the entire 0 ≤ sin θ/λ ≤ 6 Å-1 range, thus avoiding the necessity for extrapolation in the 2 ≤ sin θ/λ ≤ 6 Å-1 interval which, as was shown in the first study, may lead to inconsistencies. In contrast to the Rez et al. work [Acta Cryst. (1994), A50, 481-497], no additional approximations were introduced when calculating wavefunctions for the anions. The conventional and extended expansions were employed to produce interpolating functions for each species in both the 0 ≤ sin θ/λ ≤ 2 Å-1 and 2 ≤ sin θ/λ ≤ 6 Å-1 intervals, with the extended expansions offering a significantly better accuracy at a minimal computational overhead. The combined results of this and the previous study may be used to update the XRSFs for neutral atoms and ions listed in Vol. C of the 2006 edition of International Tables for Crystallography.
More Related Videos
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:08
Atomic Radii and Effective Nuclear Charge
 03:10
Ionic Radii
 02:18
X-ray Crystallography
Diffraction
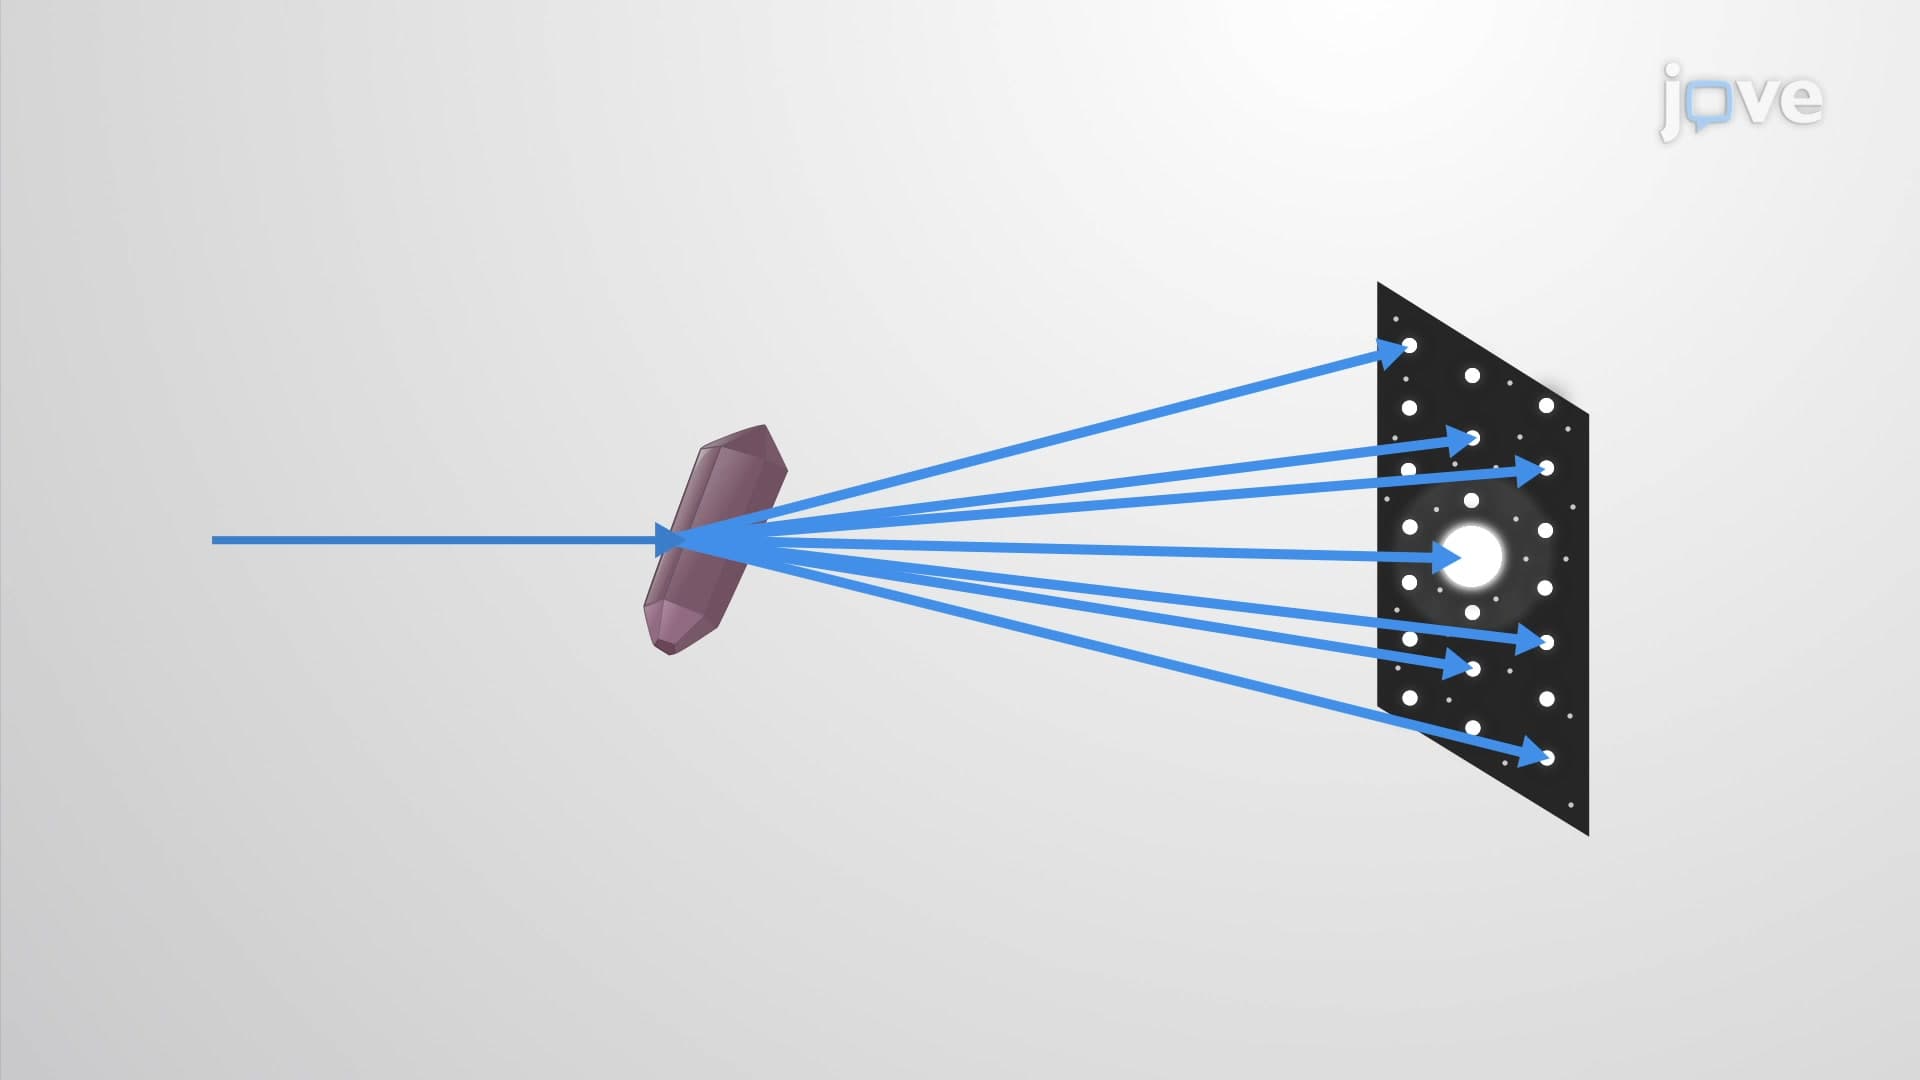
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 02:46
Electron Configurations
The relative energies of the subshells determine the order in which atomic orbitals are filled (1s, 2s, 2p, 3s, 3p,...