CO and CO2 adsorption mechanism in Fe(pz)[Pt(CN)4] probed by neutron scattering and density-functional theory calculations
Ángel Fernández-Blanco1,2, Lucía Piñeiro-López3, Mónica Jiménez-Ruiz1
1Institut Laue Langevin, 71 Avenue des Martyrs, Grenoble, CS 20156-38042, France. velamazan@ill.eu.

Abstract:
We study the binding mechanism of CO and CO2 in the porous spin-crossover compound Fe(pz)[Pt(CN)4] by combining neutron diffraction (ND), inelastic neutron scattering (INS) and density-functional theory (DFT) calculations. Two adsorption sites are identified, above the open-metal site and between the pyrazine rings. For CO adsorption, the guest molecules are parallel to the neighboring gas molecules and perpendicular to the pyrazine planes. For CO2, the molecules adsorbed on-top of the open-metal site are perpendicular to the pyrazine rings and those between the pyrazines are almost parallel to them. These configurations are consistent with the INS data, which are in good agreement with the computed generalized phonon density of states. The most relevant signatures of the binding occur in the spectral region around 100 cm-1 and 400 cm-1. The first peak blue-shifts for both CO and CO2 adsorption, while the second red-shifts for CO and remains nearly unchanged for CO2. These spectral changes depend both from steric effects and the nature of the interaction. The interpretation of the INS data as supported by the computed binding energy and the molecular orbital analysis are consistent with a physisorption mechanism for both gases. This work shows the strength of the combination of neutron techniques and DFT calculations to characterize in detail the gas adsorption mechanism in this type of materials.
Related Concept Videos
 02:42
Valence Bond Theory
 02:40
MO Theory and Covalent Bonding
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:20
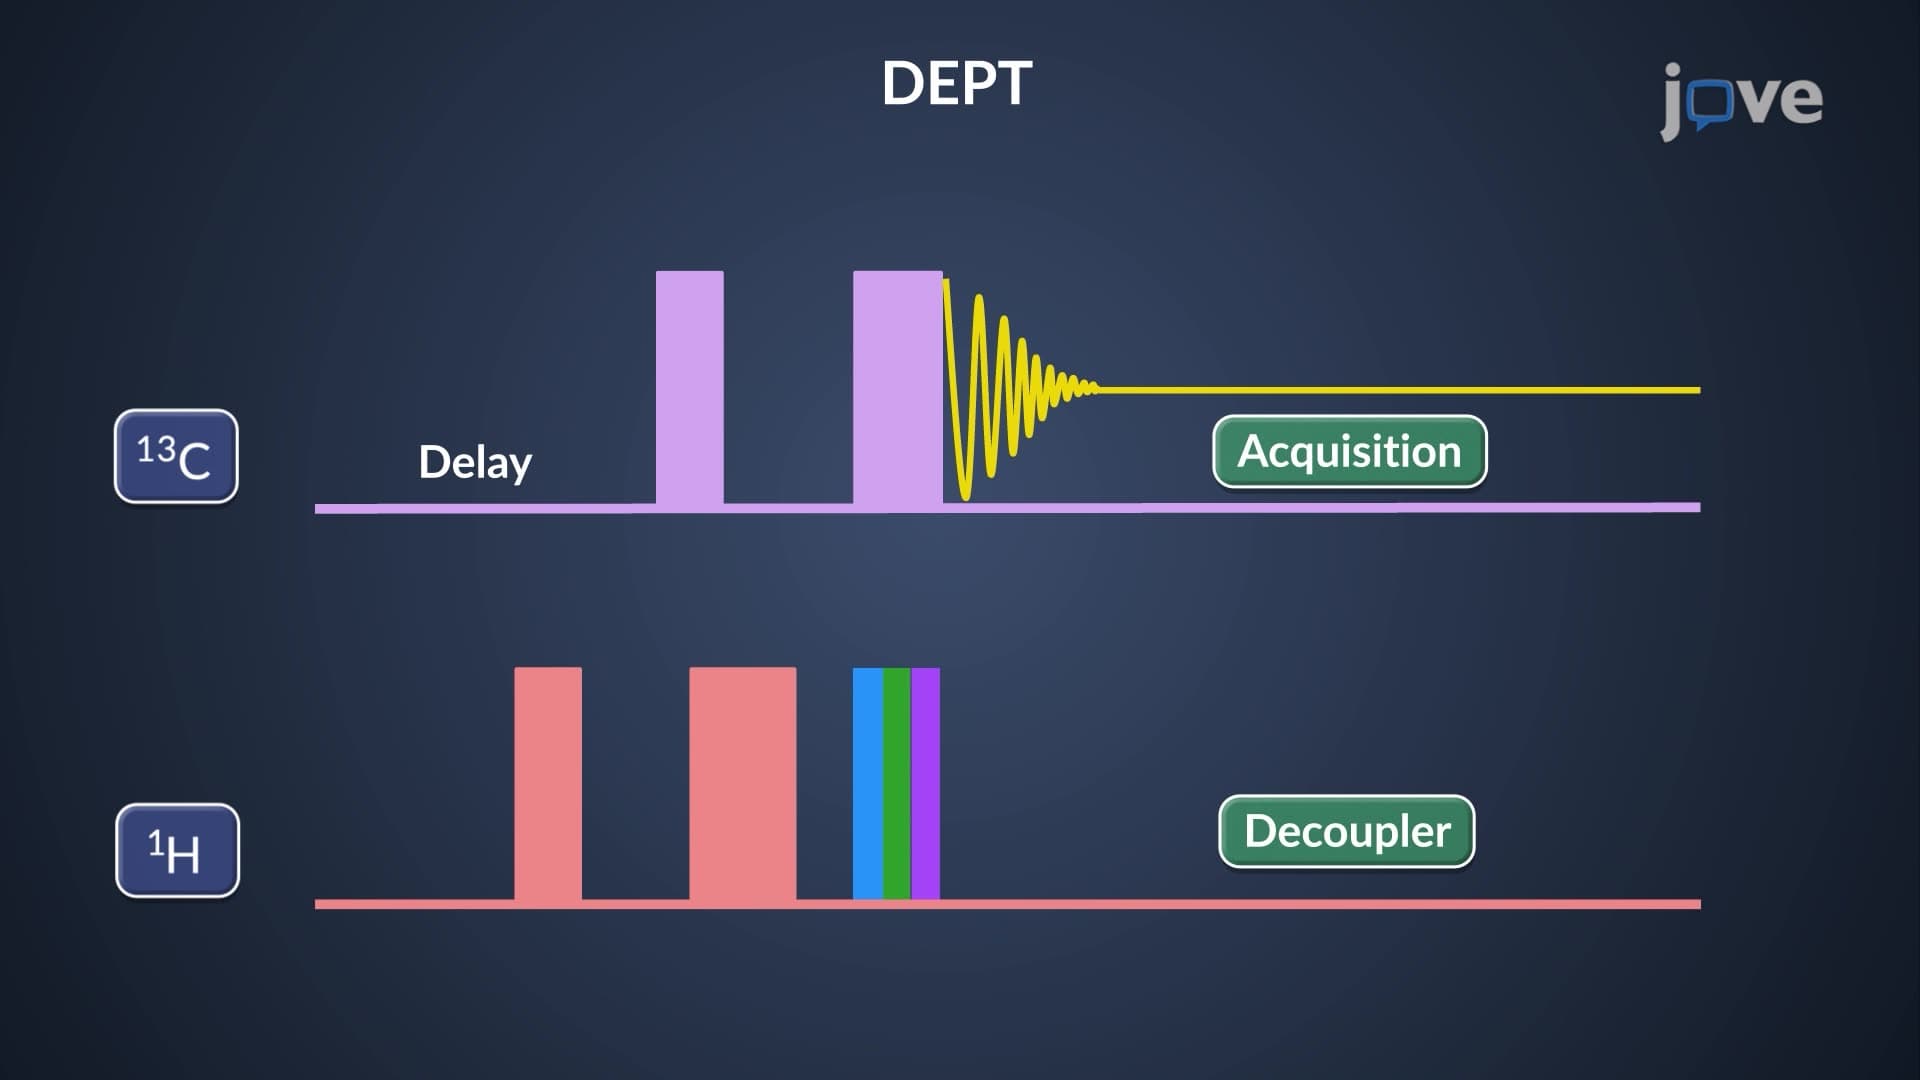
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)