Related Experiment Video
Updated: Aug 1, 2025

Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Enabling large-scale quantum path integral molecular dynamics simulations through the integration of Dcdftbmd and
Yoshifumi Nishimura1, Hiromi Nakai1,2
1Waseda Research Institute for Science and Engineering, Waseda University, 3-4-1 Okubo, Shinjuku-ku, Tokyo 169-8555, Japan.

Abstract:
A large-scale quantum chemical calculation program, Dcdftbmd, was integrated with a Python-based advanced atomistic simulation program, i-PI. The implementation of a client-server model enabled hierarchical parallelization with respect to replicas and force evaluations. The established framework demonstrated that quantum path integral molecular dynamics simulations can be executed with high efficiency for systems consisting of a few tens of replicas and containing thousands of atoms. The application of the framework to bulk water systems, with and without an excess proton, demonstrated that nuclear quantum effects are significant for intra- and inter-molecular structural properties, including oxygen-hydrogen bond distance and radial distribution function around the hydrated excess proton.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:36
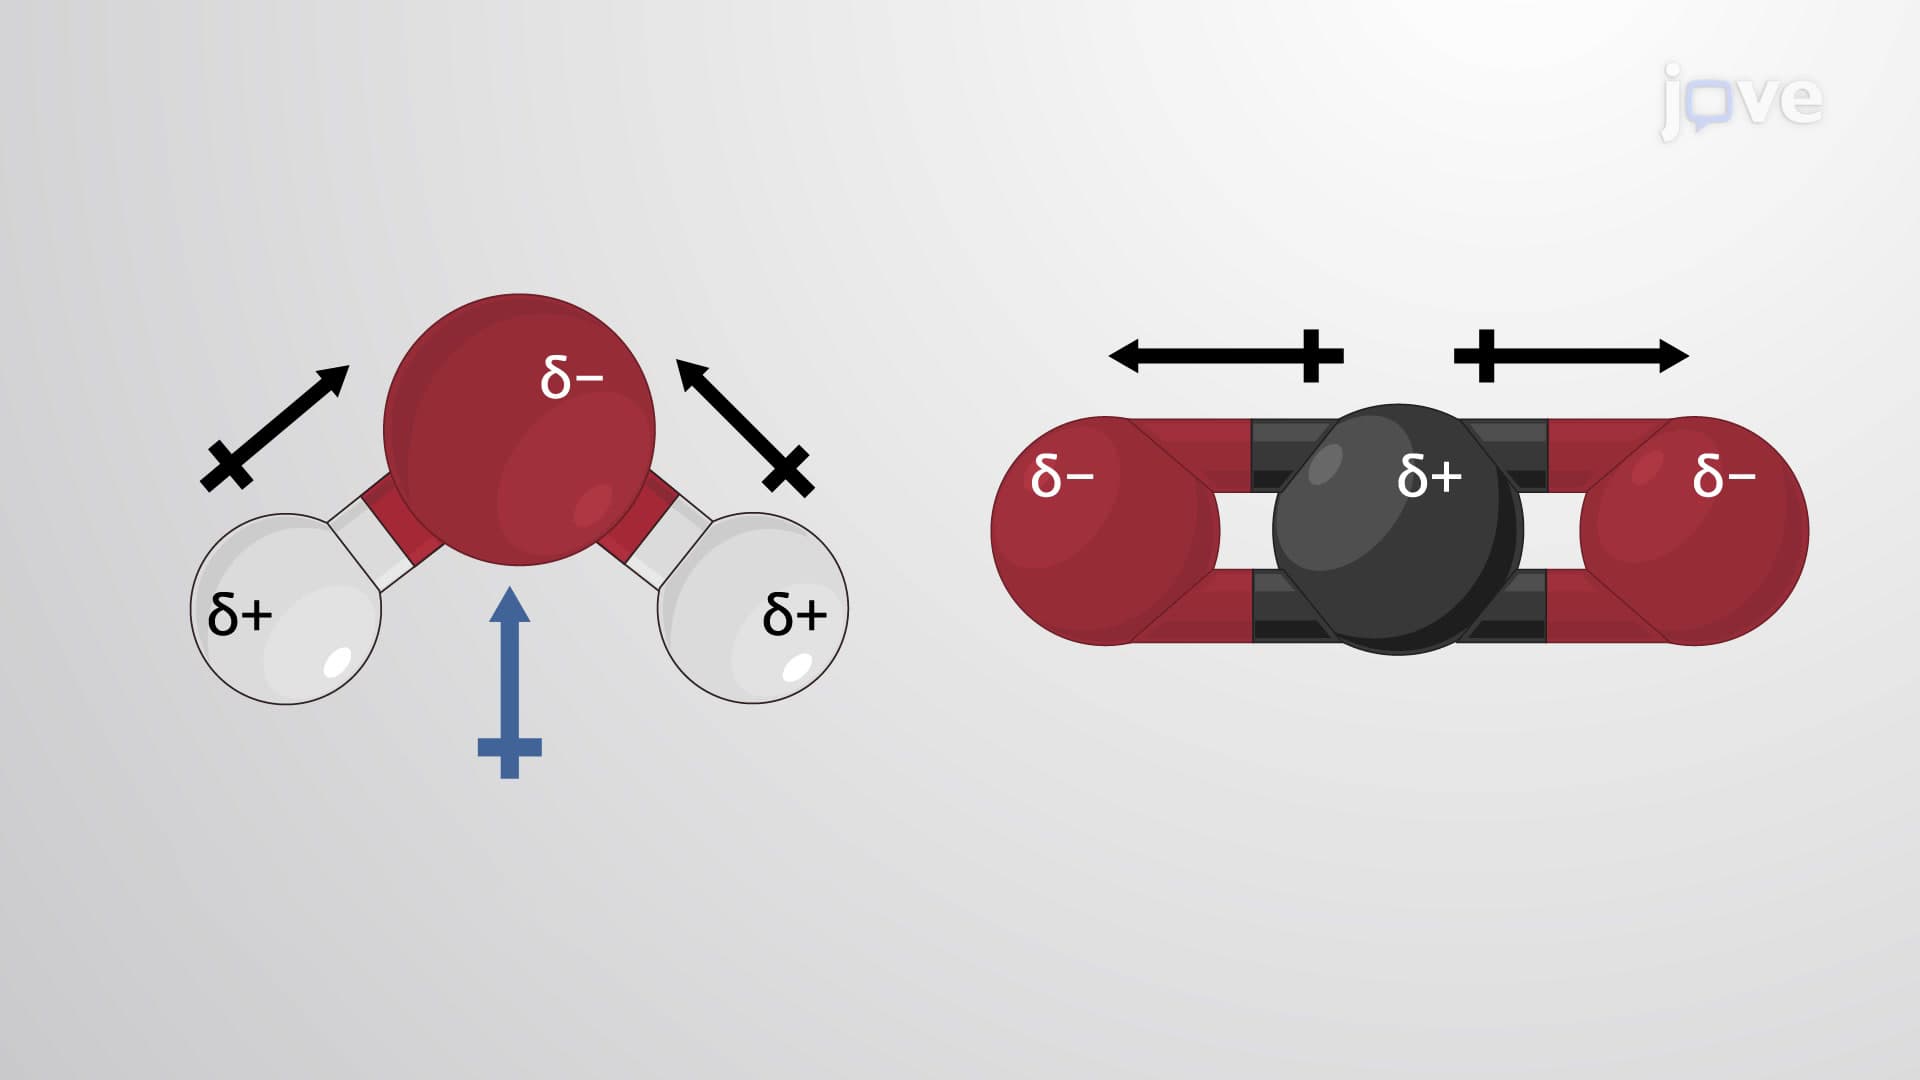
Molecular Geometry and Dipole Moments
 03:35
Hybridization of Atomic Orbitals II
 01:15
Determination of Pi Terms
The theorem indicates that...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:12
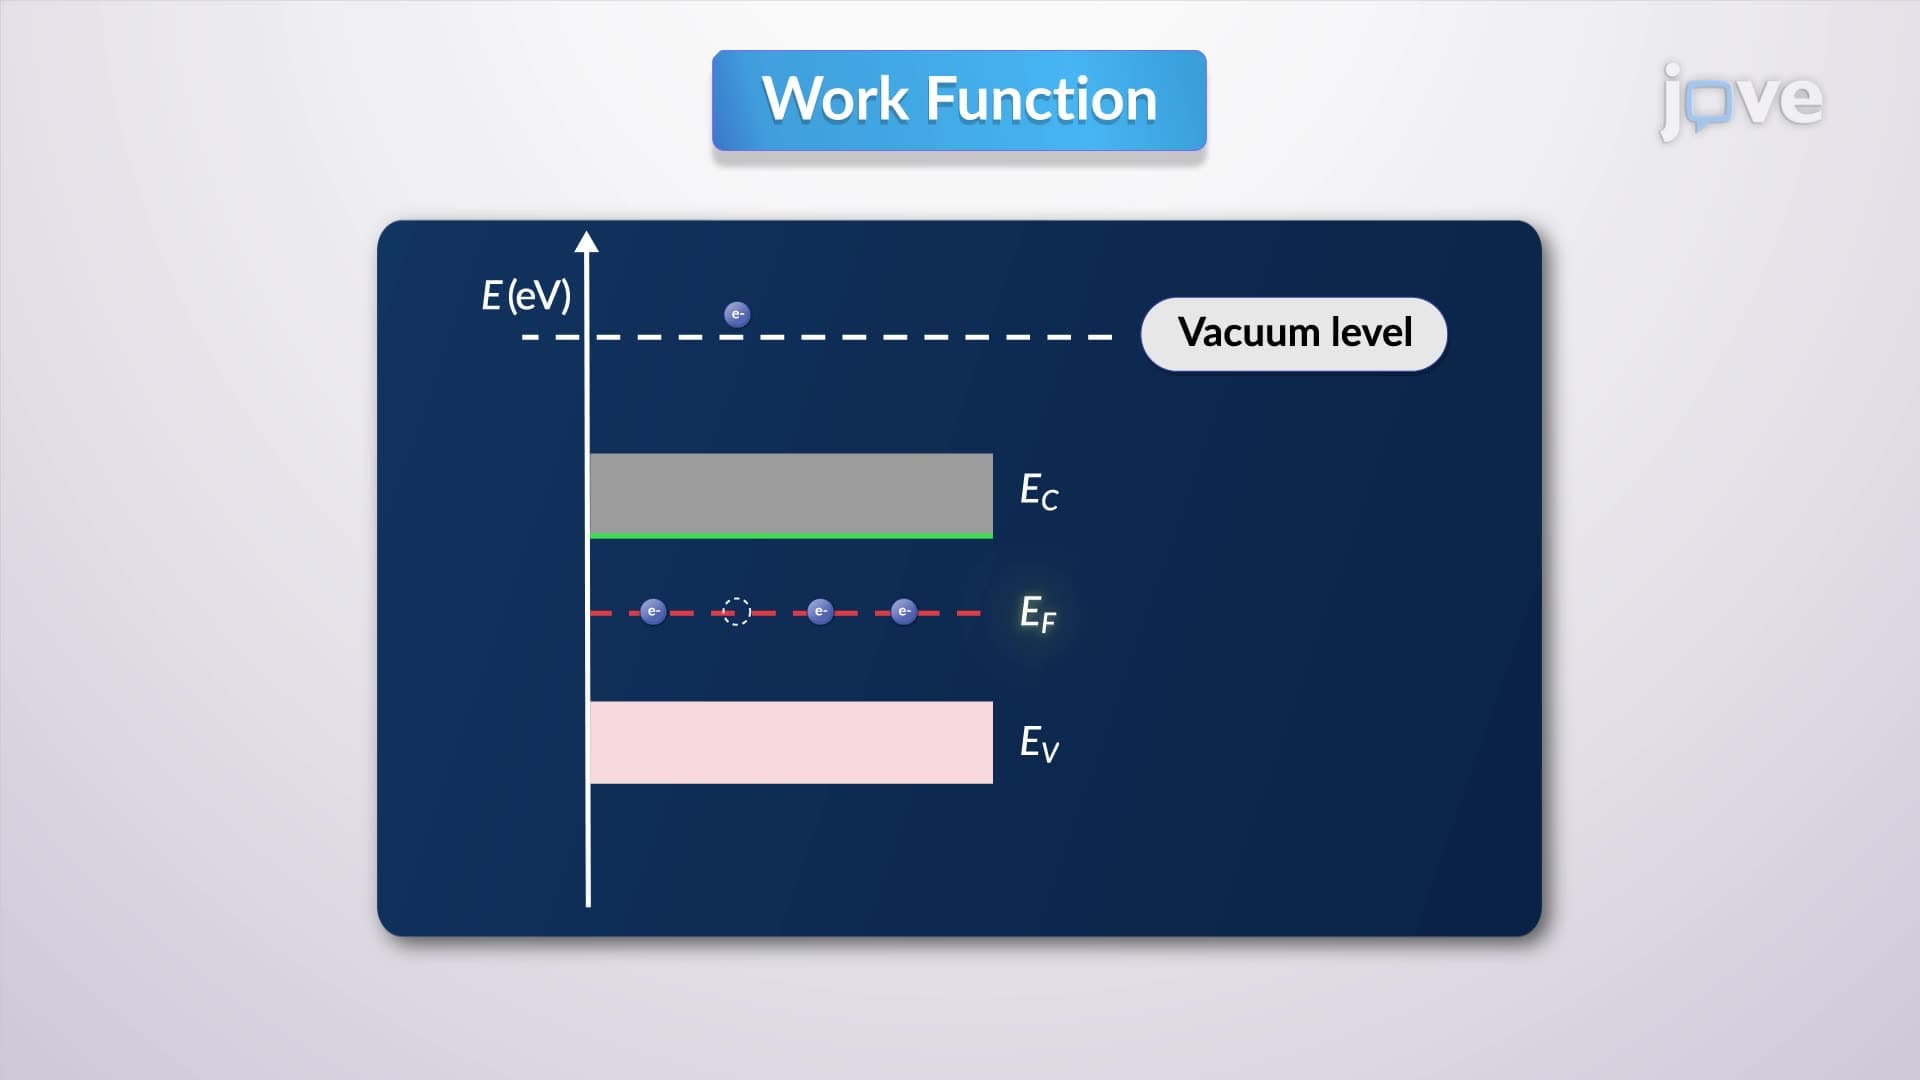
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...