Related Experiment Video
Updated: Jul 31, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Strengths and limitations of the adiabatic exact-exchange kernel for total energy calculations
Maria Hellgren1, Lucas Baguet2,3
1Sorbonne Université, MNHN, UMR CNRS 7590, IMPMC, 4 place Jussieu, 75005 Paris, France.

Abstract:
We investigate the adiabatic approximation to the exact-exchange kernel for calculating correlation energies within the adiabatic-connection fluctuation-dissipation framework of time-dependent density functional theory. A numerical study is performed on a set of systems having bonds of different character (H2 and N2 molecules, H-chain, H2-dimer, solid-Ar, and the H2O-dimer). We find that the adiabatic kernel can be sufficient in strongly bound covalent systems, yielding similar bond lengths and binding energies. However, for non-covalent systems, the adiabatic kernel introduces significant errors around equilibrium geometry, systematically overestimating the interaction energy. The origin of this behavior is investigated by studying a model dimer composed of one-dimensional, closed-shell atoms, interacting via soft-Coulomb potentials. The kernel is shown to exhibit a strong frequency dependence at small to intermediate atomic separation that affects both the low-energy spectrum and the exchange-correlation hole obtained from the corresponding diagonal of the two-particle density matrix.
Related Concept Videos
 01:18
Adiabatic Processes for an Ideal Gas
 01:20
Work Done in an Adiabatic Process
 01:16
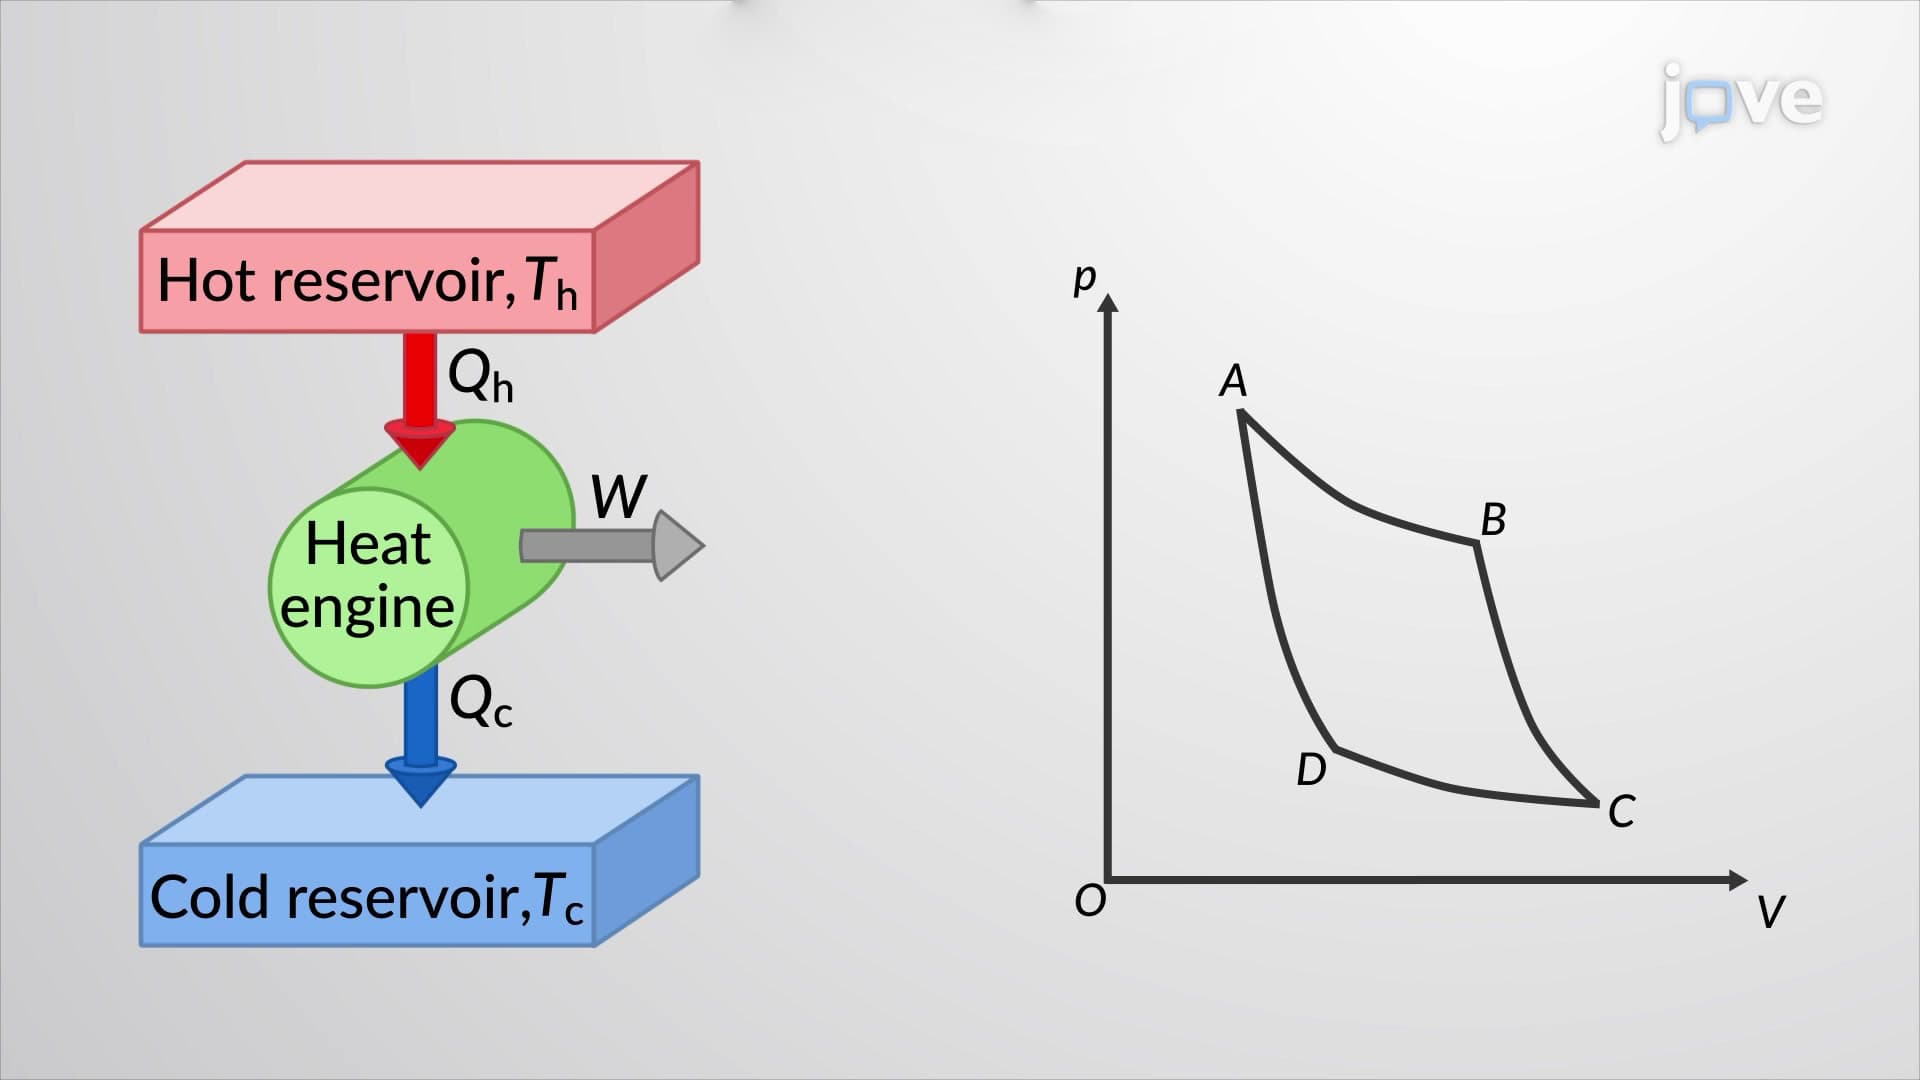
Efficiency of The Carnot Cycle
 01:27
Pressure and Volume in an Adiabatic Process
 01:26
Thermodynamic Potentials
 01:23
Heat Capacities of an Ideal Gas II