Related Experiment Video
Updated: Jul 30, 2025

Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
Mechanistic roles of metal- and ligand-protonated species in hydrogen evolution with [Cp*Rh] complexes
Wade C Henke1, Yun Peng1, Alex A Meier1
1Department of Chemistry, University of Kansas, Lawrence, KS 66045.

Abstract:
Protonation reactions involving organometallic complexes are ubiquitous in redox chemistry and often result in the generation of reactive metal hydrides. However, some organometallic species supported by η5-pentamethylcyclopentadienyl (Cp*) ligands have recently been shown to undergo ligand-centered protonation by direct proton transfer from acids or tautomerization of metal hydrides, resulting in the generation of complexes bearing the uncommon η4-pentamethylcyclopentadiene (Cp*H) ligand. Here, time-resolved pulse radiolysis (PR) and stopped-flow spectroscopic studies have been applied to examine the kinetics and atomistic details involved in the elementary electron- and proton-transfer steps leading to complexes ligated by Cp*H, using Cp*Rh(bpy) as a molecular model (where bpy is 2,2'-bipyridyl). Stopped-flow measurements coupled with infrared and UV-visible detection reveal that the sole product of initial protonation of Cp*Rh(bpy) is [Cp*Rh(H)(bpy)]+, an elusive hydride complex that has been spectroscopically and kinetically characterized here. Tautomerization of the hydride leads to the clean formation of [(Cp*H)Rh(bpy)]+. Variable-temperature and isotopic labeling experiments further confirm this assignment, providing experimental activation parameters and mechanistic insight into metal-mediated hydride-to-proton tautomerism. Spectroscopic monitoring of the second proton transfer event reveals that both the hydride and related Cp*H complex can be involved in further reactivity, showing that [(Cp*H)Rh] is not necessarily an off-cycle intermediate, but, instead, depending on the strength of the acid used to drive catalysis, an active participant in hydrogen evolution. Identification of the mechanistic roles of the protonated intermediates in the catalysis studied here could inform design of optimized catalytic systems supported by noninnocent cyclopentadienyl-type ligands.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 10:01
Protein Film Infrared Electrochemistry Demonstrated for Study of H2 Oxidation by a [NiFe] Hydrogenase
Published on: December 4, 2017
Related Concept Videos
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 02:17
Reduction of Alkenes: Asymmetric Catalytic Hydrogenation
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
 02:13
Reduction of Alkenes: Catalytic Hydrogenation
Metals like palladium, platinum, and nickel are commonly used in their solid forms — fine powder on an inert surface. As these catalysts remain insoluble in the reaction mixture, they are referred to as heterogeneous catalysts.
The hydrogenation process takes place on the...
 01:07
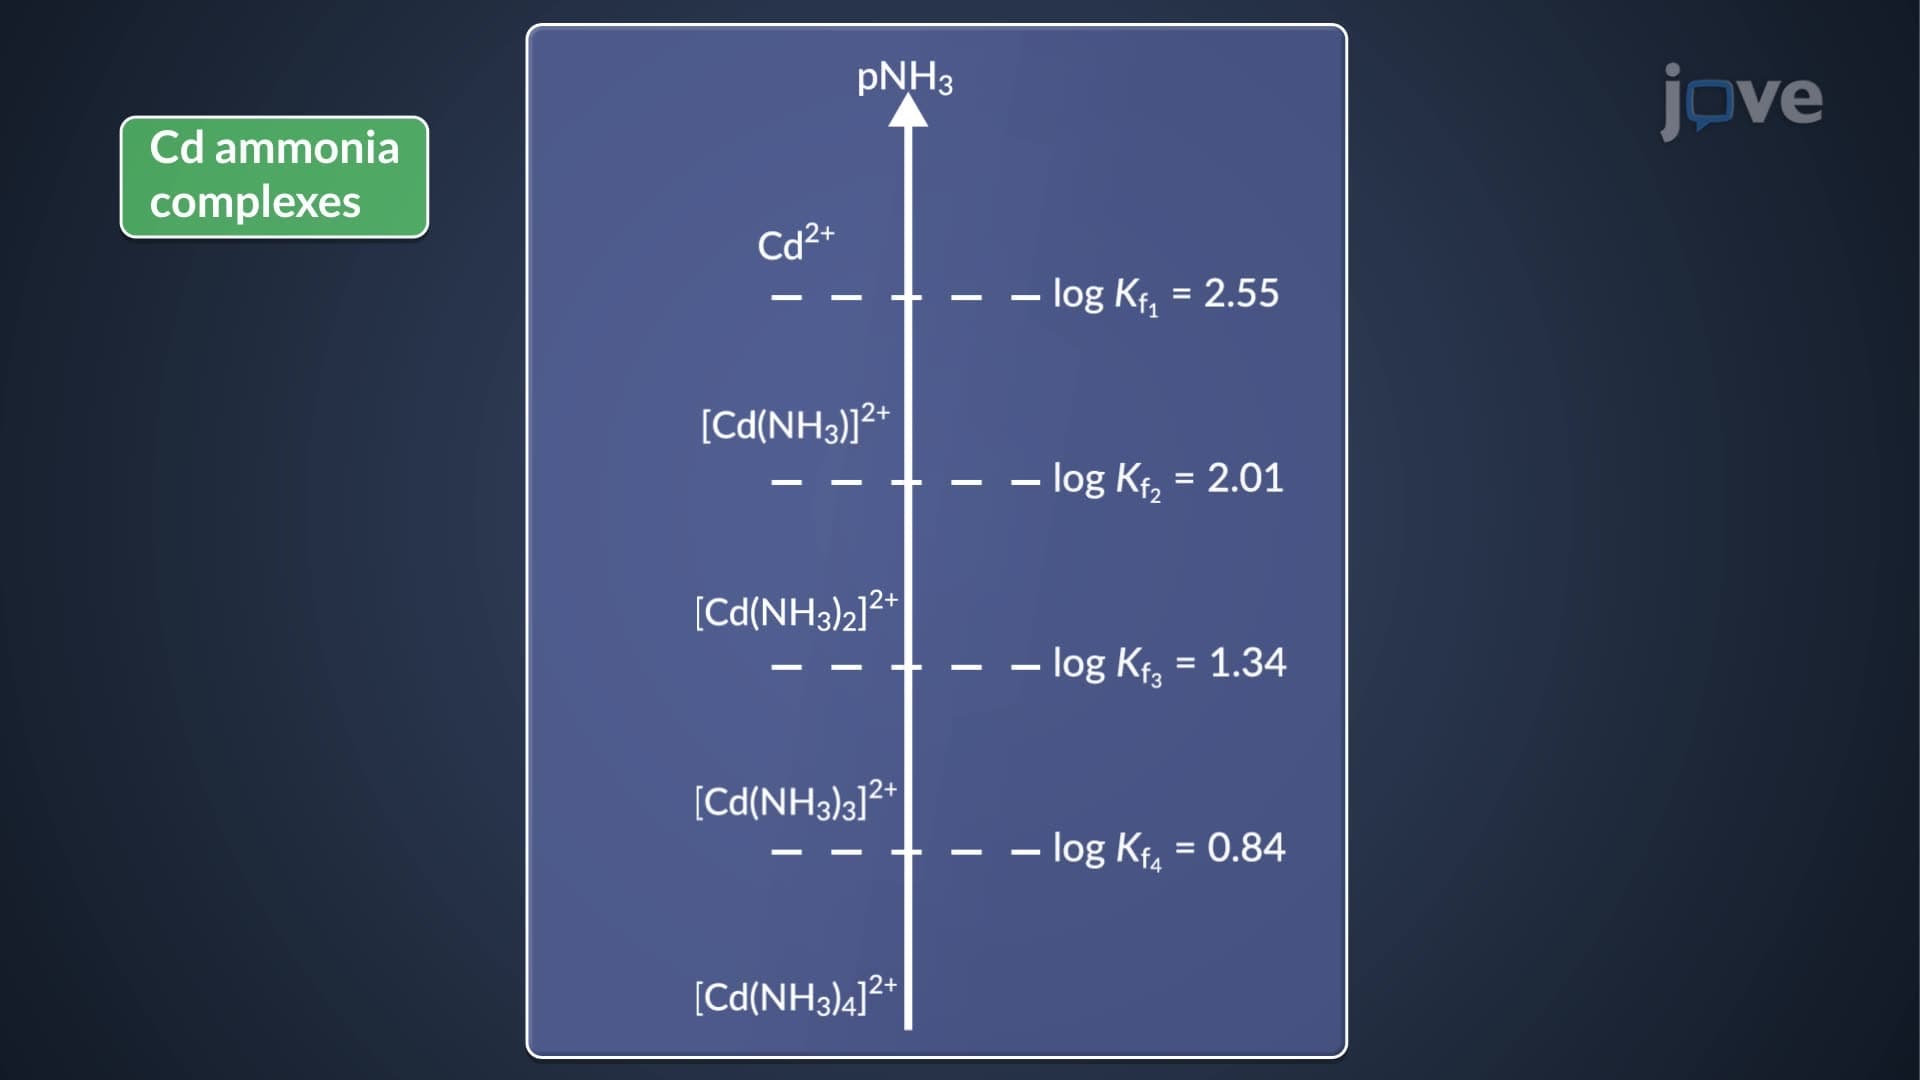
Ladder Diagrams: Complexation Equilibria
The formation constant, K1, for the formation of Cd(NH3)2+ complex from cadmium and ammonia is 3.55 × 10². Log K1 (i.e. pNH3) is 2.55, and...
 01:09
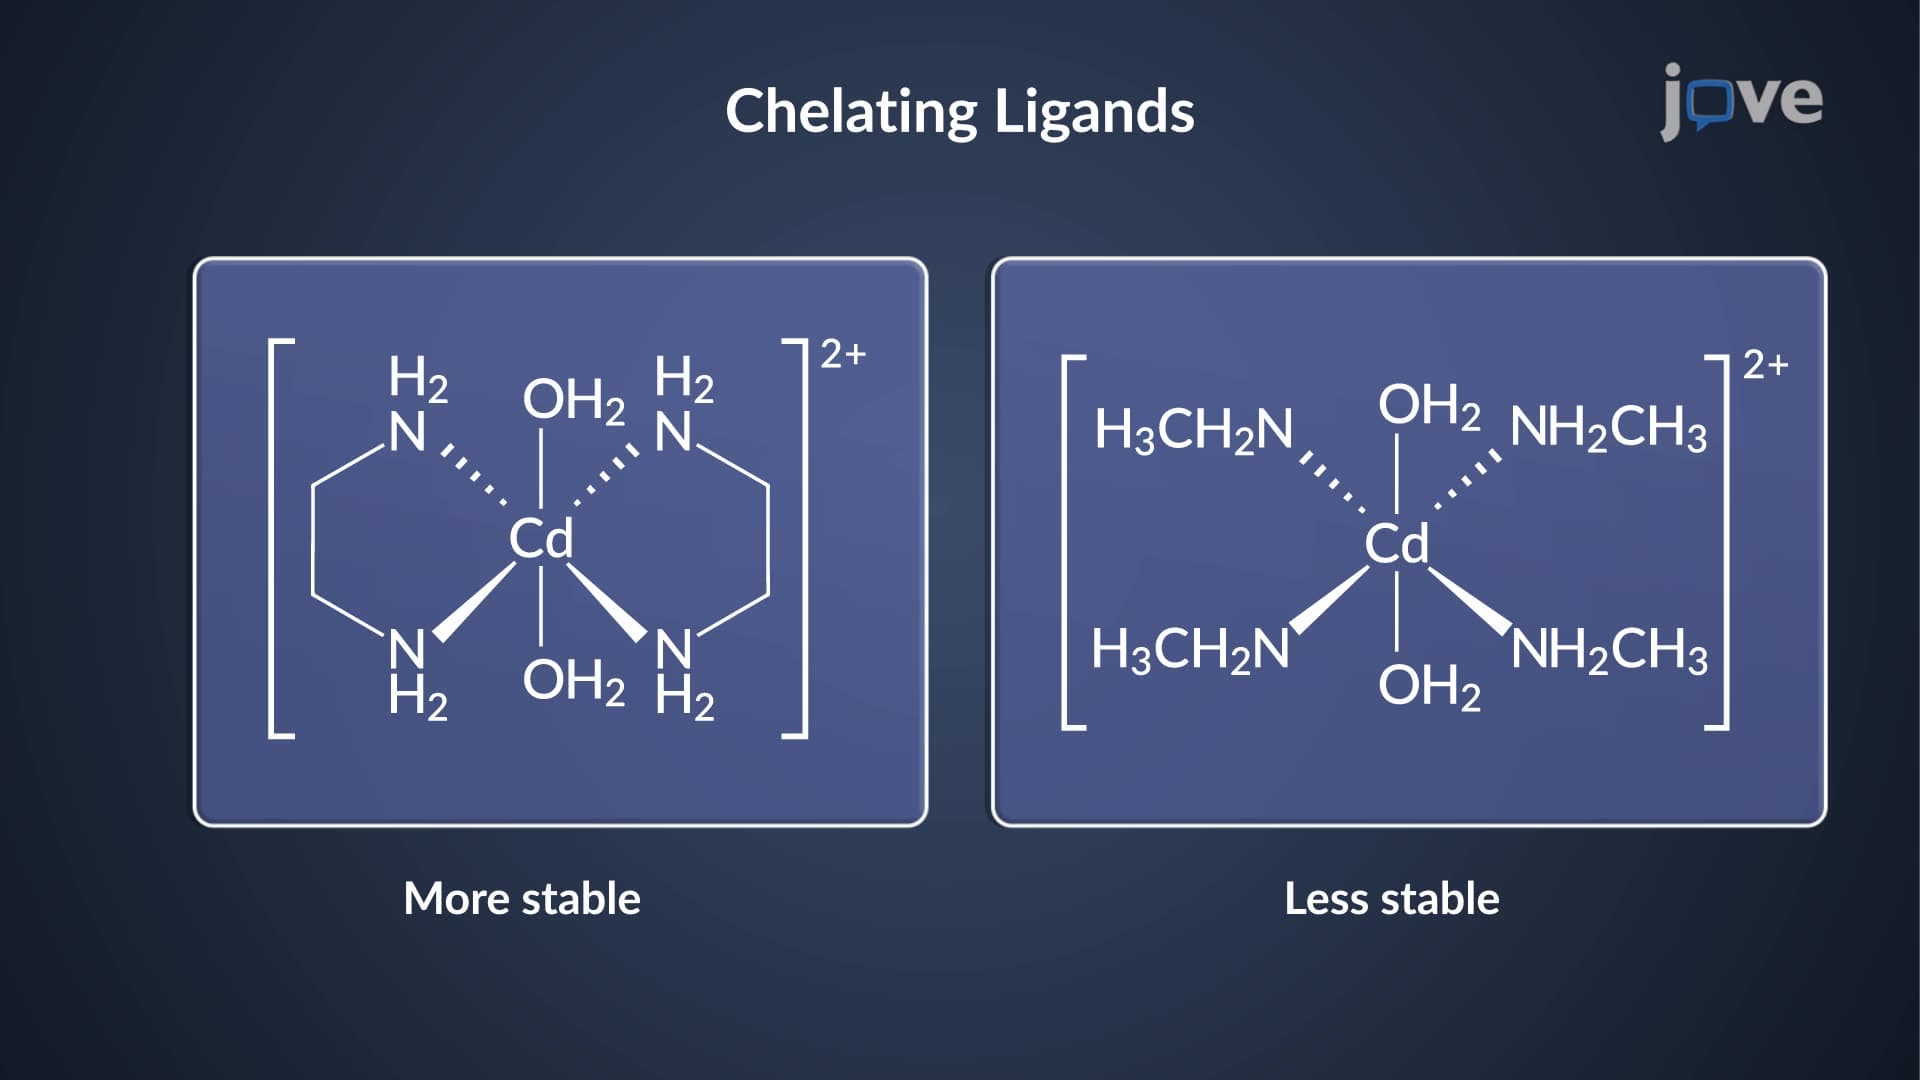
Complexation Equilibria: Factors Influencing Stability of Complexes
 02:24
Reduction of Alkynes to cis-Alkenes: Catalytic Hydrogenation
Like alkenes, alkynes can be reduced to alkanes in the presence of transition metal catalysts such as Pt, Pd, or Ni. The reaction involves two sequential syn additions of hydrogen via a cis-alkene intermediate.