Related Experiment Video
Updated: Jul 29, 2025


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Molecular Docking of Natural Compounds for Potential Inhibition of AhR
Deborah Giordano1, Angelo Facchiano1, Stefania Moccia1
1National Research Council, Institute of Food Sciences, 83100 Avellino, Italy.

Abstract:
The aryl hydrocarbon receptor (AhR) is a highly conserved environmental sensor, historically known for mediating the toxicity of xenobiotics. It is involved in numerous cellular processes such as differentiation, proliferation, immunity, inflammation, homeostasis, and metabolism. It exerts a central role in several conditions such as cancer, inflammation, and aging, acting as a transcription factor belonging to the basic helix-loop-helix/Per-ARNT-Sim (bHLH-PAS) protein family. A key step in the canonical AhR activation is AhR-ARNT heterodimerization followed by the binding to the xenobiotic-responsive elements (XREs). The present work aims to investigate the potential AhR inhibitory activity of selected natural compounds. Due to the absence of a complete structure of human AhRs, a model consisting of the bHLH, the PAS A, and the PAS B domains was constructed. Blind and focused docking simulations revealed the presence of further binding pockets, different from the canonical one presented in the PAS B domain, which could be important for AhR inhibition due to the possibility to impede AhR:ARNT heterodimerization, either preventing conformational changes or masking crucial sites necessary for protein-protein interaction. Two of the compounds retrieved from the docking simulations, i.e., β-carotene and ellagic acid, confirmed their capacity of inhibiting benzo[a]pyrene (BaP)-induced AhR activation in in vitro tests on the human hepatoma cell line HepG2, validating the efficacy of the computational approach.
More Related Videos
 10:33
Development of Inhibitors of Protein-protein Interactions through REPLACE: Application to the Design and Development Non-ATP Competitive CDK Inhibitors
Published on: October 26, 2015
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:29
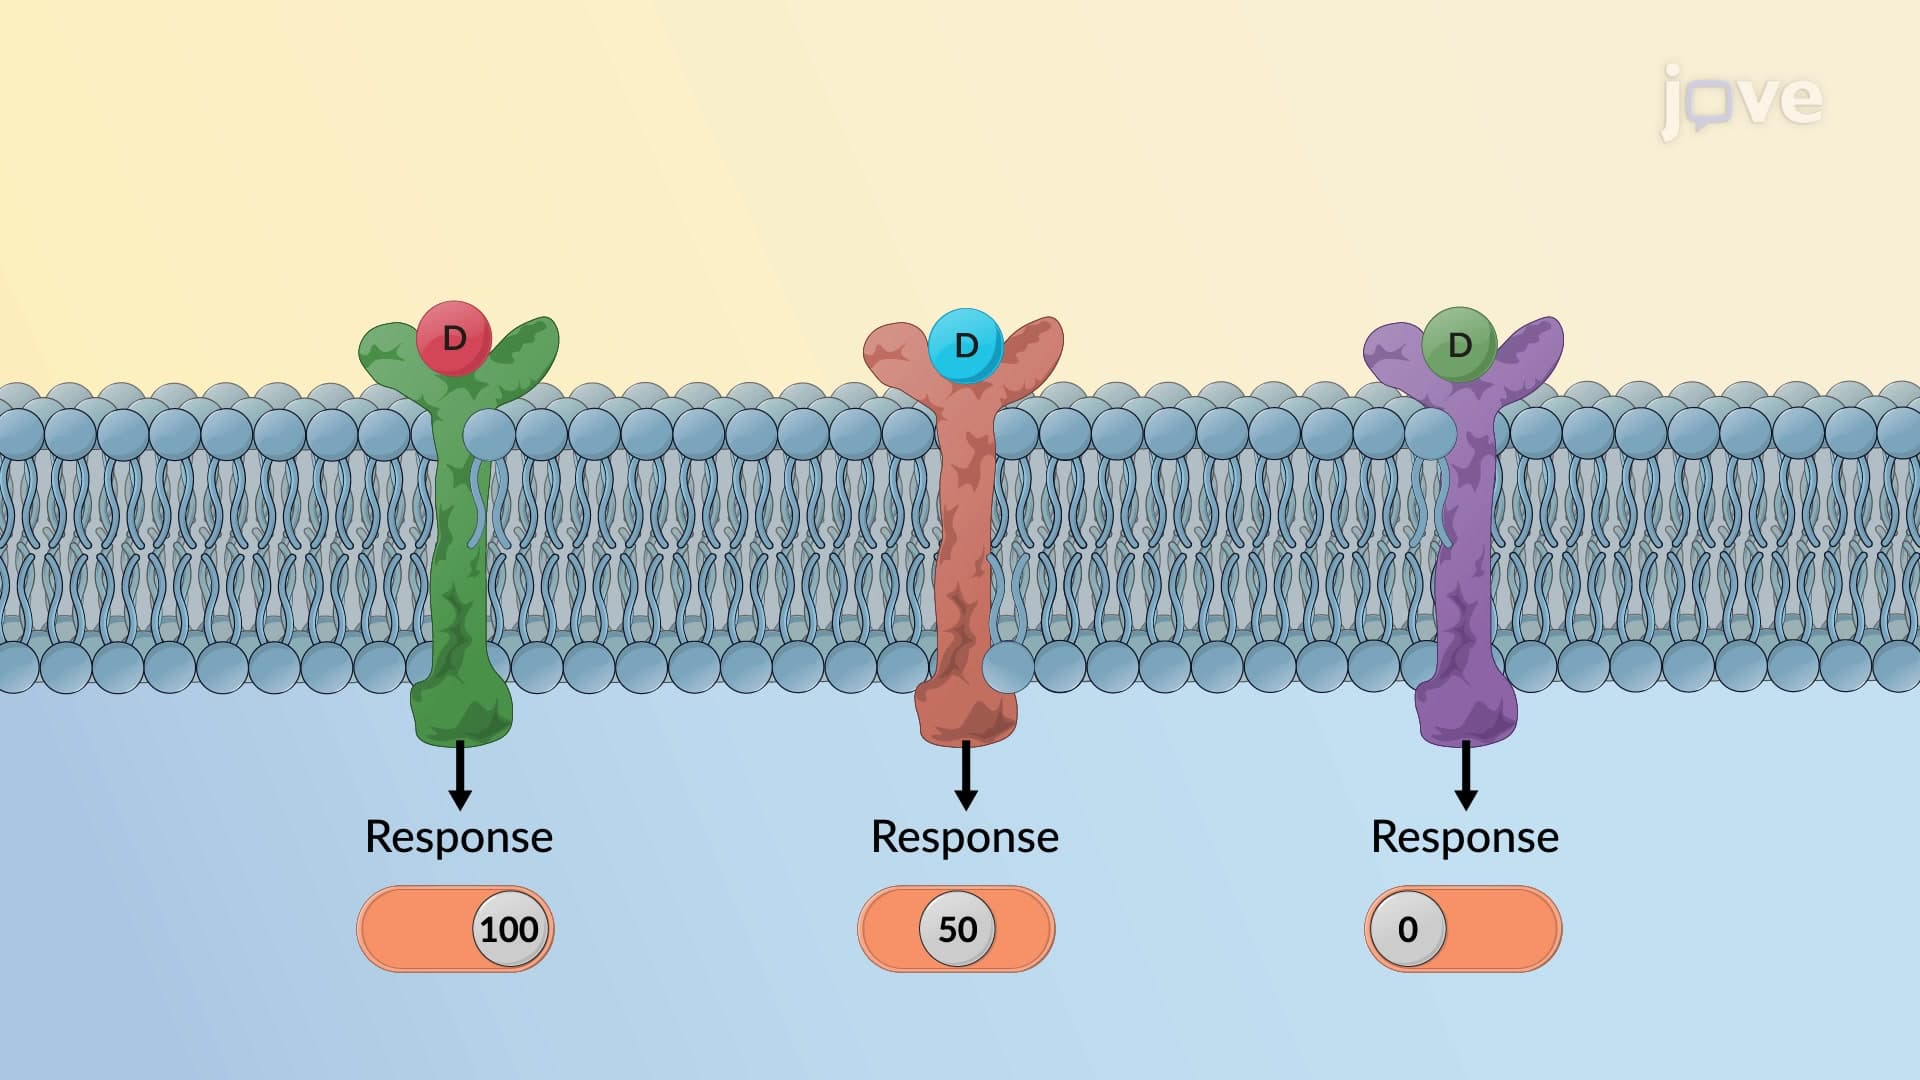
Drug-Receptor Interactions
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue....
 01:25
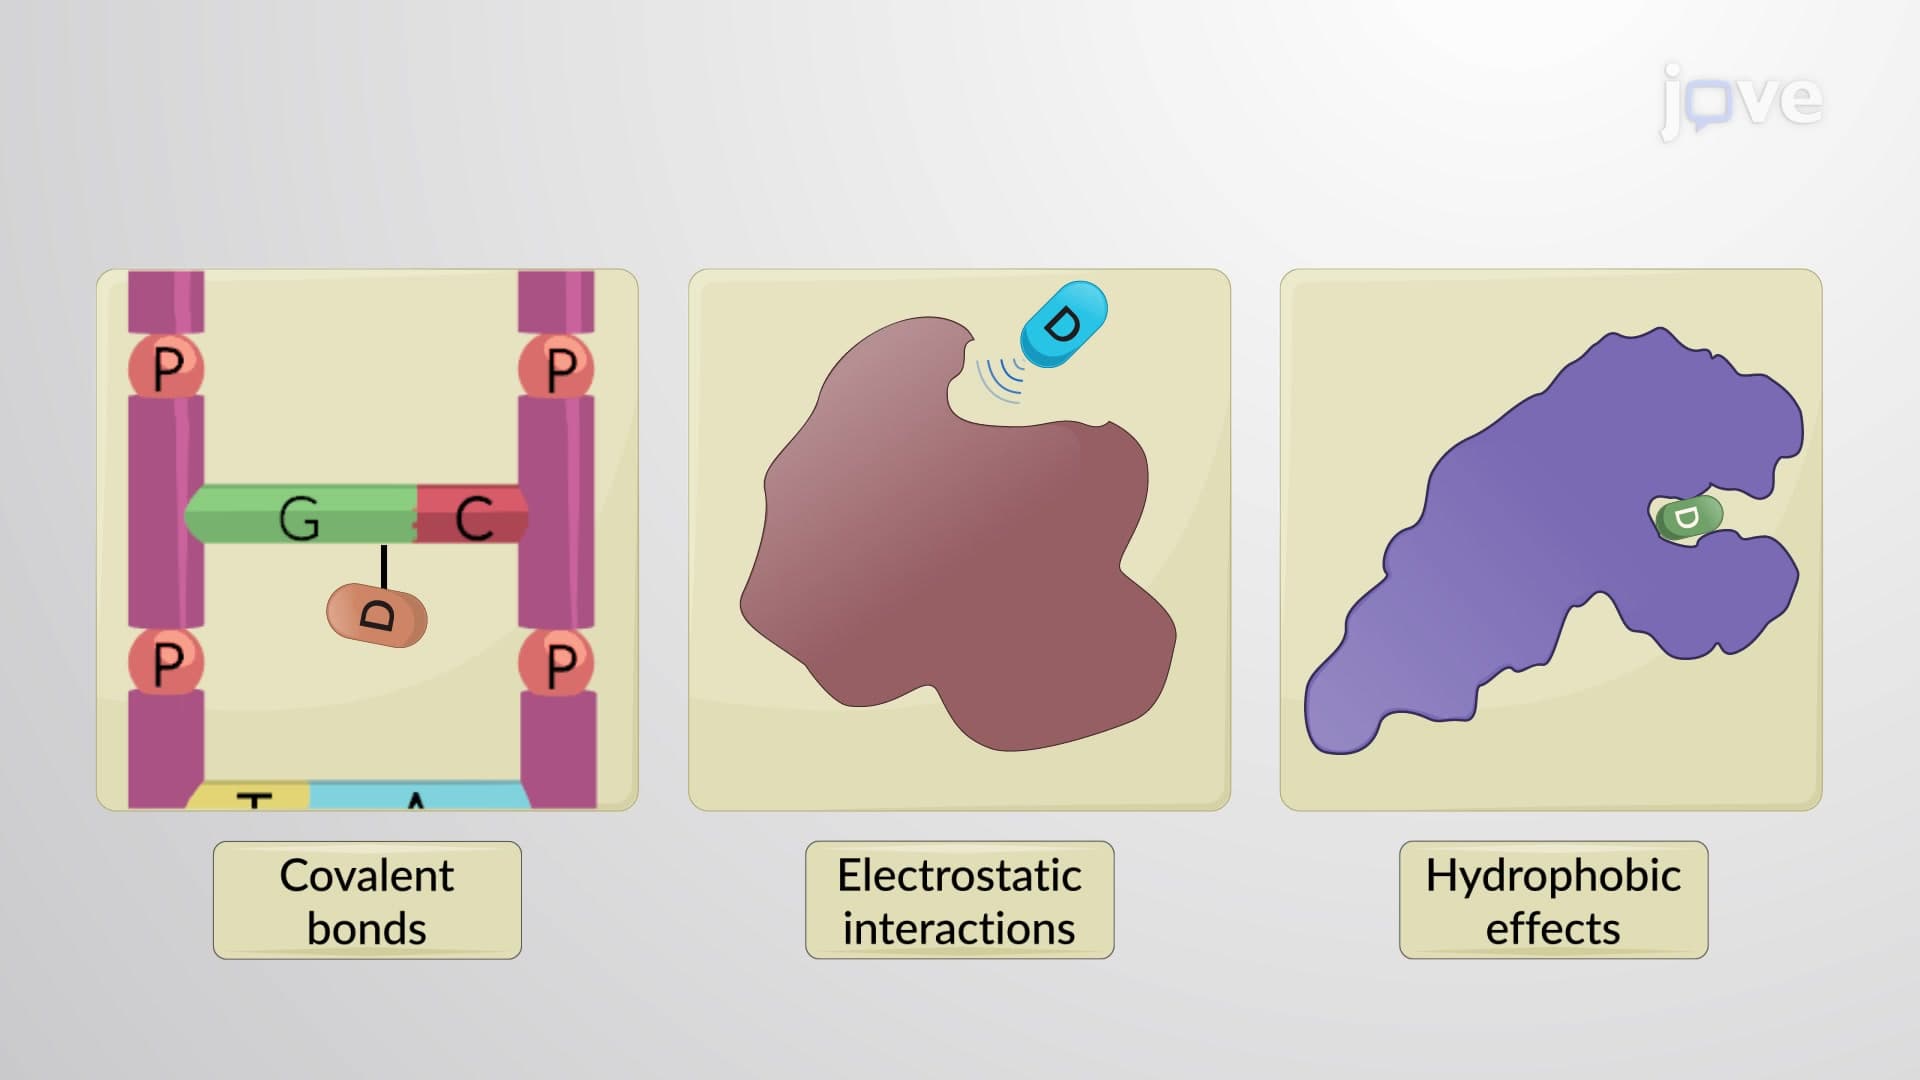
Drug-Receptor Bonds
In...
 01:26
Targets for Drug Action: Overview
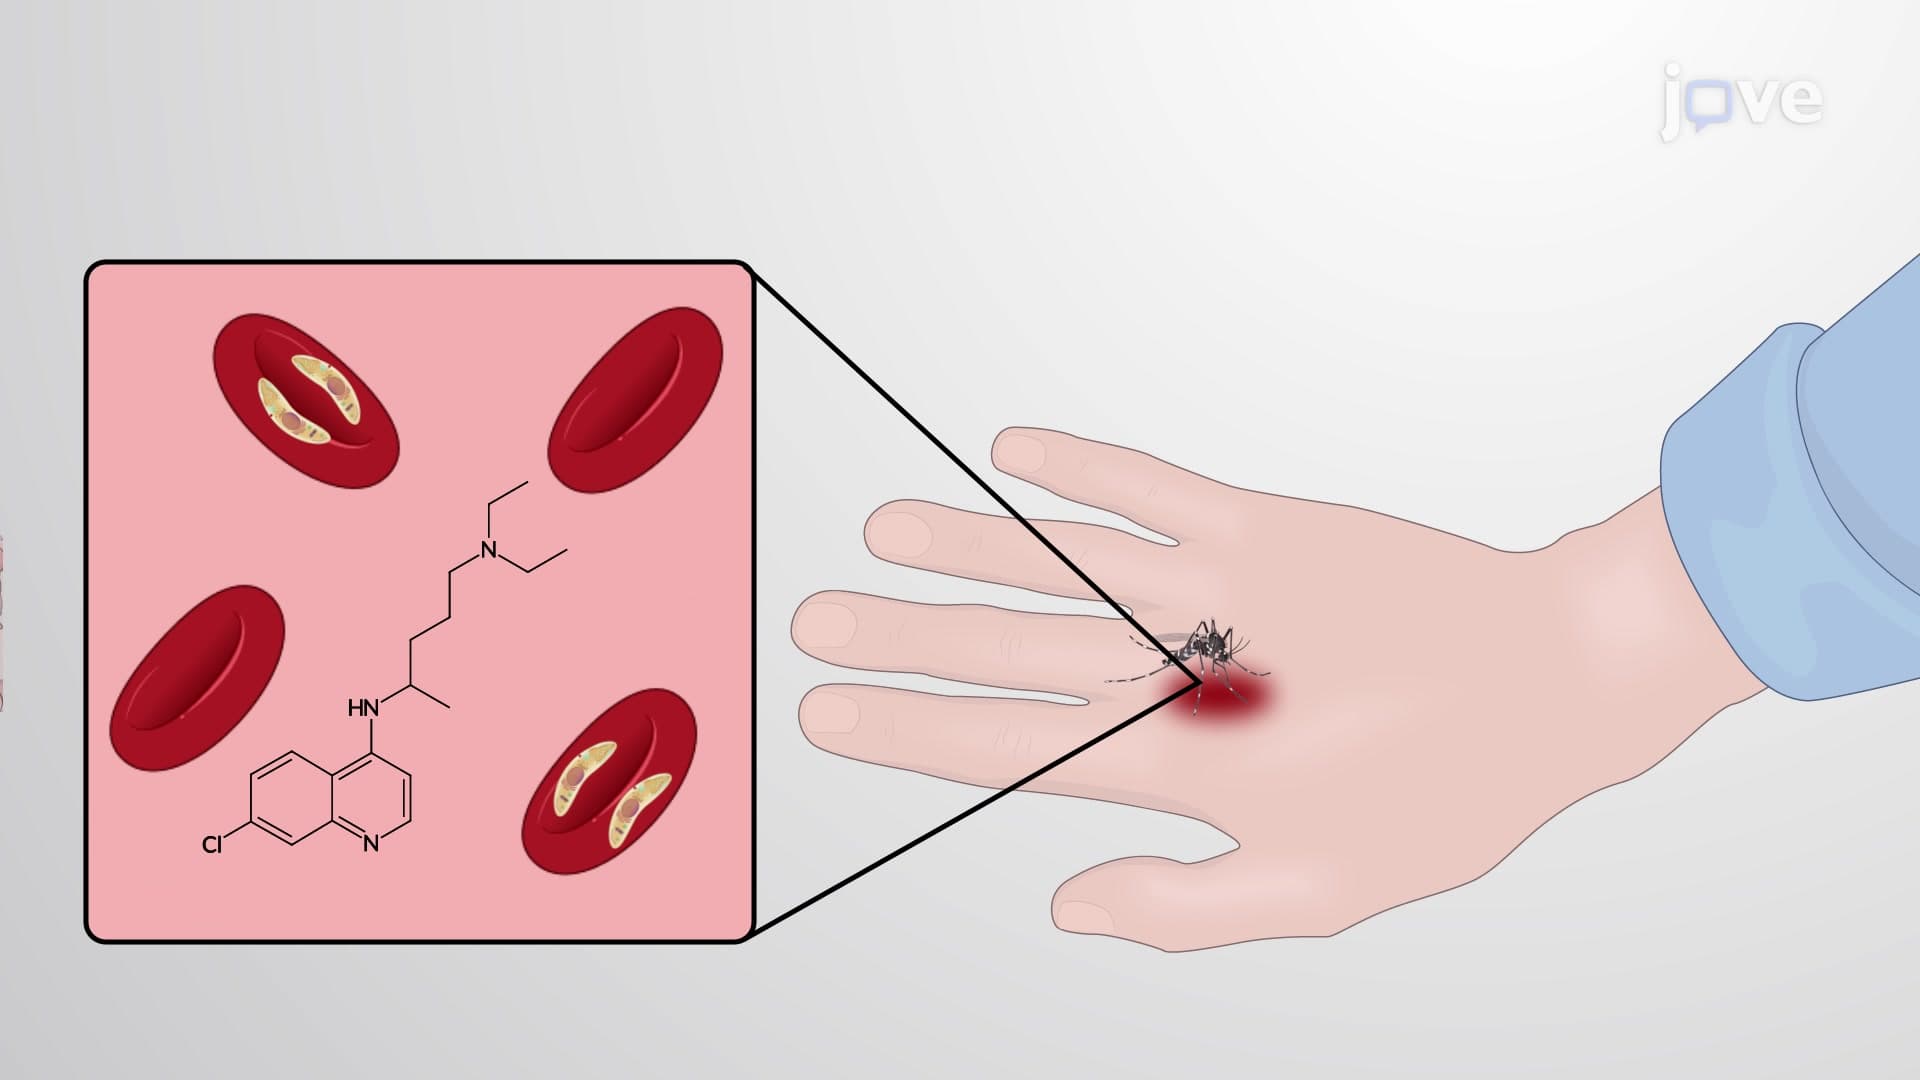
Receptors are either membrane-spanning or intracellular proteins, which upon binding a ligand, get activated and transmit the signal downstream to elicit a response. Drugs bind receptors, either mimicking the action of endogenous ligands or blocking the receptor activity to bring about a modified response. Nearly 35% of approved drugs target the G...
 01:24
Principles of Drug Action
Drugs can be agonists or antagonists. Like the endogenous ligands, agonists always bind and activate the target to produce a cellular response. Agonist binding induces a conformational change which in turn...
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...