Related Experiment Video
Updated: Jul 26, 2025

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Accuracy Meets Feasibility for the Structures and Rotational Constants of the Molecular Bricks of Life: A Joint
1Scuola Normale Superiore di Pisa, Piazza dei Cavalieri 7, 56126 Pisa, Italy.

Abstract:
A fully unsupervised computational protocol is proposed with the aim of obtaining reliable structural properties for molecular bricks of life in the gas phase. The results of the new composite scheme approach spectroscopic accuracy at a moderate cost without any empirical parameter in addition to those of the underlying electronic structure method. The whole workflow is fully automated and provides optimized geometries and equilibrium rotational constants. Direct comparison with experimental ground state rotational constants can be performed thanks to the effective computation of vibrational corrections in the framework of second-order vibrational perturbation theory. The results for all the nucleic acid bases and several flexible molecules of biological or medicinal interest show that the accuracy of the new tool is close to that delivered by state-of-the-art composite wave function methods for small semirigid molecules.
More Related Videos

Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 02:00
Molecular Models
 02:18
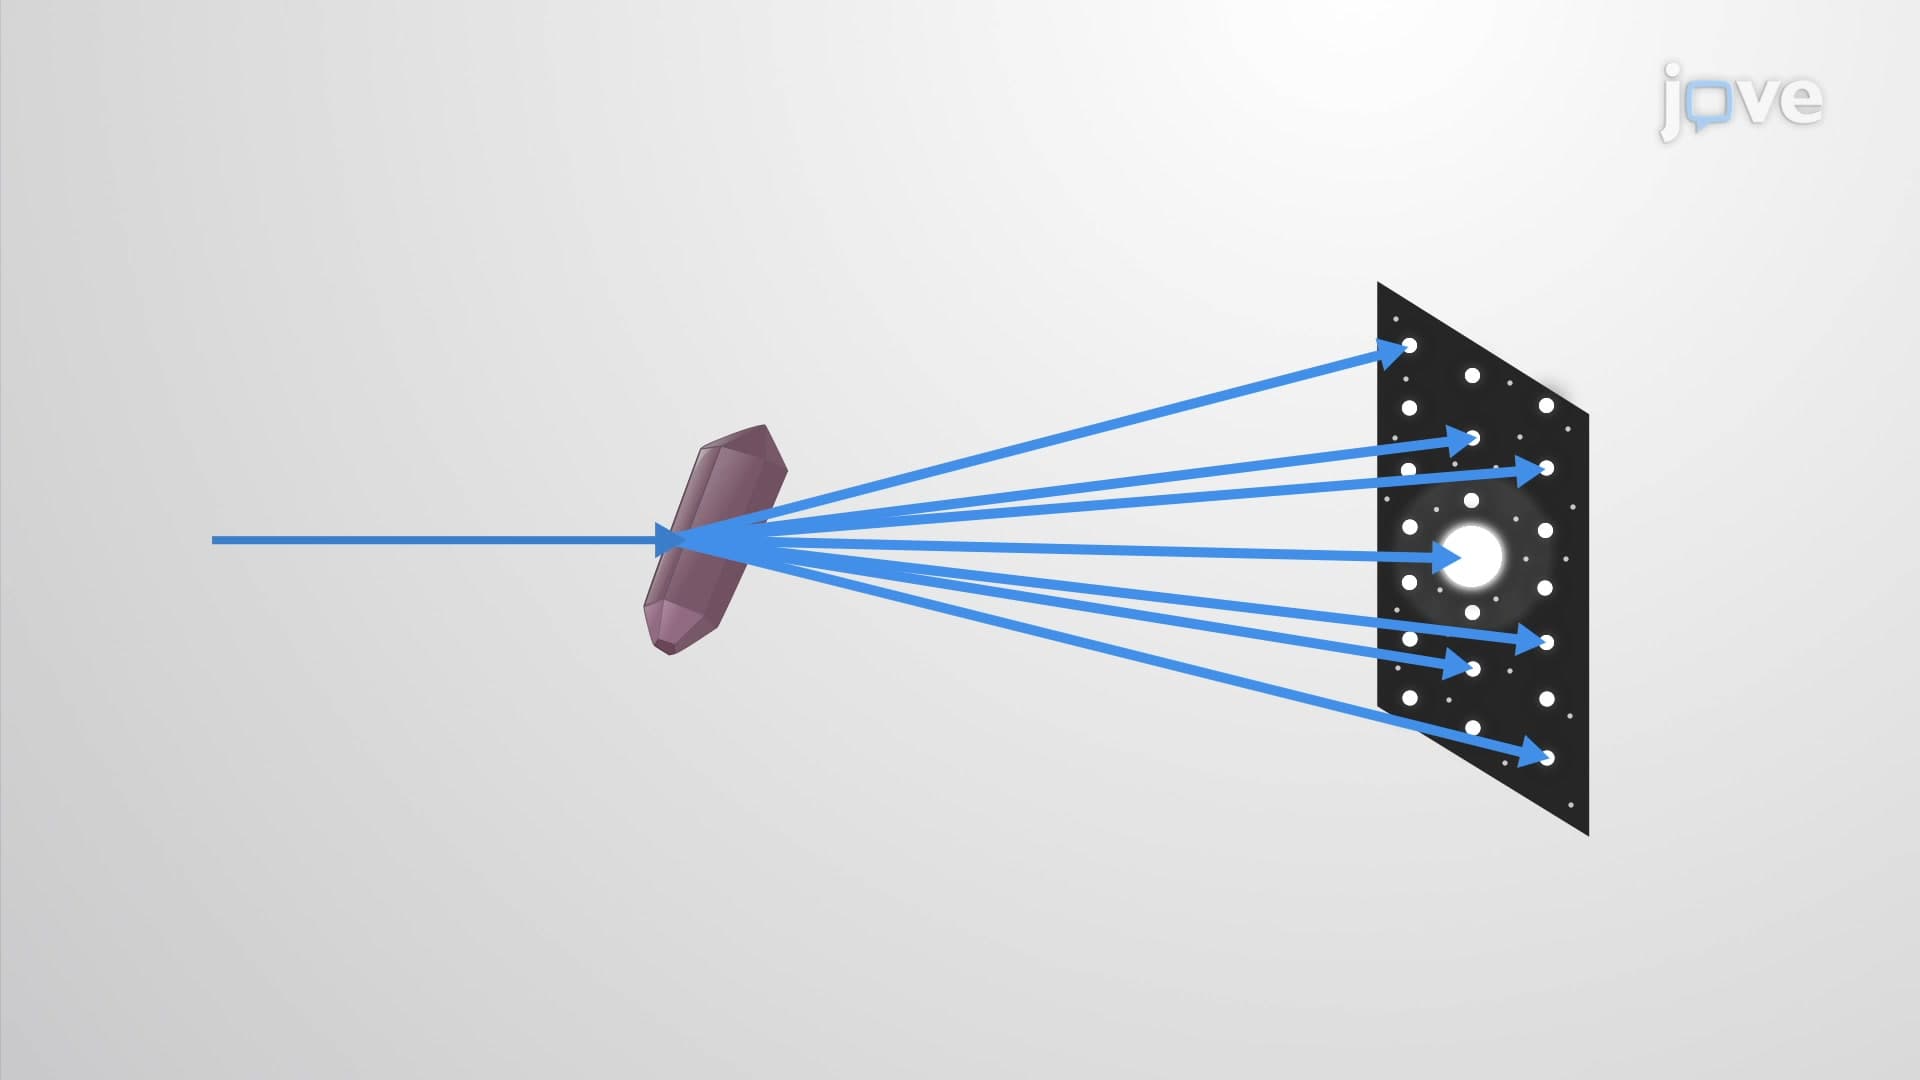
X-ray Crystallography
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:36
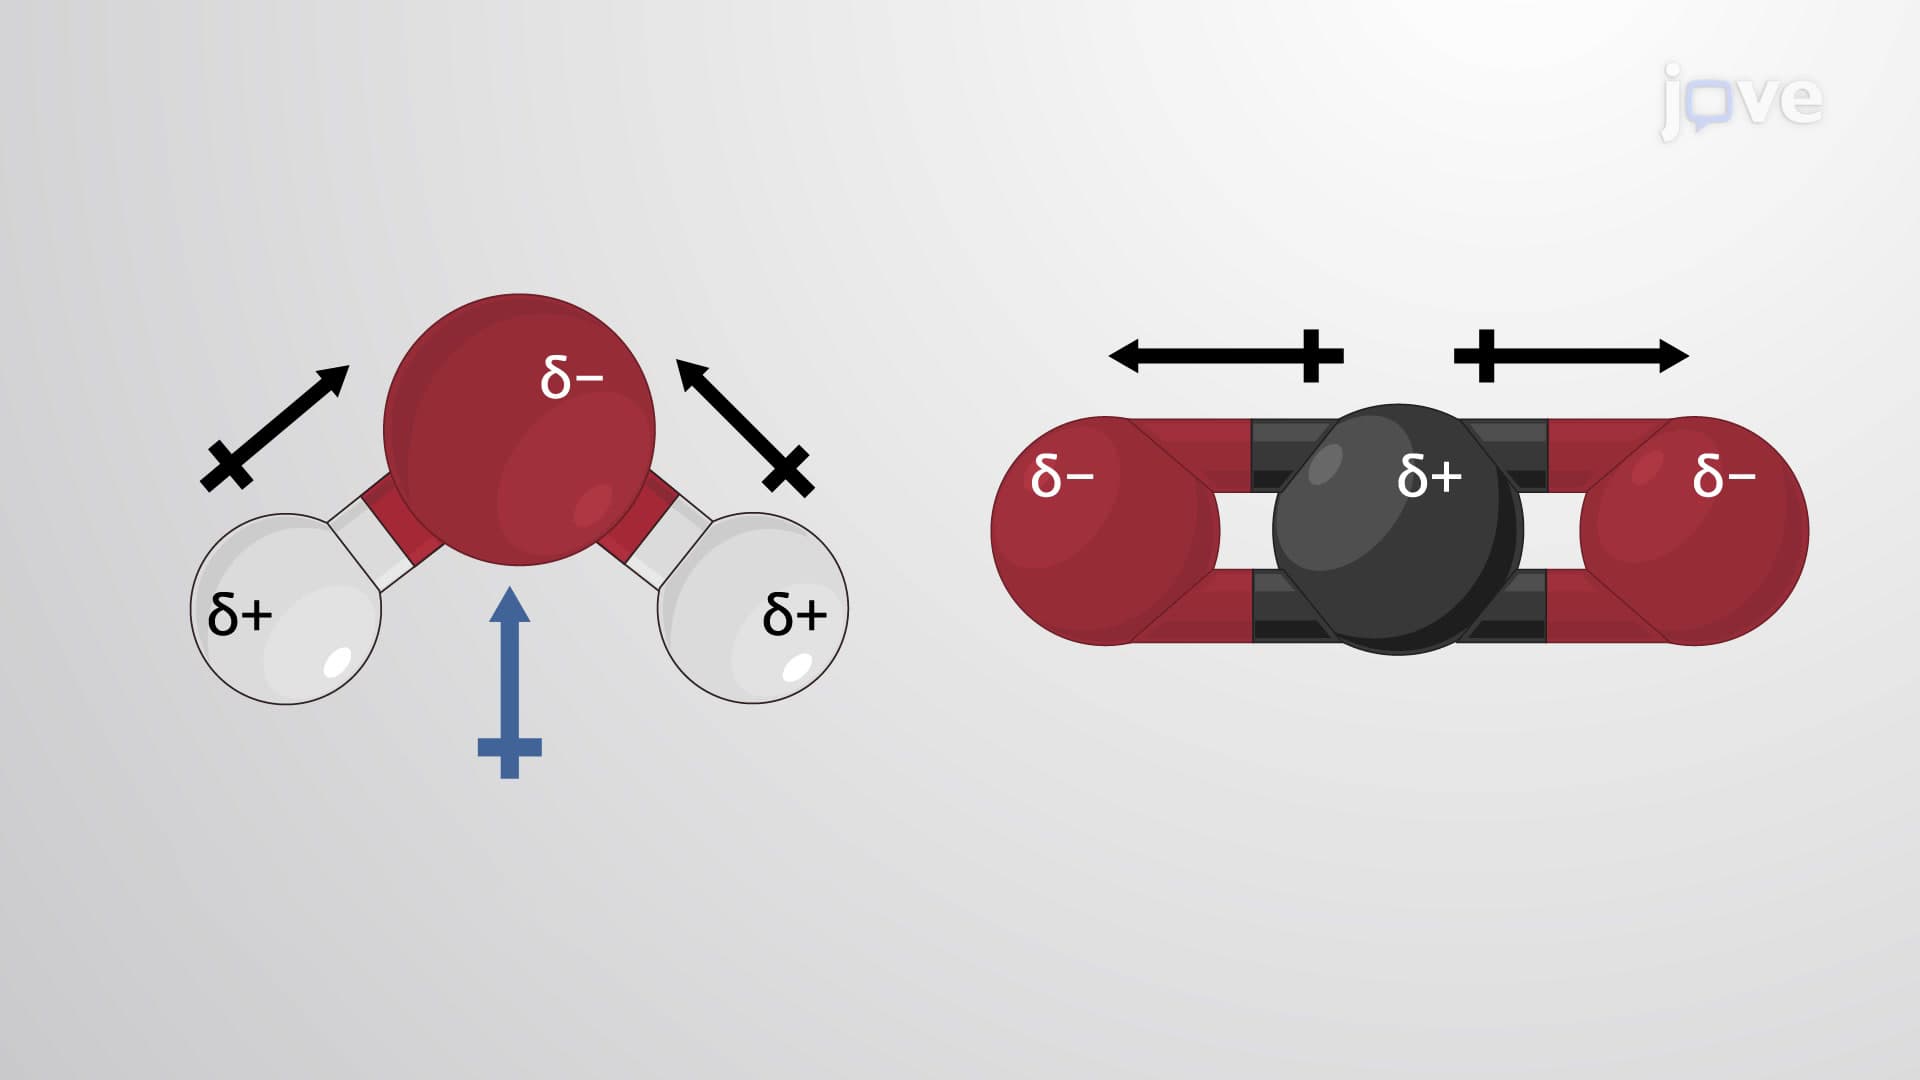
Molecular Geometry and Dipole Moments