Related Experiment Video
Updated: Jul 24, 2025
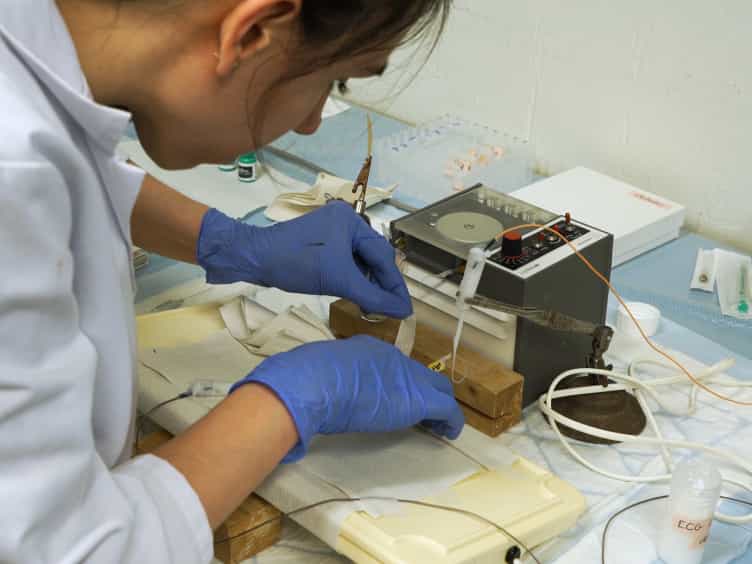
Nasal Potential Difference to Quantify Trans-epithelial Ion Transport in Mice
Published on: July 4, 2018
Nonsense mutations accelerate lung disease and decrease survival of cystic fibrosis children
Annalisa Orenti1, Iwona Pranke2, Caroline Faucon3
1Department of Clinical Sciences and Community Health, Laboratory of Medical Statistics, Biometry and Epidemiology "G. A. Maccacaro", University of Milan, Milan, Italy.

Insights
People with Cystic Fibrosis (pwCF) with two nonsense mutations (PTC/PTC) experience faster lung function decline and higher mortality. This study highlights the severe impact of PTC/PTC mutations on CF progression and survival.
Area of Science:
- Pulmonology
- Genetics
- Medical Research
Background:
- Limited data exists on the clinical status of individuals with Cystic Fibrosis (CF) who carry two nonsense mutations (PTC/PTC).
- Understanding the disease severity associated with different CFTR mutation types is crucial for patient management and therapeutic development.
Purpose of the Study:
- To compare disease severity in people with Cystic Fibrosis (pwCF) with two nonsense mutations (PTC/PTC) against those with compound heterozygous (F508del/PTC) and homozygous (F508del/F508del) F508del mutations.
- To assess CFTR mRNA and protein activity in nasal epithelial cells of pwCF with PTC/PTC mutations.
Main Methods:
- Retrospective analysis of clinical data from the European CF Society Patient Registry for pwCF in high and middle-income countries.
- Comparison of disease progression metrics (e.g., FEV1 decline, mortality, Pseudomonas aeruginosa infection rates) across different CFTR mutation groups.
- Assessment of CFTR mRNA and protein activity in primary human nasal epithelial (HNE) cells from 22 pwCF with PTC/PTC mutations.
Main Results:
- pwCF with PTC/PTC and F508del/PTC mutations showed a significantly faster decline in Forced Expiratory Volume in 1 second (FEV1) from childhood into adulthood compared to F508del/F508del.
- Mortality rates were significantly higher in pediatric pwCF with one or two PTC alleles versus F508del homozygous pairs.
- Pseudomonas aeruginosa infection was more frequent in PTC/PTC individuals, and CFTR activity in their nasal cells was minimal (0-3% of wild-type).
Conclusions:
- Nonsense mutations, particularly in the PTC/PTC genotype, significantly decrease survival and accelerate respiratory disease progression in children and adolescents with Cystic Fibrosis.
- These findings underscore the severe clinical impact of PTC/PTC mutations in CF.
- Targeted therapies for PTC/PTC mutations are warranted to improve outcomes.
Rationale:
Limited information is available on the clinical status of people with Cystic Fibrosis (pwCF) carrying 2 nonsense mutations (PTC/PTC). The main objective of this study was to compare disease severity between pwCF PTC/PTC, compound heterozygous for F508del and PTC (F508del/PTC) and homozygous for F508del (F508del+/+).
Methods:
Based on the European CF Society Patient Registry clinical data of pwCF living in high and middle income European and neighboring countries, PTC/PTC (n = 657) were compared with F508del+/+ (n = 21,317) and F508del/PTC(n = 4254).CFTR mRNA and protein activity levels were assessed in primary human nasal epithelial (HNE) cells sampled from 22 PTC/PTC pwCF.
Main Results:
As compared to F508del+/+ pwCF; both PTC/PTC and F508del/PTC pwCF exhibited a significantly faster rate of decline in Forced Expiratory Volume in 1 s (FEV1) from 7 years (-1.33 for F508del +/+, -1.59 for F508del/PTC; -1.65 for PTC/PTC, p < 0.001) until respectively 30 years (-1.05 for F508del +/+, -1.23 for PTC/PTC, p = 0.048) and 27 years (-1.12 for F508del +/+, -1.26 for F508del/PTC, p = 0.034). This resulted in lower FEV1 values in adulthood. Mortality of pediatric pwCF with one or two PTC alleles was significantly higher than their F508del homozygous pairs. Infection with Pseudomonas aeruginosa was more frequent in PTC/PTC versus F508del+/+ and F508del/PTC pwCF. CFTR activity in PTC/PTC pwCF's HNE cells ranged between 0% to 3% of the wild-type level.
Conclusions:
Nonsense mutations decrease the survival and accelerate the course of respiratory disease in children and adolescents with Cystic Fibrosis.
More Related Videos
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:39
Mutations
 02:27
Nonsense-mediated mRNA Decay
Usually, Upf3 binds to an Exon Junction Complex (EJC) at mRNA splice sites. If a ribosome fully translates the mRNA,...
 02:46
Genome Copying Errors
 02:07
Tumor Progression
Colon cancer is one of the best-documented examples of tumor progression. Early mutation in the APC gene in colon cells causes a small growth on the colon wall called a polyp. With time, this polyp grows into a benign, pre-cancerous tumor. Further...