Related Experiment Video
Updated: Jul 22, 2025

Fabricating van der Waals Heterostructures with Precise Rotational Alignment
Published on: July 5, 2019
First-principles Calculations Reveal Frictional Advantage for C2 N/C6 N6 van der Waals Heterostructures
Moumita Mukherjee1, Sucharita Mandal1, Ayan Datta1
1School of Chemical Sciences, Indian Association for the Cultivation of Science, Jadavpur, Kolkata, 700032, India.

Abstract:
Friction at the atomic scale is determined for three different carbon nitride structures namely C2 N/C2 N, C6 N6 /C6 N6 and C6 N6 /C2 N employing ab-initio density functional theory (DFT). The sliding path along the lowest energy corrugations determines the static frictional forces. Both the homo-layer structures (C2 N/C2 N and C6 N6 /C6 N6 ) have higher corrugation energy and correspondingly higher static lateral forces with respect to the hetero-layer structure (C2 N/C6 N6 ). The corrugation energy for the C2 N/C6 N6 heterostructure ( =0.29 meV/atom) is one-order lower than C2 N/C2 N ( =2.08 meV/atom) and C6 N6 /C6 N6 ( =4.37 meV/atom). Such a significantly lower corrugation energy for the heterostructure arises due to the reduced fluctuation in the interfacial charge density along the sliding pathway. Moreover, the change in the interlayer distance along the sliding pathway is only 0.2 Å for the heterostructure while its 0.3 Å and 0.4 Å for C2 N and C6 N6 homo-layers respectively. The friction coefficients (FL /FN , FL =static lateral force; FN =normal force) decrease with increasing load for all the systems with the lowest value (0.04) for C2 N/C6 N6 at 2 GPa. The van der Waals heterostructures are, therefore, predicted to be highly efficient lubricant materials for reducing friction at the atomic scale.
More Related Videos


Related Concept Videos
 02:18
Network Covalent Solids
To break or to melt a covalent network solid, covalent bonds must be broken. Because covalent bonds are relatively strong, covalent network solids are typically...
 01:24
Van der Waals Interactions
 01:10
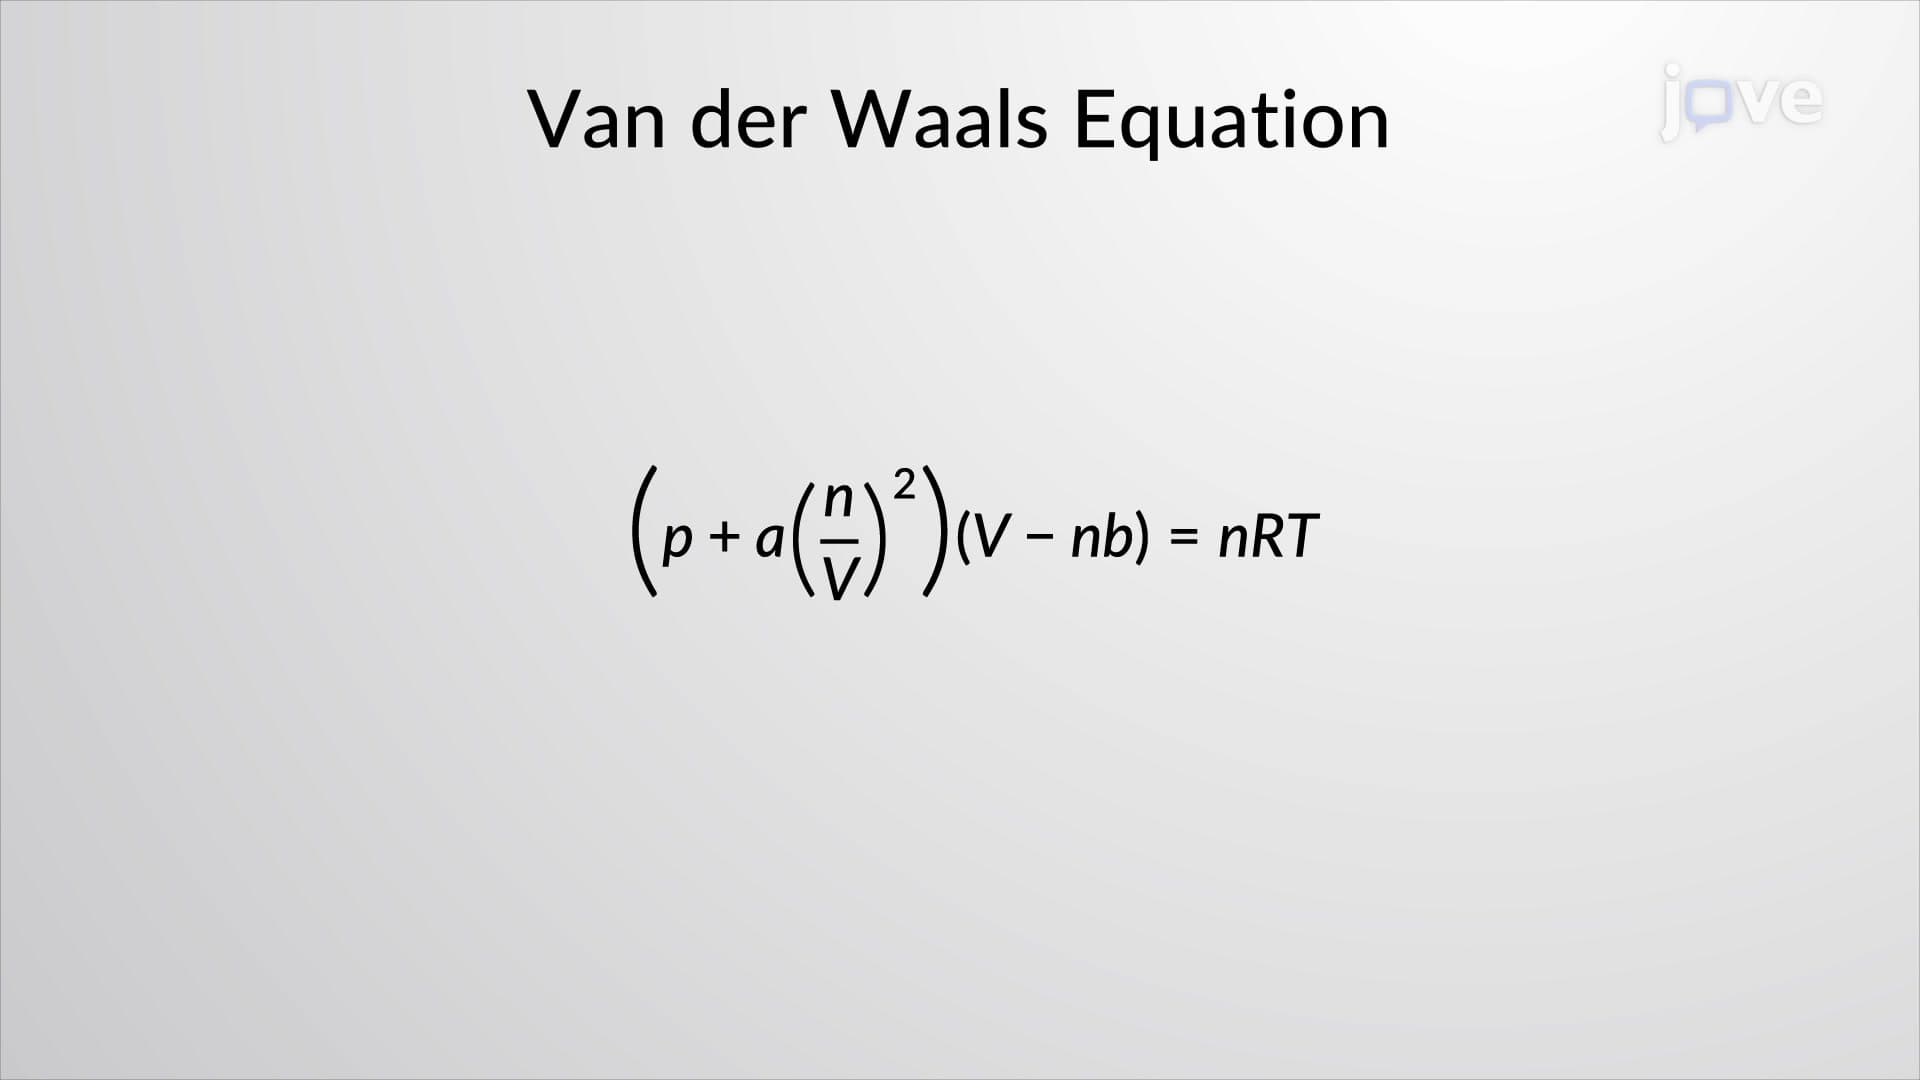
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:22
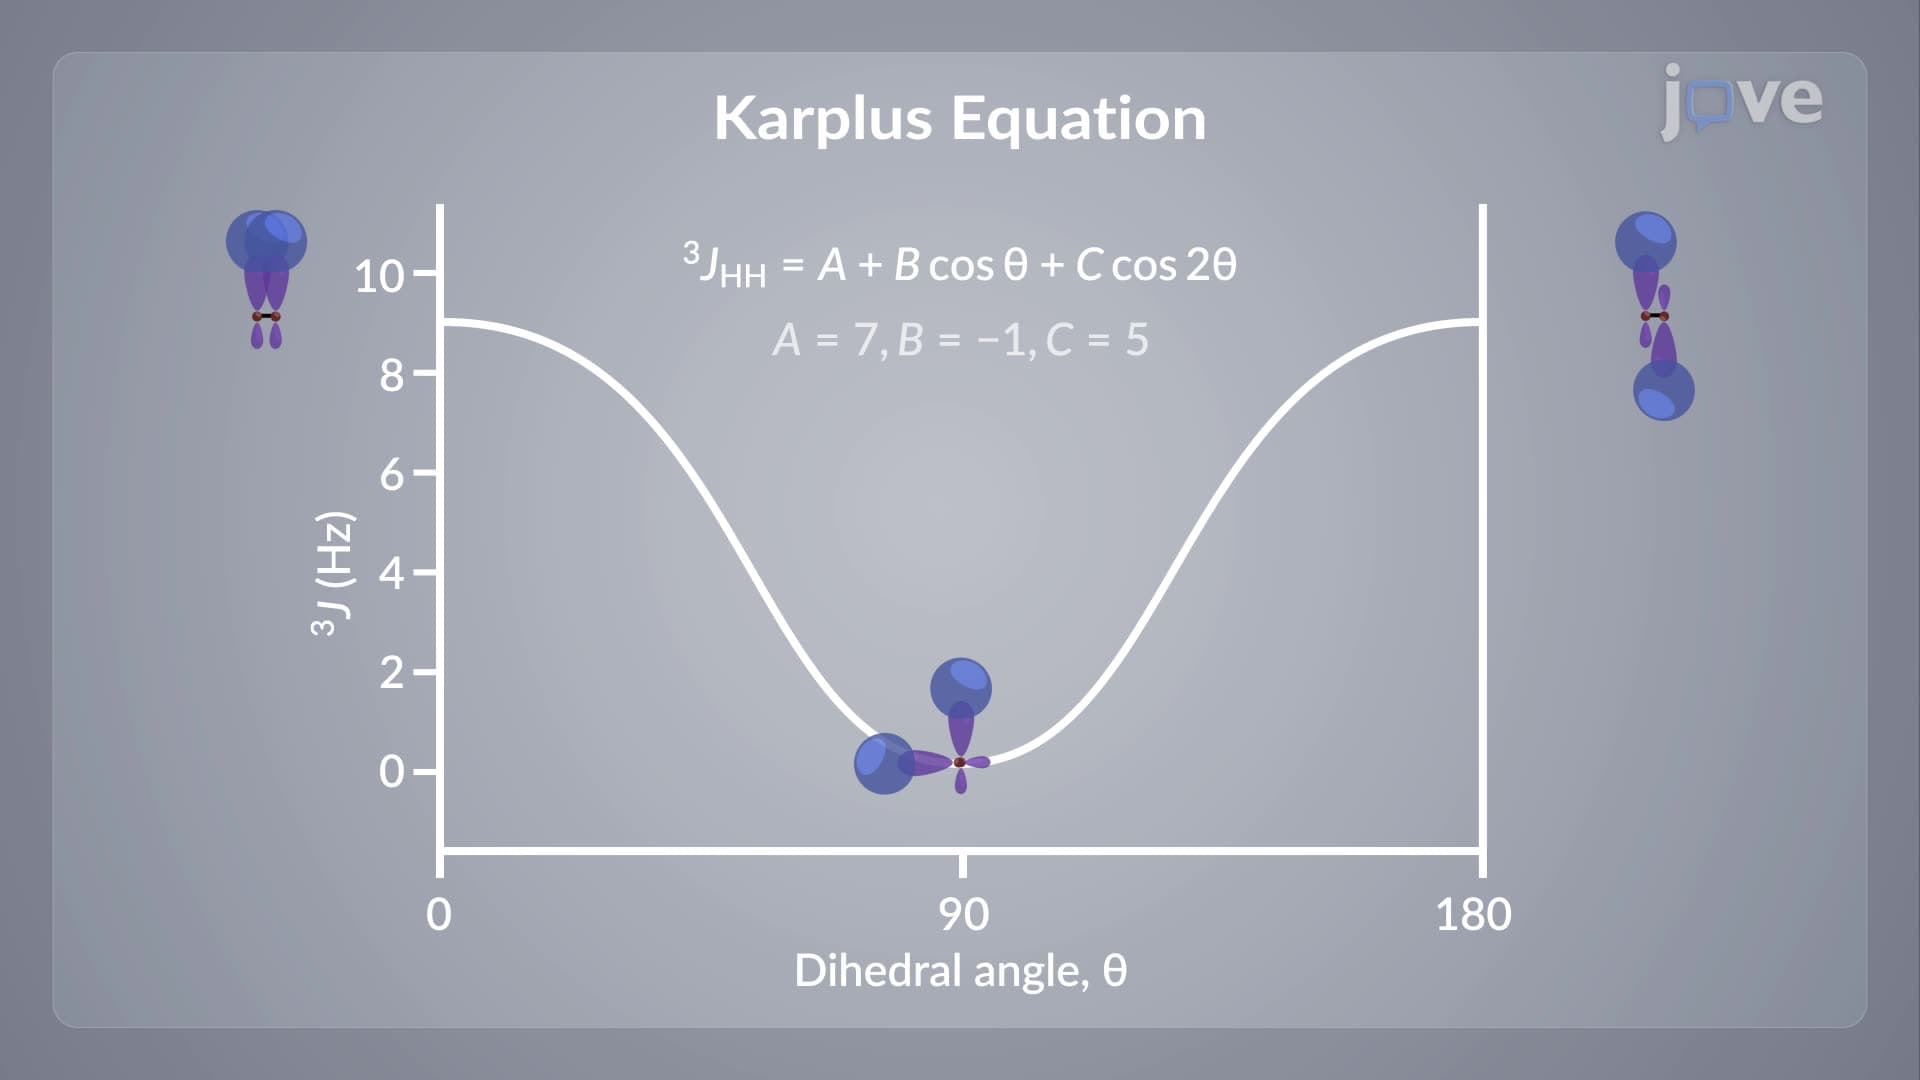
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 03:35
Hybridization of Atomic Orbitals II
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...