

Assessment of DFT functionals for a minimal nitrogenase [Fe(SH)4H]- model employing state-of-the-art ab initio
Victor P Vysotskiy1, Magne Torbjörnsson1, Hao Jiang1
1Department of Computational Chemistry, Lund University, Chemical Centre, SE-221 00 Lund, Sweden.

Abstract:
We have designed a [Fe(SH)4H]- model with the fifth proton binding either to Fe or S. We show that the energy difference between these two isomers (∆E) is hard to estimate with quantum-mechanical (QM) methods. For example, different density functional theory (DFT) methods give ∆E estimates that vary by almost 140 kJ/mol, mainly depending on the amount of exact Hartree-Fock included (0%-54%). The model is so small that it can be treated by many high-level QM methods, including coupled-cluster (CC) and multiconfigurational perturbation theory approaches. With extrapolated CC series (up to fully connected coupled-cluster calculations with singles, doubles, and triples) and semistochastic heat-bath configuration interaction methods, we obtain results that seem to be converged to full configuration interaction results within 5 kJ/mol. Our best result for ∆E is 101 kJ/mol. With this reference, we show that M06 and B3LYP-D3 give the best results among 35 DFT methods tested for this system. Brueckner doubles coupled cluster with perturbaitve triples seems to be the most accurate coupled-cluster approach with approximate triples. CCSD(T) with Kohn-Sham orbitals gives results within 4-11 kJ/mol of the extrapolated CC results, depending on the DFT method. Single-reference CC calculations seem to be reasonably accurate (giving an error of ∼5 kJ/mol compared to multireference methods), even if the D1 diagnostic is quite high (0.25) for one of the two isomers.
More Related Videos
 06:45
Author Spotlight: Characterizing Porous Materials for Aiding the Development of Robust Metal-Organic Frameworks with Adsorption Behavior
Published on: March 8, 2024
 10:01
Protein Film Infrared Electrochemistry Demonstrated for Study of H2 Oxidation by a [NiFe] Hydrogenase
Published on: December 4, 2017
Related Concept Videos
 04:01
VSEPR Theory and the Effect of Lone Pairs
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:22
Inorganic Nitrogen Assimilation
 01:16
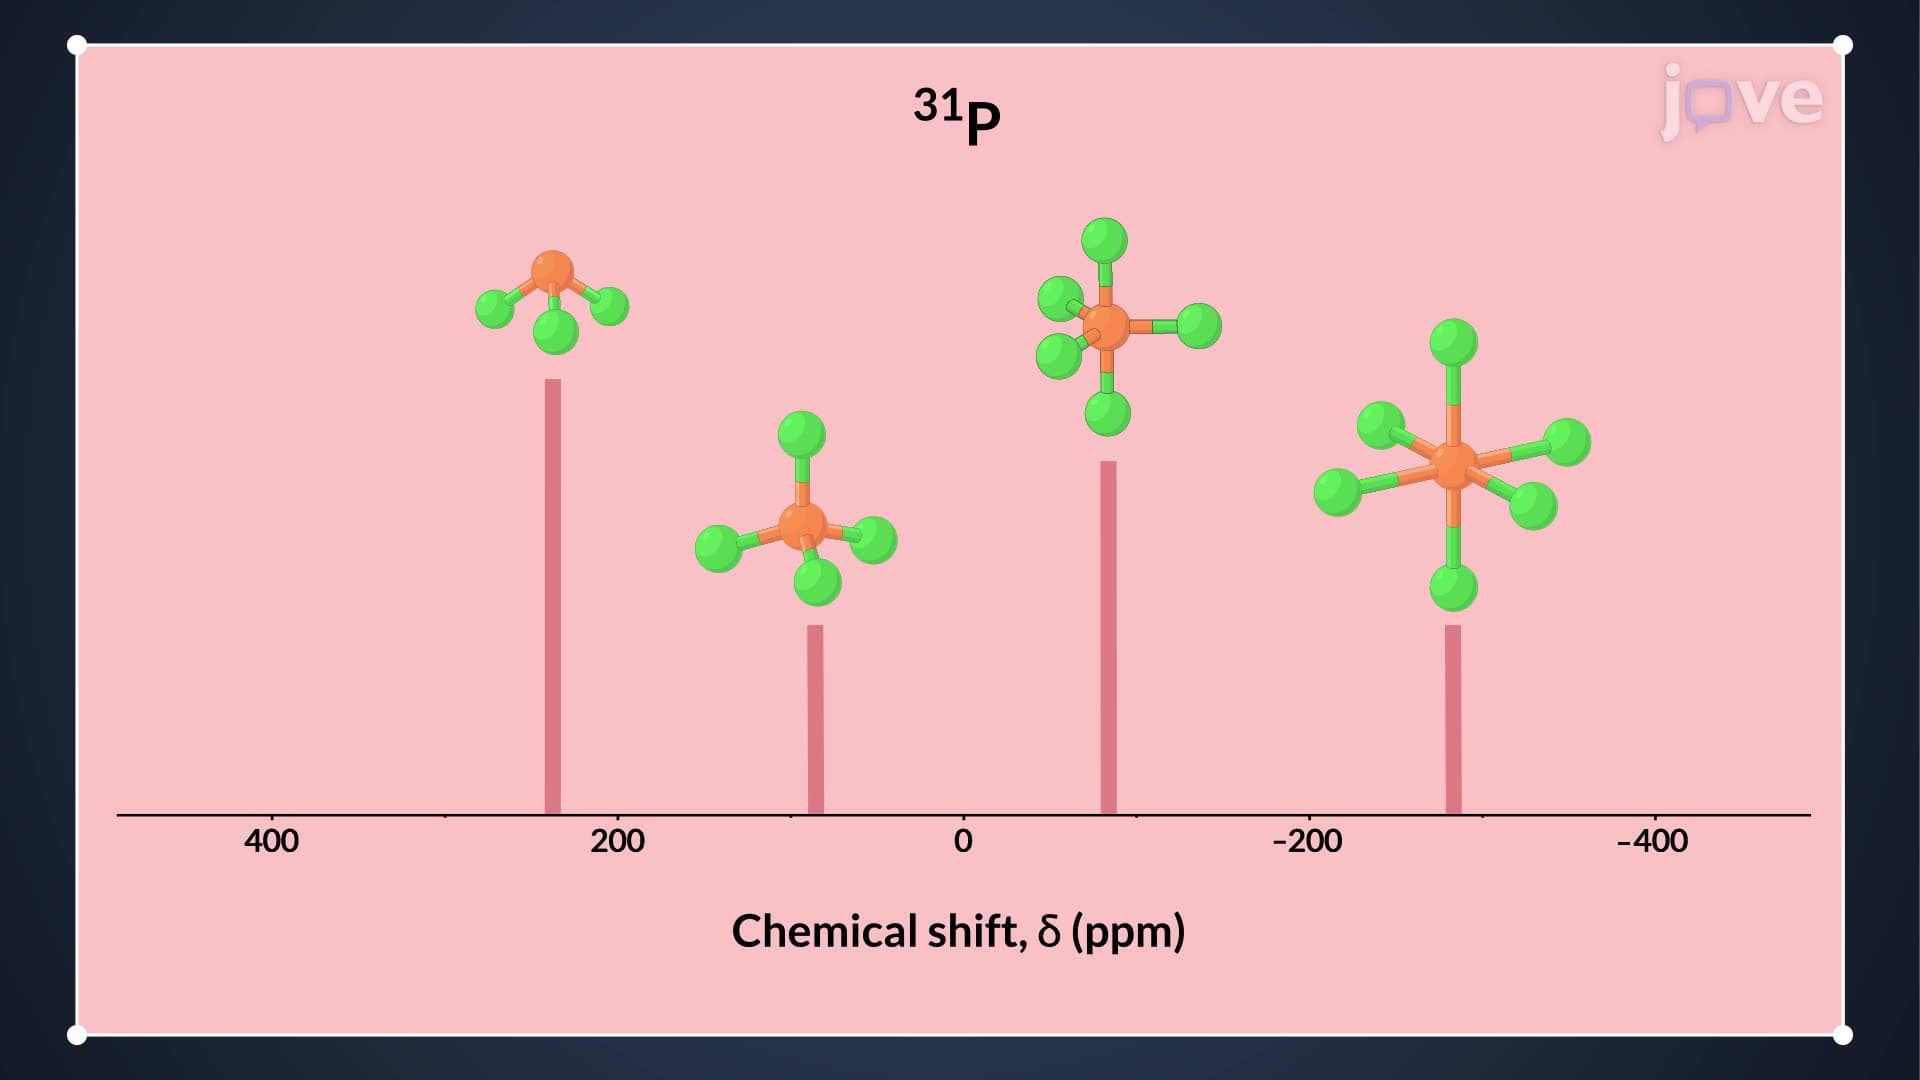
Other Nuclides: 31P, 19F, 15N NMR
While fluorine-19 and phosphorous-31 have high natural abundances (100%) and positive gyromagnetic ratios, nitrogen-15 has a low natural abundance and a negative gyromagnetic ratio. However, nitrogen-15 is still preferred over nitrogen-14 (which has a...
 01:19
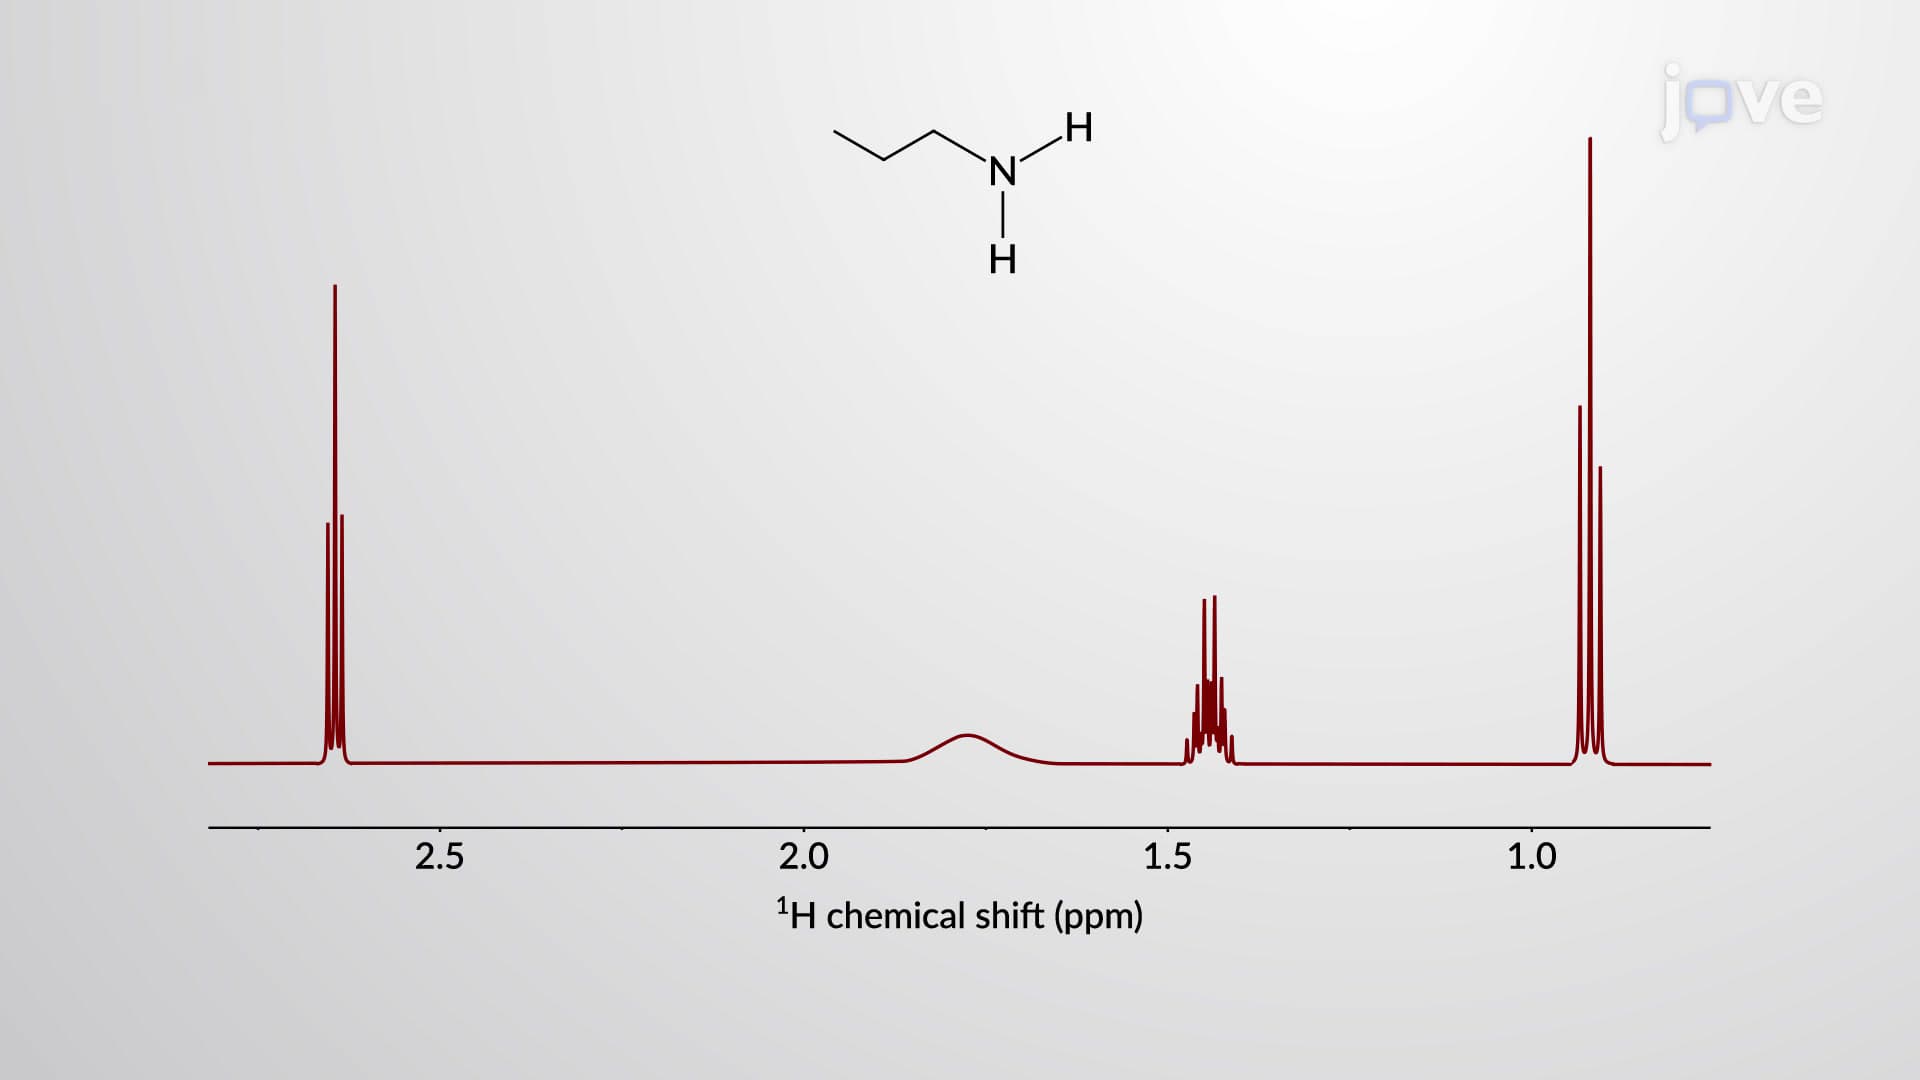
NMR Spectroscopy Of Amines
 01:19
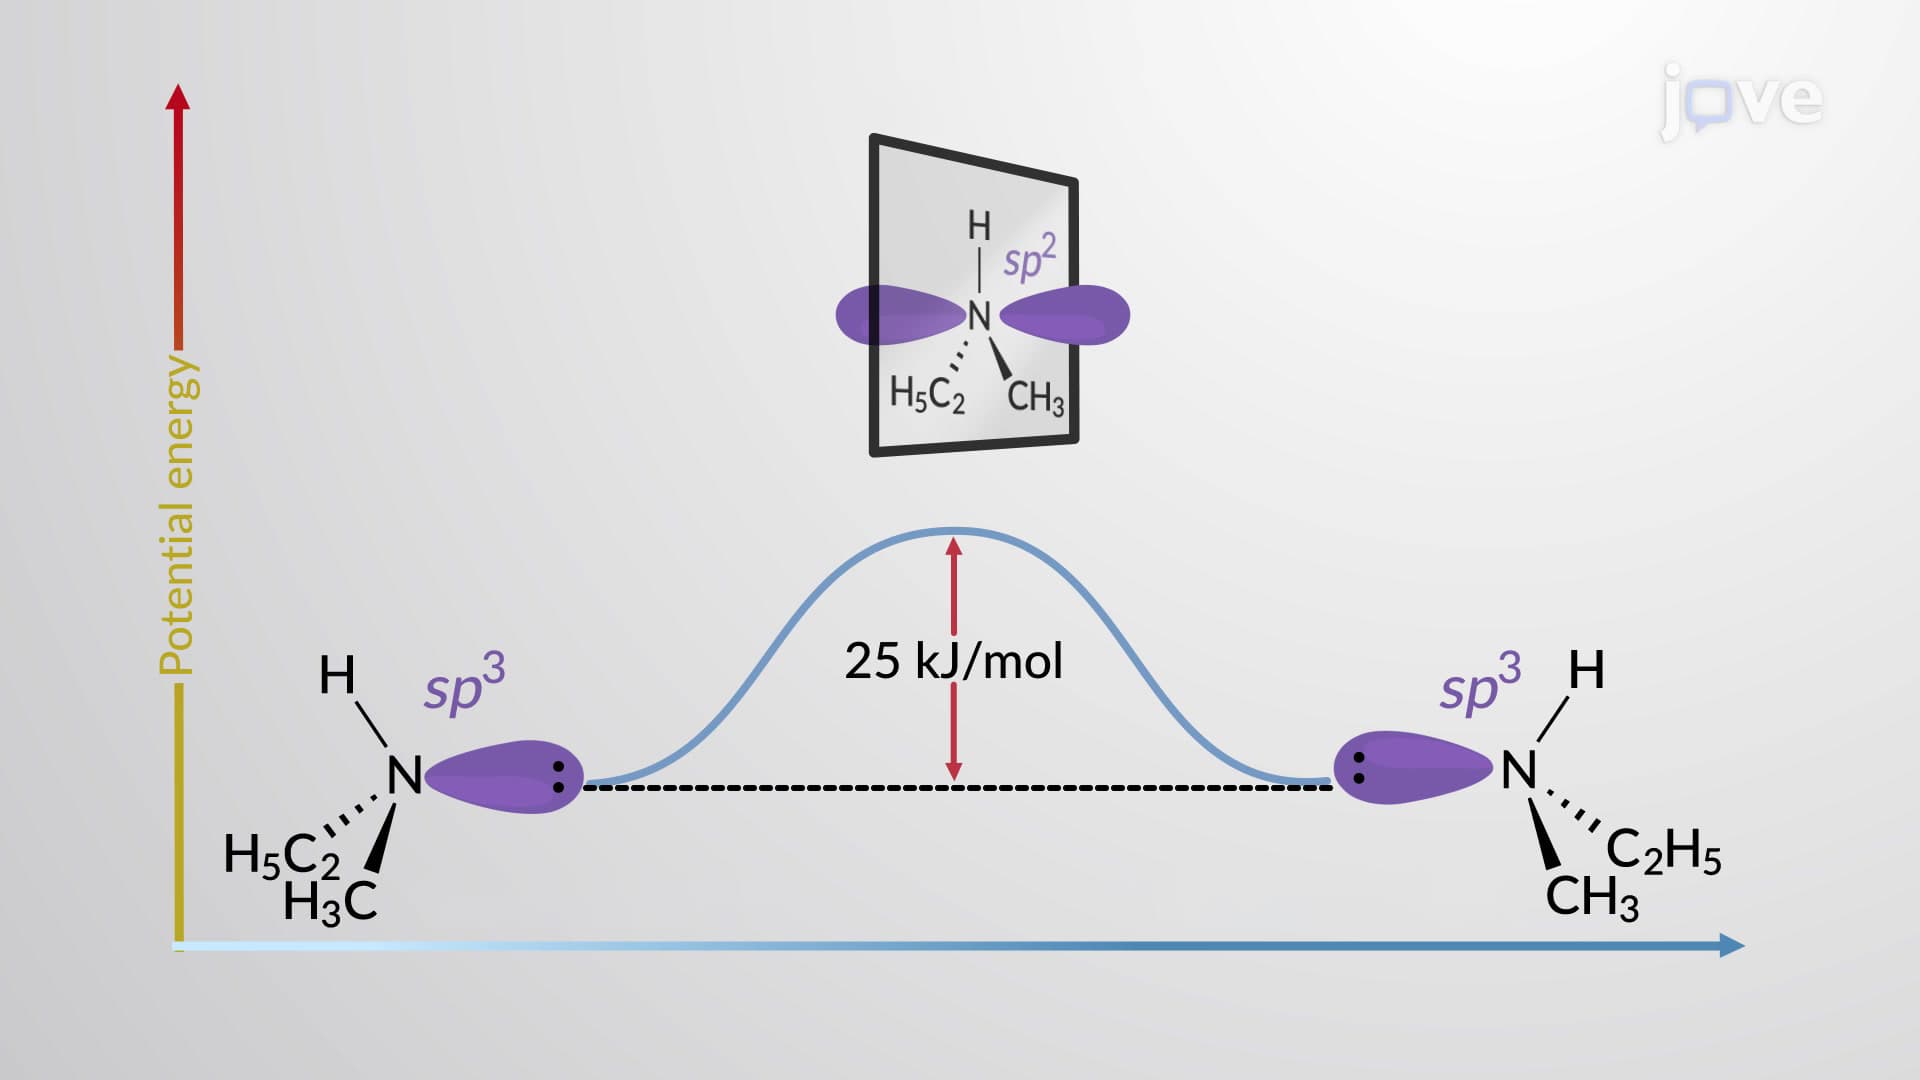
Structure of Amines