Related Experiment Video
Updated: Jul 21, 2025
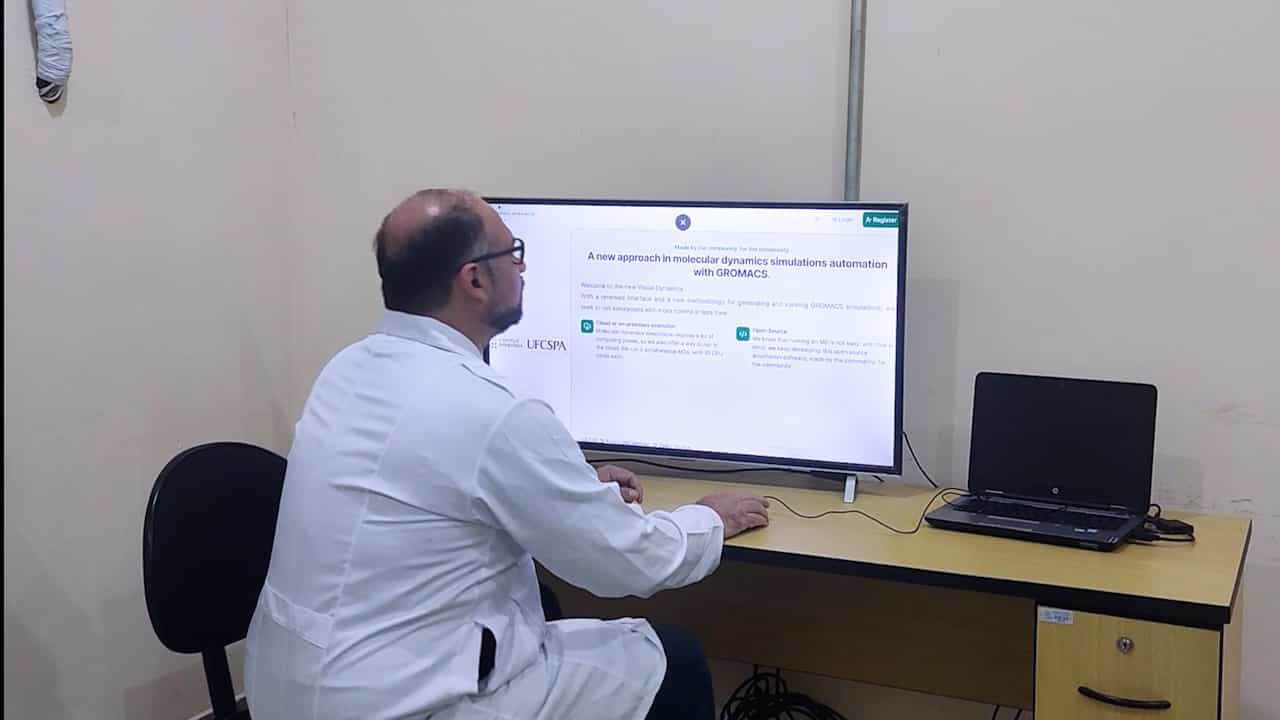

Author Spotlight: Streamlining Visual Dynamics to Simplify Molecular Dynamics Simulations Using Gromacs
Published on: August 9, 2024
VMD as a Platform for Interactive Small Molecule Preparation and Visualization in Quantum and Classical Simulations
Mariano Spivak1, John E Stone1, João Ribeiro1
1Theoretical and Computational Biophysics Group, NIH Resource for Macromolecular Modeling and Visualization, Beckman Institute for Advanced Science and Technology, University of Illinois at Urbana-Champaign, Urbana, Illinois 61801, United States.

Abstract:
Modeling and simulation of small molecules such as drugs and biological cofactors have been both a major focus of computational chemistry for decades and a growing need among computational biophysicists who seek to investigate the interaction of different types of ligands with biomolecules. Of particular interest in this regard are quantum mechanical (QM) calculations that are used to more accurately describe such small molecules, which can be of heterogeneous structures and chemistry, either in purely QM calculations or in hybrid QM/molecular mechanics (MM) simulations. QM programs are also used to develop MM force field parameters for small molecules to be used along with established force fields for biomolecules in classical simulations. With this growing need in mind, here we report a set of software tools developed and closely integrated within the broadly used molecular visualization/analysis program, VMD, that allow the user to construct, modify, and parametrize small molecules and prepare them for QM, hybrid QM/MM, or classical simulations. The tools also provide interactive analysis and visualization capabilities in an easy-to-use and integrated environment. In this paper, we briefly report on these tools and their major features and capabilities, along with examples of how they can facilitate molecular research in computational biophysics that might be otherwise prohibitively complex.
More Related Videos

Related Concept Videos
 02:00
Molecular Models
 01:24
Van der Waals Interactions
 01:18
Molecular Shapes
Two regions of electron density in a diatomic...
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is the extent...
 02:36
Molecular Geometry and Dipole Moments