Related Experiment Video
Updated: Jul 16, 2025

Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Molecular Dynamics of Artificially Pair-Decoupled Systems: An Accurate Tool for Investigating the Importance of
Michele Gandolfi1, Michele Ceotto1
1Dipartimento di Chimica, Università degli Studi di Milano, via Golgi 19, 20133 Milano, Italy.

Abstract:
We propose a numerical technique to accurately simulate the vibrations of organic molecules in the gas phase, when pairs of atoms (or, in general, groups of degrees of freedom) are artificially decoupled, so that their motion is instantaneously decorrelated. The numerical technique we have developed is a symplectic integration algorithm that never requires computation of the force but requires estimates of the Hessian matrix. The theory we present to support our technique postulates a pair-decoupling Hamiltonian function, which parametrically depends on a decoupling coefficient α ∈ [0, 1]. The closer α is to 0, the more decoupled the selected atoms. We test the correctness of our numerical method on small molecular systems, and we apply it to study the vibrational spectroscopic features of salicylic acid at the Density Functional Theory ab initio level on a fitted potential. Our pair-decoupled simulations of salicylic acid show that decoupling hydrogen-bonded atoms do not significantly influence the frequencies of stretching modes, but enhance enormously the out-of-plane wagging and twisting motions of the hydroxyl and carboxyl groups to the point that the carboxyl and hydroxyl groups may overcome high potential energy barriers and change the salicylic acid conformation after a short simulation time. In addition, we found that the acidity of salicylic acid is more influenced by the dynamical couplings of the proton of the carboxylic group with the carbon ring than with the hydroxyl group.
Related Concept Videos
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene...
 01:20
Spin–Spin Coupling: Two-Bond Coupling (Geminal Coupling)
The central atom need not be NMR-active because its electrons are affected by the electron polarization of the spin-active atoms. However, spin information is transmitted less effectively than in one-bond coupling, and 2J values are usually weaker than 1J values. The energy of...
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 01:22
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
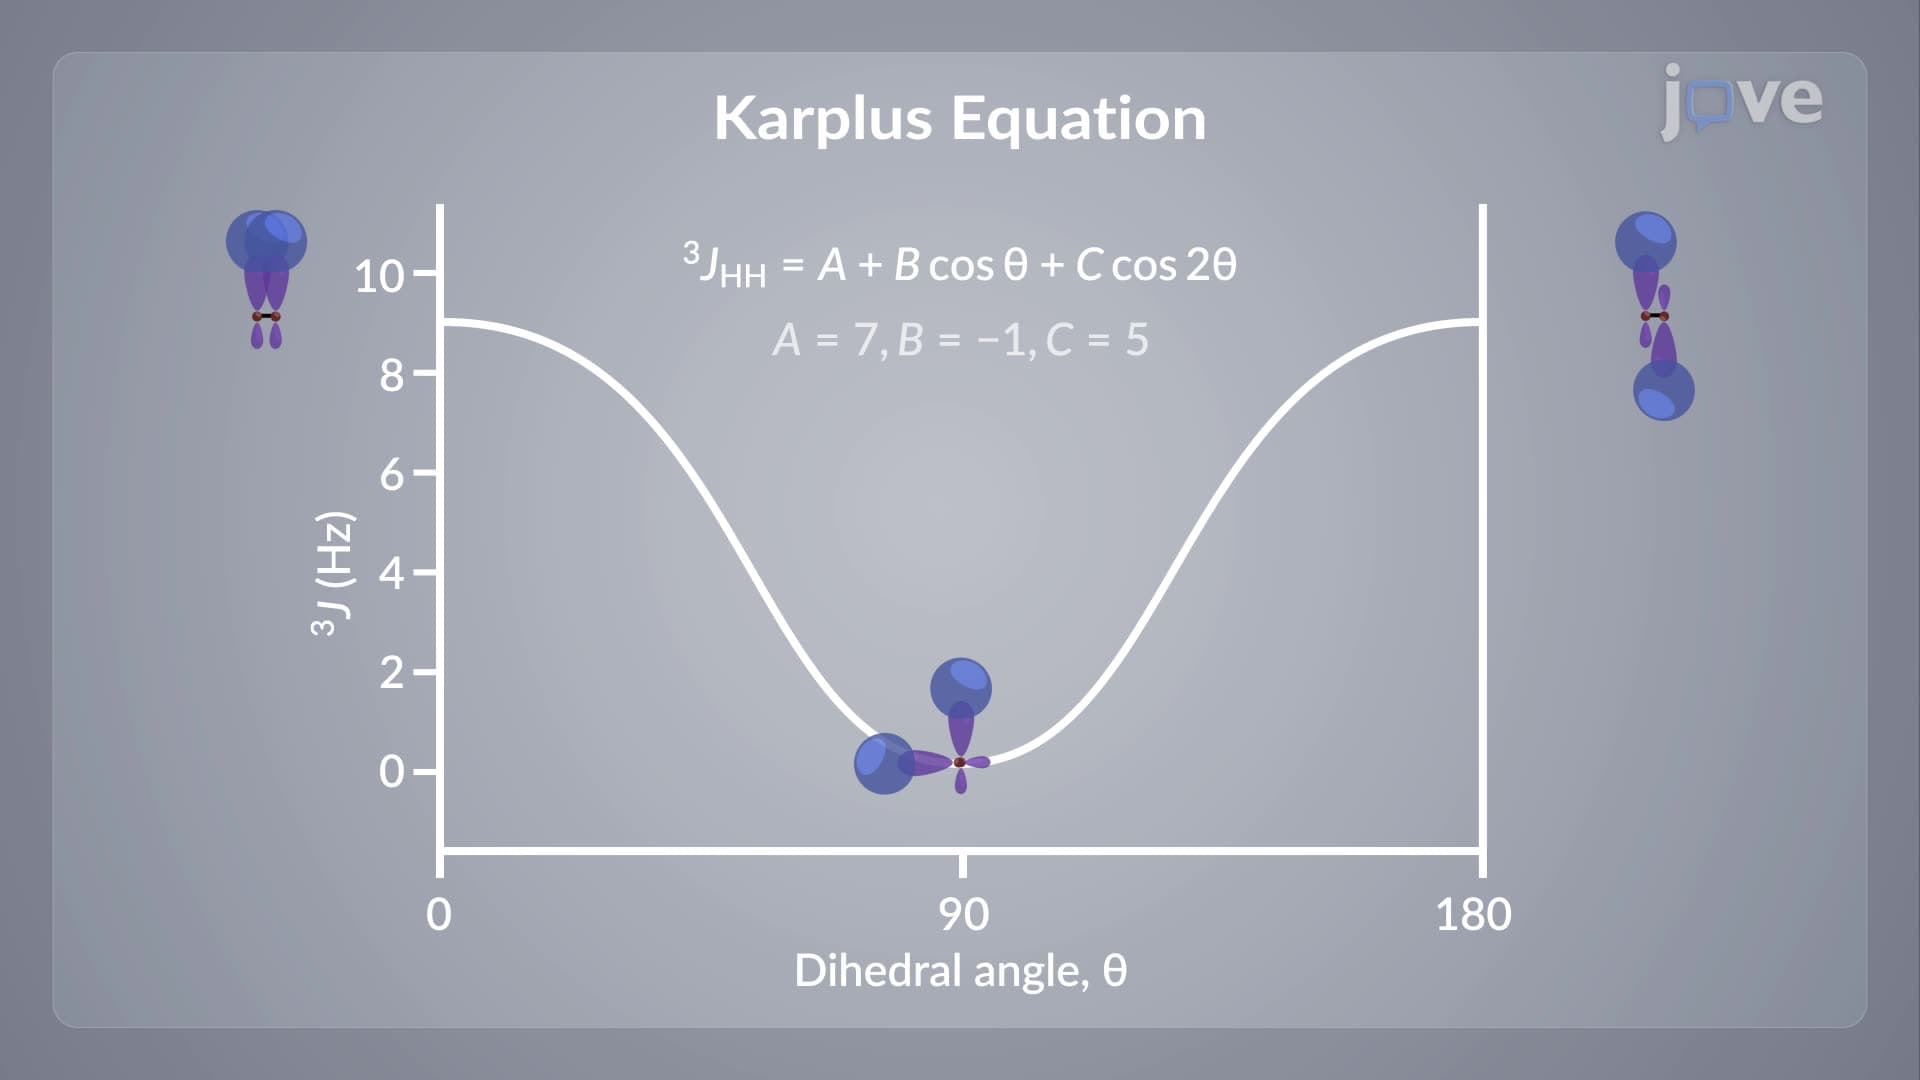
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 01:17
Spin–Spin Coupling: One-Bond Coupling
 01:04
¹³C NMR: ¹H–¹³C Decoupling
A broadband decoupling technique is used to simplify these complex, sometimes overlapping, signals. Broadband decoupling relies on a...