Related Experiment Video
Updated: Jul 16, 2025
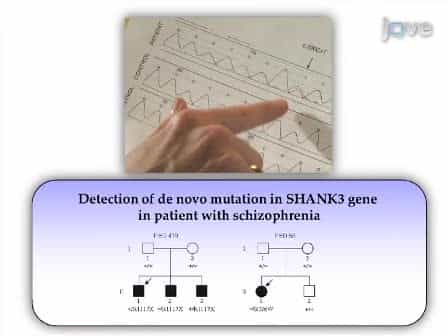

A Strategy to Identify de Novo Mutations in Common Disorders such as Autism and Schizophrenia
Published on: June 15, 2011
DEVOUR: Deleterious Variants on Uncovered Regions in Whole-Exome Sequencing
Erdem Türk1,2, Akif Ayaz3, Ayhan Yüksek1
1Department of Computer Engineering, Muğla Sıtkı Koçman University, Muğla, Turkey.

Abstract:
The discovery of low-coverage (i.e. uncovered) regions containing clinically significant variants, especially when they are related to the patient's clinical phenotype, is critical for whole-exome sequencing (WES) based clinical diagnosis. Therefore, it is essential to develop tools to identify the existence of clinically important variants in low-coverage regions. Here, we introduce a desktop application, namely DEVOUR (DEleterious Variants On Uncovered Regions), that analyzes read alignments for WES experiments, identifies genomic regions with no or low-coverage (read depth < 5) and then annotates known variants in the low-coverage regions using clinical variant annotation databases. As a proof of concept, DEVOUR was used to analyze a total of 28 samples from a publicly available Hirschsprung disease-related WES project (NCBI Bioproject: https://www.ncbi.nlm.nih.gov/bioproject/?term=PRJEB19327), revealing the potential existence of 98 disease-associated variants in low-coverage regions. DEVOUR is available from https://github.com/projectDevour/DEVOUR under the MIT license.
More Related Videos
 11:35
Screening for Functional Non-coding Genetic Variants Using Electrophoretic Mobility Shift Assay EMSA and DNA-affinity Precipitation Assay DAPA
Published on: August 21, 2016
 09:34
Targeted Next-generation Sequencing and Bioinformatics Pipeline to Evaluate Genetic Determinants of Constitutional Disease
Published on: April 4, 2018
Related Concept Videos
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
 03:00
Next-generation Sequencing
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
 01:05
Single Nucleotide Polymorphisms-SNPs
 01:57
Sanger Sequencing
 02:46
Genome Copying Errors