Related Experiment Video
Updated: Jul 15, 2025
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
Exploring π-π interactions and electron transport in complexes involving a hexacationic host and PAH guest: a
Haobam Kisan Singh1, Upasana Nath1, Niharika Keot1
1Department of Chemistry, Indian Institute of Technology Guwahati, North-Guwahati, Guwahati-781039, India. msarma@iitg.ac.in.

Abstract:
Single isolated molecules and supramolecular host-guest systems, which consist of π-π stacking interactions, are emerging as promising building blocks for creating molecular electronic devices. In this article, we have investigated the noncovalent π-π interaction and intermolecular electron charge transport involved in a series of host-guest complexes formed between a cage-like host (H6+) and polycyclic aromatic hydrocarbon (PAH) guests (G1-G7) using different quantum chemical approaches. The host (H6+) consists of two triscationic π-electron-deficient trispyridiniumtriazine (TPZ3+) units that are bridged face-to-face by three ethylene-triazole-ethylene. Our theoretical calculations show that the perylene and naphthalene inclusion complexes G7⊂H and G1⊂H have the highest and lowest interaction energies, respectively. In addition, energy decomposition analysis (EDA) indicated that the dispersion interaction term, ΔEdisp, significantly contributes to the host-guest interaction and is correlated with the existence of π-π van der Waals interaction. Using the nonequilibrium Greens function (NEGF) method in combination with density functional theory (DFT), the current-voltage (I-V) curves of the complexes were estimated. The conductance values increased when the guests were embedded inside the host cavity. Notably, the complex G7⊂H has the maximum conductance value. Overall, this study provided the electron transport of the PAH inclusion host-guest complex through π-π interaction and provided a direction for the fabrication of future supramolecular molecular devices.
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
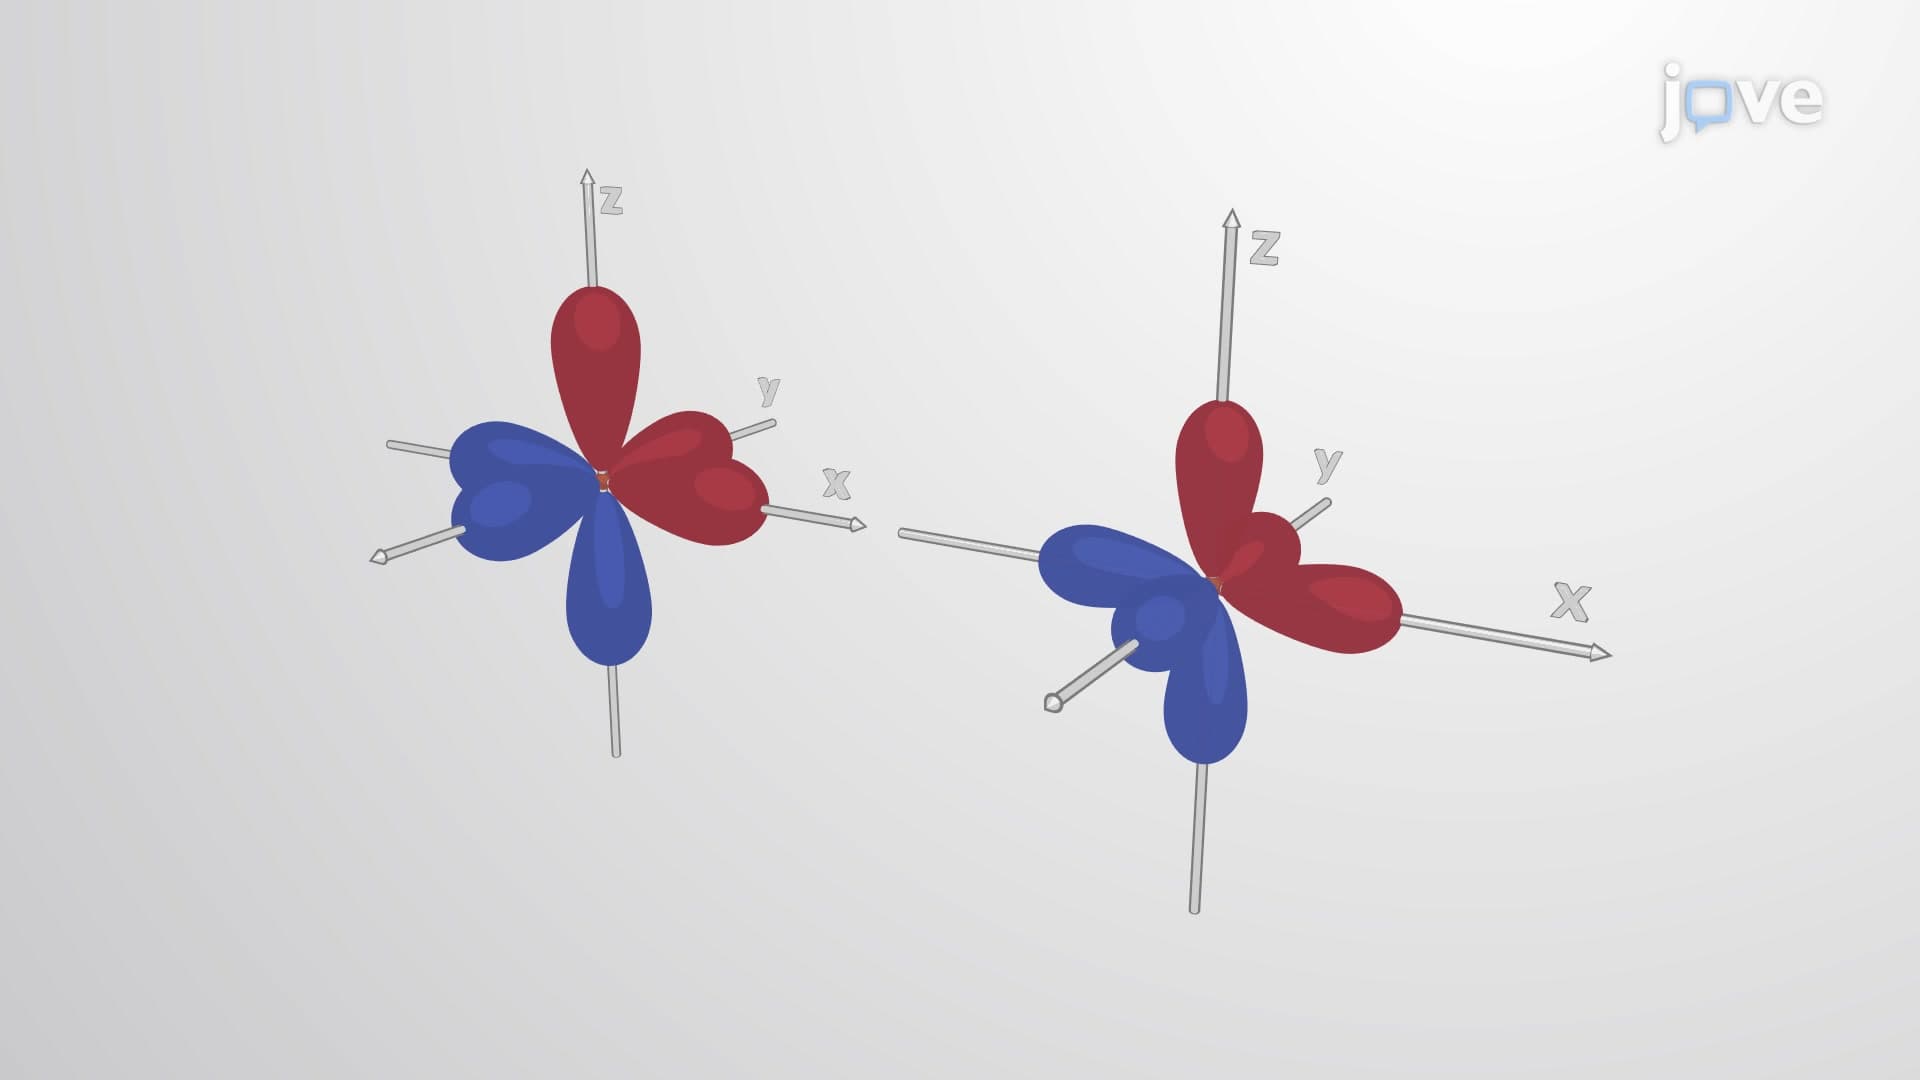
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:35
Molecular Orbital Theory I
 02:42
Valence Bond Theory
 01:27
π Electron Effects on Chemical Shift: Overview
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...