One bead per residue can describe all-atom protein structures
Related Experiment Video
Updated: Jul 10, 2025

A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
1Department of Biochemistry and Molecular Biology, Michigan State University, East Lansing, MI 48824, USA.

Abstract:
Atomistic resolution is the standard for high-resolution biomolecular structures, but experimental structural data are often at lower resolution. Coarse-grained models are also used extensively in computational studies to reach biologically relevant spatial and temporal scales. This study explores the use of advanced machine learning networks for reconstructing atomistic models from reduced representations. The main finding is that a single bead per amino acid residue allows construction of accurate and stereochemically realistic all-atom structures with minimal loss of information. This suggests that lower resolution representations of proteins may be sufficient for many applications when combined with a machine learning framework that encodes knowledge from known structures. Practical applications include the rapid addition of atomistic detail to low-resolution structures from experiment or computational coarse-grained models. The application of rapid, deterministic all-atom reconstruction within multi-scale frameworks is further demonstrated with a rapid protocol for the generation of accurate models from cryo-EM densities close to experimental structures.

Related Concept Videos
 01:24
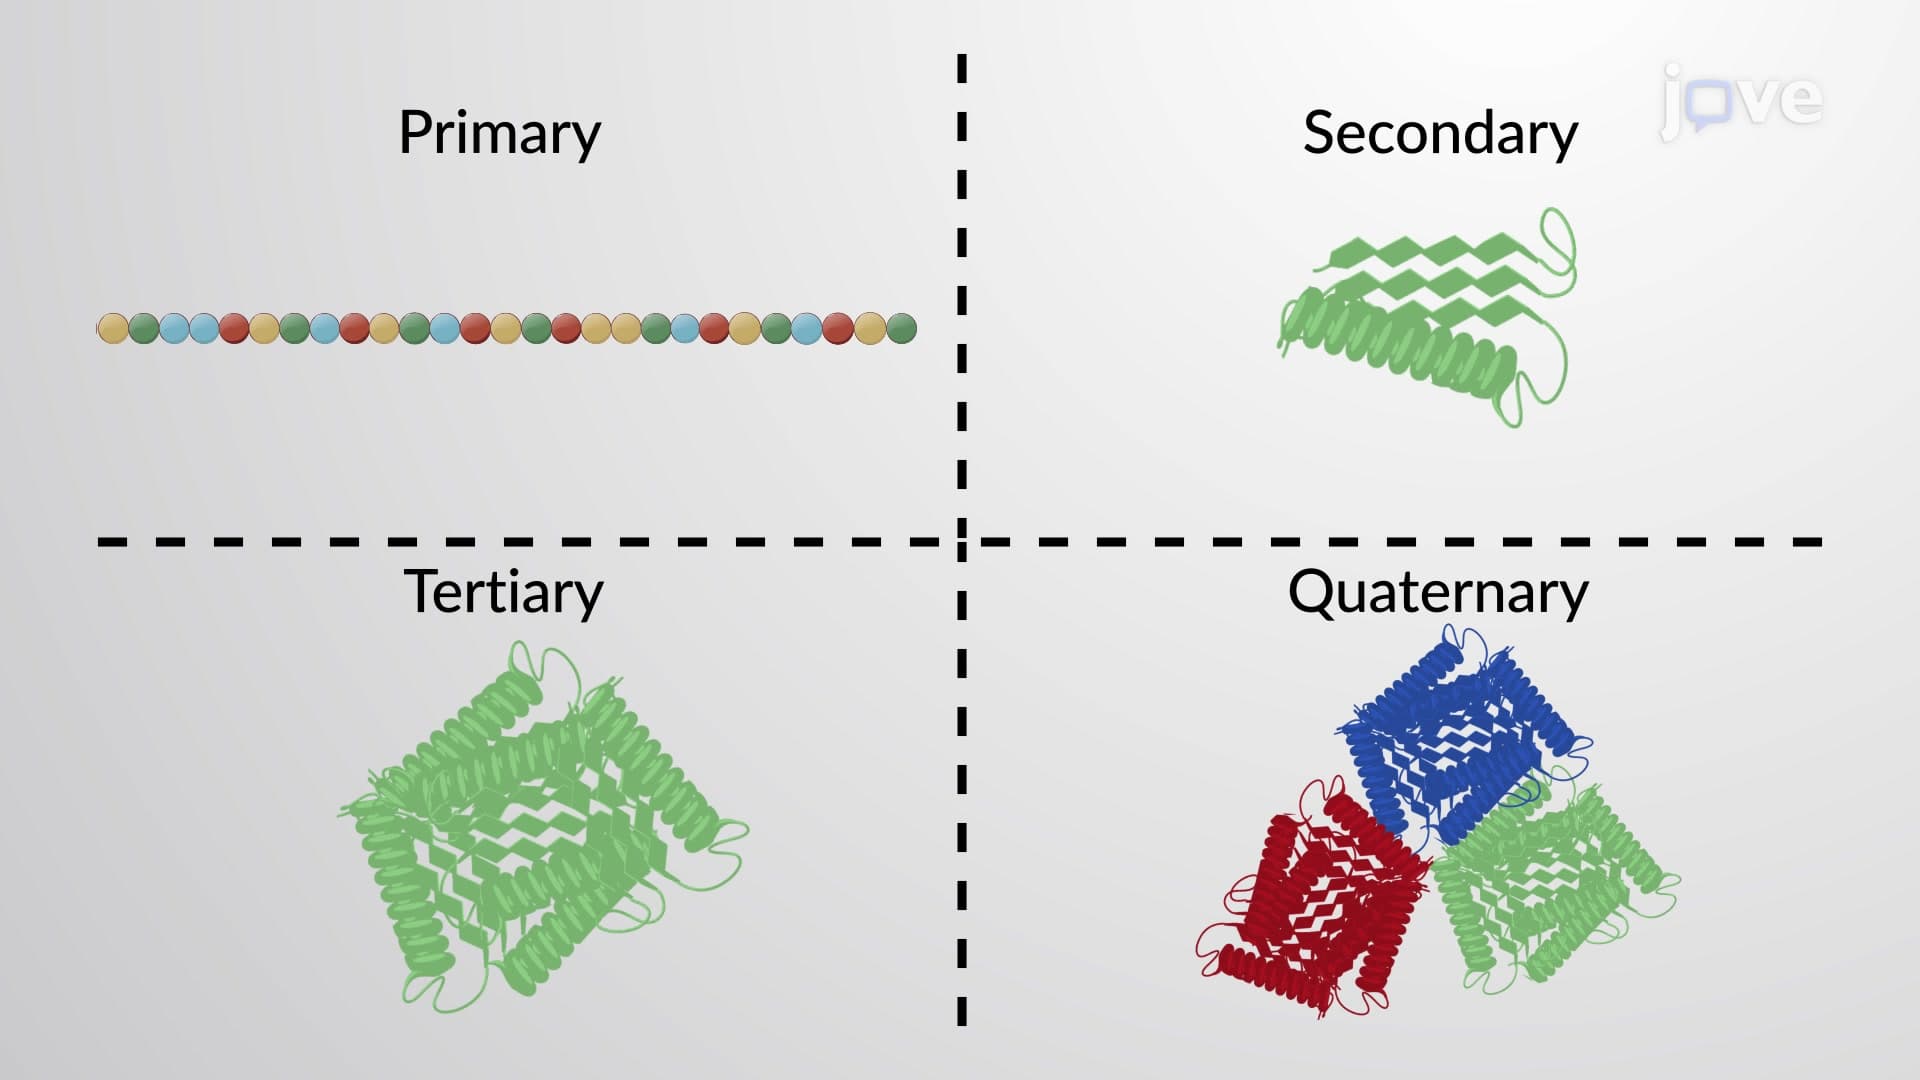
Protein Organization
The primary structure of a protein is its amino acid sequence....
 02:00
Molecular Models
 02:06
Newman Projections
The organic molecules rotate across the single bonds leading to numerous temporary three-dimensional structures of varying energy known as...
 01:22
Protein Folding
 01:18
Structure of Benzene: Molecular Orbital Model
 02:15
Protein and Protein Structures