Related Experiment Video
Updated: Jul 7, 2025

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Error of relativistic effective core potentials for closed-shell diatomic molecules of p-block heavy and superheavy
Yanzhao Lu1, Zhifan Wang2, Fan Wang1
1Institute of Atomic and Molecular Physics, Key Laboratory of High Energy Density Physics and Technology, Ministry of Education, Sichuan University, Chengdu 610065, People's Republic of China.

Abstract:
Pseudopotentials (PP) are extensively used in electronic structure calculations, particularly for molecules containing heavy elements. Parameters in PPs are mainly determined from ab initio results, and errors of such PPs in density functional theory (DFT) calculations have been studied previously. However, PP errors on results with spin-orbit coupling and those in time-dependent DFT (TDDFT) calculations have not been reported previously. In this work, we investigate the error of the small-core energy-consistent Stuttgart/Koln pseudopotentials in DFT and TDDFT calculations with and without spin-orbit coupling. Ground state bond lengths, harmonic frequencies, dissociation energies, and vertical excitation energies for a series of closed-shell diatomic heavy and superheavy p-block molecules are calculated using several popular exchange-correlation functionals. PP errors are estimated by comparing with results using the all-electron Dirac-Coulomb (-Gaunt) Hamiltonian. Our results show that the difference between ground state properties and most excitation energies in scalar-relativistic calculations with the PP and those of all-electron calculations is quite small. This difference becomes somewhat larger when spin-orbit coupling (SOC) is present, especially for properties that are affected by SOC to some extent. In addition, the errors of the PPs are insensitive to the employed exchange-correlation functionals in most cases. Our results indicate that reasonable DFT and TDDFT results can be obtained using the small-core energy-consistent Stuttgart/Koln pseudopotentials for heavy and super-heavy p-block molecules.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 03:08
Atomic Radii and Effective Nuclear Charge
 01:10
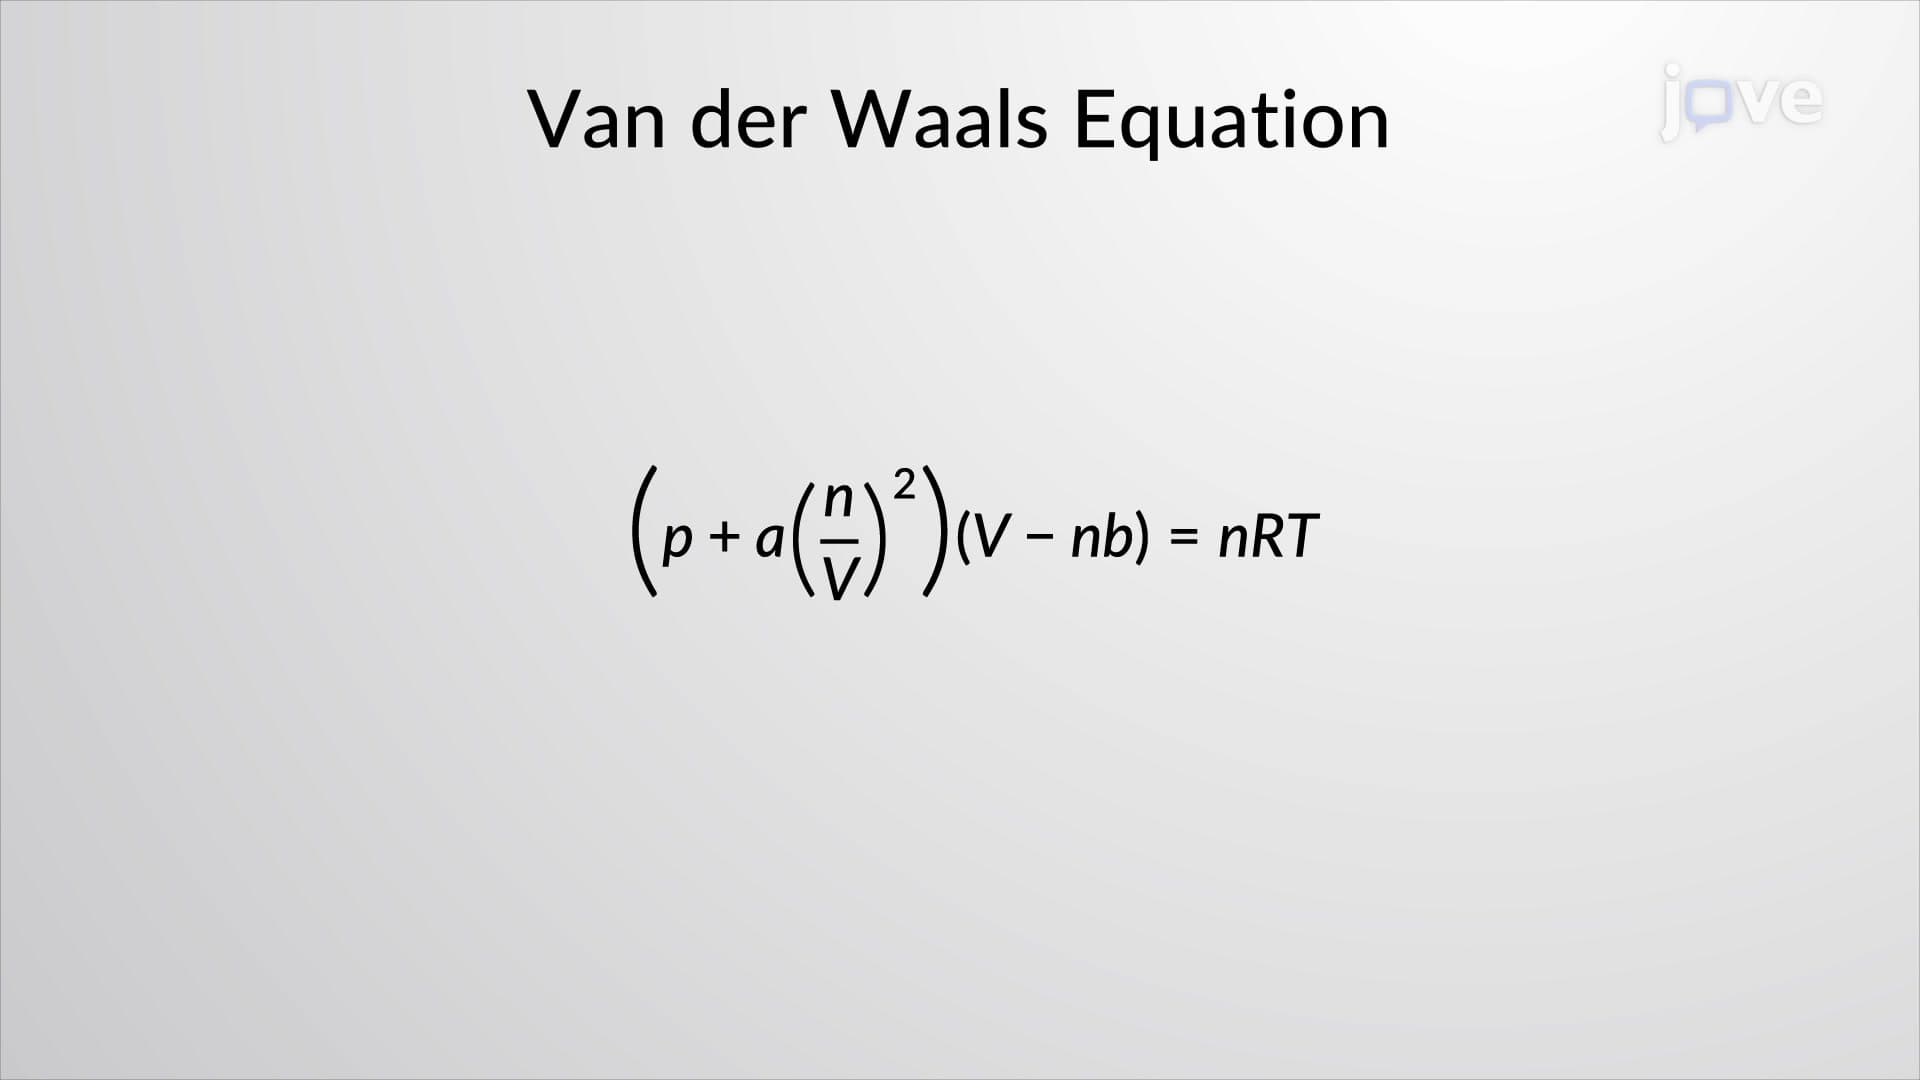
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:26
Thermodynamic Potentials
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I