Related Experiment Video
Updated: Jul 6, 2025

A Precision Medicine Tool for Measurement and Monitoring of Hemoglobin S in Sickle Cell Disease Patients Receiving Transfusion Therapy
Αlpha-thalassemia: A practical overview
Khaled M Musallam1, M Domenica Cappellini2, Thomas D Coates3
1Center for Research on Rare Blood Disorders (CR-RBD), Burjeel Medical City, Abu Dhabi, United Arab Emirates.

α-Thalassemia is an inherited blood disorder characterized by decreased synthesis of α-globin chains that results in an imbalance of α and β globin and thus varying degrees of ineffective erythropoiesis, decreased red blood cell (RBC) survival, chronic hemolytic anemia, and subsequent comorbidities. Clinical presentation varies depending on the genotype, ranging from a silent or mild carrier state to severe, transfusion-dependent or lethal disease. Management of patients with α-thalassemia is primarily supportive, addressing either symptoms (eg, RBC transfusions for anemia), complications of the disease, or its transfusion-dependence (eg, chelation therapy for iron overload). Several novel therapies are also in development, including curative gene manipulation techniques and disease modifying agents that target ineffective erythropoiesis and chronic hemolytic anemia. This review of α-thalassemia and its various manifestations provides practical information for clinicians who practice beyond those regions where it is found with high frequency.
α-Thalassemia is an inherited blood disorder characterized by decreased synthesis of α-globin chains that results in an imbalance of α and β globin and thus varying degrees of ineffective erythropoiesis, decreased red blood cell (RBC) survival, chronic hemolytic anemia, and subsequent comorbidities. Clinical presentation varies depending on the genotype, ranging from a silent or mild carrier state to severe, transfusion-dependent or lethal disease. Management of patients with α-thalassemia is primarily supportive, addressing either symptoms (eg, RBC transfusions for anemia), complications of the disease, or its transfusion-dependence (eg, chelation therapy for iron overload). Several novel therapies are also in development, including curative gene manipulation techniques and disease modifying agents that target ineffective erythropoiesis and chronic hemolytic anemia. This review of α-thalassemia and its various manifestations provides practical information for clinicians who practice beyond those regions where it is found with high frequency.
More Related Videos
 11:59
Detection of Residual Donor Erythroid Progenitor Cells after Hematopoietic Stem Cell Transplantation for Patients with Hemoglobinopathies
Published on: September 6, 2017
 05:23
Continuous Manual Exchange Transfusion for Patients with Sickle Cell Disease: An Efficient Method to Avoid Iron Overload
Published on: March 14, 2017
Related Concept Videos
 01:15
Blood Transfusion
Blood Transfusion Overview
A blood transfusion is a medical procedure used to replace blood lost due to injury, surgery, or to treat conditions such as anemia or cancer. During a transfusion, donor blood is...
 01:49
Multiple Allele Traits
 01:24
Hemoglobin
When all four heme groups are bound to oxygen, the resulting molecule is called oxyhemoglobin. As a result, arterial blood...
 01:27
Disorders of Erythrocytes
Erythrocyte disorders can be broadly categorized into two main types: anemic and polycythemic conditions.
A low oxygen-carrying capacity of the blood due to the loss, lower production, or destruction of erythrocytes is termed anemia. Hemorrhagic anemia, for example, occurs when bleeding from an external wound or internal ulcer reduces erythrocyte counts.
On the other...
 01:10
Blood Typing
Antigens are protein molecules that reside on the surface of red blood cells (RBCs). The ABO and Rh blood typing systems target...
 01:12
The ABO Blood Group
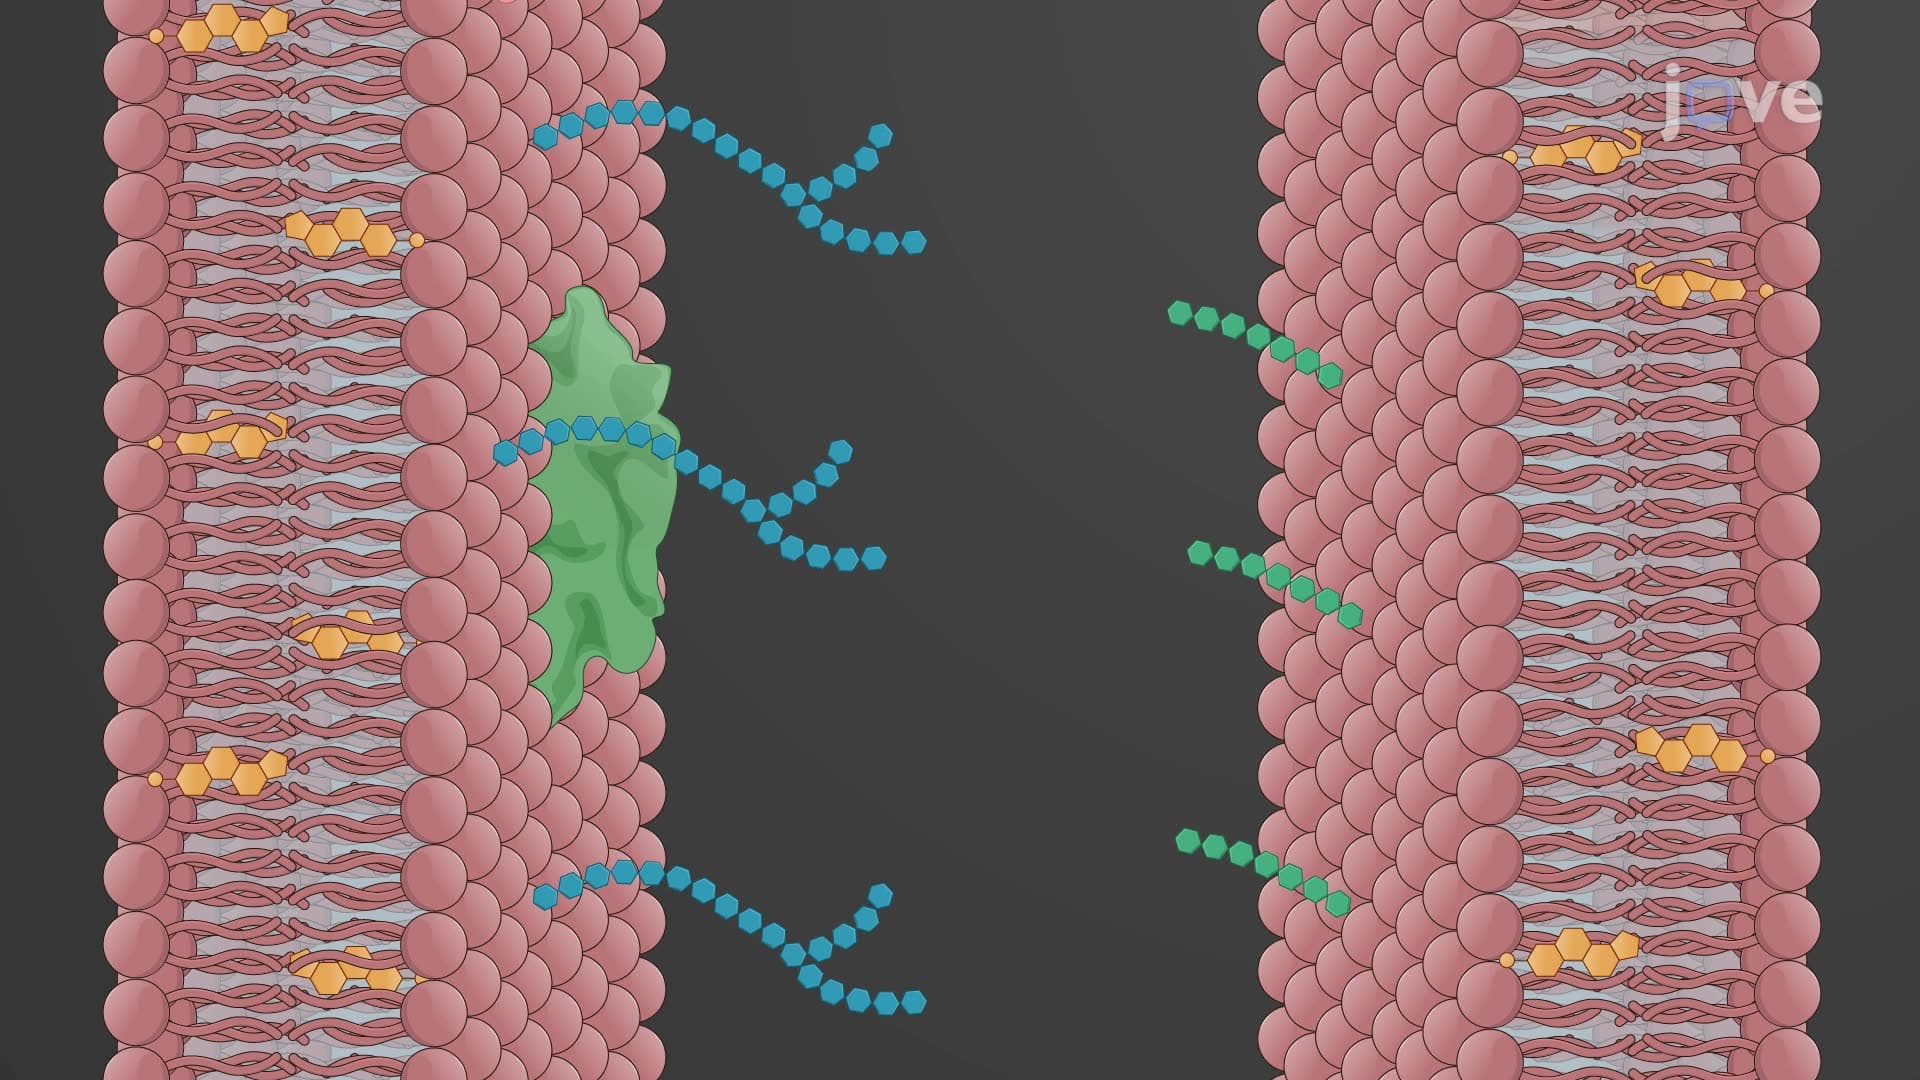
Antigens in the ABO Blood Group System
Antigens are substances that can trigger an immune response, leading to the production of antibodies. In the ABO blood group system,...