Related Experiment Video
Updated: Jul 6, 2025


A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Unexplored regions of the protein sequence-structure map revealed at scale by a library of foldtuned language models
Arjuna M Subramanian1, Zachary A Martinez1, Alec L Lourenço1
1Division of Biology and Biological Engineering, California Institute of Technology, Pasadena, CA.

Abstract:
The combinatorial scale of amino-acid sequence-space has traditionally precluded substantive study of the full protein sequence-structure map. It remains unknown, for instance, how much of the vast uncharted landscape of far-from-natural sequences encodes the familiar ensemble of natural folds in a fashion consistent with the laws of biophysics but seemingly untouched by evolution on Earth. The scale of sequence perturbations required to access these spaces exceeds the reach of even gold-standard experimental approaches such as directed evolution. We surpass this limitation guided by the innate capacity of protein language models (PLMs) to explore sequences outside their natural training data through generation and self-feedback. We recast PLMs as probes that explore into regions of protein "deep space" that possess little-to-no detectable homology to natural examples, while enforcing core structural constraints, in a novel sequence design approach that we term "foldtuning." We build a library of foldtuned PLMs for >700 natural folds in the SCOP database, covering numerous high-priority targets for synthetic biology, including GPCRs and small GTPases, composable cell-surface-receptor and DNA-binding domains, and small signaling/regulatory domains. Candidate proteins generated by foldtuned PLMs reflect distinctive new "rules of language" for sequence innovation beyond detectable homology to any known protein and sample subtle structural alterations in a manner reminiscent of natural structural evolution and diversification. Experimental validation of three markedly different fold targets; the tyrosine-kinase- and small-GTPase-regulating SH3 domain, the bacterial RNase inhibitor barstar, and the peptide hormone insulin demonstrates that foldtuning proposes protein variants that express and fold stably in vitro and function in vivo. Foldtuning reveals protein sequence-structure information at scale outside of the context of evolution and promises to push forward the redesign and reconstitution of novel-to-nature synthetic biological systems for applications in health and catalysis.
More Related Videos


Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:47
Protein Families
 01:22
Protein Folding
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...
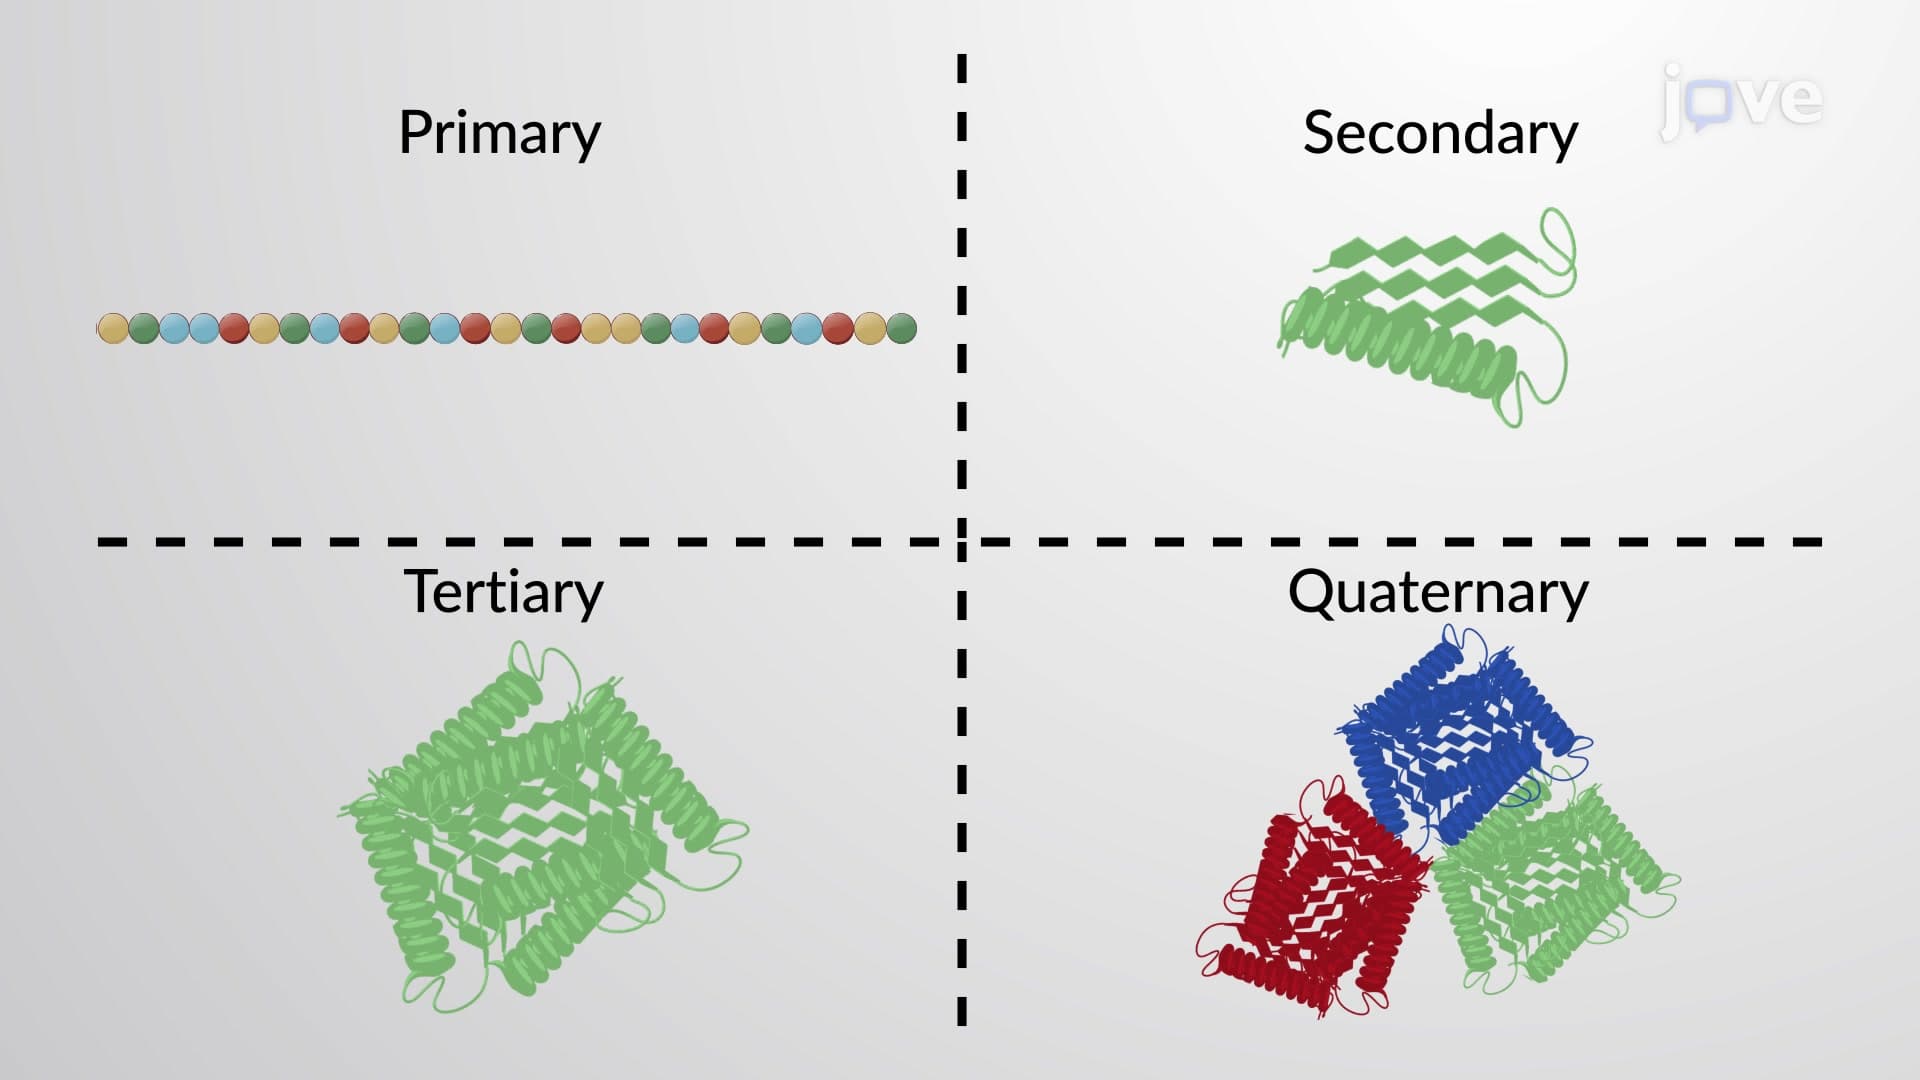 02:15
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...
02:26Conservation of Protein Domains