Structure-Energy Relationship Prediction of the HZSM-5 Zeolite with Different Acid Site Distributions by the Neural Network Model
Related Experiment Video
Updated: Jul 4, 2025
Adsorption Device Based on a Langatate Crystal Microbalance for High Temperature High Pressure Gas Adsorption in Zeolite H-ZSM-5
Published on: August 25, 2016
Yang Yang1, Wenming Zhang1, Shengbin Chen1
1School of New Energy, North China Electric Power University, Beijing 102206, People's Republic of China.

Abstract:
Zeolites are a very important family of catalysts. The catalytic activity of zeolites depends on the distribution of acid sites, which has been extensively studied. However, the relationship between the acid site distribution and catalytic efficiency remains unestablished. An onerous computational burden can be imposed when static calculations are applied to analyze the relationship between a catalyst structure and its energy. To resolve this issue, the current work uses neural network (NN) models to evaluate the relationship. By taking the typical HZSM-5 zeolite as an example, we applied the provided atomic coordinates to predict the energy. The network performances of the artificial neural network (ANN) and high-dimensional neural network (HDNN) are compared using the trained results from a dataset containing the identical number of acid sites. Furthermore, the importance of the feature is examined with the aid of a random forest model to identify the pivotal variables influencing the energy. In addition, the HDNN is employed to forecast the energy of an HZSM-5 system with varying numbers of acid sites. This study emphasizes that the energy of zeolites can be rapidly and accurately predicted through the NN, which can assist our understanding of the relationship between the structure and properties, thereby providing more accurate and efficient methods for the application of zeolite materials.
Related Concept Videos
 02:54
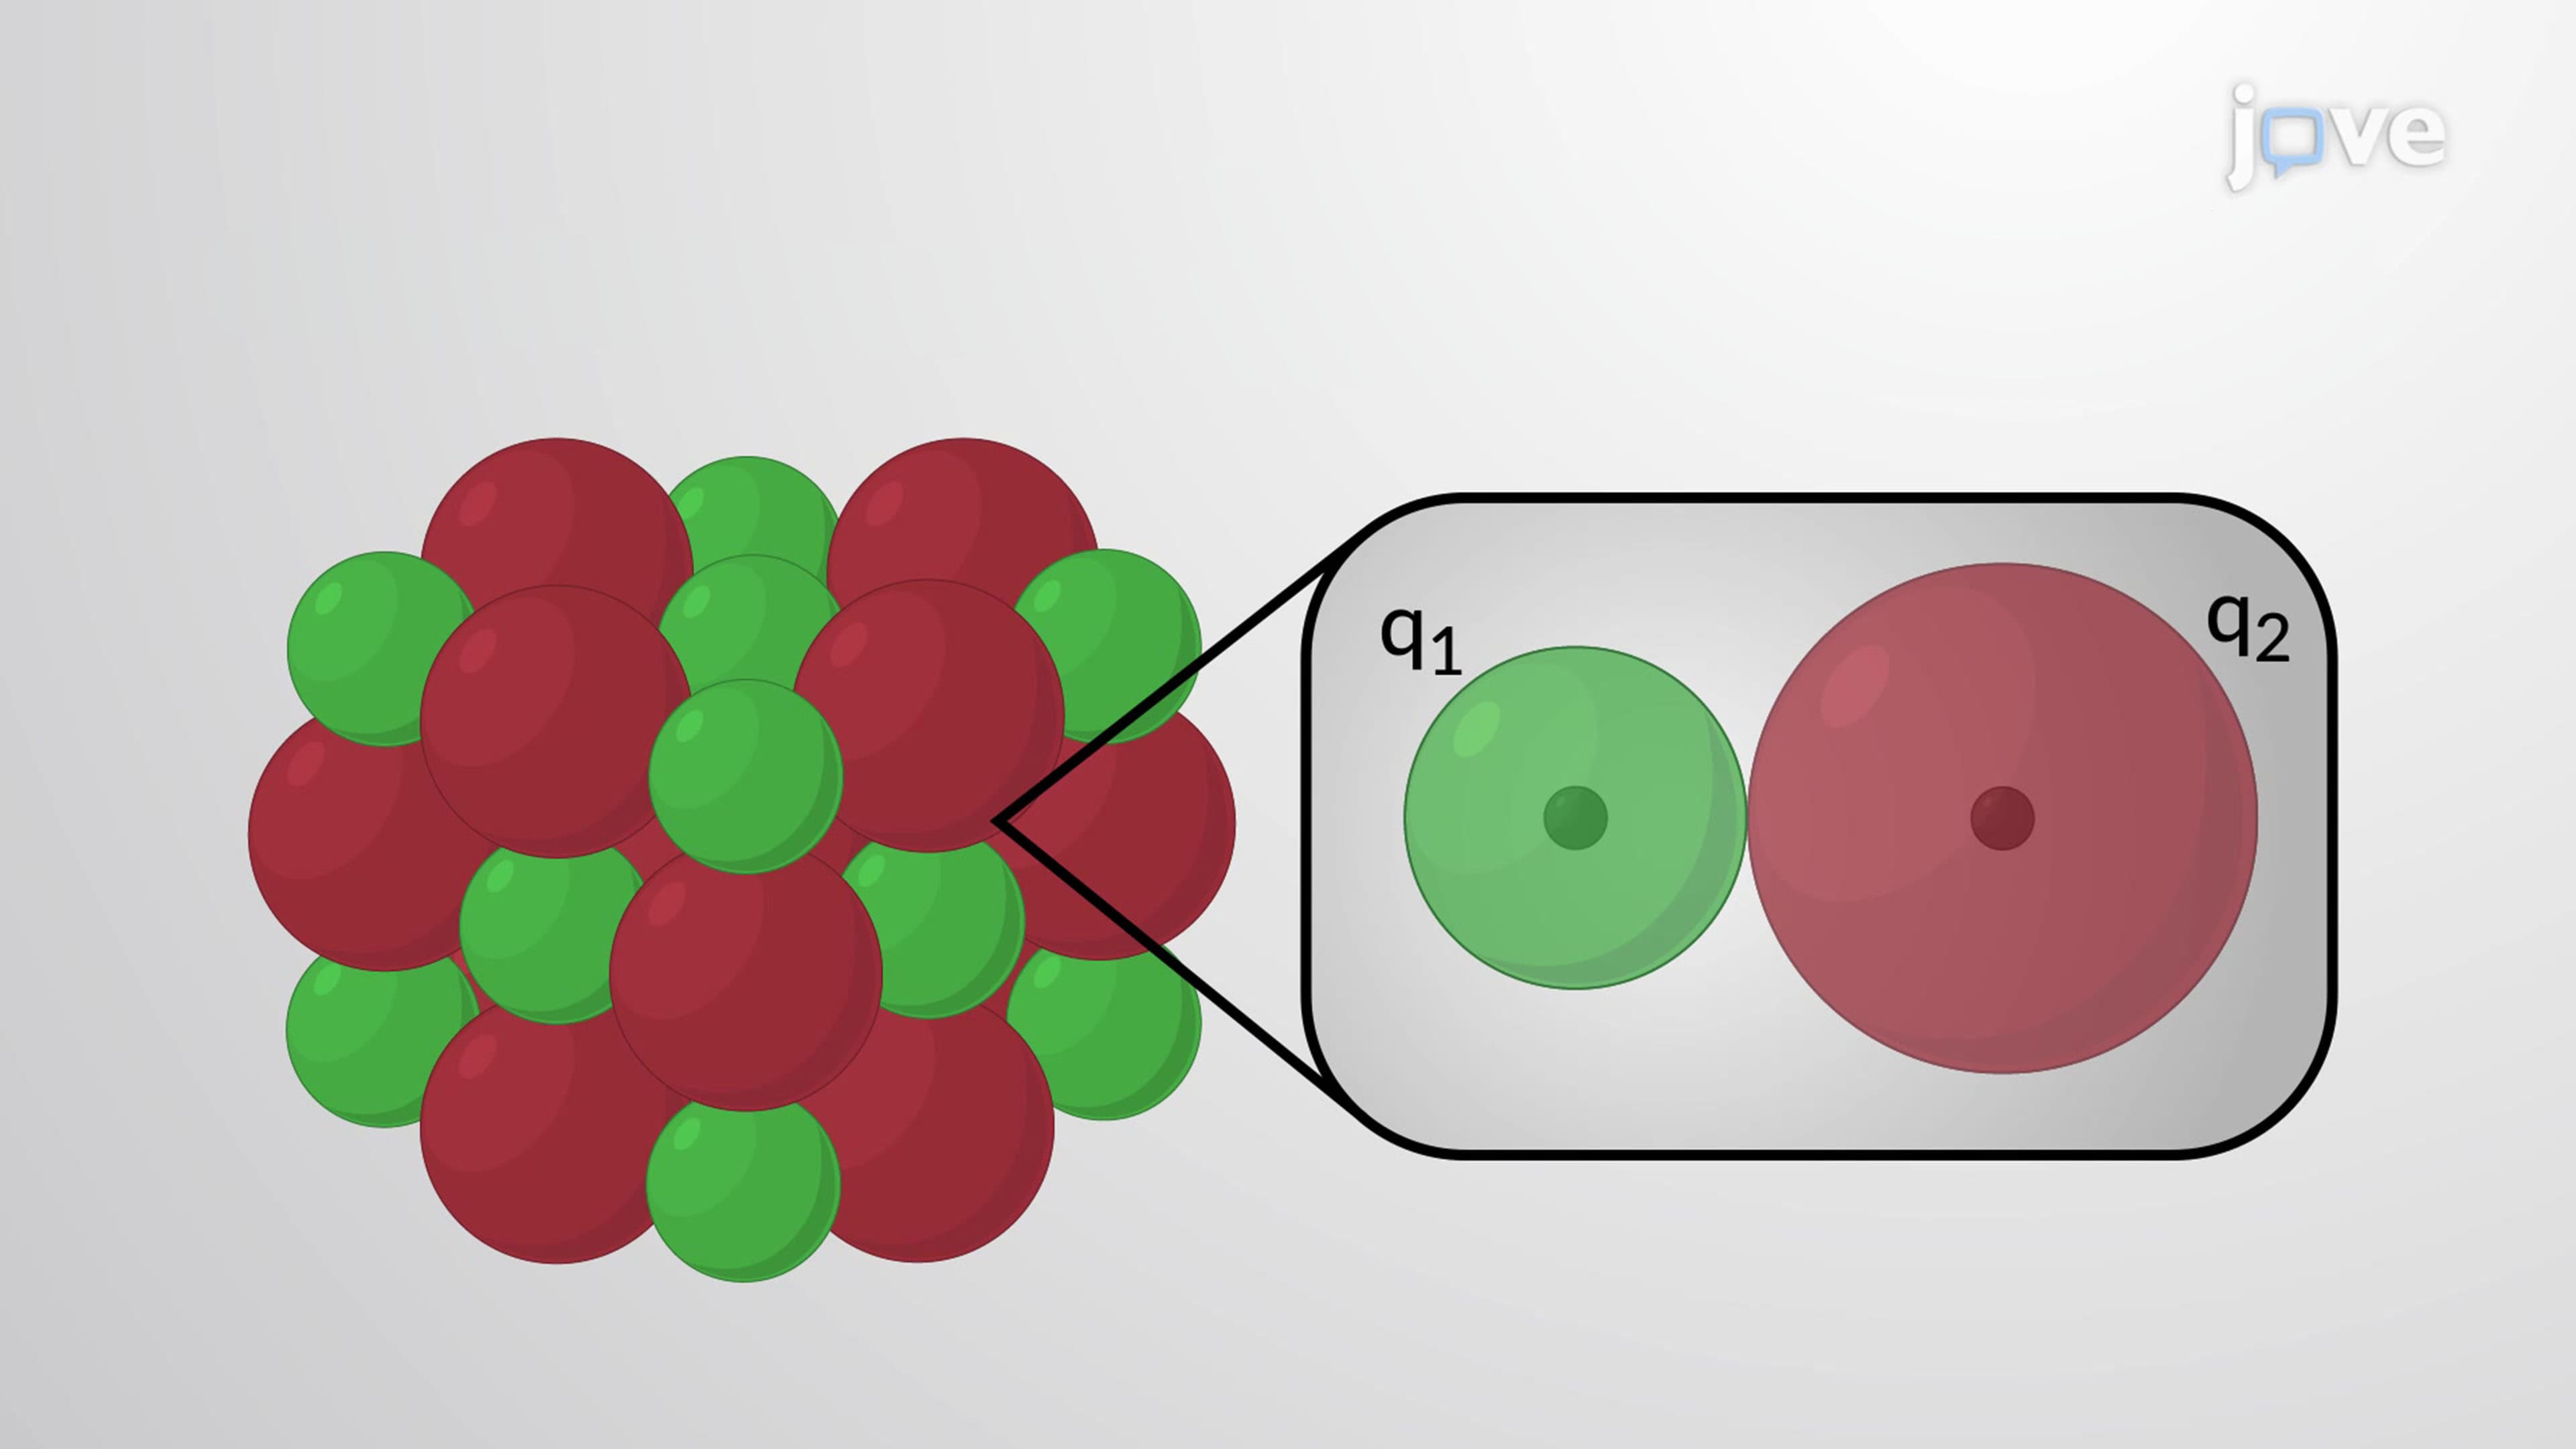
Trends in Lattice Energy: Ion Size and Charge
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
 01:25
Basicity of Heterocyclic Aromatic Amines
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:44
The Born-Haber Cycle