Related Experiment Video
Updated: Jul 3, 2025


Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Coupled 3D (J ≥ 0) Time-Dependent Wave Packet Calculation for the F + H2 Reaction on Accurate Ab Initio Multi-State
Koushik Naskar1, Soumya Mukherjee1, Sandip Ghosh1,2
1School of Chemical Sciences, Indian Association for the Cultivation of Science 2A & 2B Raja S. C. Mullick Road, Jadavpur, Kolkata 700032, West Bengal, India.

Abstract:
We had calculated adiabatic potential energy surfaces (PESs), nonadiabatic, and spin-orbit (SO) coupling terms among the lowest three electronic states (12A', 22A', and 12A″) of the F + H2 system using the multireference configuration interaction (MRCI) level of theory, and the adiabatic-to-diabatic transformation equations were solved to formulate the diabatic Hamiltonian matrix [J. Chem. Phys. 2020, 153, 174301] for the entire region of the nuclear configuration space. The accuracy of such diabatic PESs is explored by performing scattering calculations to evaluate integral cross sections (ICSs) and rate constants. The nonadiabatic and SO effects are studied by utilizing coupled 3D time-dependent wave packet formalism with zero and nonzero total angular momentum on multiple adiabatic/diabatic surfaces calculation. We depict the convergence profiles of reaction probabilities for the reactive as well as nonreactive processes on various electronic states at different collision energies with respect to total angular momentum including all helicity quantum numbers. Finally, total ICSs are calculated as functions of collision energies for the initial rovibrational state (v = 0, j = 0) of the H2 molecule along with the temperature-dependent rate coefficient, where those quantities are compared with previous theoretical and experimental results.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I
 02:45
The Quantum-Mechanical Model of an Atom
 03:51
Molecular Orbital Theory II
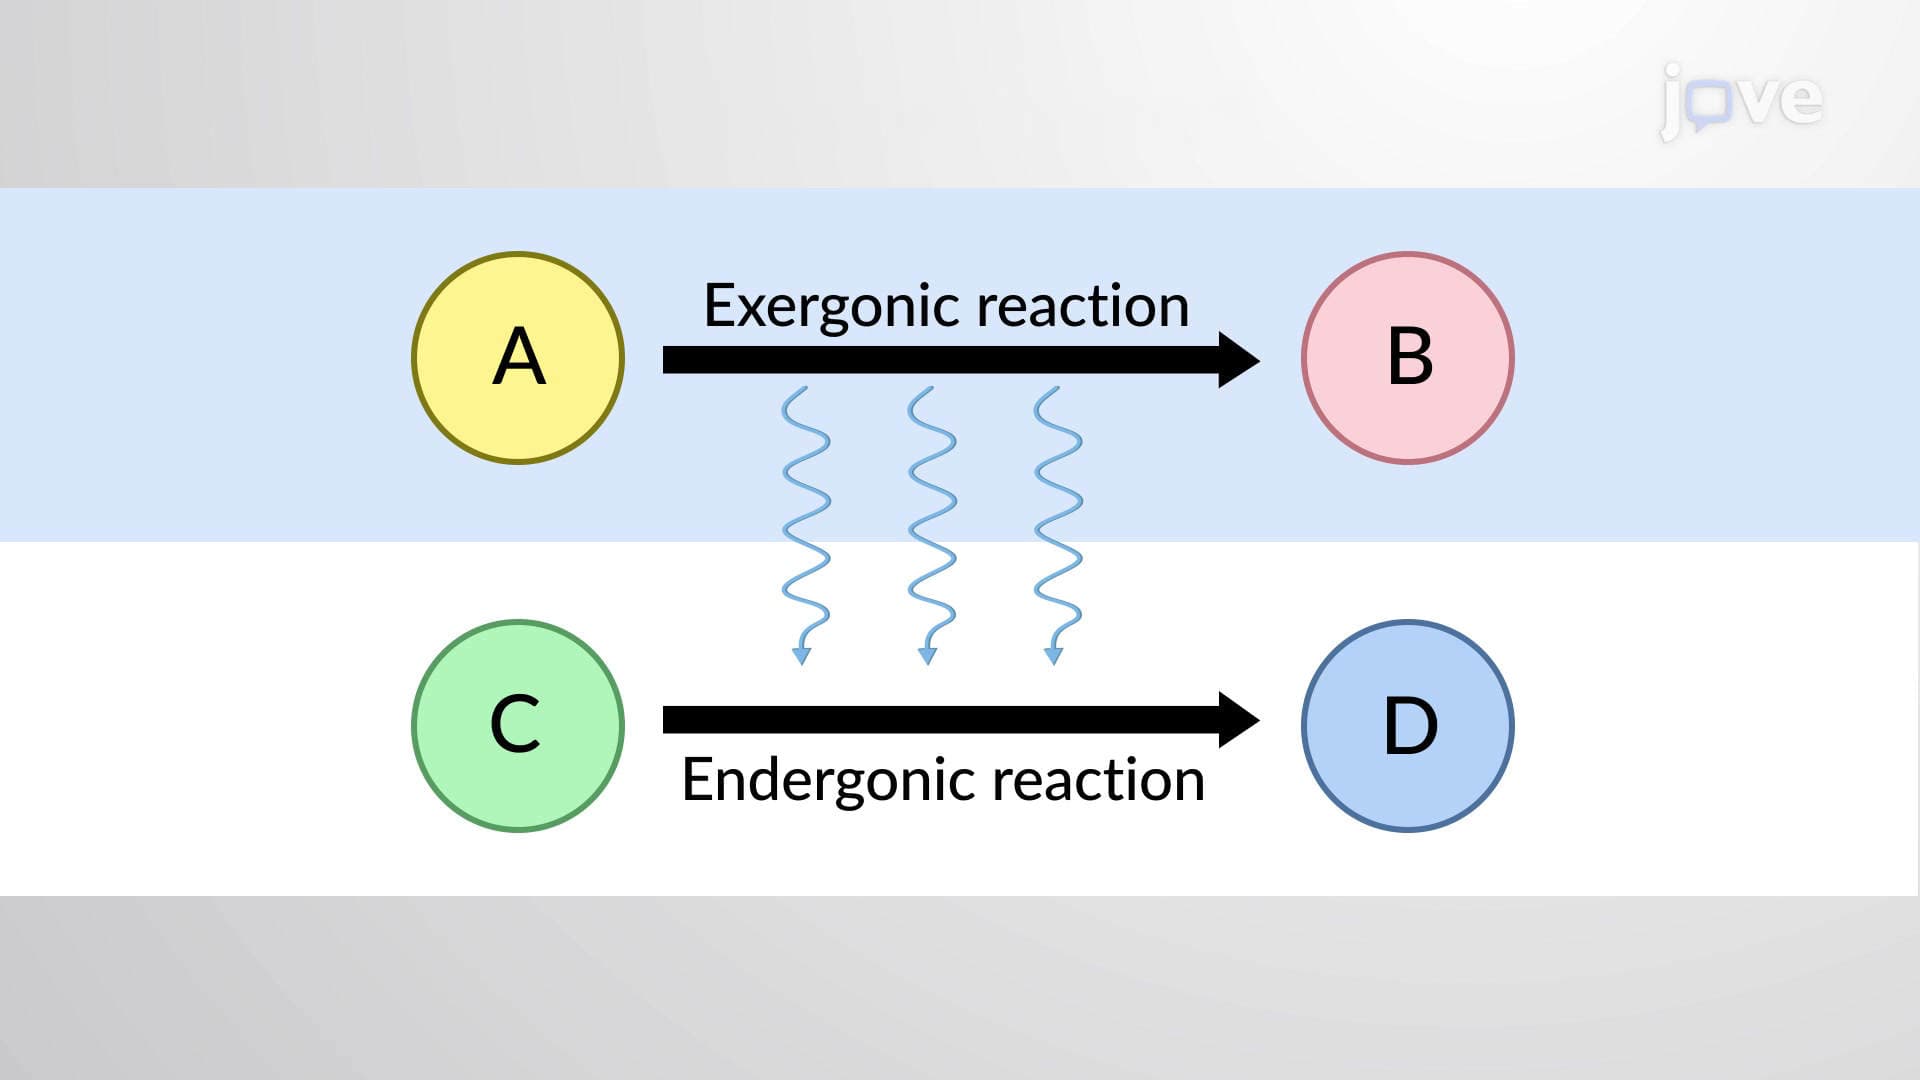 01:17
Coupled Reactions
Energy in adenosine triphosphate or ATP molecules is easily accessible to do work. ATP powers the majority of energy-requiring cellular reactions....
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...