Related Experiment Video
Updated: Jul 2, 2025
Use of Microscale Thermophoresis to Measure Protein-Lipid Interactions
Published on: February 10, 2022
Influence of Desialylation on the Drug Binding Affinity of Human Alpha-1-Acid Glycoprotein Assessed by Microscale
Tino Šeba1, Robert Kerep1, Tin Weitner1
1Department of General and Inorganic Chemistry, Faculty of Pharmacy and Biochemistry, University of Zagreb, A. Kovačića 1, 10000 Zagreb, Croatia.

Abstract:
Human serum alpha-1-acid glycoprotein (AAG) is an acute-phase plasma protein involved in the binding and transport of many drugs, especially basic and lipophilic substances. The sialic acid groups that terminate the N-glycan chains of AAG have been reported to change in response to numerous health conditions and may have an impact on the binding of drugs to AAG. In this study, we quantified the binding between native and desialylated AAG and seven drugs from different pharmacotherapeutic groups (carvedilol, diltiazem, dipyridamole, imipramine, lidocaine, propranolol, vinblastine) using microscale thermophoresis (MST). This method was chosen due to its robustness and high sensitivity, allowing precise quantification of molecular interactions based on the thermophoretic movement of fluorescent molecules. Detailed glycan analysis of native and desialylated AAG showed over 98% reduction in sialic acid content for the enzymatically desialylated AAG. The MST results indicate that desialylation generally alters the binding affinity between AAG and drugs, leading to either an increase or decrease in Kd values, probably due to conformational changes of AAG caused by the different sialic acid content. This effect is also reflected in an increased denaturation temperature of desialylated AAG. Our findings indicate that desialylation impacts free drug concentrations differently, depending on the binding affinity of the drug with AAG relative to human serum albumin (HSA). For drugs such as dipyridamole, lidocaine, and carvedilol, which have a higher affinity for AAG, desialylation significantly changes free drug concentrations. In contrast, drugs such as propranolol, imipramine, and vinblastine, which have a strong albumin binding, show only minimal changes. It is noteworthy that the free drug concentration of dipyridamole is particularly sensitive to changes in AAG concentration and glycosylation, with a decrease of up to 15% being observed, underscoring the need for dosage adjustments in personalized medicine.
Related Concept Videos
 01:18
Factors Affecting Protein-Drug Binding: Drug-Related Factors
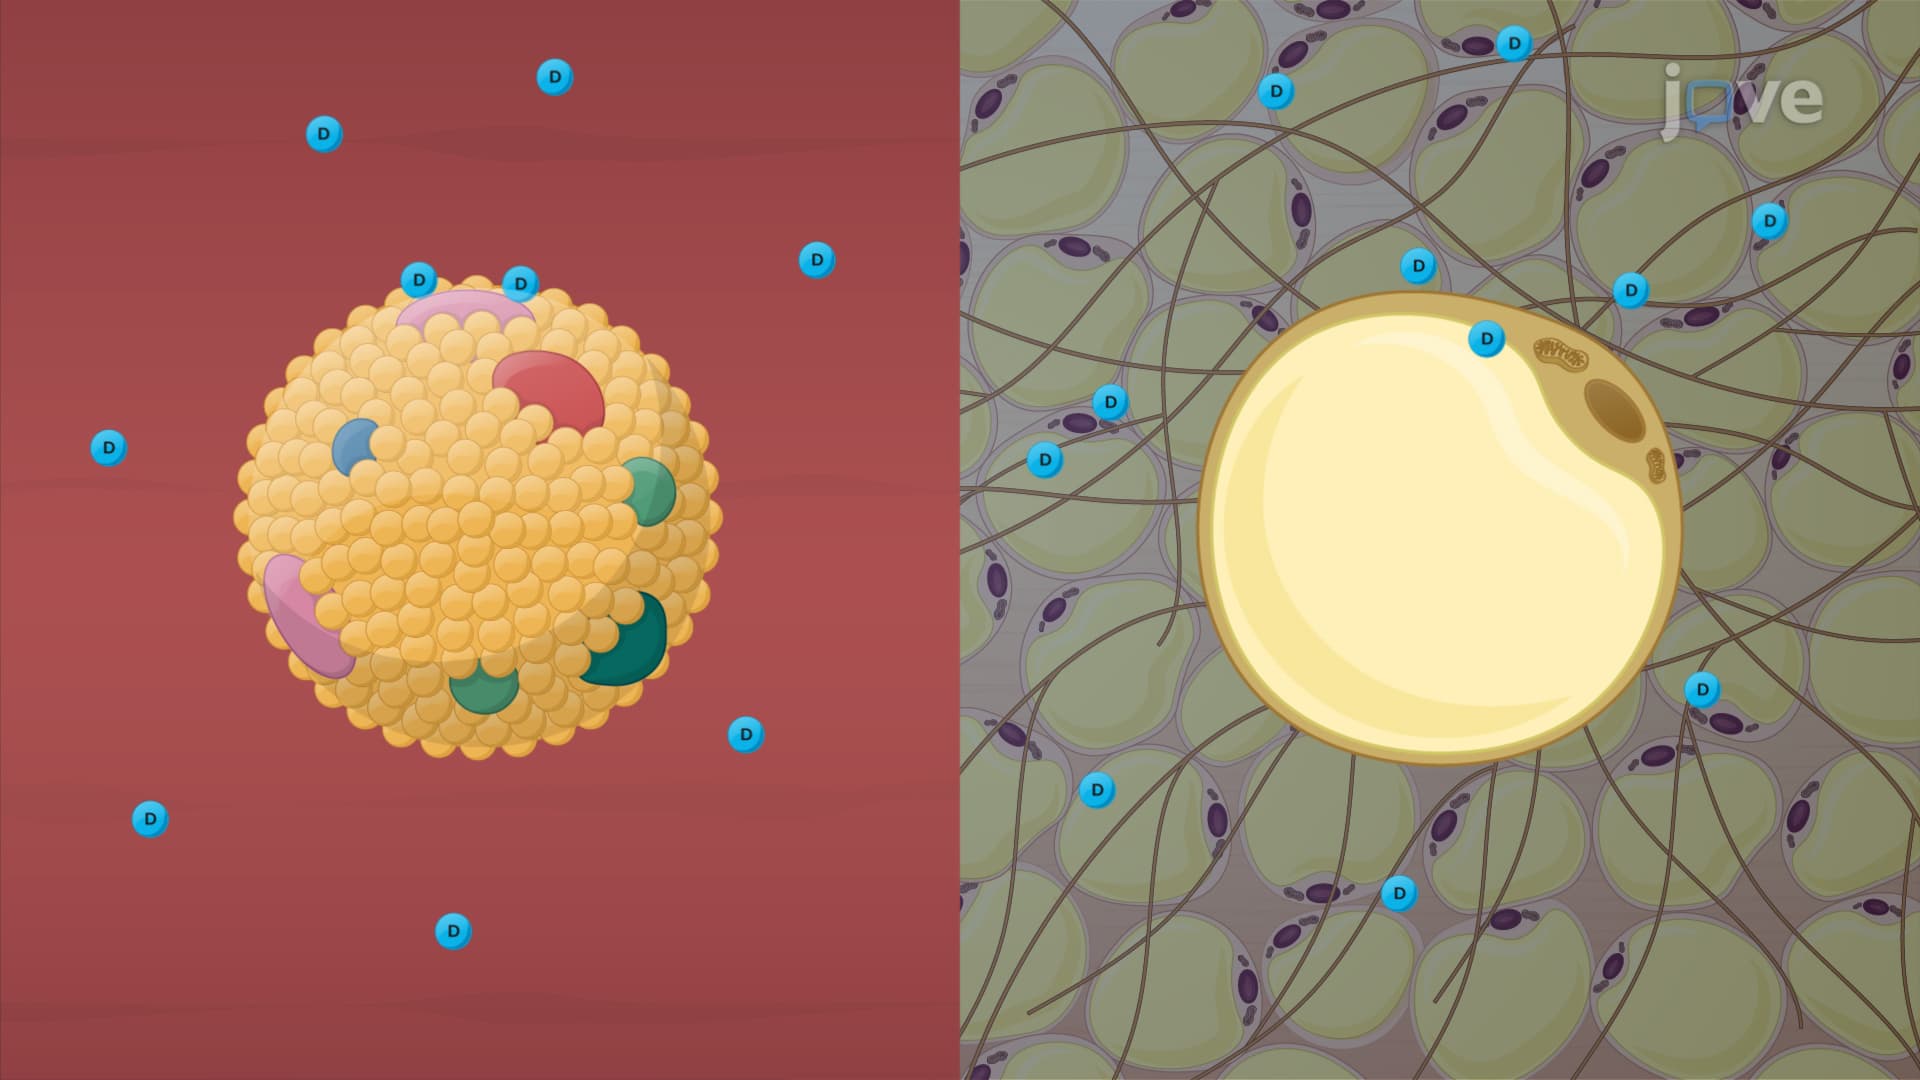
One crucial factor in drug-protein binding is the drug's lipophilicity or its affinity for fat. More lipophilic drugs tend to have higher binding extents. For example, highly lipophilic drugs like cloxacillin exhibit substantial protein binding, with as much as 95% of the drug binding to proteins. In...
 01:20
Factors Affecting Protein-Drug Binding: Protein-Related Factors
The physicochemical properties of a drug play a significant role in its ability to bind to proteins. Lipophilic drugs, which dissolve in fats, oils, and lipids, can be...
 01:22
Protein-Drug Binding: Determination Methods
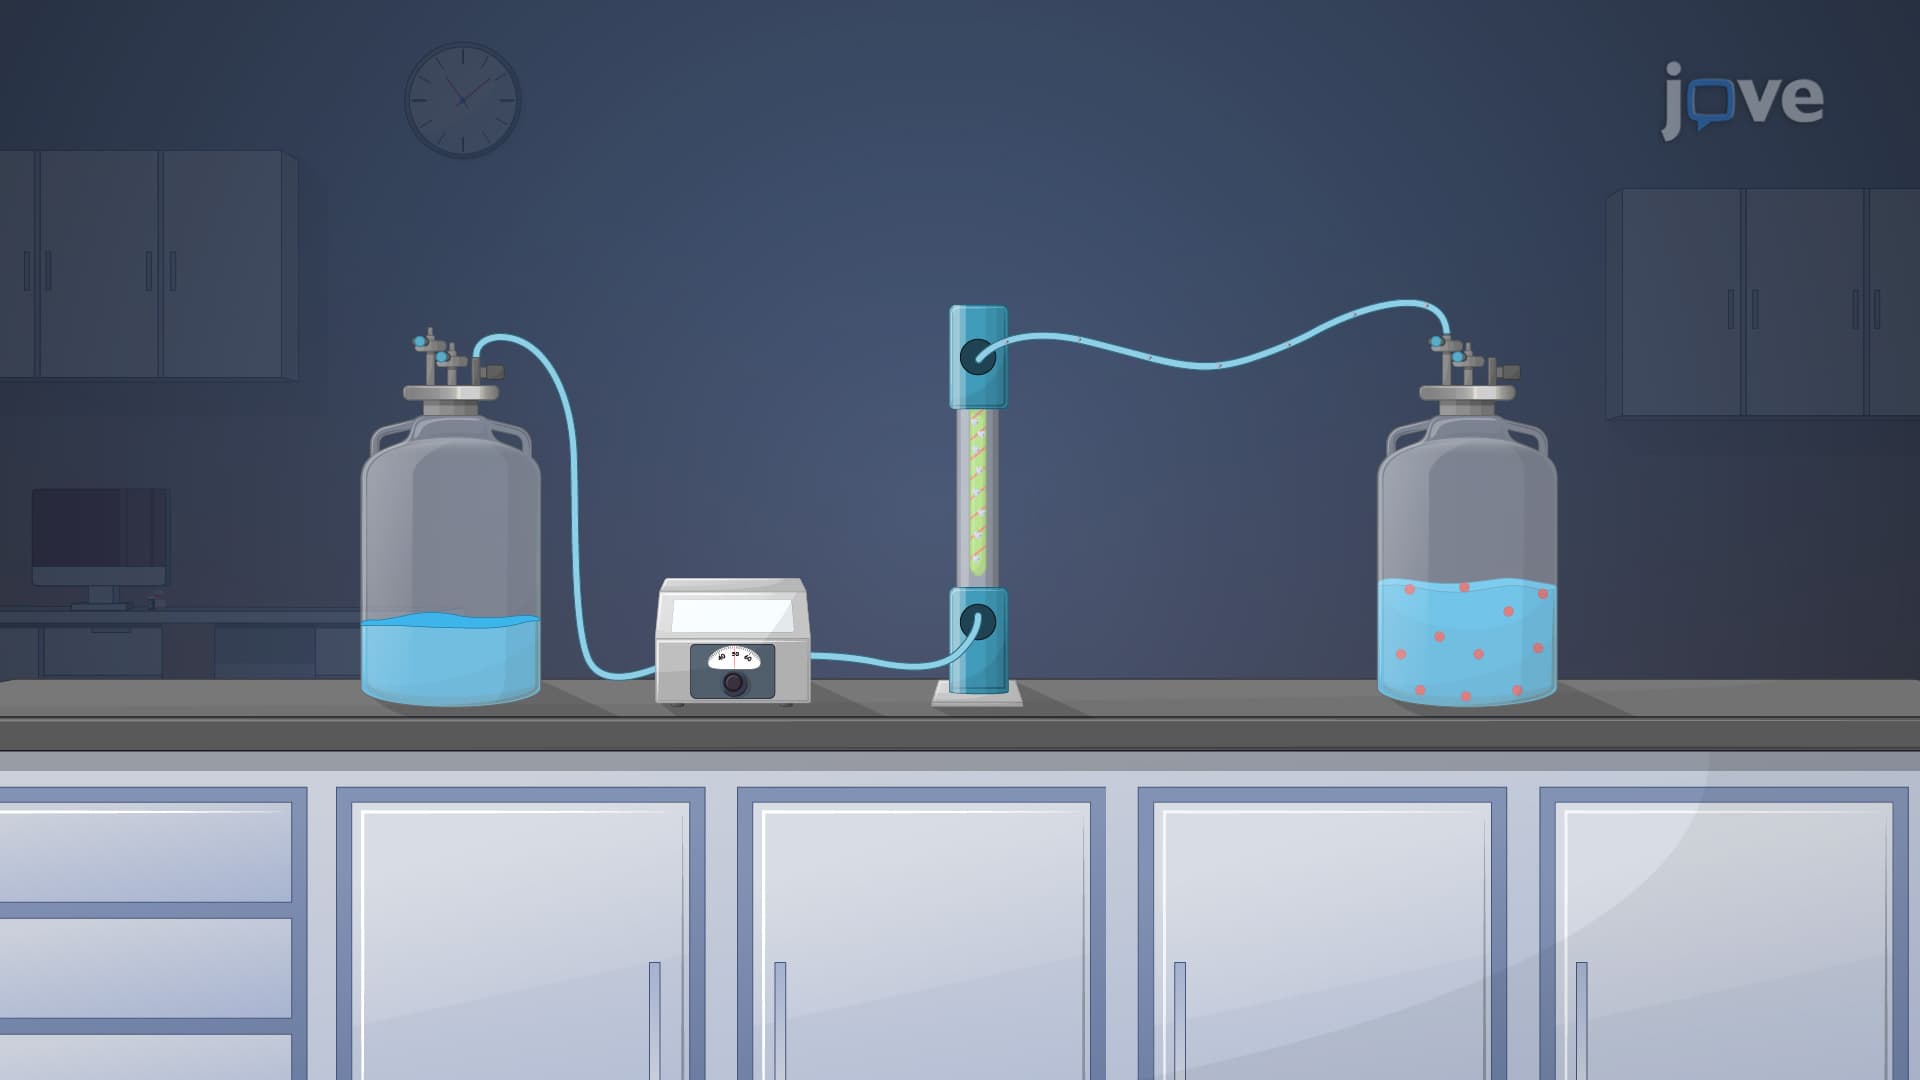
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:29
Factors Affecting Protein-Drug Binding: Patient-Related Factors
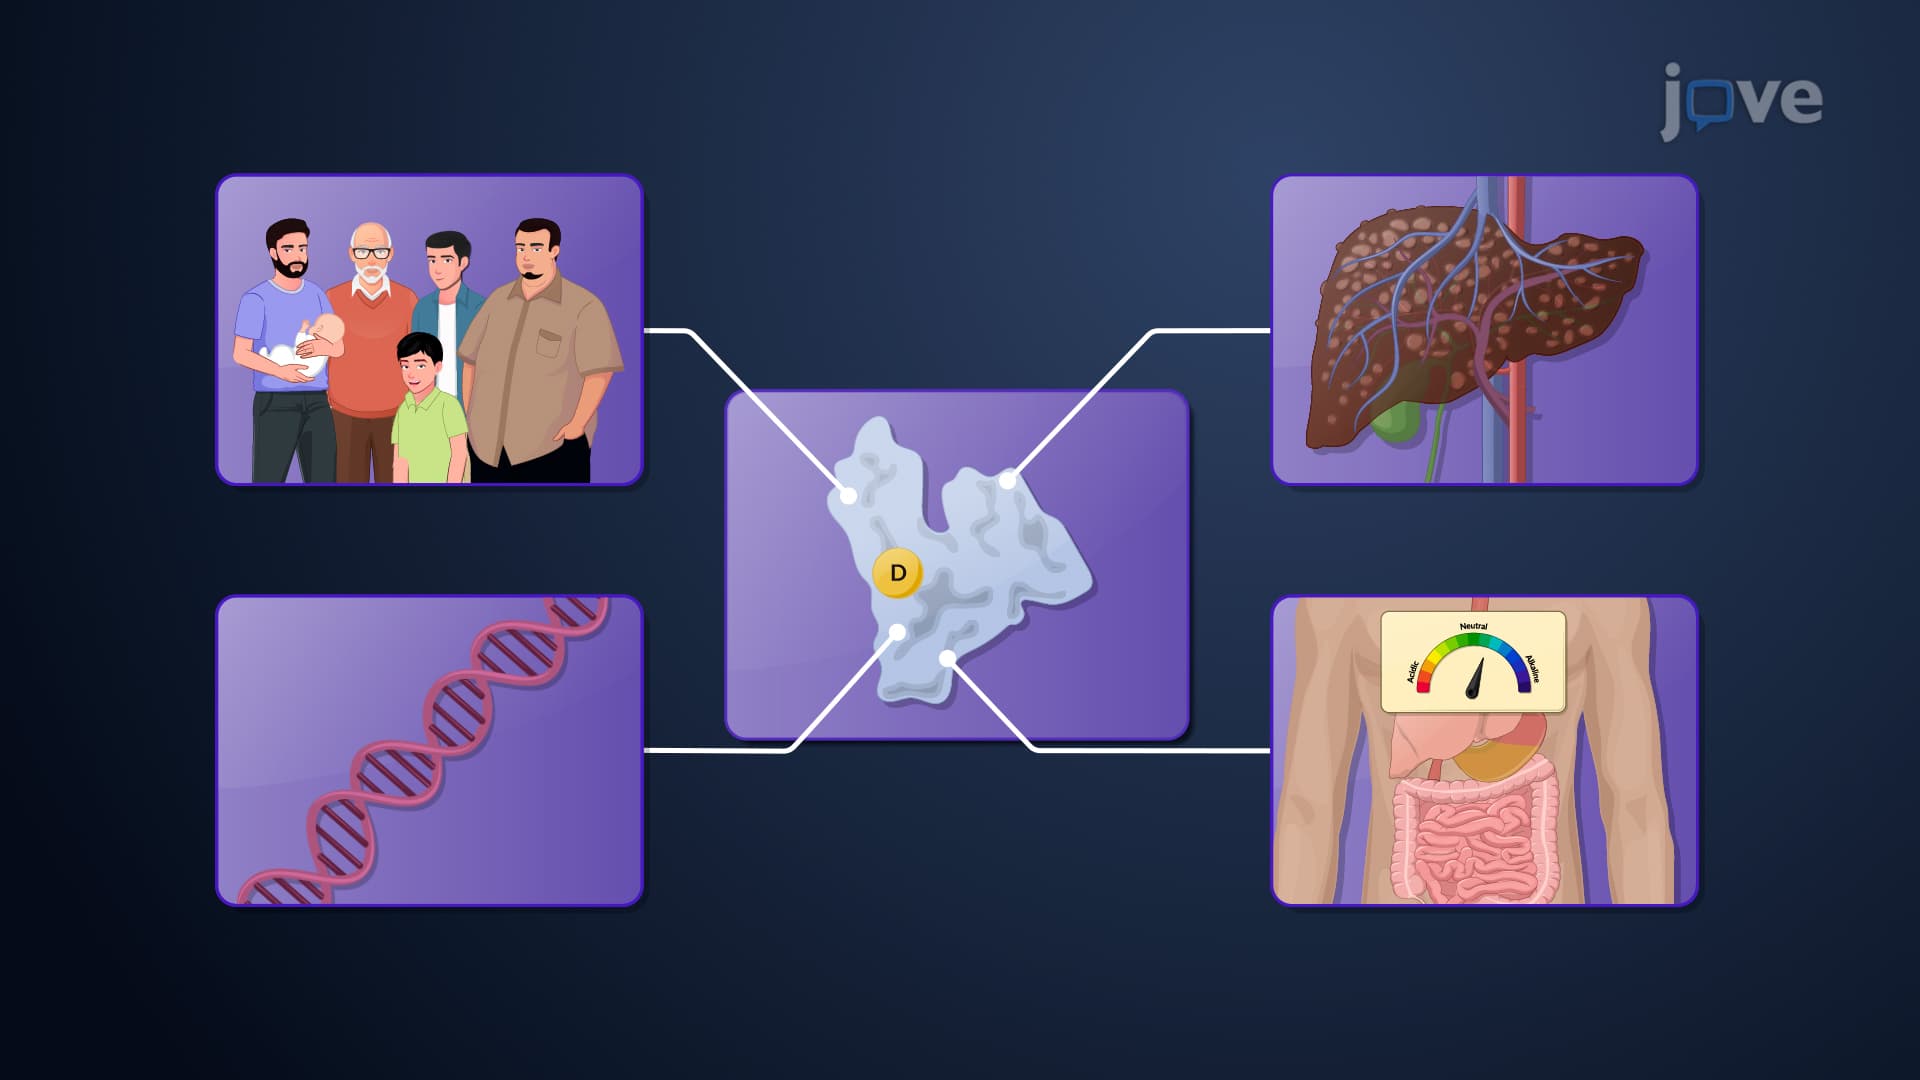
Age stands as a key determinant in protein-drug binding. Neonates, characterized by low albumin content, experience heightened concentrations of unbound drugs such as phenytoin and...
 01:30
Drug Binding to Blood Components
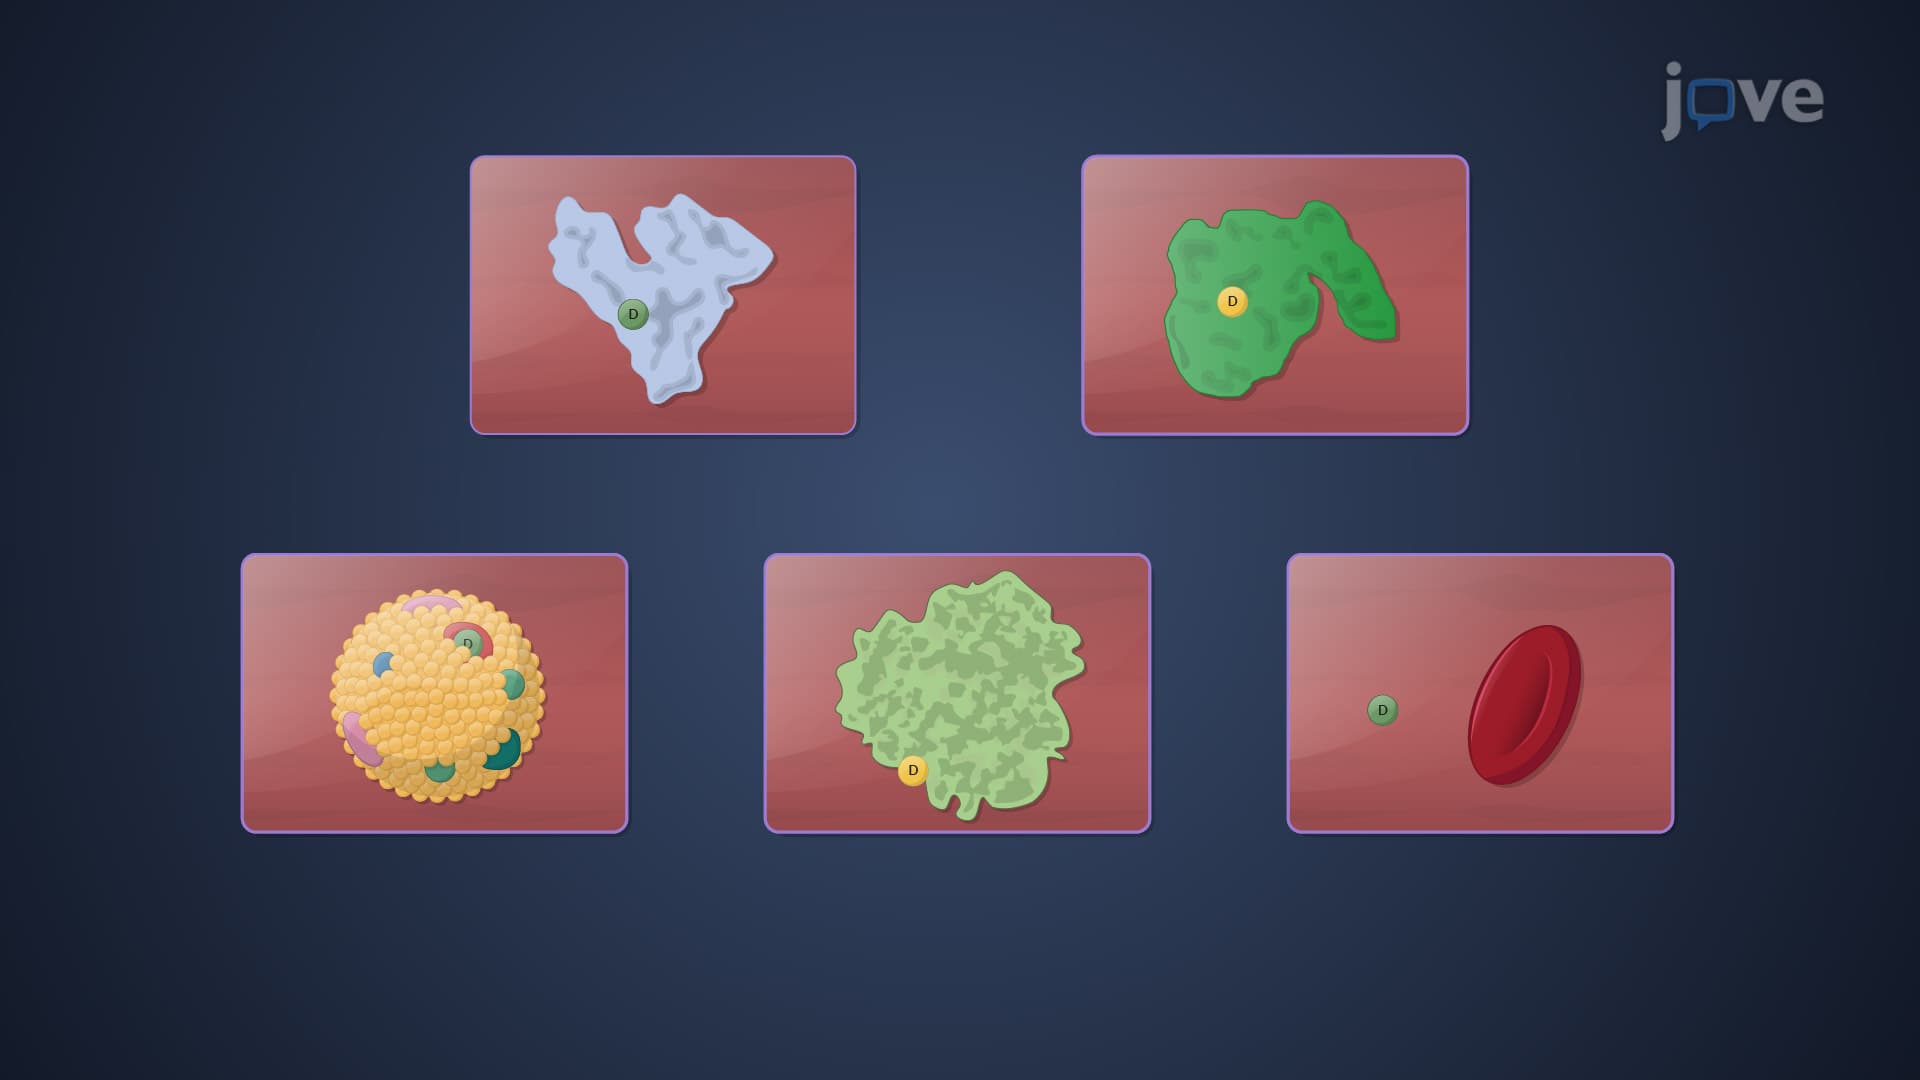
HSA is the most abundant plasma protein and is vital in drug binding. It contains distinct drug-binding sites, with different drugs exhibiting affinity for specific sites. There are three main drug-binding domains for HSA: sites I, II, and III. These domains are...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...