Related Experiment Video
Updated: Jun 28, 2025
Author Spotlight: Exploring Self-Assembled MOF-Polymer Composites
Published on: June 14, 2024
Understanding the effect of density functional choice and van der Waals treatment on predicting the binding
Jonathan R Owens1, Bojun Feng2, Jie Liu1
1Material Chemistry and Physics Lab, GE Vernova Advanced Research, Niskayuna, New York 12309, USA.

Abstract:
Metal organic frameworks (MOFs) are crystalline, three-dimensional structures with high surface areas and tunable porosities. Made from metal nodes connected by organic linkers, the exact properties of a given MOF are determined by node and linker choice. MOFs hold promise for numerous applications, including gas capture and storage. M2(4,4'-dioxidobiphenyl-3,3'-dicarboxylate)-henceforth simply M2(dobpdc), with M = Mg, Mn, Fe, Co, Ni, Cu, or Zn-is regarded as one of the most promising structures for CO2 capture applications. Further modification of the MOF with diamines or tetramines can significantly boost gas species selectivity, a necessity for the ultra-dilute CO2 concentrations in the direct-air capture of CO2. There are countless potential diamines and tetramines, paving the way for a vast number of potential sorbents to be probed for CO2 adsorption properties. The number of amines and their configuration in the MOF pore are key drivers of CO2 adsorption capacity and kinetics, and so a validation of computational prediction of these quantities is required to suitably use computational methods in the discovery and screening of amine-functionalized sorbents. In this work, we study the predictive accuracy of density functional theory and related calculations on amine loading and configuration for one diamine and two tetramines. In particular, we explore the Perdew-Burke-Ernzerhof (PBE) functional and its formulation for solids (PBEsol) with and without the Grimme-D2 and Grimme-D3 pairwise corrections (PBE+D2/3 and PBEsol+D2/3), two revised PBE functionals with the Grimme-D2 and Grimme-D3 pairwise corrections (RPBE+D2/3 and revPBE+D2/3), and the nonlocal van der Waals correlation (vdW-DF2) functional. We also investigate a universal graph deep learning interatomic potential's (M3GNet) predictive accuracy for loading and configuration. These results allow us to identify a useful screening procedure for configuration prediction that has a coarse component for quick evaluation and a higher accuracy component for detailed analysis. Our general observation is that the neural network-based potential can be used as a high-level and rapid screening tool, whereas PBEsol+D3 gives a completely qualitatively predictive picture across all systems studied, and can thus be used for high accuracy motif predictions. We close by briefly exploring the predictions of relative thermal stability for the different functionals and dispersion corrections.
More Related Videos
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 02:42
Valence Bond Theory
 01:09
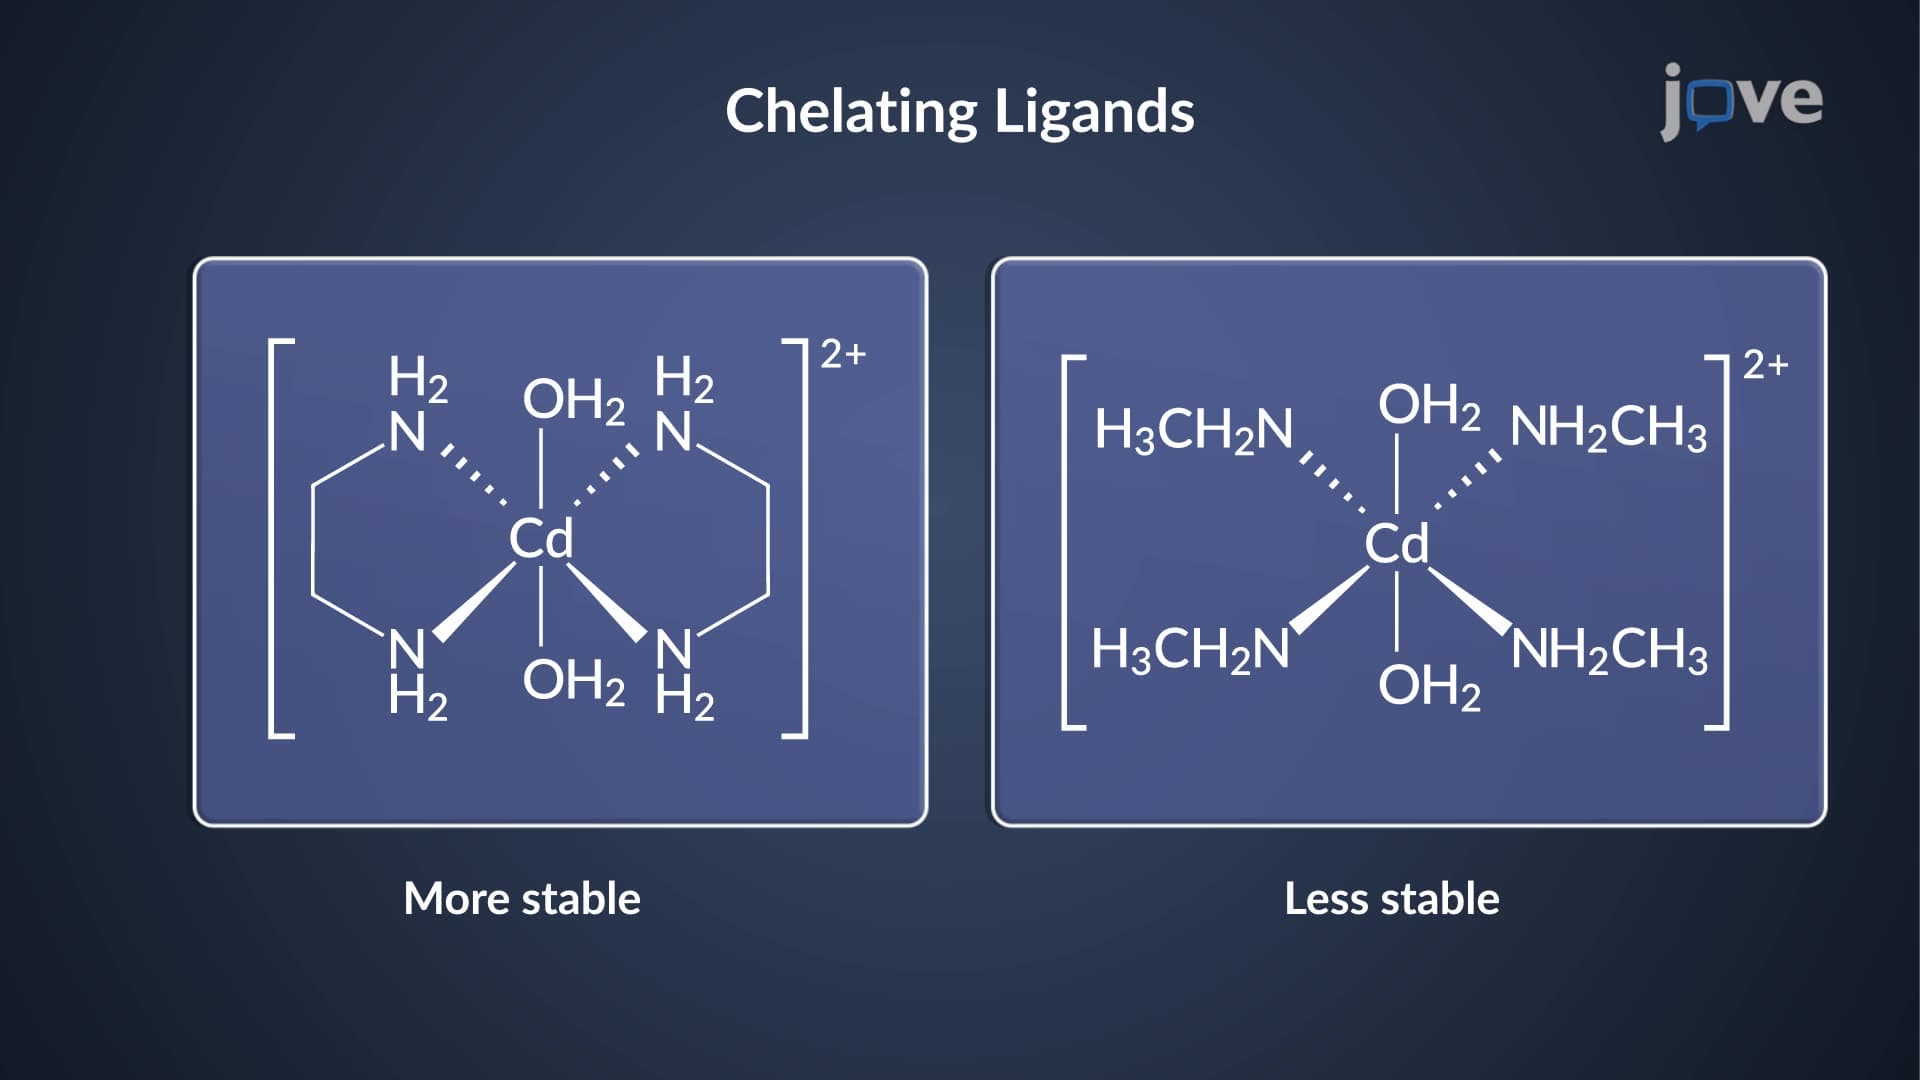
Complexation Equilibria: Factors Influencing Stability of Complexes
 01:19
Complexation Equilibria: The Chelate Effect
 01:19
Structure of Amines