Related Experiment Video
Updated: Jun 26, 2025

Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Quasi-degenerate extension of local N-electron valence state perturbation theory with pair-natural orbital method
Manami Hayashi1, Masaaki Saitow1, Kazuma Uemura1
1Department of Chemistry, Graduate School of Science, Nagoya University, Furocho, Chikusa Ward, Nagoya, Aichi 464-8601, Japan.

Abstract:
Chemical phenomena involving near-degenerate electronic states, such as conical intersections or avoided crossing, can be properly described using quasi-degenerate perturbation theory. This study proposed a highly scalable quasi-degenerate second-order N-electron valence state perturbation theory (QD-NEVPT2) using the local pair-natural orbital (PNO) method. Our recent study showed an efficient implementation of the PNO-based state-specific NEVPT2 method using orthonormal localized virtual molecular orbitals (LVMOs) as an intermediate local basis. This study derived the state-coupling (or off-diagonal) terms to implement QD-NEVPT2 in an alternative manner to enhance efficiency based on the internally contracted basis and PNO overlap matrices between different references. To facilitate further acceleration, a local resolution-of-the-identity (RI) three-index integral generation algorithm was developed using LMOs and LVMOs. Although the NEVPT2 theory is considered to be less susceptible to the intruder-state problem (ISP), this study revealed that it can easily suffer from ISP when calculating high-lying excited states. We ameliorated this instability using the imaginary level shift technique. The PNO-QD-NEVPT2 calculations were performed on small organic molecules for the 30 lowest-lying states, as well as photoisomerization involving the conical intersection of 1,1-dimethyldibenzo[b,f] silepin with a cis-stilbene skeleton. These calculations revealed that the PNO-QD-NEVPT2 method yielded negligible errors compared to the canonical QD-NEVPT2 results. Furthermore, we tested its applicability to a large photoisomerization system using the green fluorescent protein model and the ten-state calculation of the large transition metal complex, showcasing that off-diagonal elements can be evaluated at a relatively low cost.
More Related Videos
 09:00
Experimental Methods for Spin- and Angle-Resolved Photoemission Spectroscopy Combined with Polarization-Variable Laser
Published on: June 28, 2018
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:38
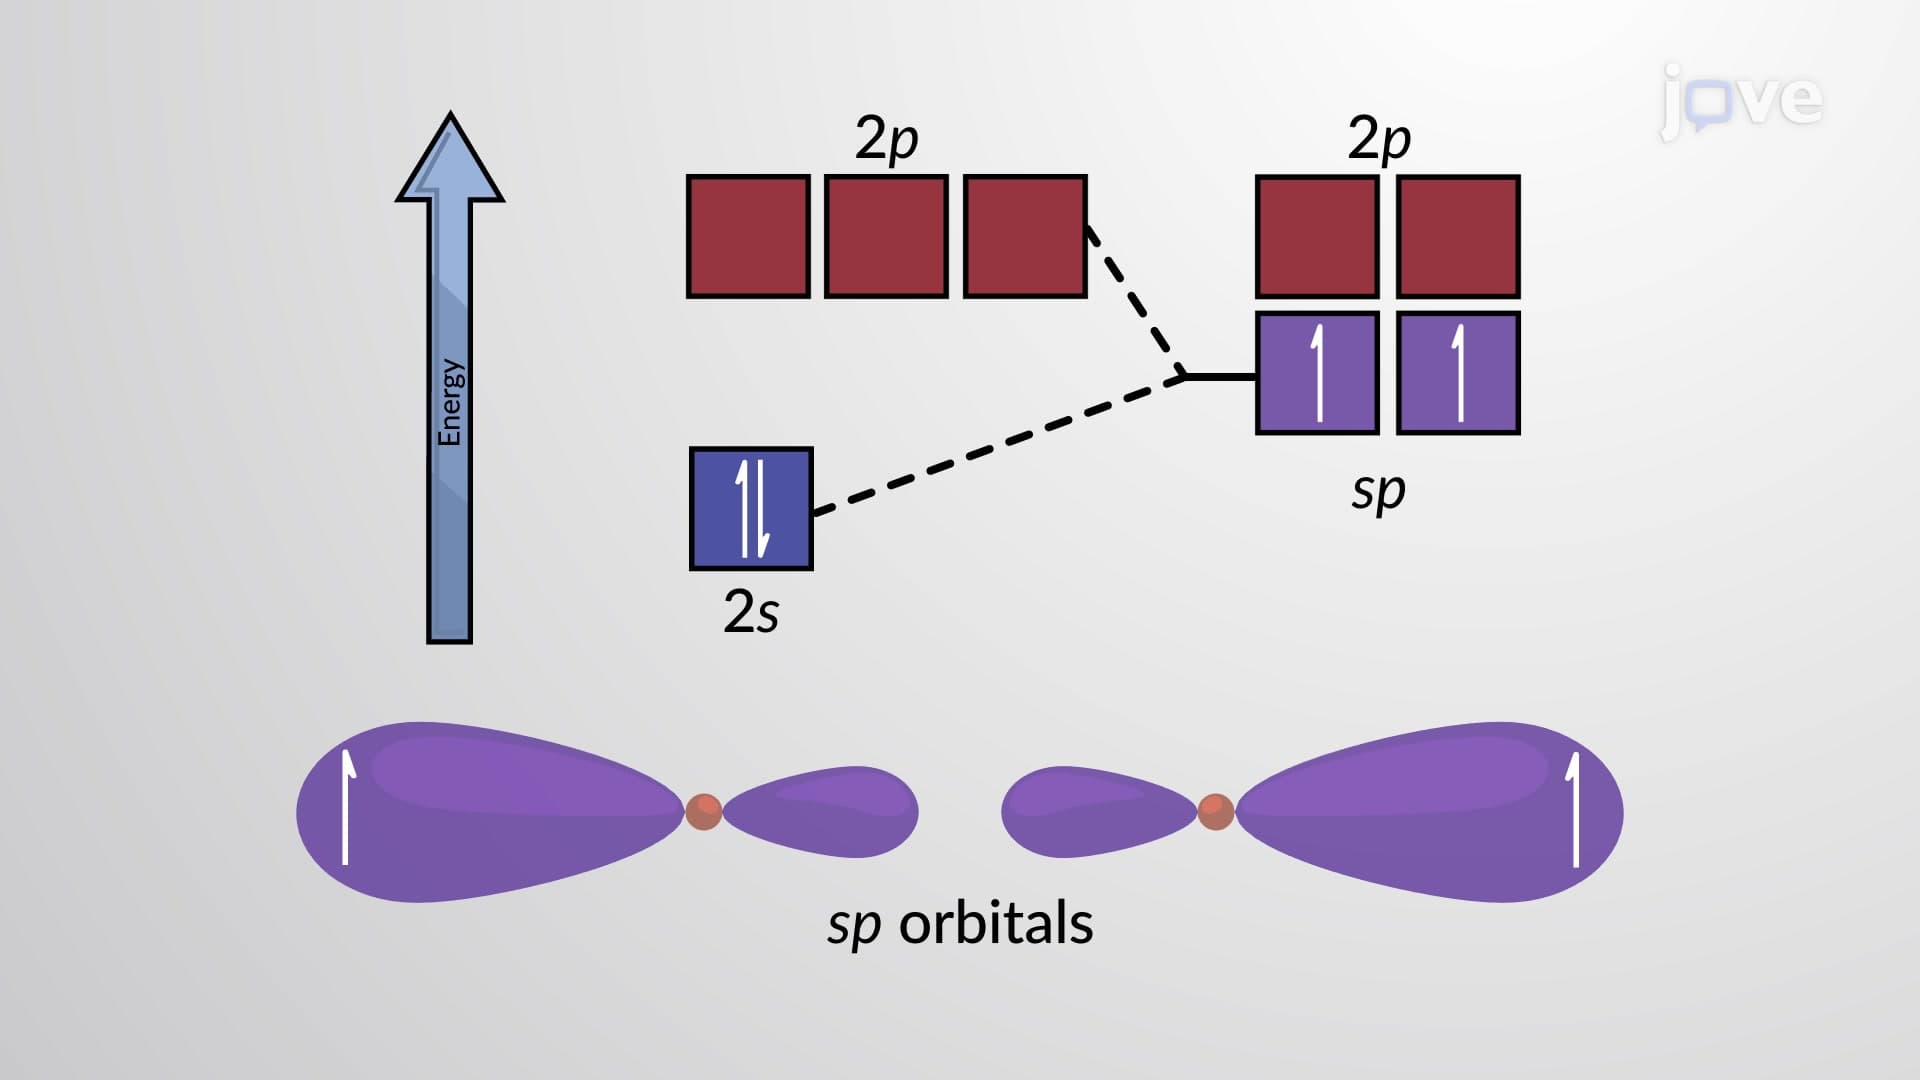
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:35
Molecular Orbital Theory I
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I